Related Experiment Video
Updated: Jan 9, 2026


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Benchmarking all-atom biomolecular structure prediction with FoldBench
Sheng Xu1, Qiantai Feng1, Lifeng Qiao2
1Research Institute of Intelligent Complex Systems, Fudan University, Shanghai, China.

Abstract:
Accurate prediction of biomolecular complex structures is fundamental for understanding biological processes and rational therapeutic design. Recent advances in deep learning methods, particularly all-atom structure prediction models, have significantly expanded their capabilities to include diverse biomolecular entities, such as proteins, nucleic acids, ligands, and ions. However, comprehensive benchmarks covering multiple interaction types and molecular diversity remain scarce, limiting fair and rigorous assessment of model performance and generalizability. To address this gap, we introduce FoldBench, an extensive benchmark dataset consisting of 1522 biological assemblies categorized into nine distinct prediction tasks. Our evaluations reveal critical performance dependencies, showing that ligand docking accuracy notably diminishes as ligand similarity to the training set decreases, a pattern similarly observed in protein-protein interaction modeling. Furthermore, antibody-antigen predictions remain particularly challenging, with current methods exhibiting failure rates exceeding 50%. Among evaluated models, AlphaFold 3 consistently demonstrates superior accuracy across the majority of tasks. In summary, our results highlight significant advancements yet reveal persistent limitations within the field, providing crucial insights and benchmarks to inform future model development and refinement.


Related Concept Videos
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:22
Protein Folding
 01:29
Protein Folding Quality Check in the RER
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:21
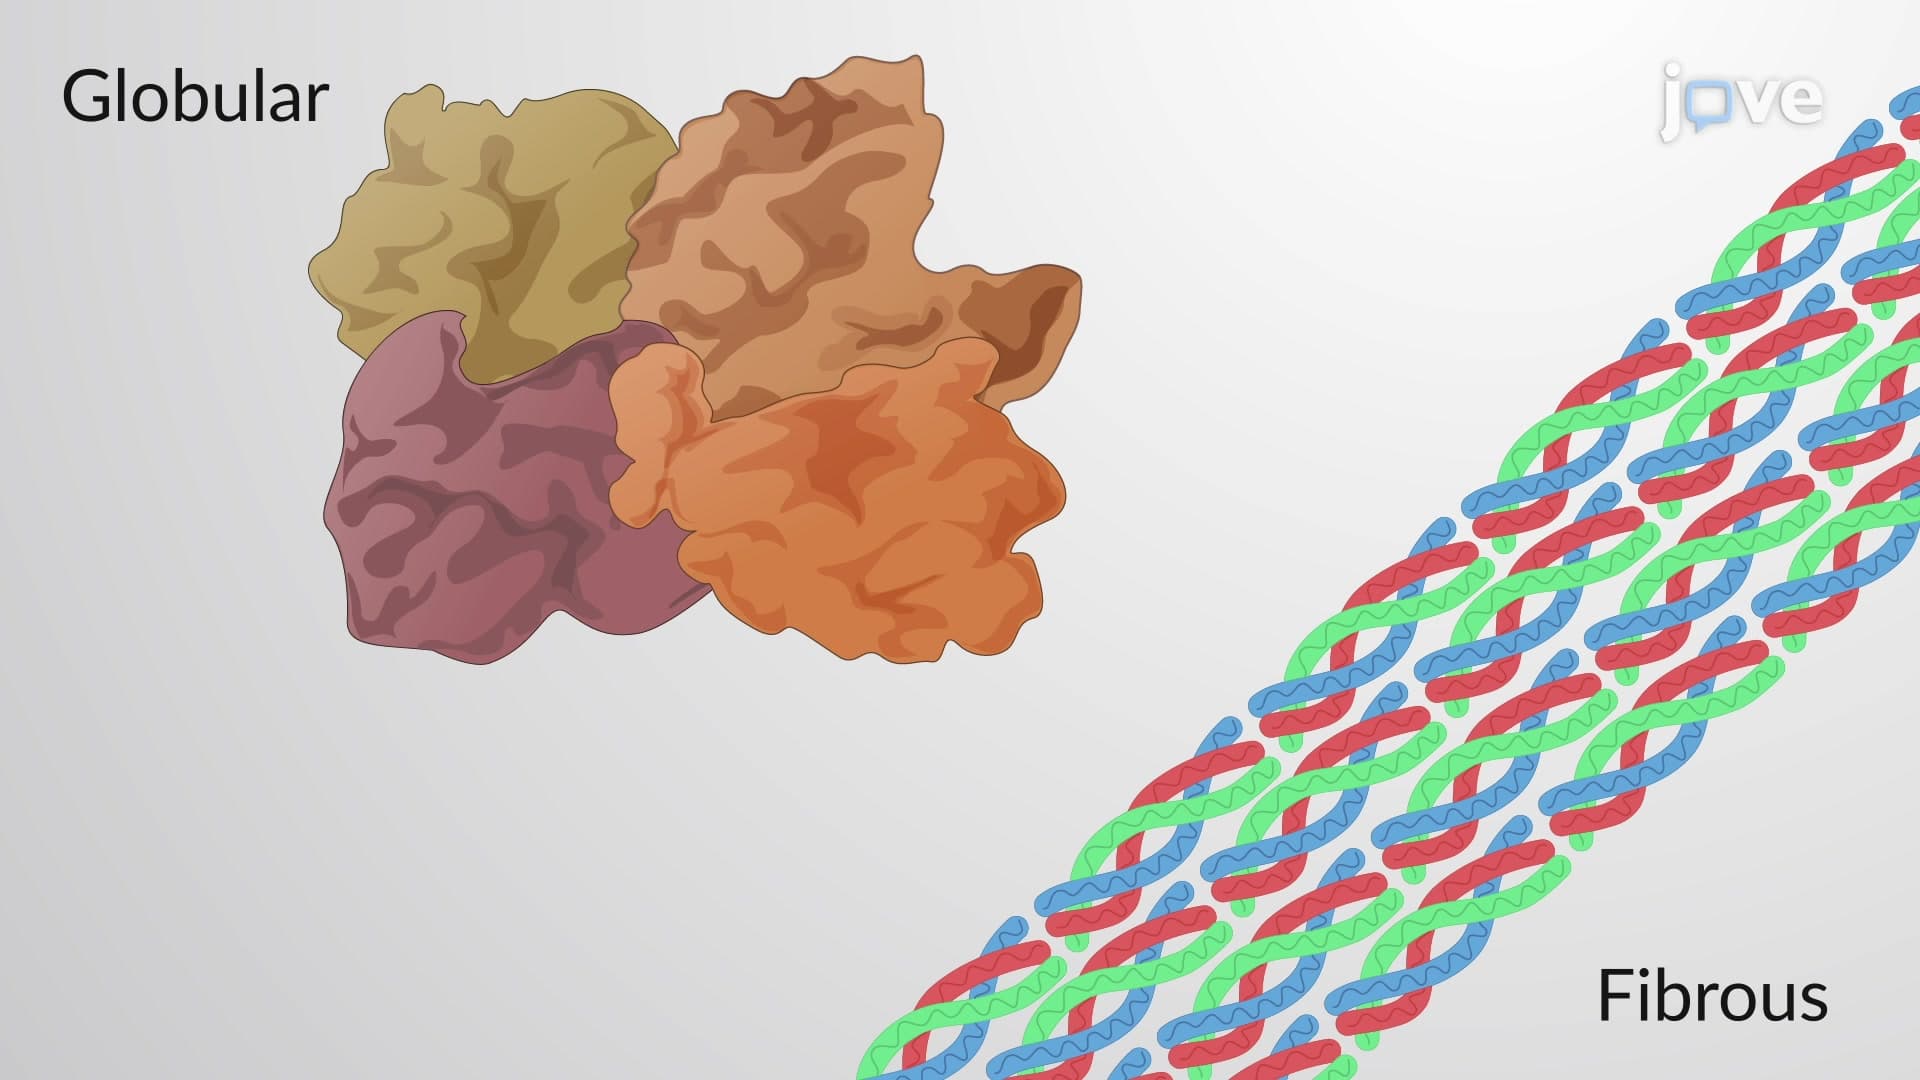
Globular and Fibrous Proteins
Globular proteins are also known as spheroproteins and typically are approximately round in shape. They contain a mix of amino acid types and contain differing sequences in their primary structures. Globular proteins have many different functions, such as enzymes, cellular messengers, and molecular transporters. These roles often require the proteins to be...
 02:18
The Equilibrium Binding Constant and Binding Strength