
Understanding the Carbon Dioxide Binding Potential Landscape near Open Metal Sites in M-MOF-74 (M = Mg, Mn, Fe, Ni,
Ishan Pandey1, Chau-Chyun Chen1, Joshua D Howe1
1Department of Chemical Engineering, Texas Tech University, P.O. Box 43121, Lubbock, Texas 79409-3121, United States.

Abstract:
Metal-organic frameworks (MOFs), especially those with undercoordinated open metal sites (OMS), are attractive candidates for applications involving adsorption of small molecules, including CO2. Among MOFs with OMS, MOF-74 variants have drawn considerable interest due to a high gravimetric density of such adsorption sites. The electronic nature of adsorbate-adsorbent interactions at OMS typically results in the accuracy of conventional force fields being insufficient to describe adsorption energetics, motivating the need for more accurate predictive methods. Toward this goal, we have utilized density functional theory (DFT) to explore the binding characteristics of CO2 in M-MOF-74s. We focus specifically on examining how various parameters, such as binding distance, variation in adsorption angles, and dihedral (precession) angles influence adsorption energetics. We consider M-MOF-74 variants across a range of d-band occupations, including Mg (3d0), Mn (3d5), Fe(3d6), Ni (3d8), and Zn (3d10) in this work, illustrating the impact of electronic structure of the OMS metal on the energetics of CO2 adsorption both at and in the vicinity of the primary adsorption sites at the OMS. Our findings, particularly at the minimum energy binding configurations, closely align with experimental structures and isosteric heats of adsorption. Our exploration of potential energy surfaces (PES) at OMS in off-minimum binding configurations uncovers more complicated interaction mechanisms, highlighting local minima and maxima within the PES. Developing these comprehensive energy profiles provides vital insights, potentially aiding in the creation of force fields that can accurately capture adsorbate-adsorbent interactions in MOFs with OMS.
More Related Videos
 10:31
Detection and Recovery of Palladium, Gold and Cobalt Metals from the Urban Mine Using Novel Sensors/Adsorbents Designated with Nanoscale Wagon-wheel-shaped Pores
Published on: December 6, 2015
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 01:09
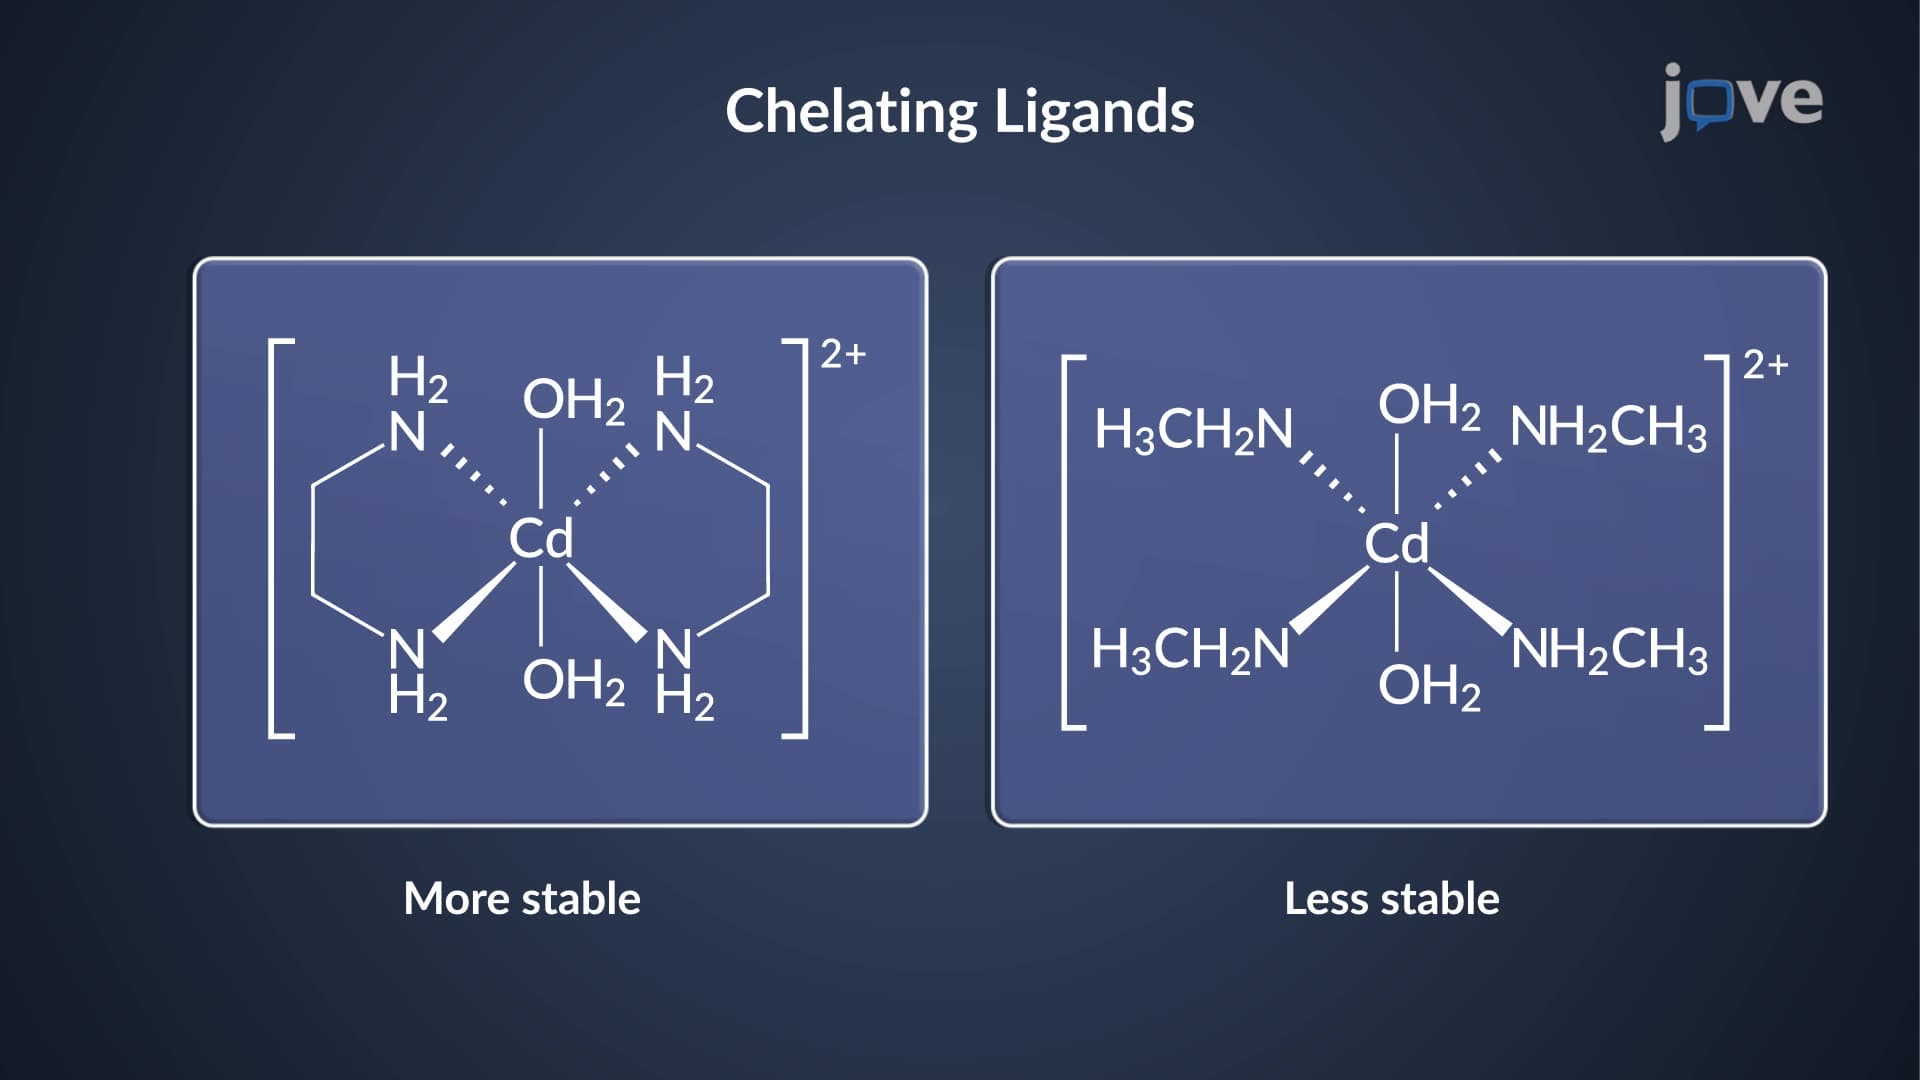
Complexation Equilibria: Factors Influencing Stability of Complexes
 01:07
Ladder Diagrams: Complexation Equilibria
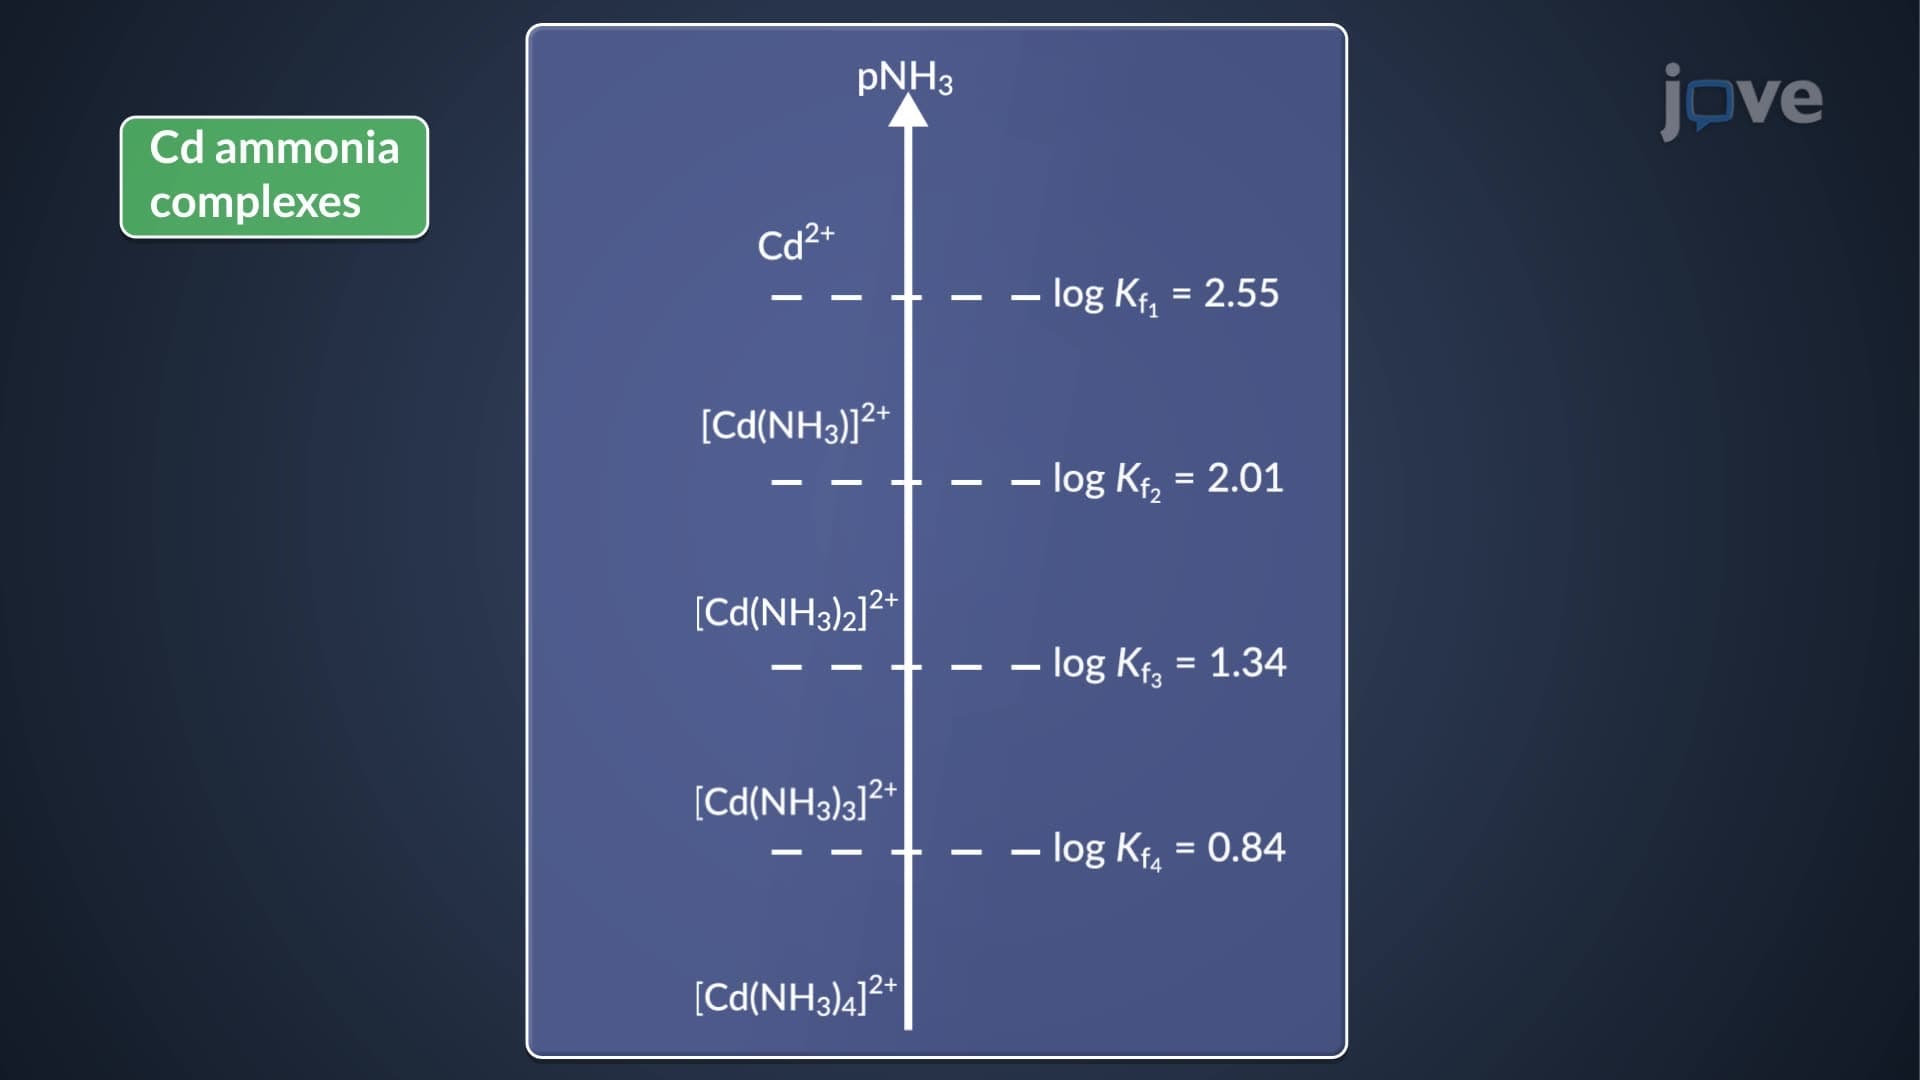
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...