Related Experiment Video
Updated: Jan 7, 2026

Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
Reliable Redox-Potential Simulations of Proteins
Michael Stier1, Johannes Kästner1
1Institute for Theoretical Chemistry, University of Stuttgart, Pfaffenwaldring 55, 70569 Stuttgart, Germany.

Abstract:
The computation of protein redox potentials is crucial for understanding various biological processes but is challenging due to very large system sizes precluding full quantum mechanical (QM) treatment. Commonly applied hybrid quantum mechanical/molecular mechanics (QM/MM) methods are often unsuitable due to nonpolarizable force fields, making pure QM calculations on cluster models a better choice. However, truncating a solvated protein with several thousand atoms to a small cluster model introduces potential sources of error. In the present study, redox properties were computed with density functional theory (DFT) for a diverse set of ten proteins using cluster models of varying sizes, ranging from 200 to 1500 atoms, including full protein calculations. Our results show that cluster models containing approximately 500 atoms are sufficient, as truncation errors become smaller than the typical DFT errors beyond that size. Additionally, we computed the standard redox potentials of ten proteins using the 500-atom cluster models within a snapshot-based approach, grounded in Marcus theory. In the proposed snapshot approach, QM/MM geometry optimizations were performed on snapshots from a classical molecular dynamics trajectory. The QM/MM geometries were truncated to 500-atom cluster models, and redox potentials were computed from vertical reduction energies using DFT. Comparison with experimental redox potentials revealed a mean absolute error (MAE) of 0.12 V, achieved without any reference or calibration calculations, which is only slightly larger than the previously estimated intrinsic DFT error. In contrast, QM/MM energies yielded rather unreliable results, with an MAE of 0.36 V for standard redox potentials. These findings are not only significant for improving redox potential predictions but also broadly relevant for electronic structure calculations in complex systems involving charge changes, such as protonation/deprotonation processes, electron transfer processes, and electrocatalysis.

Related Concept Videos
 01:24
Redox Reactions
 01:27
Redox Reactions
 01:23
Redox Equilibria: Overview
 01:30
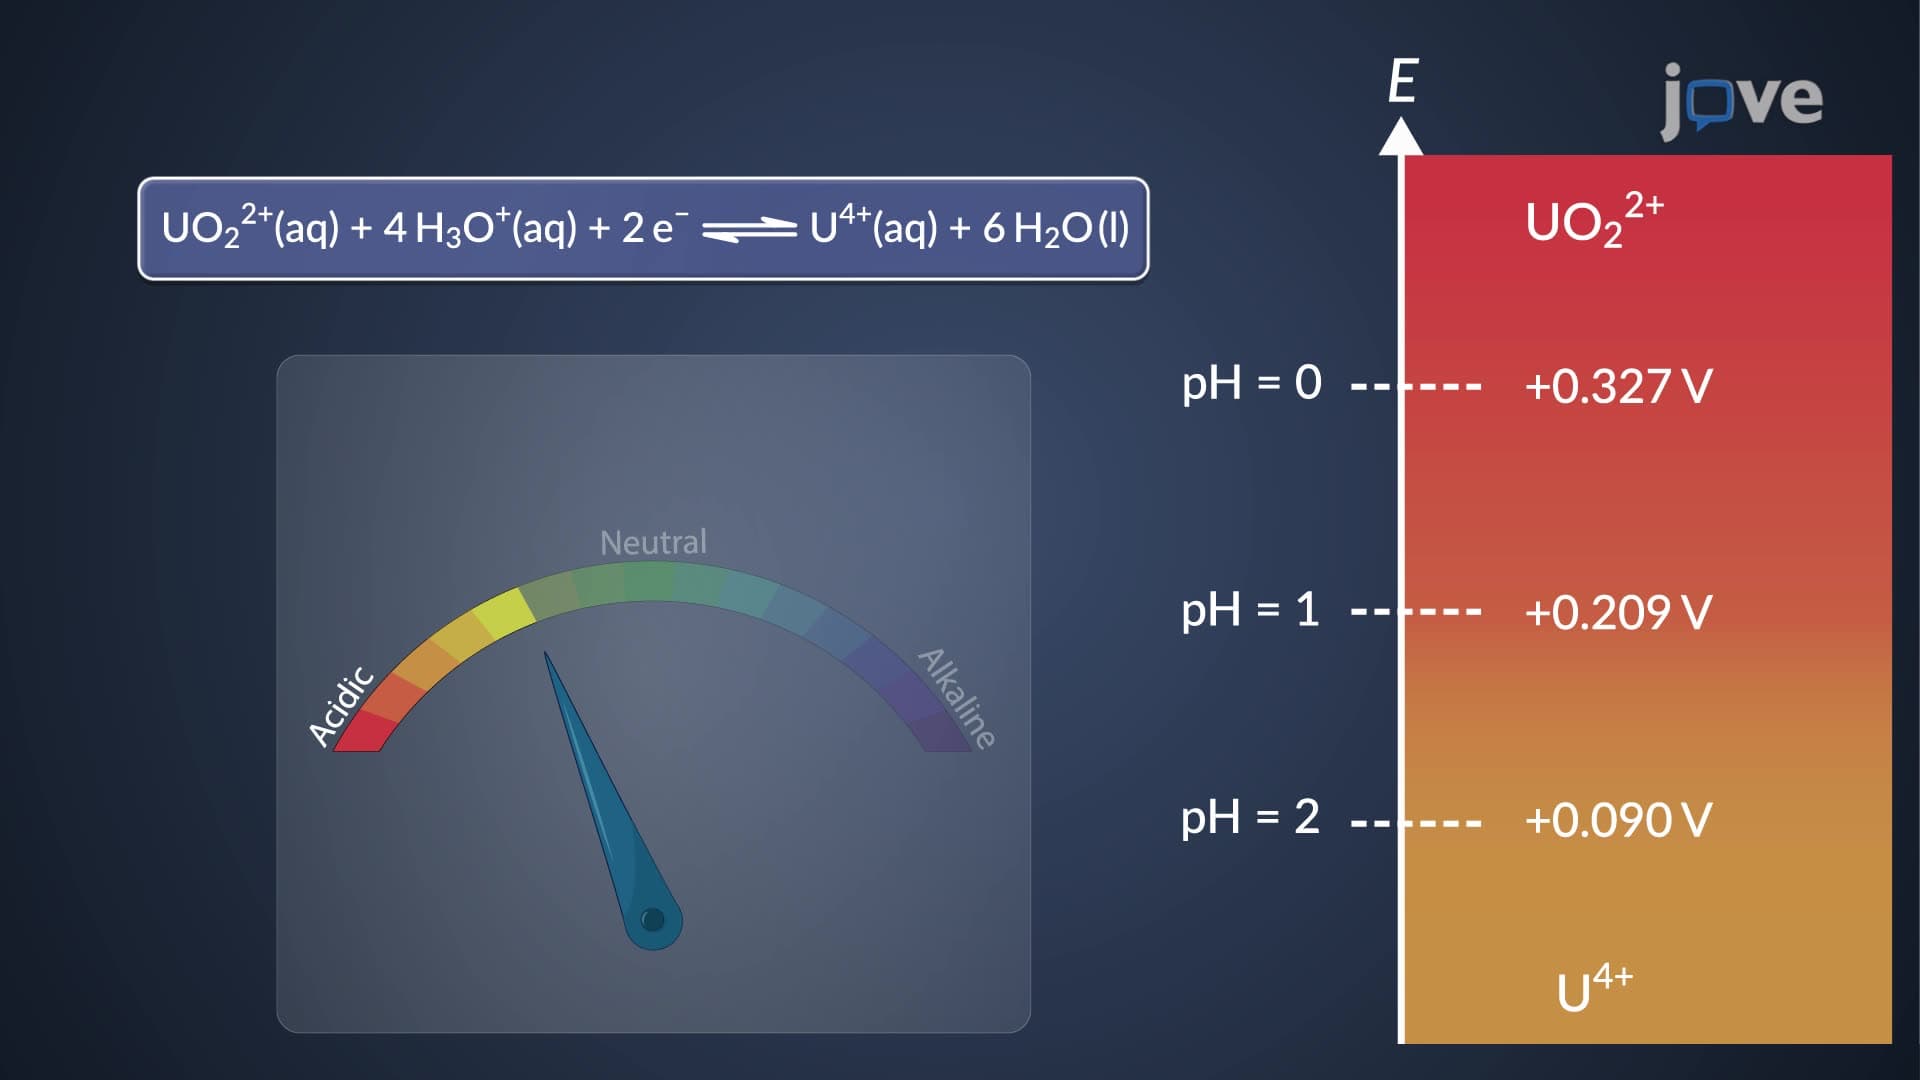
Ladder Diagrams: Redox Equilibria
Consider the Fe3+/Fe2+ half-reaction, which has a standard-state potential of +0.771 V. At potentials more positive than +0.771 V, Fe3+ predominates, whereas Fe2+...
 01:19
Protein Dynamics in Living Cells
Fluorescent recovery after photobleaching (FRAP) is a fluorescent-protein-based detection technique used to quantify protein movement rates within the cell. This method exposes a small portion of the cell to an intense laser beam. The laser beam causes permanent photobleaching of the fluorophore-tagged proteins in the exposed region. As the bleached...
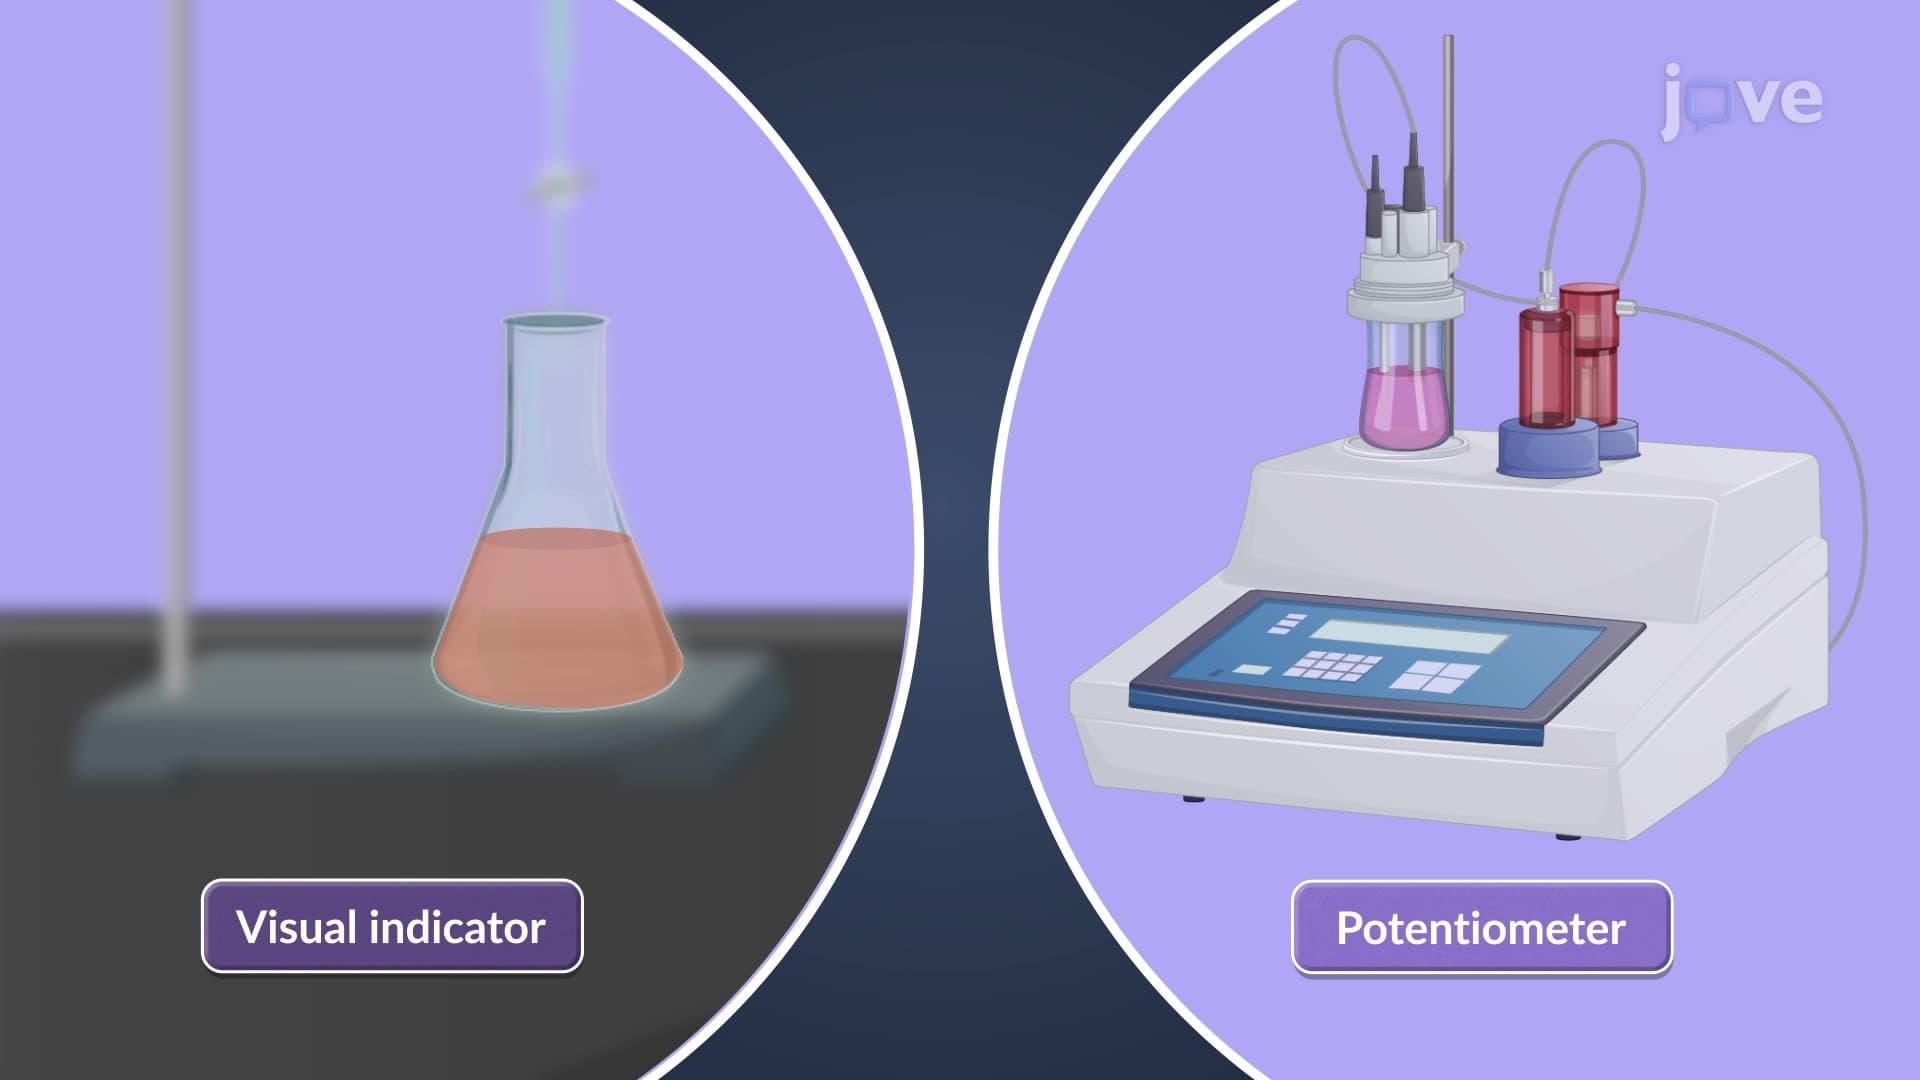 01:21
Redox Titration: Overview