Related Experiment Video
Updated: Jan 7, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
The covalent docking software landscape: features and applications in drug design
Natesh Singh1, Philippe Vayer2, Bruno O Villoutreix2
1Evotec SE, Molecular Architects, Integrated Drug Discovery, Campus Curie, 195 Rte d'Espagne, 31100 Toulouse, France.

Abstract:
Covalent small-molecule ligands have re-emerged as powerful tools in drug discovery, offering prolonged target engagement, enhanced potency, and the ability to modulate proteins once considered undruggable. However, the rational design and virtual screening (VS) of covalent ligands remain challenging. Many docking tools cannot accurately model the energetics of covalent bond formation, often requiring more rigorous quantum mechanical (QM) or semi-empirical QM calculations for reliable predictions. Despite these limitations, the computational landscape is rapidly evolving. An increasing number of open-source, commercial, and web-based platforms now support binding mode exploration, lead optimization, and structure-based VS of covalent ligands. Alongside traditional approaches, new artificial intelligence (AI) and machine learning (ML) tools are assisting in prioritizing candidate molecules. This review introduces the fundamental principles and mechanisms of covalent inhibition, then provides a comprehensive overview of computational tools including covalent docking, warhead placement algorithms, and pharmacophore modeling, supporting early-stage drug discovery and chemical biology. Case studies highlight practical applications. We also cover curated databases of covalent binders and experimental 3D protein-ligand complexes, plus tools for assessing nucleophilic residue reactivity, all essential for robust covalent modeling. Finally, we briefly address risks associated with covalent chemistry. While progress is notable, further advances are needed. Nonetheless, today's covalent docking and AI-driven tools already make a meaningful impact by enabling rational design, generating new ideas, refining hypotheses, and expanding the boundaries of druggability.
More Related Videos
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
Related Concept Videos
 01:25
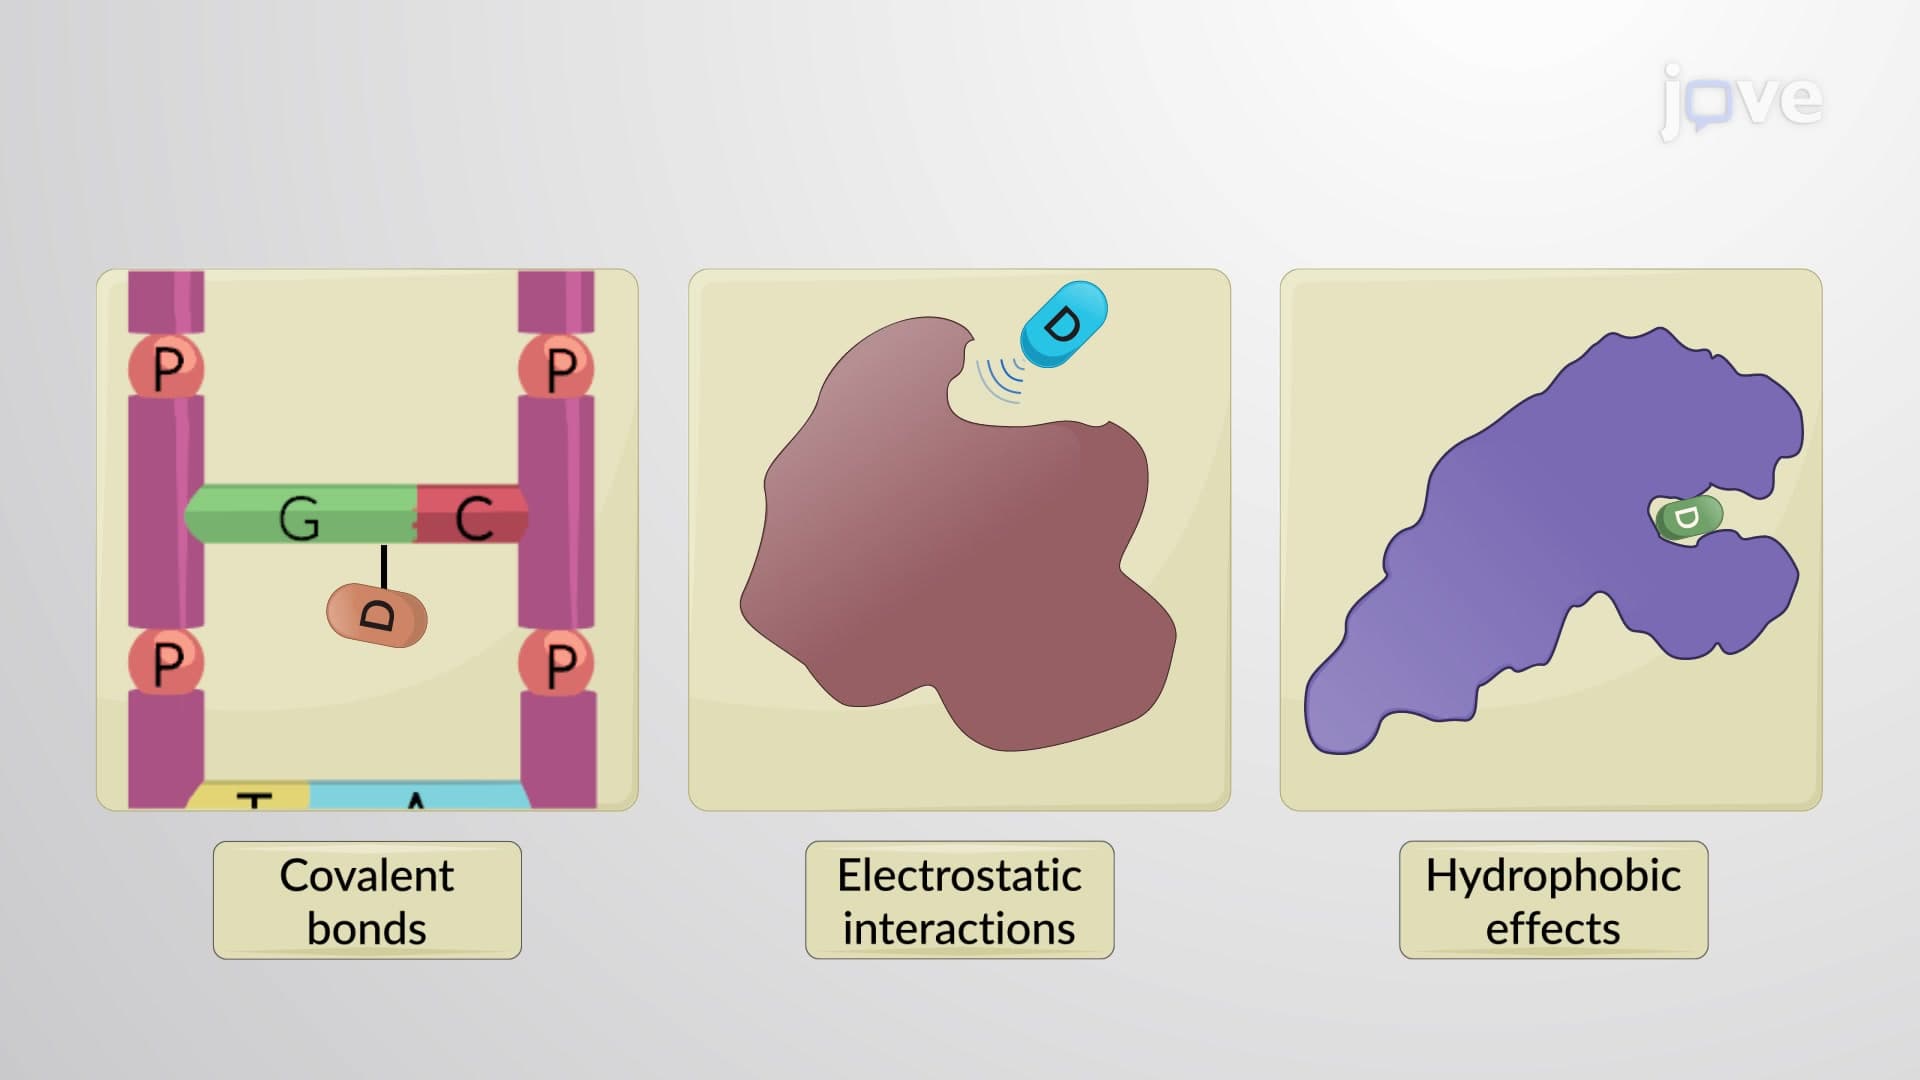
Drug-Receptor Bonds
In...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
 02:04
Protein-protein Interfaces
 01:26
Drug Discovery: Overview
 01:22
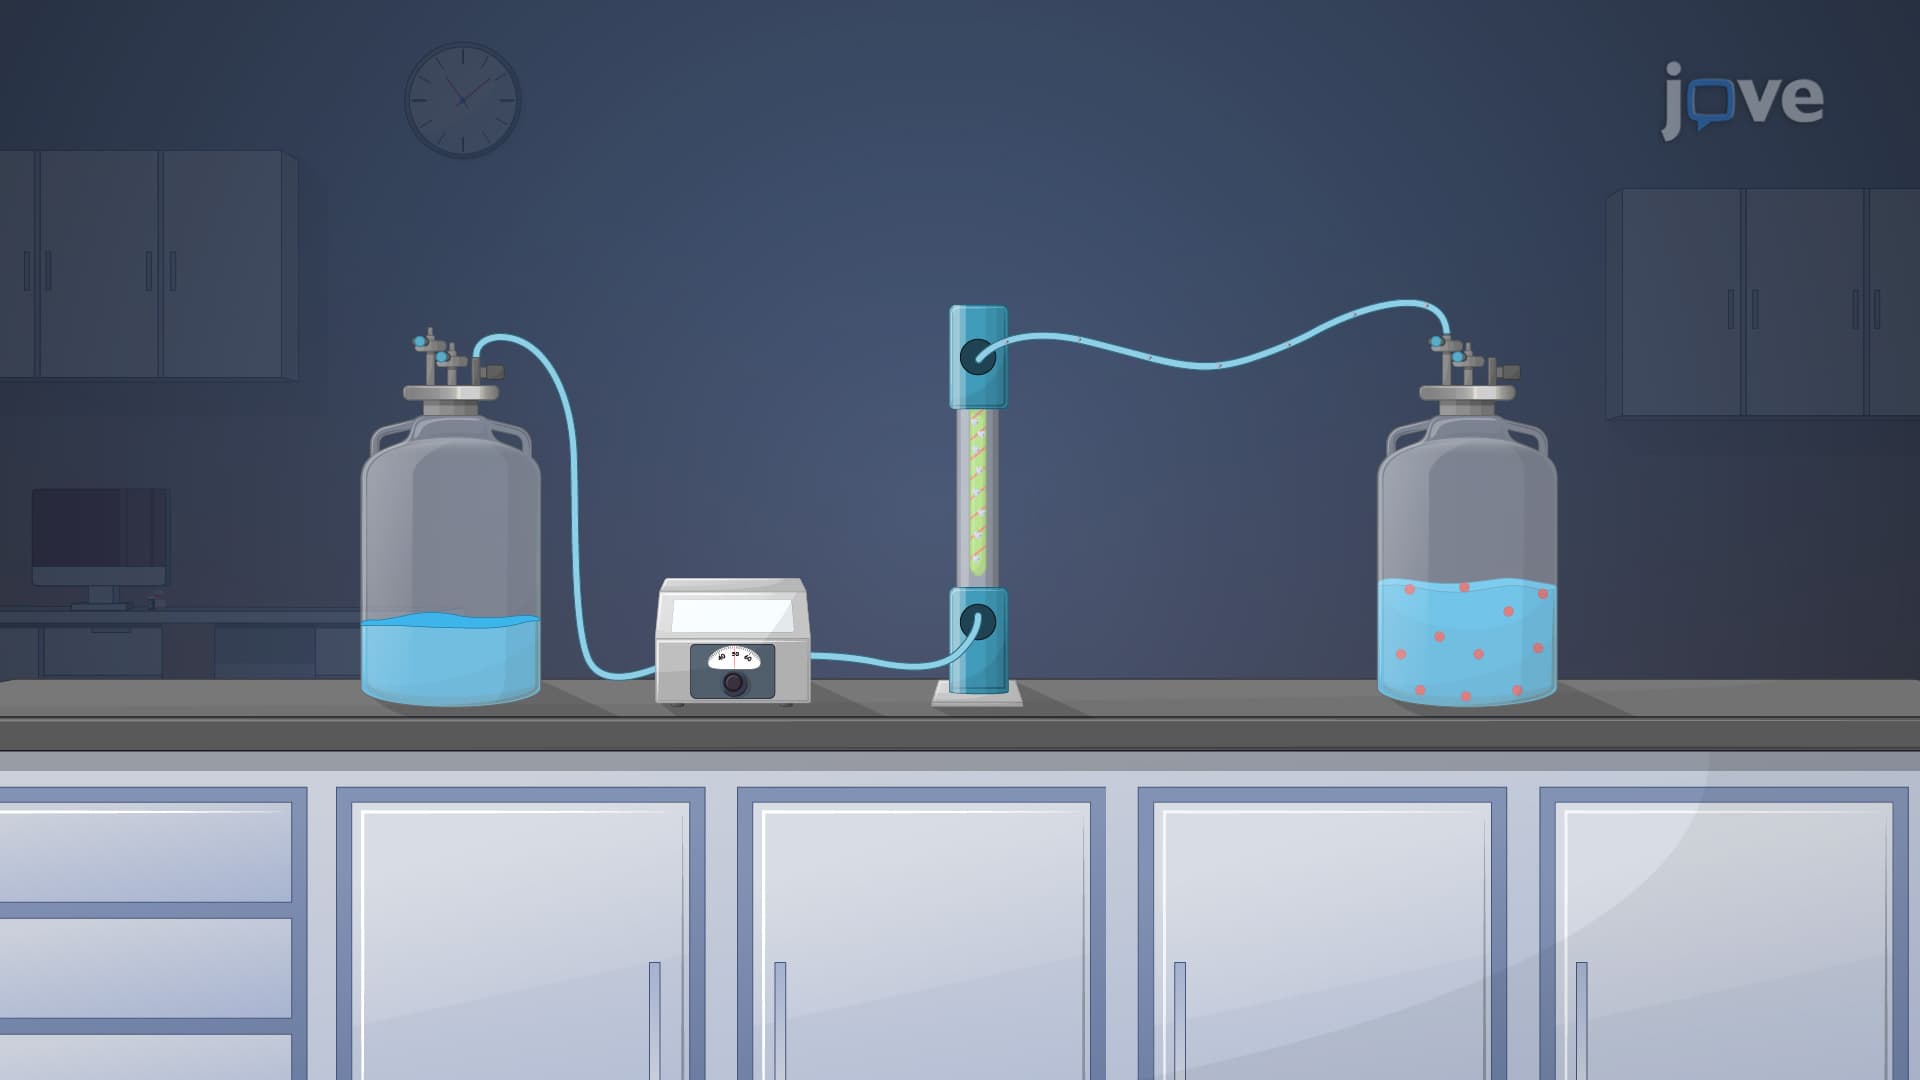
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...