Transcriptional profiling of Hutchinson-Gilford progeria patients identifies primary target pathways of progerin
Related Experiment Video
Updated: Jan 7, 2026
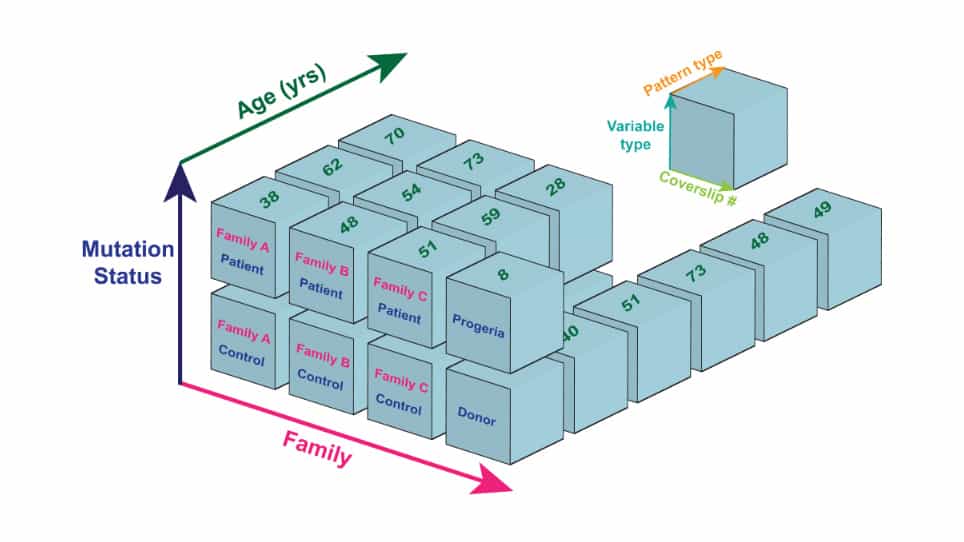
Databases to Efficiently Manage Medium Sized, Low Velocity, Multidimensional Data in Tissue Engineering
Published on: November 22, 2019
Sandra Vidak1, Sohyoung Kim2, Tom Misteli1
1Cell Biology of Genomes, National Cancer Institute, National Institutes of Health, Bethesda, MD, USA.

Insights
Hutchinson Gilford Progeria Syndrome (HGPS), a rare premature aging disorder, involves the LMNA gene mutation. Transcriptomic analysis reveals consistent pathway misregulation across patients, identifying progerin
Area of Science:
- Genetics and Molecular Biology
- Cellular Biology
- Aging Research
Background:
- Hutchinson Gilford Progeria Syndrome (HGPS) is a rare genetic disorder causing rapid premature aging.
- It stems from a mutation in the LMNA gene, producing toxic progerin.
- Current understanding relies heavily on cellular and mouse models.
Purpose of the Study:
- To investigate the clinical relevance of cellular pathways in HGPS patients.
- To assess gene expression variability among HGPS patients.
- To identify progerin's primary target pathways in vivo.
Main Methods:
- Transcriptomic analysis of a comprehensive patient cohort.
- Comparison with an inducible HGPS cellular model.
- Analysis of gene expression patterns and pathway misregulation.
Main Results:
- Consistent misregulation of key cellular pathways, including signaling, Unfolded Protein Response (UPR), and mesodermal cell fate, was observed across HGPS patients.
- Limited variability in gene expression heterogeneity among patients.
- Identification of primary target pathways affected by progerin.
Conclusions:
- Transcriptomic analysis in HGPS patients confirms the clinical relevance of specific cellular pathways.
- Progerin's effects on gene expression are largely consistent across patients.
- This study provides insights into HGPS pathogenesis and potential therapeutic targets.
Abstract:
Hutchinson Gilford Progeria Syndrome (HGPS) is an ultra-rare pediatric premature aging disorder. It is caused by a point mutation in the LMNA gene leading to the production of the dominant-negative progerin isoform of the nuclear envelope protein lamin A. Most of the mechanistic insights into the disease have come from studies using cellular or mouse models of HGPS. To probe the clinical relevance of previously implicated cellular pathways and to address the extent of gene expression heterogeneity between patients, we performed transcriptomic analysis of a comprehensive set of HGPS patients. We find misexpression of several cellular pathways, including multiple signaling pathways, the Unfolded Protein Response (UPR) and mesodermal cell fate specification. Variability amongst individual patients was limited, with misregulation of the major pathways observed in most patients. Comparing the transcriptome of patients with an inducible HGPS cell model, we also identified the primary target pathways of the disease-causing progerin protein.
More Related Videos
 06:23
Detection of Nuclear Blebbing and DNA Leakage in Mammalian Cells by Immunofluorescence
Published on: January 17, 2025
 10:25
Monitoring of Ubiquitin-proteasome Activity in Living Cells Using a Degron dgn-destabilized Green Fluorescent Protein GFP-based Reporter Protein
Published on: November 10, 2012
