Related Experiment Video
Updated: Jan 23, 2026

A Simple and Low-cost Assay for Measuring Ambulation in Mouse Models of Muscular Dystrophy
Published on: December 29, 2017
LAMA2 variants associated with muscular dystrophy, brain structural abnormalities, and epilepsy: a genotype-phenotype
Jian Zha1, Ying Yu2, Fangfang Cao1
1Department of Neurology, Jiangxi Provincial Children's Hospital, Nanchang, Jiangxi, China.

Background:
LAMA2-related congenital muscular dystrophy (LAMA2-MD) is a genetically heterogeneous disorder defined by progressive muscle weakness, brain structural abnormalities, epilepsy, and multisystem involvement. The primary goal of this study was to characterize the clinical features, temporal progression, and genotype-phenotype correlations of LAMA2-MD.
Methods:
Medical records of patients with genetically confirmed LAMA2-MD were extracted from a clinical data repository and analyzed retrospectively. Clinical manifestations, laboratory findings, and neuroimaging features were systematically reviewed and compared across different age groups. Variant data were retrieved from public databases to perform comprehensive genetic analyses.
Results:
A total of five patients (two males and three females) were enrolled, delayed motor milestones and varying degrees of ankle contractures and persistent motor impairment in all patients were the initial presenting symptom at diagnosis in all cases, and two patients also exhibited cognitive delays. Laboratory analysis of muscle enzymes showed varying degrees of abnormalities, with creatine kinase (CK) levels displaying the most significant elevation. Cranial magnetic resonance imaging (MRI) revealed symmetrical white matter abnormalities in four patients. Seizures were documented in three school-aged patients. All patients carried compound heterozygous variants in the LAMA2 gene. A literature review indicated that the most common variant types were stop-gain and missense variants: stop-gain variants were predominantly associated with complete merosin deficiency (MDC1A), whereas missense variants typically correlated with late-onset limb-girdle muscular dystrophy.
Conclusion:
LAMA2-MD exhibits a broad phenotypic spectrum and a progressive disease course. Early manifestations include muscle weakness, delayed achievement of developmental milestones, joint contractures, seizures and characteristic intracranial abnormalities.
More Related Videos
 10:30
Multi-exon Skipping Using Cocktail Antisense Oligonucleotides in the Canine X-linked Muscular Dystrophy
Published on: May 24, 2016
 09:18
Measurements of Motor Function and Other Clinical Outcome Parameters in Ambulant Children with Duchenne Muscular Dystrophy
Published on: January 12, 2019
Related Concept Videos
 01:21
Satellite Stem Cells and Muscular Dystrophy
 01:18
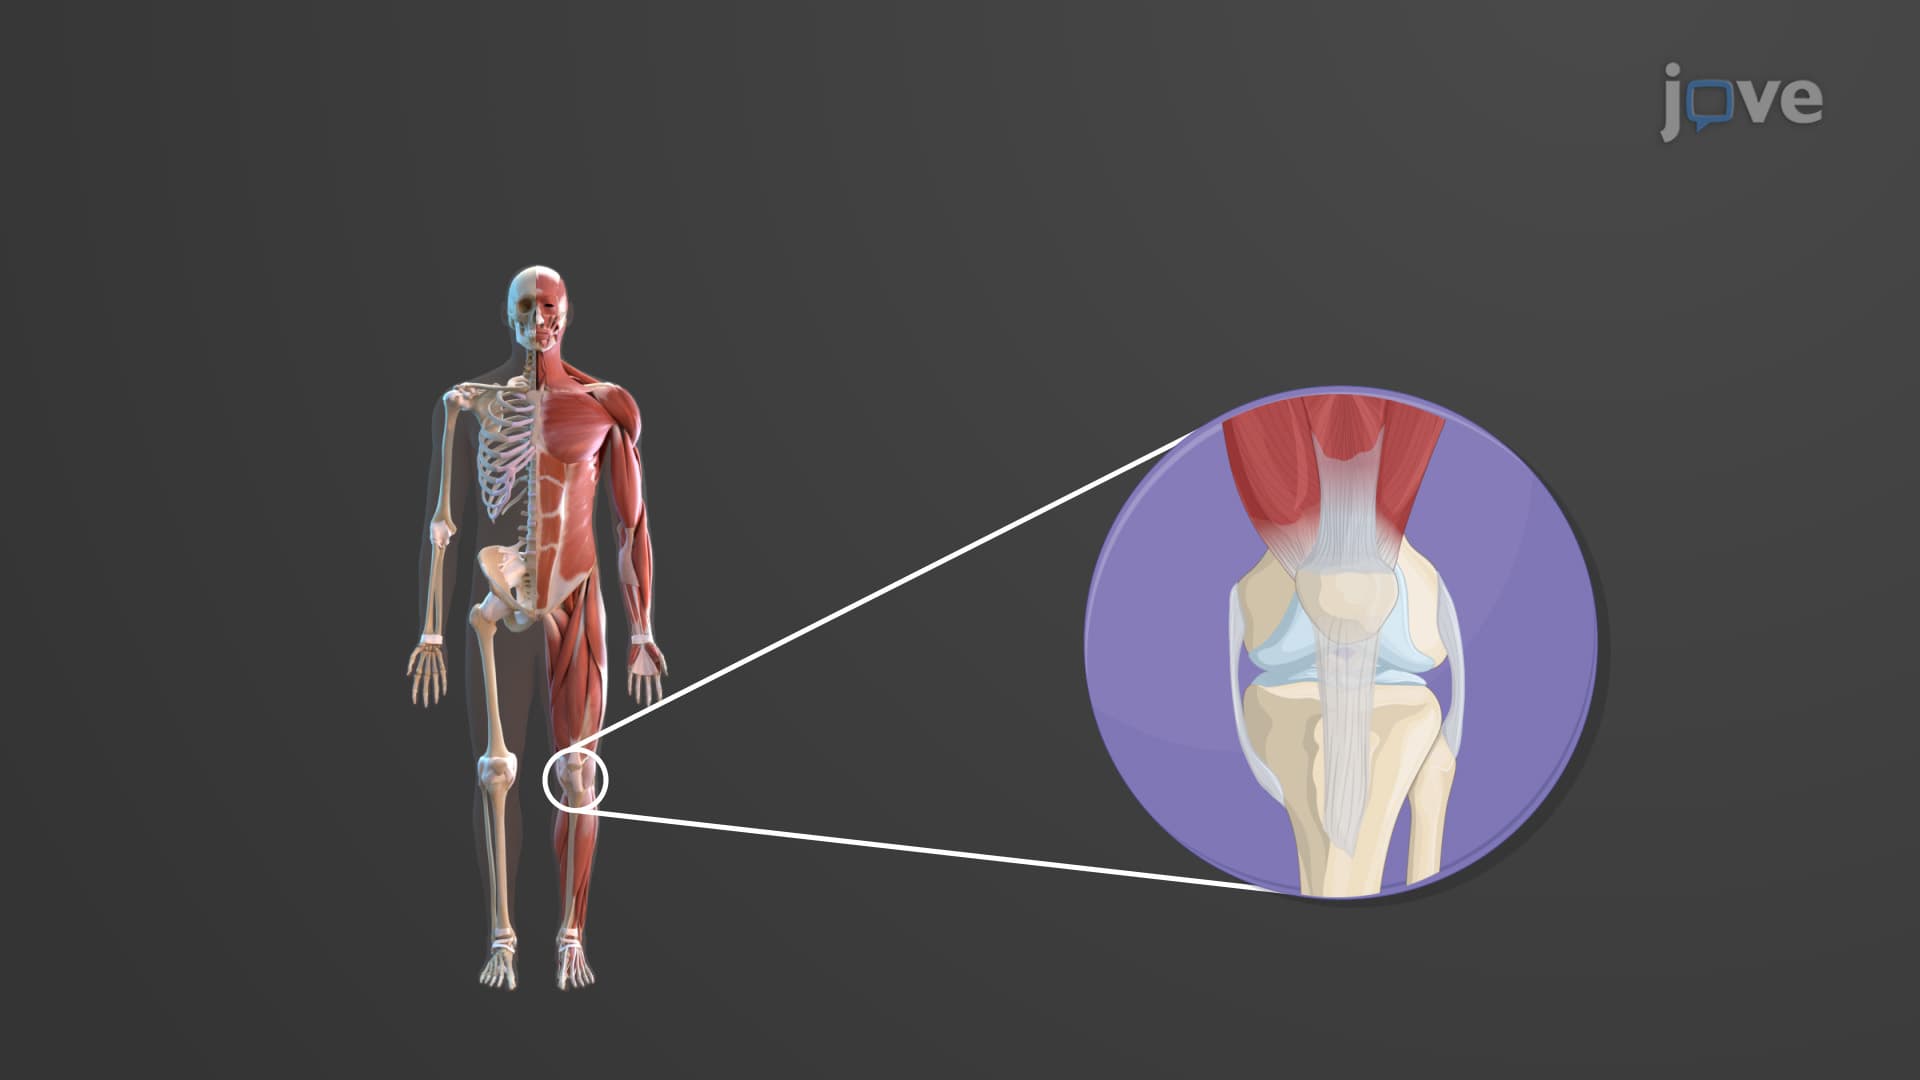
The Muscular System
 02:30
Histone Variants at the Centromere
 02:23
Abnormal Proliferation
 03:17
Inheritance of Chromatin Structures
 01:30
Respiratory System Abnormal Finding I: Inspection and Percussion
Inspection Findings
During an inspection, several findings may suggest the presence of respiratory distress or disease. Pursed-lip breathing, where exhalation is slowed by...