

Exploring the in vitro urease inhibitory potential, molecular docking and dynamics simulations investigation of
Obaid Ullah1, Aftab Alam2, Muhammad Ayaz1
1Department of Chemistry, University of Malakand, Khyber Pakhtunkhwa, Pakistan.

Aims:
To biologically evaluate a series of hydrazide derivatives of mefenamic acid as anti-urease agents, elucidating their mechanism of action through integrated computational and experimental approaches.
Method:
The inhibitory mechanism was interpreted through comprehensive computational studies: molecular docking to identify binding modes within the catalytic site of H. pylori urease (PDB: 4UBP), Density Functional Theory (DFT) calculations to evaluate global chemical reactivity and frontier molecular orbitals (FMOs), and Molecular Dynamics (MD) simulations to confirm complex stability. Preliminary pharmacokinetic and toxicity profiles were predicted in silico.
Results:
Compound 2a, featuring a para-nitro substituent, emerged as the most potent inhibitor (IC50 = 21.33 ± 0.01 µM), outperforming the standard thiourea (IC50 = 22.36 ± 0.30 µM).
Discussion:
DFT analysis revealed 2a possessed the lowest energy gap (Δε = 2.417 eV) and highest electrophilicity index (ω = 8.658 eV), correlating with superior reactivity and charge-transfer capacity. Molecular docking confirmed competitive inhibition with 2a forming key hydrogen bonds with Arg339 and His323.
Conclusion:
This work identifies 2a as a potent, and competitive urease inhibitor. The integrated experimental, ADMET and computational strategy provides a robust molecular blueprint for its activity, positioning it as a promising lead candidate for developing new therapeutic agents against urease-related pathologies.
More Related Videos
 05:57
Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 03:05
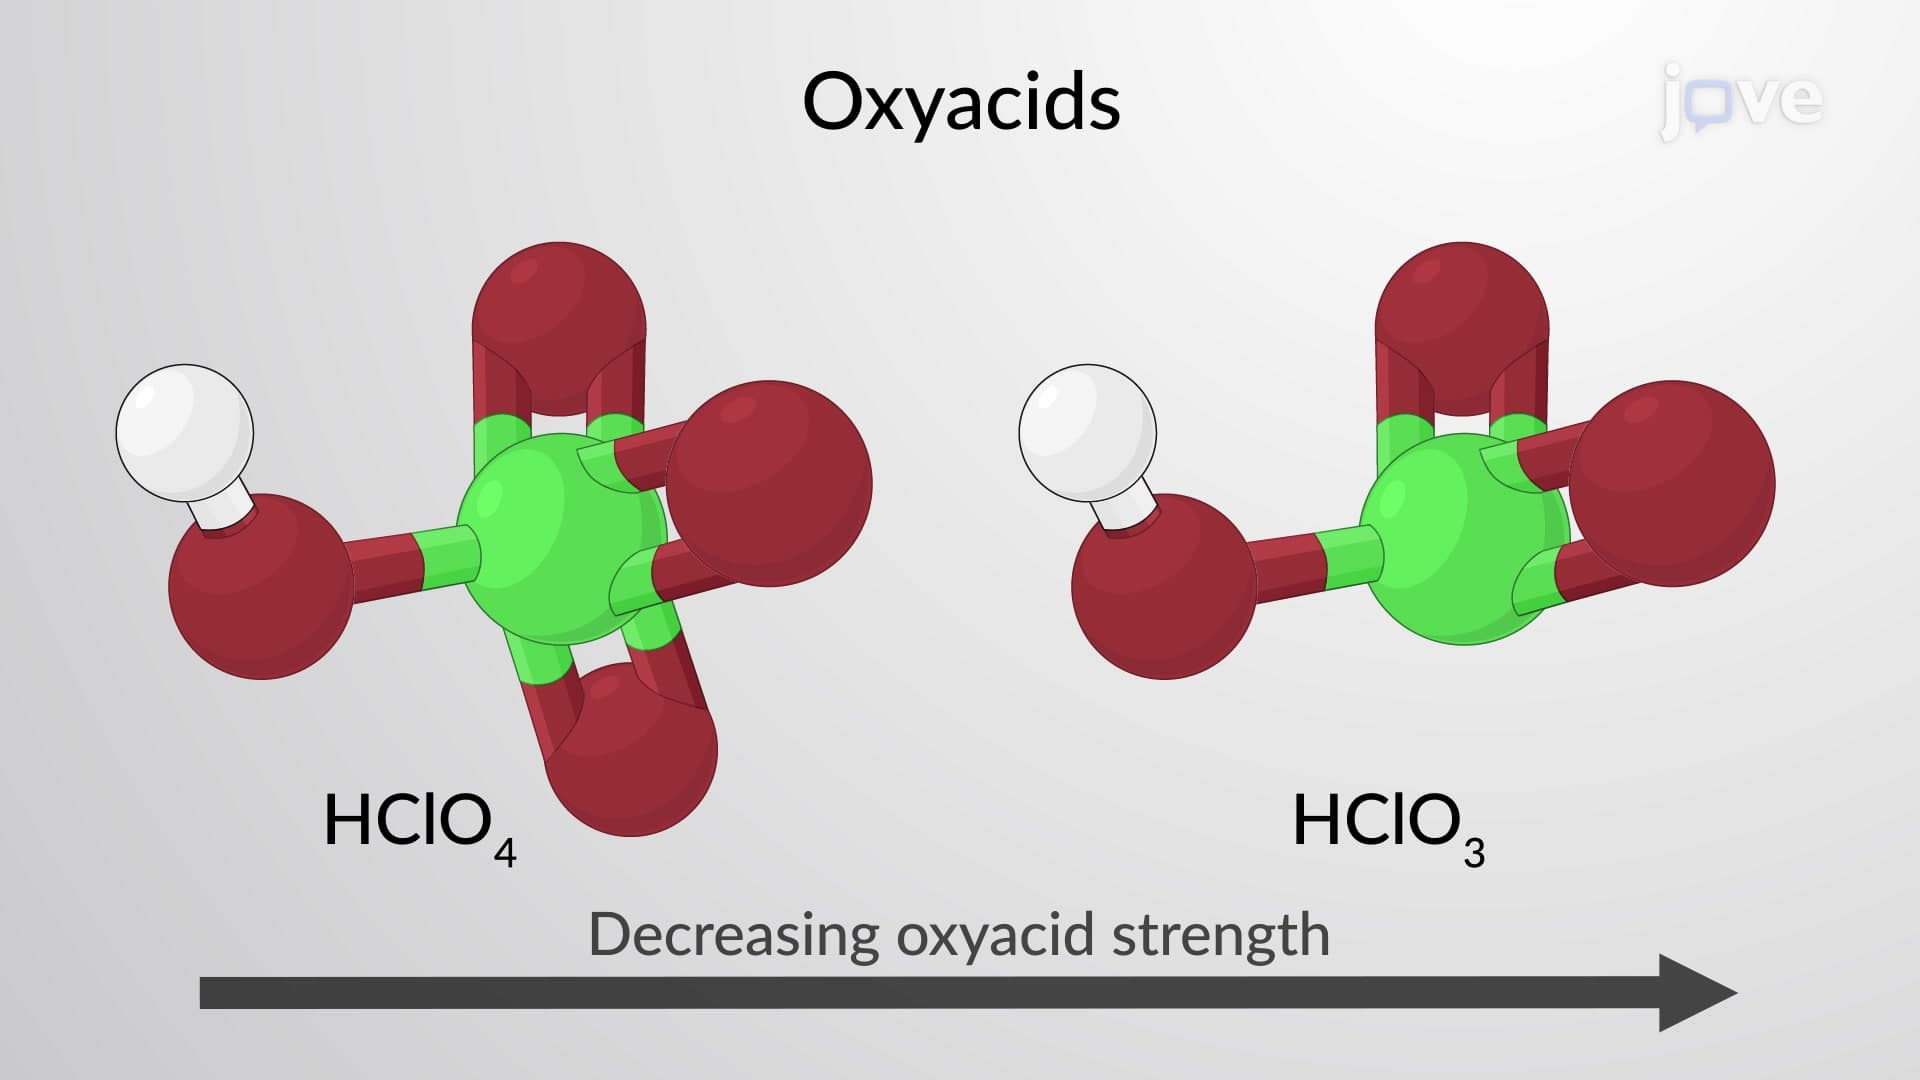
Acid Strength and Molecular Structure
In the absence of any leveling effect, the acid strength of binary compounds of hydrogen with nonmetals (A) increases as the H-A bond strength decreases down a group in the periodic table. For group 17, the order of increasing acidity is HF < HCl < HBr < HI. Likewise, for group 16, the order of increasing acid strength is H2O < H2S < H2Se < H2Te. Across a row in the periodic table, the acid strength of binary hydrogen compounds increases with increasing...
 02:34
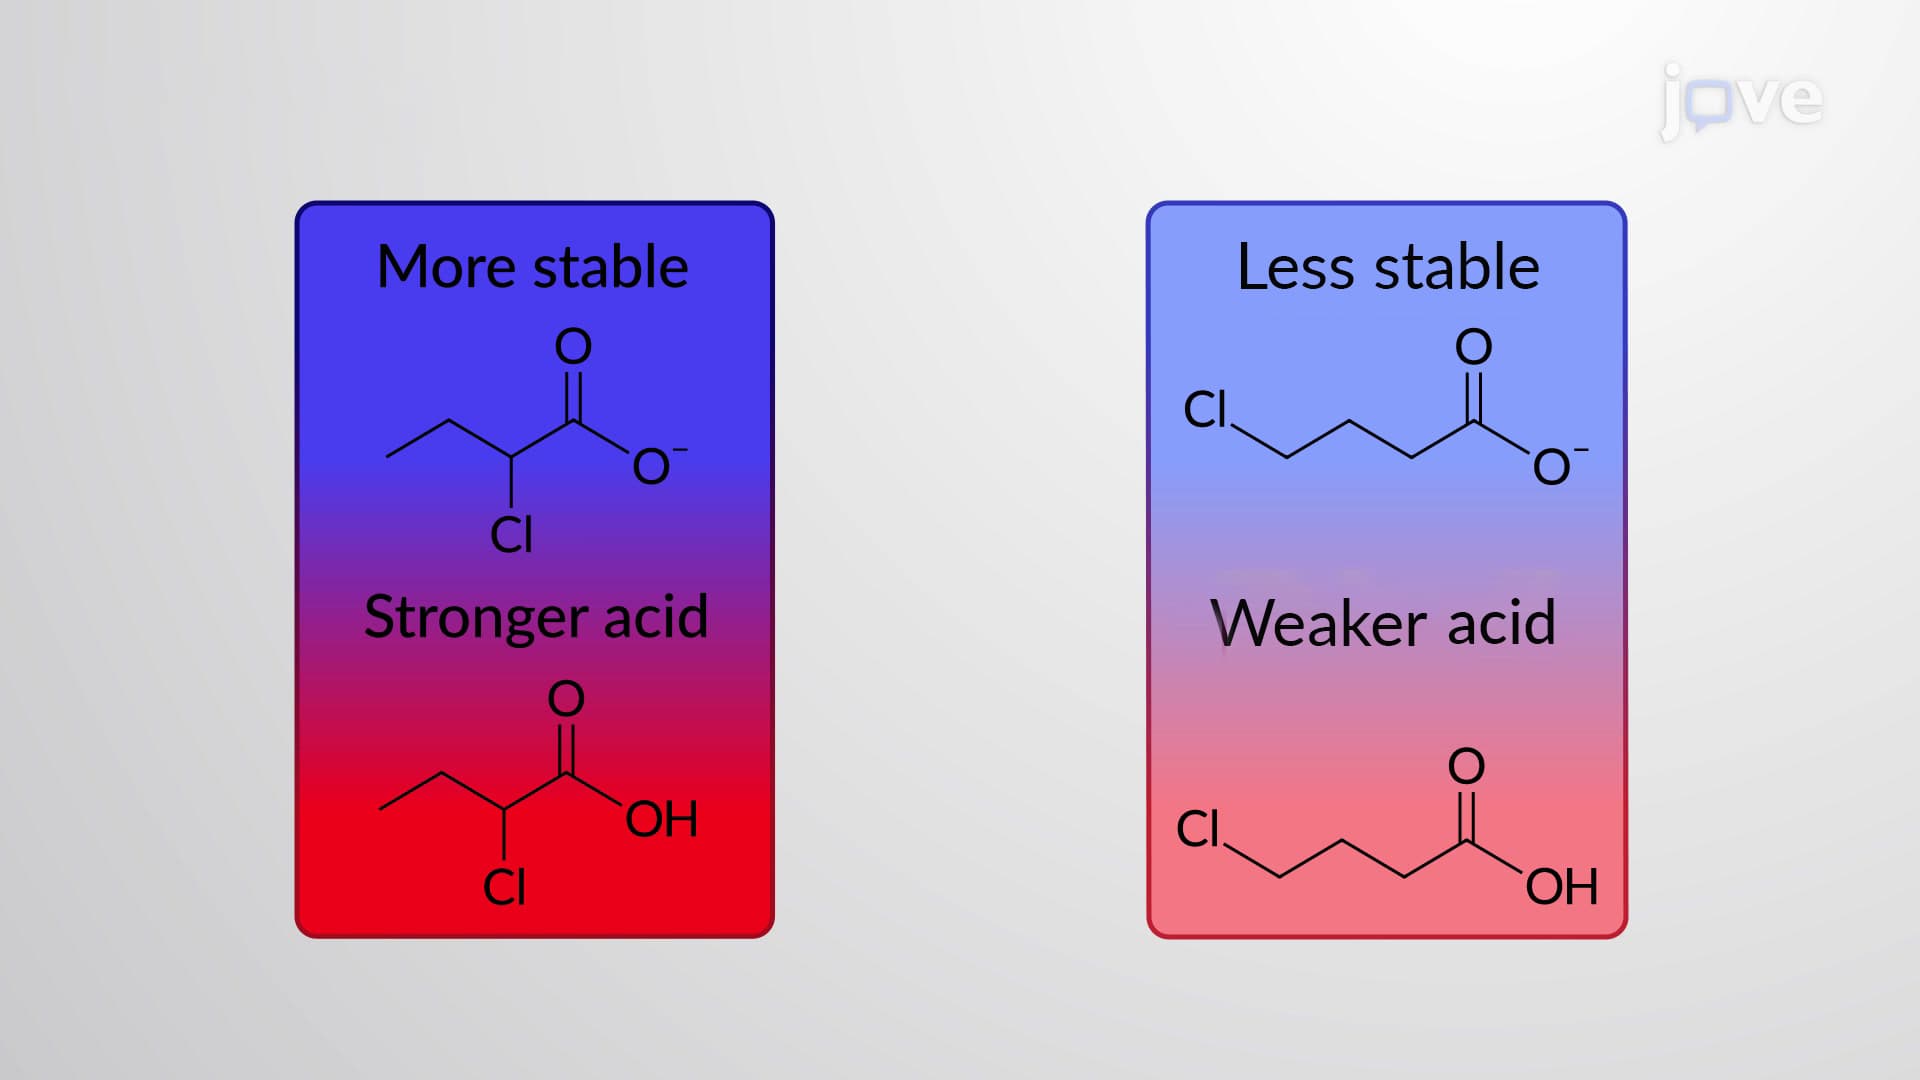
Molecular Structure and Acidity
The size effect explains the change in atomic size on acidity. When comparing the acids formed from elements that belong to the same column in the periodic table, their atomic sizes...
 01:25
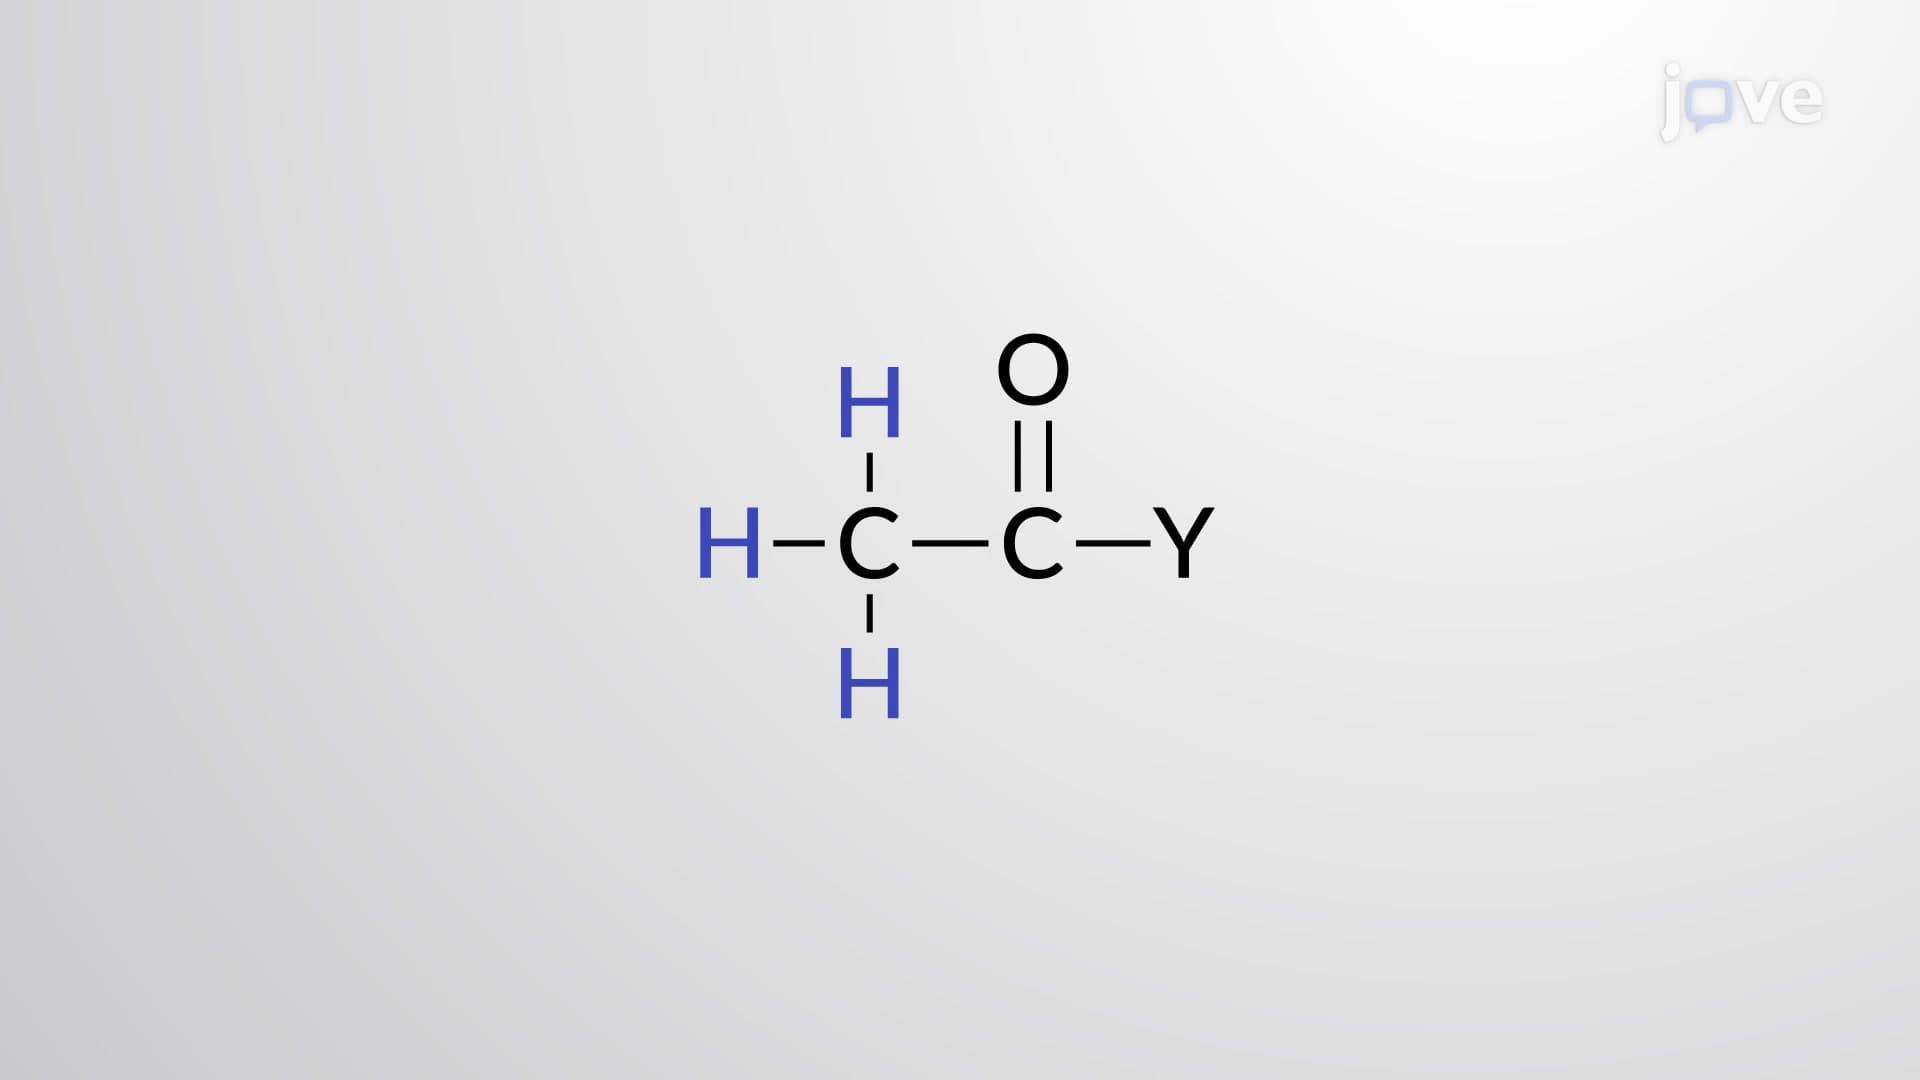
Acidity and Basicity of Carboxylic Acid Derivatives
The relative acidic strength of the derivatives can be explained based on the extent of resonance stabilization of the conjugate base. The...
 01:28
Structures of Carboxylic Acid Derivatives
Carboxylic acid derivatives contain an acyl group attached to a heteroatom such as chlorine, oxygen, or nitrogen. The carbonyl carbon and oxygen are both sp2-hybridized with an unhybridized p orbital.
The three sp2 orbitals of the carbonyl carbon form three σ bonds, one each with the carbonyl oxygen, the α carbon, and the heteroatom, whereas the other two sp2 orbitals of the carbonyl oxygen are occupied by the lone pairs. Further, the unhybridized p...
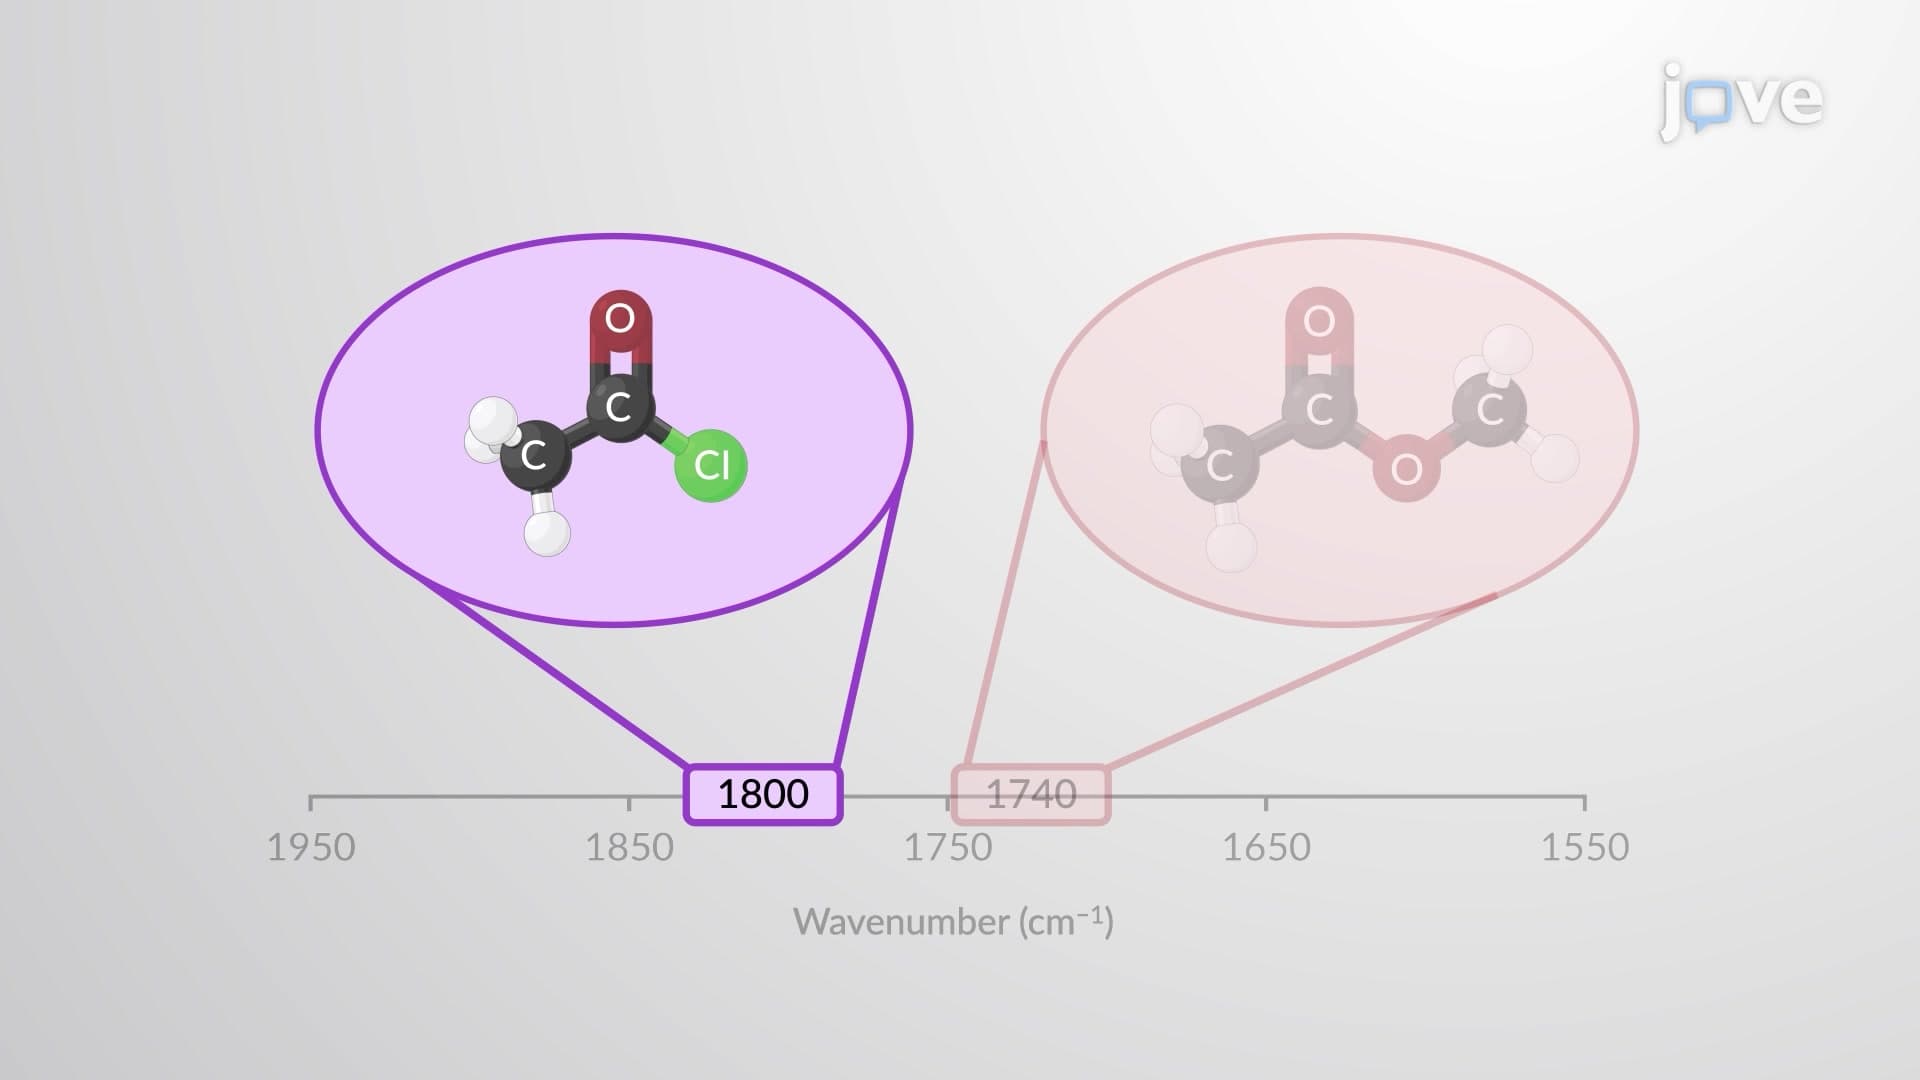 01:26
Spectroscopy of Carboxylic Acid Derivatives
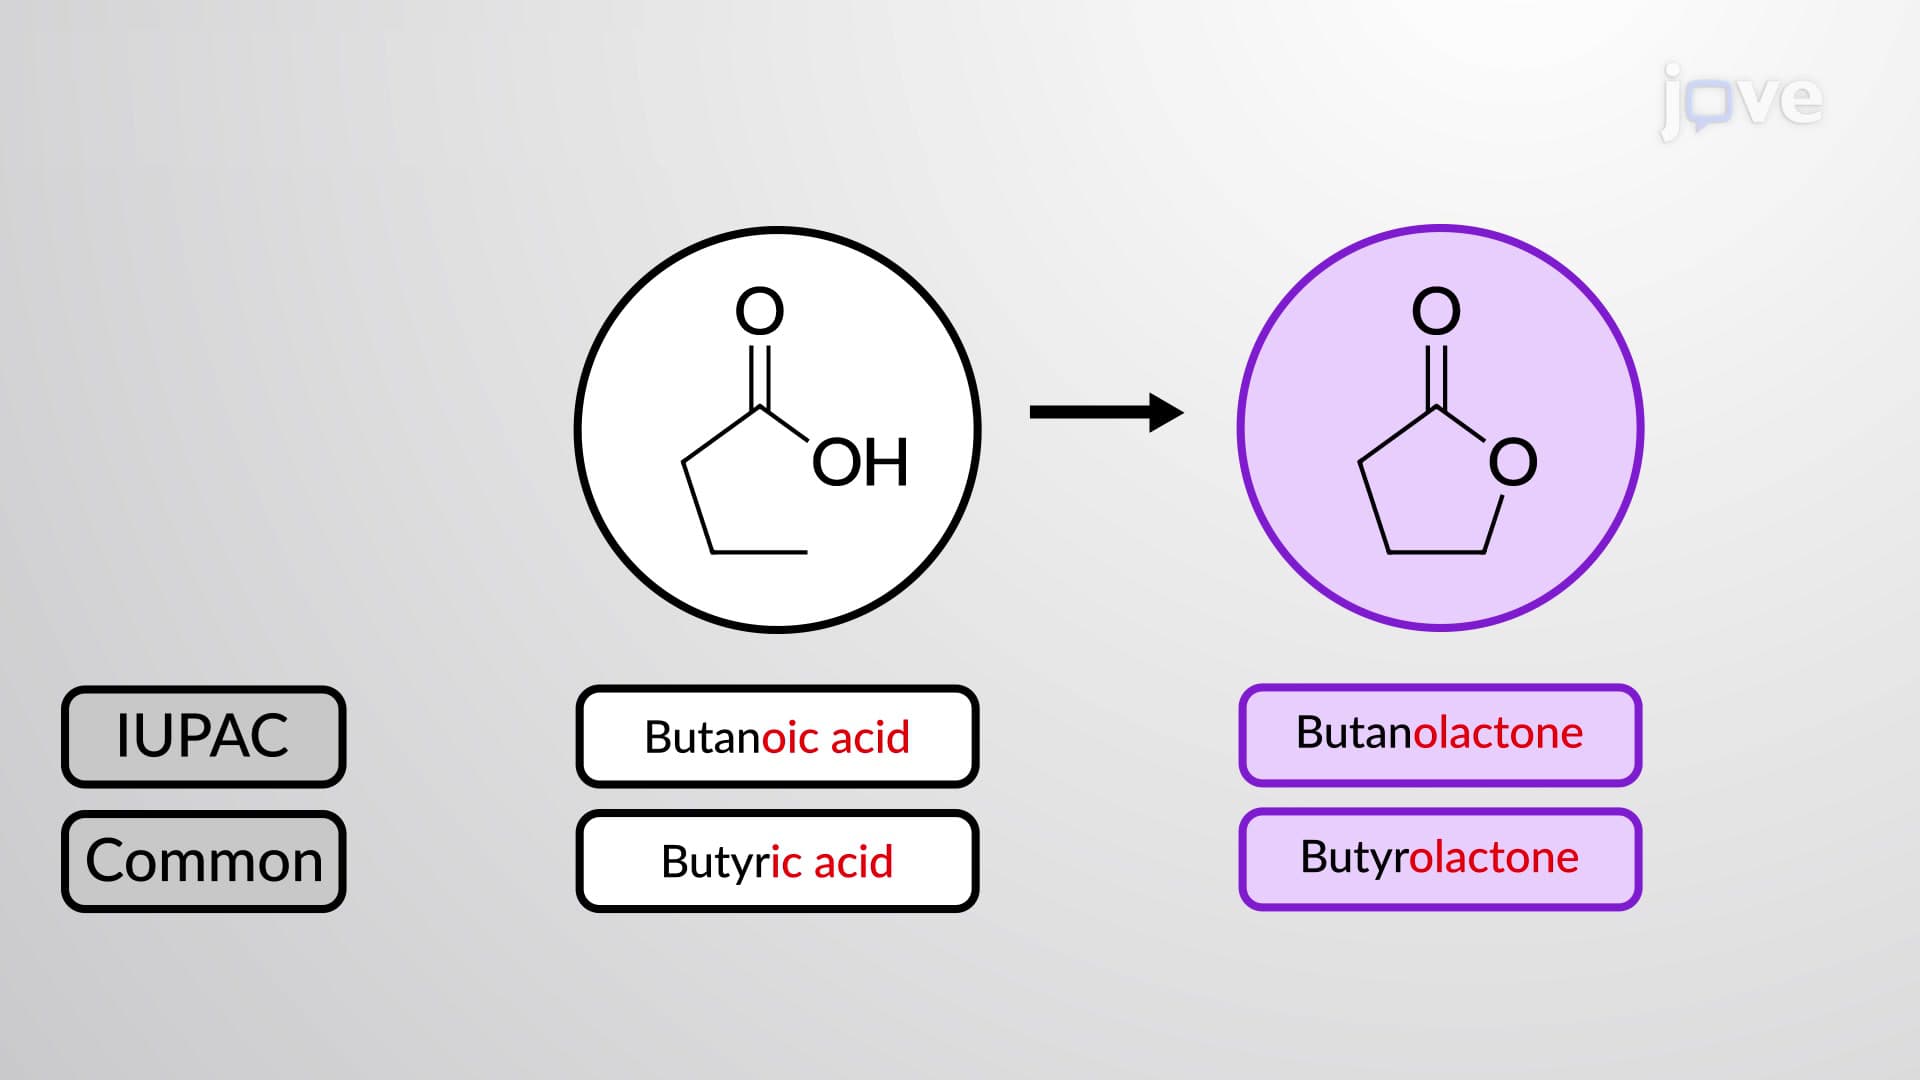 01:16
Nomenclature of Carboxylic Acid Derivatives: Acid Halides, Esters, and Acid Anhydrides
The IUPAC and common names of acid halides are derived from the corresponding carboxylic acids, by changing “ic acid” to “yl halide.” For example, as shown below, the IUPAC name ethanoyl chloride is derived from ethanoic acid, and the common name, acetyl chloride, is obtained from acetic acid.