
Long-term Outcomes of Pediatric CFSPID: A 15-Year Clinical and Genomic Study Across Newborn Screening Cystic Fibrosis
Ana Morales-Tirado1, Enrique Blitz-Castro1, Ana Tabares-González2
1Cystic Fibrosis Unit, Department of Pediatrics, Hospital Universitario Ramón y Cajal, M-607, Km 9.100, 28034 Madrid, Spain; Department of Medicine and Medical Specialties, Faculty of Medicine and Health Sciences, Universidad de Alcalá, 28801 Alcalá de Henares, Spain; Instituto Ramón y Cajal de Investigación Sanitaria (IRYCIS), M-607, Km 9.100, 28034 Madrid, Spain.

Background:
Newborn blood spot screening (NBS) for cystic fibrosis (CF) increasingly identifies infants with inconclusive results, classified as CF screen-positive inconclusive diagnosis (CFSPID). However, long-term outcome data remain limited.
Objective:
To analyze the clinical and biochemical course of children with CFSPID.
Methods:
We conducted a bi-center observational cohort study including all children designated as CFSPID through the Madrid NBS program from July 2009 to June 2024. Follow-up assessments included serial sweat chloride (SC) testing, respiratory and gastrointestinal (GI) evaluations, spirometry from age 5 or older, nasopharyngeal cultures, and fecal elastase measurements.
Results:
A total of 100 children were enrolled. After a median follow-up of 3.59 years [IQR, 1.73-5.43], 25% were reclassified as unaffected carriers following CFTR variant reinterpretation. The remaining 75 were categorized into 3 genotype groups: 66 with CF-causing (CFc)/Variant of Varying Clinical Consequence (VVCC), 4 with VVCC/VVCC, and 5 with CFc/Variant of Uncertain Significance (VUS). By the end of follow-up, 42.67% developed at least 1 intermediate or positive SC value. Three children (4%), all carrying the CFc/VVCC genotype, converted to cystic fibrosis (mean age at conversion, 4.23 years), and 1 child (1.3%) developed a CFTR-related disorder. Clinical signs were mild, with normal spirometry and full pancreatic sufficiency. No Pseudomonas aeruginosa isolates were detected.
Conclusions:
Conversion from CFSPID to CF was rare (4%), but abnormal SC values were frequent (42.67%), supporting the need for structured, long-term monitoring. Continued follow-up in specialized CF centers is essential for early detection of disease progression.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 02:02
Genomics
 01:16
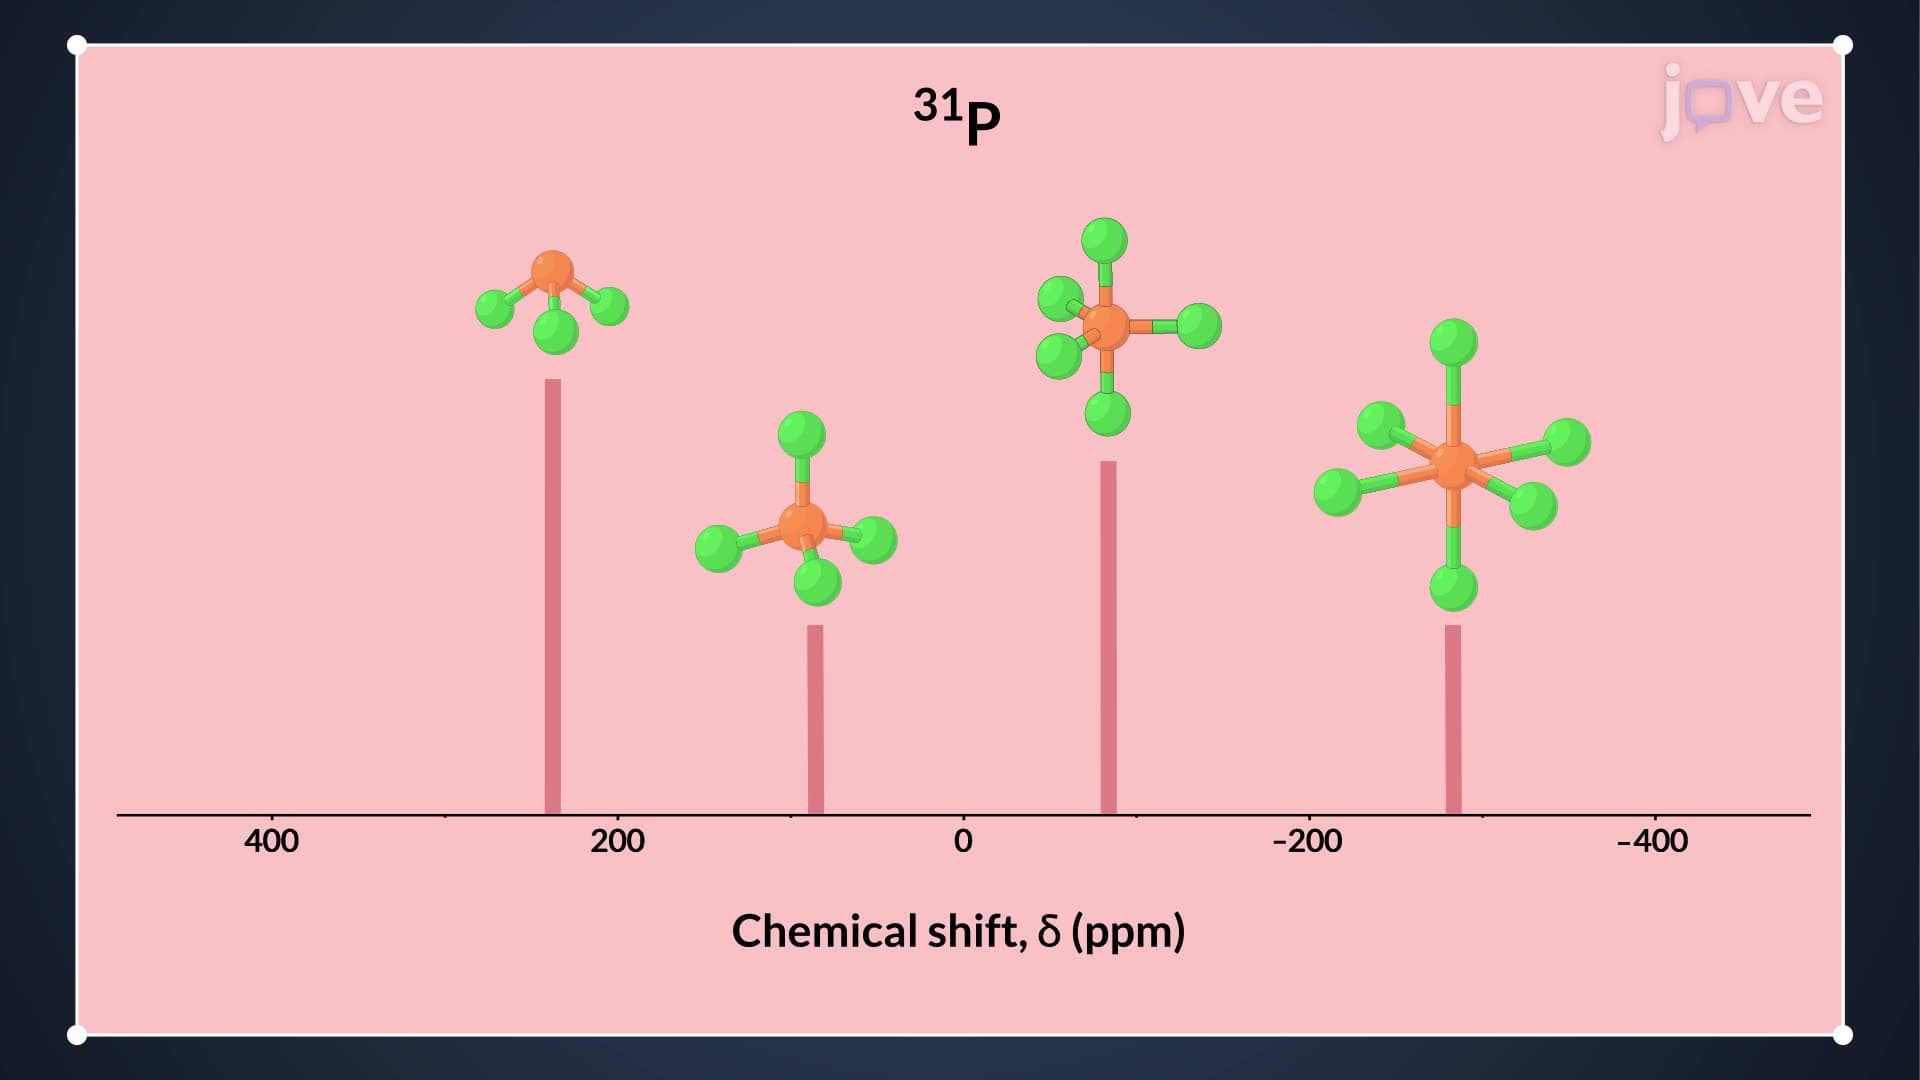
Other Nuclides: 31P, 19F, 15N NMR
While fluorine-19 and phosphorous-31 have high natural abundances (100%) and positive gyromagnetic ratios, nitrogen-15 has a low natural abundance and a negative gyromagnetic ratio. However, nitrogen-15 is still preferred over nitrogen-14 (which has a...
 03:38
Measurement: Standard Units
 03:02
Measurement: Derived Units