Related Experiment Video
Updated: Feb 26, 2026
Shotgun Lipidomics of Rodent Tissues
Published on: November 18, 2022
p19ARF deficiency disrupts lung and lipid homeostasis resembling the human alveolar proteinosis
Lidia Jiménez-Garcia1, Andrea Pérez-Montero1,2, Sandra Herranz1
1Unidad de Terapias Farmacológicas, Área de Genética Humana, Instituto de Investigación de Enfermedades Raras, Instituto de Salud Carlos III, Madrid, Spain.

Abstract:
The alternative reading frame (ARF) protein, encoded by the CDKN2A locus, is well-recognized for its role in tumor suppression. Emerging evidence has highlighted ARF as a critical regulator of innate immunity and inflammation, with links to increased susceptibility to cardiometabolic diseases. This study investigates the role of ARF in lung homeostasis and reveals that its deficiency in mice affects lipid metabolism and leads to pulmonary abnormalities resembling pulmonary alveolar proteinosis (PAP). ARF-deficient mice exhibited abnormal surfactant clearance, characterized by lipid and protein accumulation in the alveoli, foamy alveolar macrophages (AMs) with enlarged and vacuolated morphology, and increased bronchoalveolar lavage fluid turbidity. These changes were linked to disrupted surfactant homeostasis resulting from an imbalance between increased lipid uptake (via upregulation of scavenger receptors such as SR-A1 and CD36) and impaired lipid efflux, evidenced by reduced expression of the cholesterol transporter SR-BI. These mice also display reduced AM numbers, increased eosinophil and neutrophil infiltration, consistent with secondary PAP. Additionally, a distinctive chemokine and cytokine profile (elevated Ccl12, Ccl2, Cxcl1, and IL-10) was observed, which may be associated with type 2 immune responses and alternative AM polarization. Interestingly, ARF deficiency also appears to compromise AM maintenance through effects on self-renewal and survival. Pulmonary function tests revealed increased tissue elastance and damping, suggesting early-stage lung stiffness. Collectively, these findings highlight the essential role of ARF in lung homeostasis and lipid regulation, providing insights into its potential involvement in PAP pathogenesis.
Related Concept Videos
 01:05
Breathing
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:21
Acute Respiratory Failure-II
The underlying physiological abnormalities that contribute to hypoxemic respiratory failure include:
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:26
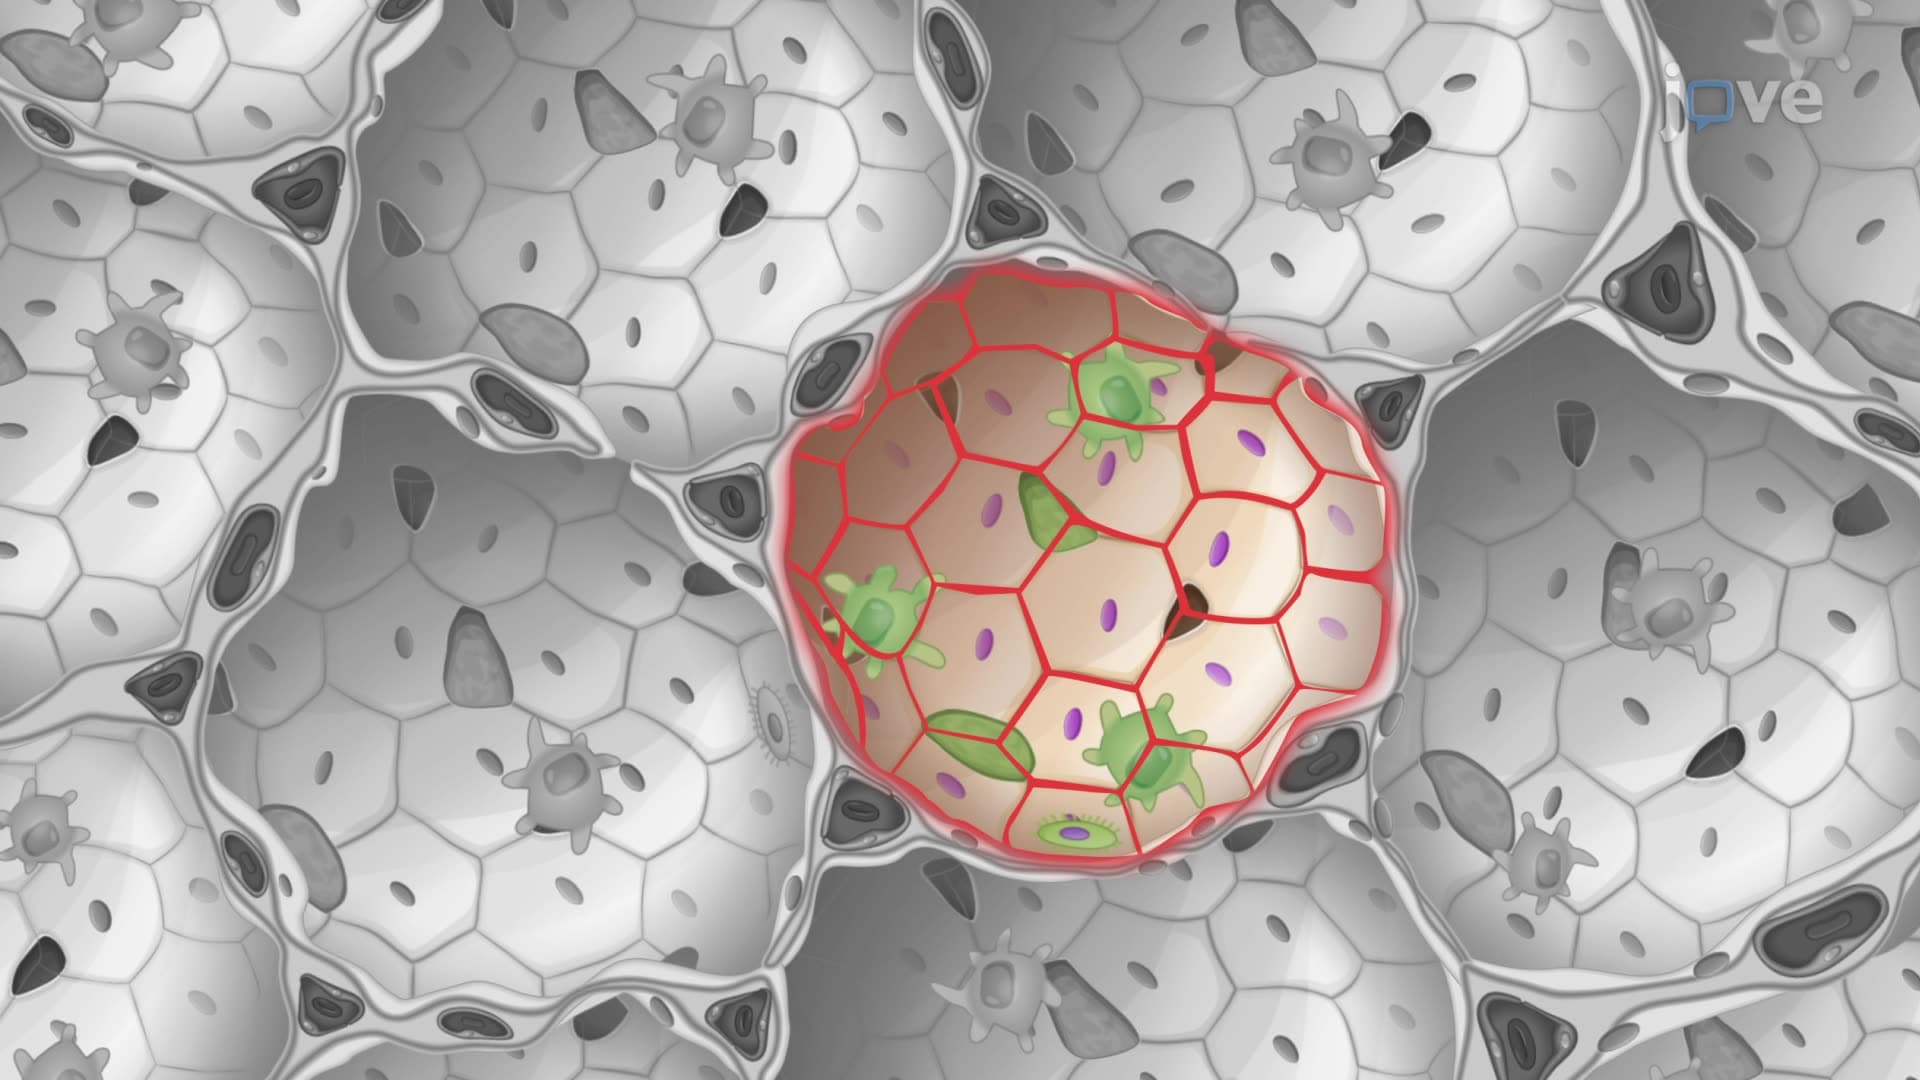
Alveoli and Alveolar Ducts
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...