Related Experiment Video
Updated: Feb 28, 2026

Nano-Differential Scanning Fluorimetry for Screening in Fragment-based Lead Discovery
Published on: May 16, 2021
Query Matters: How Selection Strategies Influence Active Learning in Drug Discovery
Huw J Williams1, Stephen D Pickett2, Andrew Baxter2
1Department of Chemistry, University of Strathclyde, 295 Cathedral Street, Glasgow, G11XL, Scotland.

Abstract:
We present SimDMTA, an in silico framework designed to simulate the Design-Make-Test-Analyze (DMTA) cycle used in preclinical drug discovery. Using docking scores as a proxy for biological assays, the simulations allow factors controlling the efficiency of the DMTA cycle to be explored in a manner that would not be feasible using traditional experiments due to time and cost constraints. In this workflow, a machine learning model predicts docking scores, selects compounds using various query strategies, docks selected molecules, and retrains iteratively. Starting from a broad chemical space, the model actively samples molecules derived from a 3,5-dimethyl-4-phenylisoxazole scaffold, an active warhead for the Bromodomain 4 (BRD4) BD1 binding site, to refine its predictions. Our results show that uncertainty-based sampling significantly outperforms greedy and hybrid approaches in both hit discovery and the ability of the model that predicts docking scores to generalize beyond its training set. Notably, by the final iteration, 37 of the top 50 ranked compounds were within the top 1% of the chemical space of all evaluated compounds. Strategies that include some random selection correct systematic biases more rapidly, but are less effective at predicting top-performing molecules. These findings underscore the value of incorporating molecular diversity and uncertainty into design strategies. While such strategies may deprioritize those molecules with the highest absolute predictions in early rounds, they markedly accelerate model refinement, ultimately leading to more effective hit identification in discovery driven by active learning.
More Related Videos
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:24
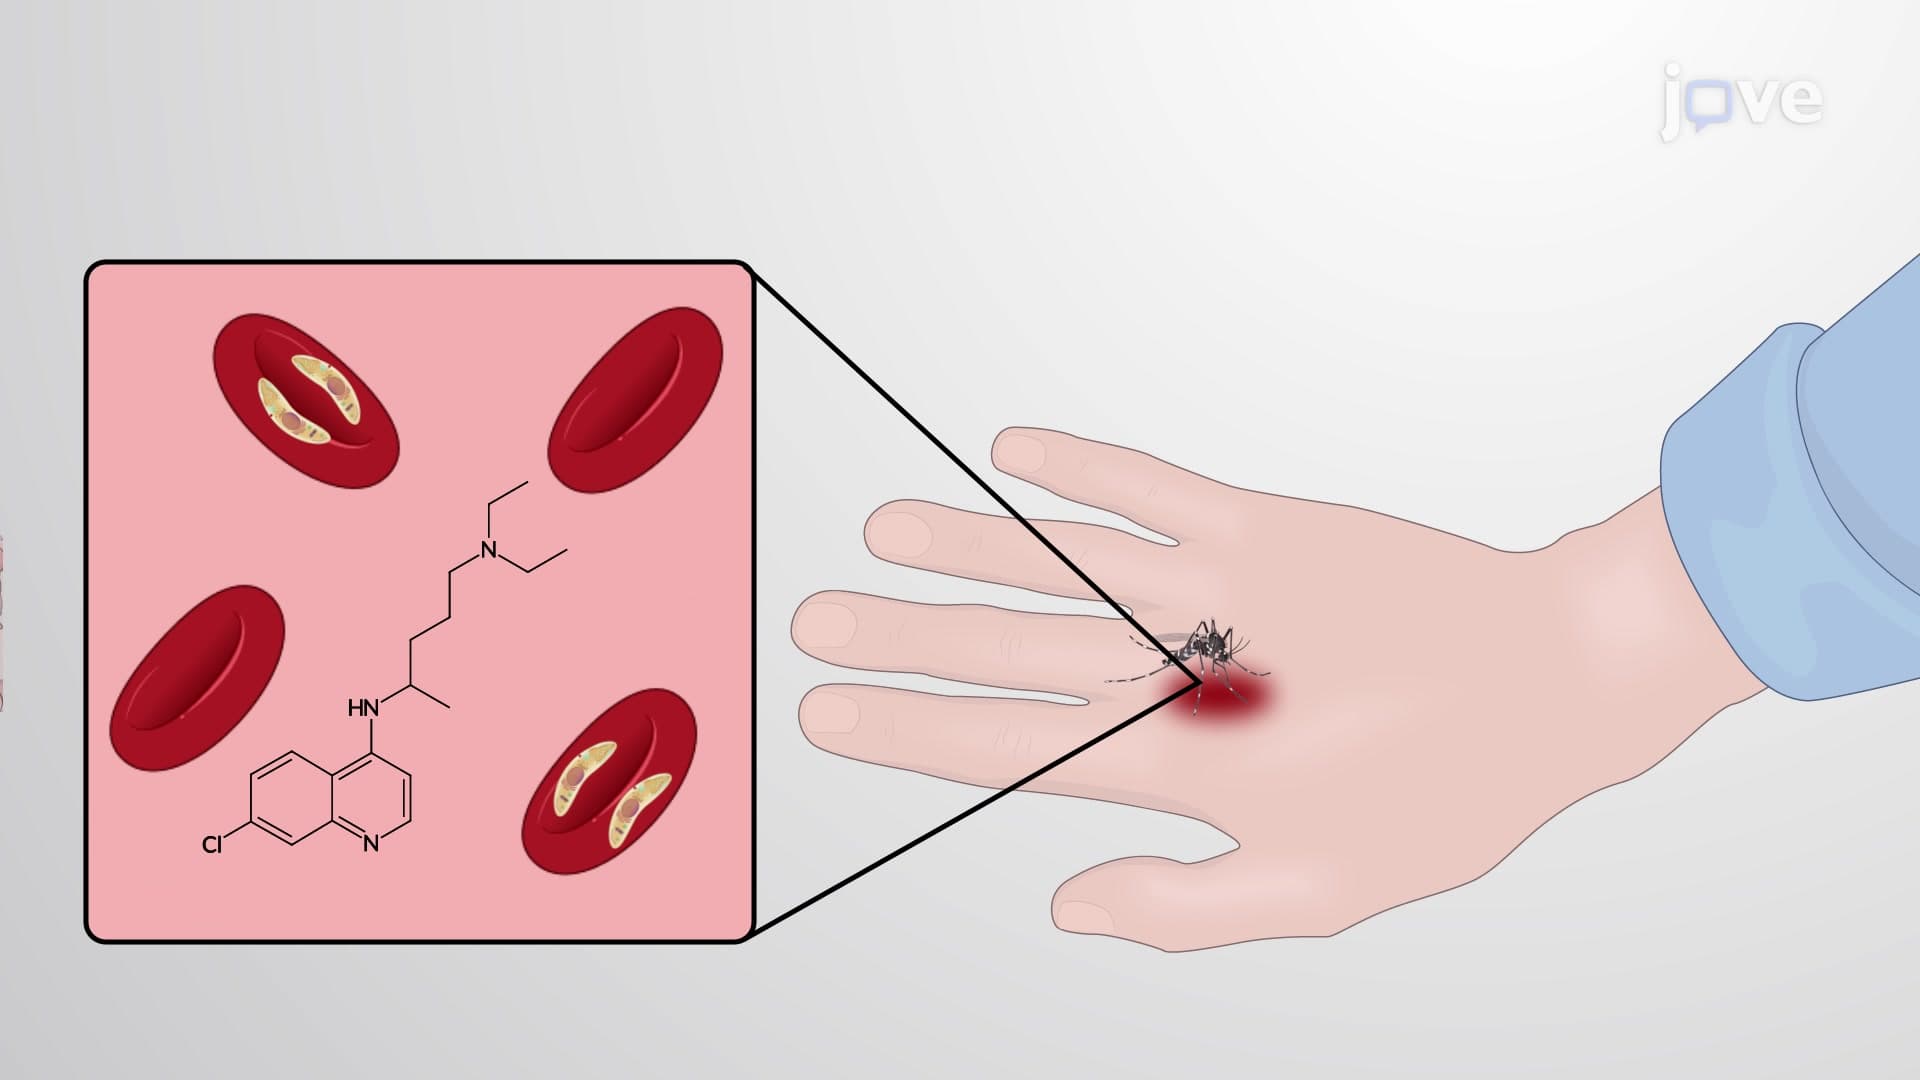
Principles of Drug Action
Drugs can be agonists or antagonists. Like the endogenous ligands, agonists always bind and activate the target to produce a cellular response. Agonist binding induces a conformational change which in turn...
 01:19
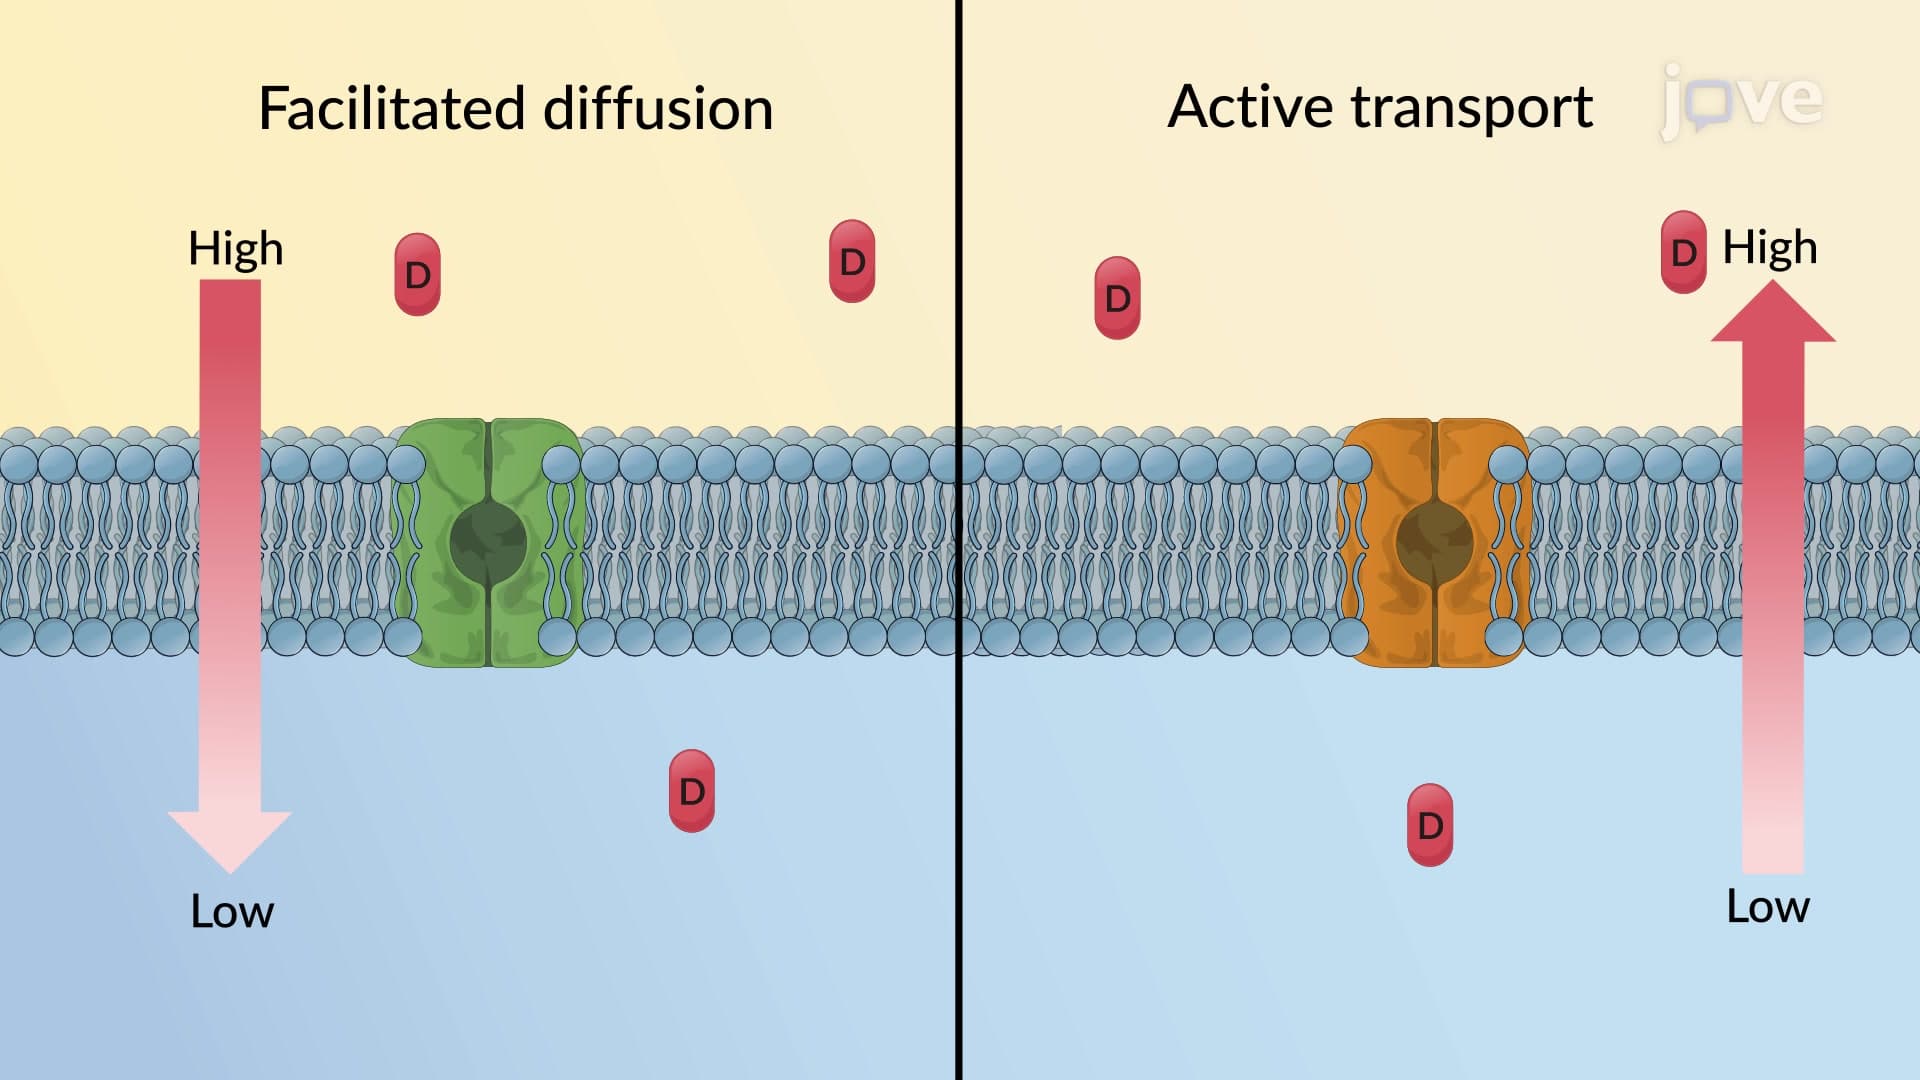
Drug Absorption Mechanism: Carrier-Mediated Membrane Transport
Facilitated diffusion is a passive process that utilizes human Solute Carrier (SLC) transporters. These transporters bind to the drug, undergo structural...
 01:26
Targets for Drug Action: Overview
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...
 01:29
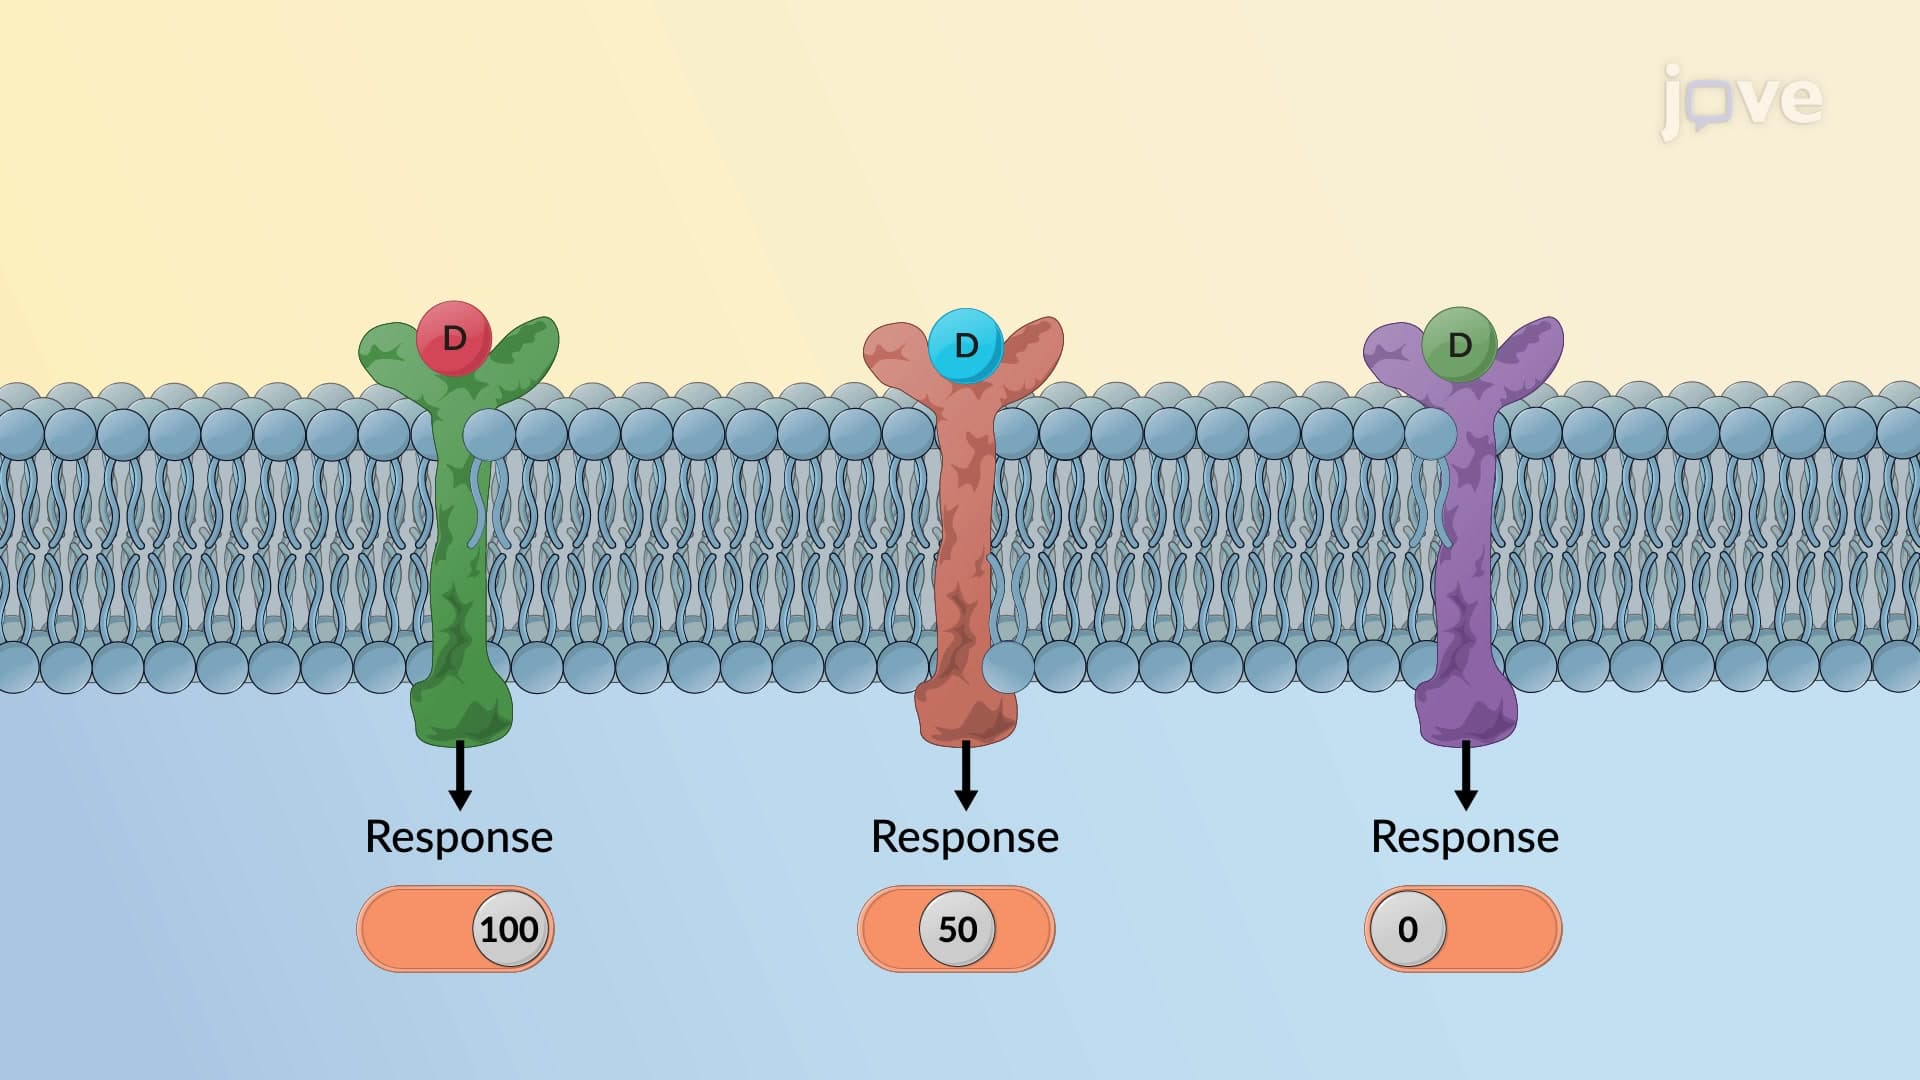
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....