Related Experiment Video
Updated: Feb 28, 2026
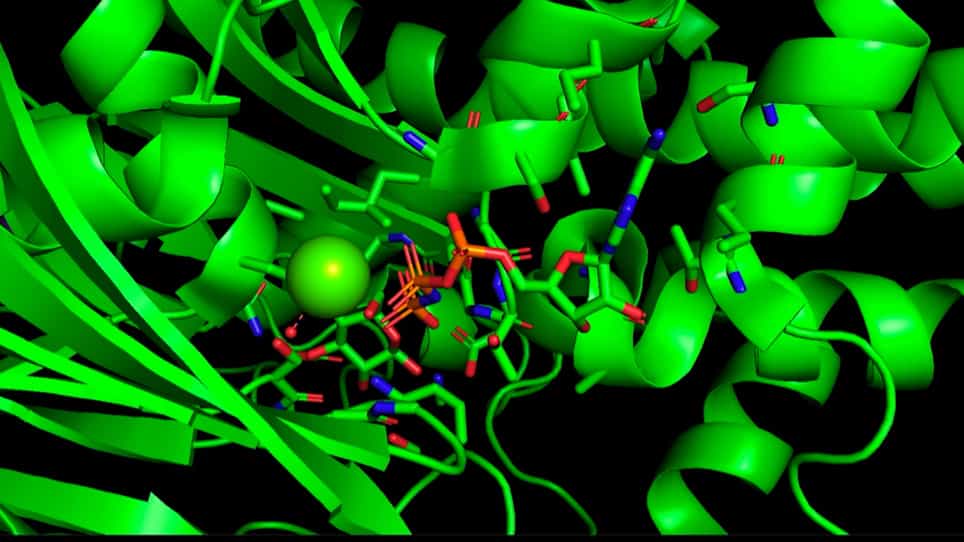
Modeling an Enzyme Active Site using Molecular Visualization Freeware
Published on: December 25, 2021
Defining the Active Conformation of Typical Protein Kinase Domains from Substrate-Bound PDB Structures Enables
Joan Gizzio1, Bulat Faezov1,2,3, Qifang Xu1
1Institute for Cancer Research, Fox Chase Cancer Center, Philadelphia PA 19111, USA.

Abstract:
Humans have 437 catalytically competent protein kinase domains with the typical kinase fold, similar to the structure of Protein Kinase A (PKA). The active form of a kinase must satisfy requirements for binding ATP, magnesium, and substrate. From structural bioinformatics analysis of 248 crystal structures of 54 unique substrate-bound kinases, we derived structural criteria for the active form of typical protein kinases. We include well-known requirements on the DFG motif of the activation loop and the N-terminal domain salt bridge, but also on the positions of the N-terminal and C-terminal segments of the activation loop that must be placed appropriately to bind substrate. With these criteria, only 130 of the 437 human catalytic protein kinases (30%) are in the Protein Data Bank in their active form. Because the active forms of catalytic kinases are needed for understanding substrate specificity and the effects of mutations on catalytic activity in cancer and other diseases, we used AlphaFold2 to produce models of all 437 human protein kinases in the active form. This was accomplished with templates from the PDB that resemble substrate-bound structures, shallow multiple sequence alignments of orthologs and close paralogs of the query protein, and application of the active-kinase criteria to the output models. We selected models for each kinase based on intramolecular ipSAE scores of the activation loop residues of these models, demonstrating that the highest scoring models have the lowest or close to the lowest RMSD to 29 non-redundant substrate-bound structures in the PDB. A larger benchmark of 117 active kinase structures with solved activation loops in the PDB shows that 71% of the highest scoring AlphaFold2 models had backbone RMSD < 1.0 Å to the benchmark structures and 92% were within 2.0 Å. Models for all 437 catalytic kinases are available at https://dunbrack.fccc.edu/kincore/activemodels. We believe they may be useful for interpreting mutations leading to constitutive catalytic activity in cancer as well as for templates for modeling substrate and inhibitor binding for molecules which bind to the active state.
Related Concept Videos
 02:54
Protein Kinases and Phosphatases
Protein kinases
Many proteins in the cell are regulated by phosphorylation, the addition of a phosphate group. A family of enzymes called kinases...
02:54Protein Kinases and Phosphatases
 01:30
Assembly of Signaling Complexes
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...
 01:13
Induced-fit Model
Enzymes exhibit substrate specificity, meaning that they can only bind to certain substrates. This is mainly determined by the shape and chemical...
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...