EMMVEP: An Ensemble Method for Protein Missense Variant Effect Prediction Based on Multi-Source Feature Fusion
Related Experiment Video
Updated: Feb 28, 2026
Identification and Classification of Position-specific GABAA Receptor Subunit Missense Variants for Their Role In Hippocampal Pyramidal Neurons
Published on: June 6, 2025
Huiling Zhang1, Junwen Huang2, Yuetong Li1
1College of Mathematics and Information, College of Software Engineering, South China Agricultural University, Guangzhou, 510642, China.

Abstract:
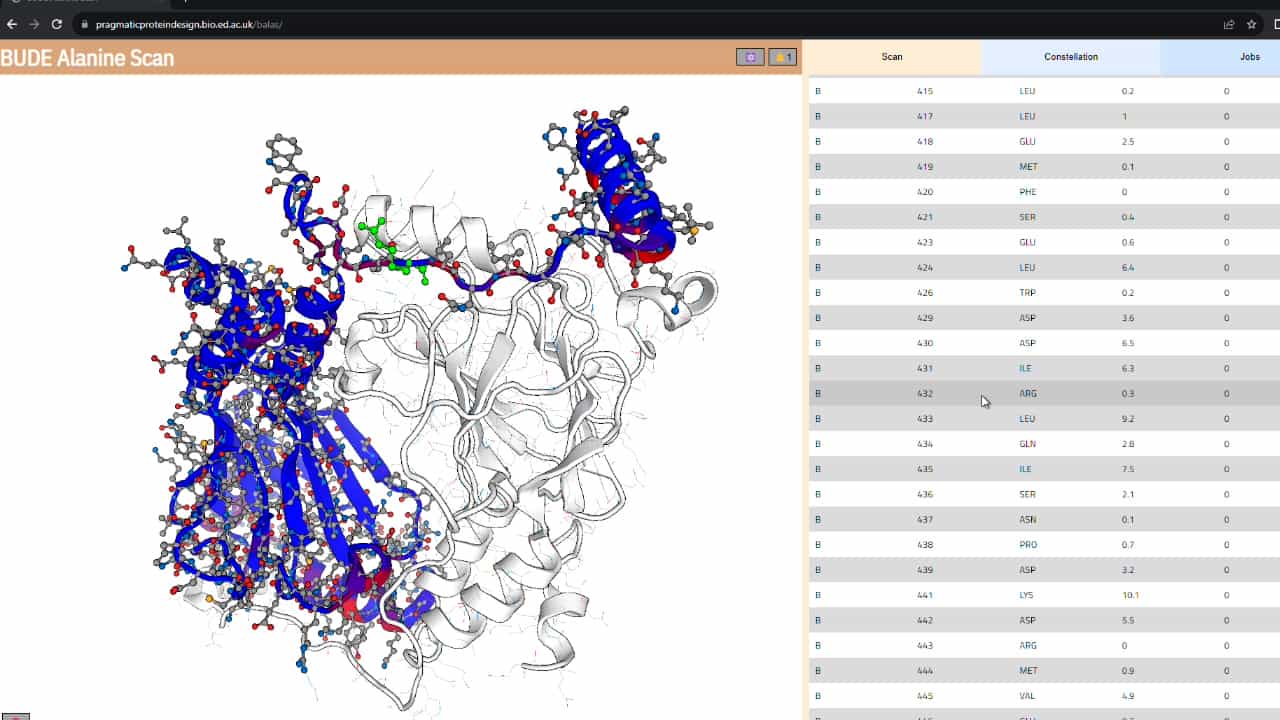
Missense mutations are common in the coding genome and can alter protein functions. Distinguishing pathogenic from benign variants remains challenging despite computational advances. In the present work, we introduce EMMVEP, an ensemble-based approach designed for predicting the effects of protein missense mutations. EMMVEP leverages categorical boosting to integrate different types of features: one-hot encoding from protein sequence, physicochemical and environment properties extracted from AlphaFold database, and allele frequency information from gnomAD. When evaluated on a benchmark dataset with 112,832 clinical significance labels, our method achieved AUC and AUPR of 0.907 and 0.879, outperforming 20 general VEP methods. To aid in the identification of pathogenic mutations among the vast number of rare variants discovered through large-scale sequencing studies, we provide the pathogenicity probabilities of 216 million potential amino acid substitutions in 19,233 human protein-encoding genes. Our work demonstrates that EMMVEP can offer valuable independent insights for missense mutation interpretation in proteins, with significant applicability in both research and clinical contexts.
More Related Videos

Related Concept Videos
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
 01:24
Tagging and Fusion Proteins
 02:51
Multi-species Conserved Sequences
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
 02:07
Improving Translational Accuracy
02:07Improving Translational Accuracy
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...