Related Experiment Video
Updated: Mar 18, 2026


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
A dual-branch graph neural network architecture for drug-target binding affinity prediction
Khushnood Abbas1, Chen Hao2, Dong Shi3
1School of Computer Science and Technology, Zhoukou Normal University, Henan, China. khushnood.abbas@zknu.edu.cn.

Abstract:
Graph Neural Networks have emerged as a powerful paradigm for artificial intelligence driven drug discovery, offering molecular representation learning that surpasses many conventional approaches. Traditional experimental pipelines are both time and resource-intensive, modern computational strategies-particularly those that integrate curated libraries of FDA-approved drugs-can accelerate target identification and candidate prioritization. In this work we introduce a dual-branch GNN architecture that synergistically combines Graph Convolutional Neural Networks, the GraphSage framework, and Jumping-Knowledge modules. This network jointly encodes structural topology and functional attributes, generating enriched embeddings for molecular graphs. We evaluated the proposed model against 45 state-of-the-art drug and target encoding baselines across well known Davis and KIBA datasets.The Proposed Model demonstrates a quantitative improvement over the GCN model, achieving a reduction in MSE (33.98 vs. 35.24), a slightly higher Pearson index (76.49 vs. 76.19), and a better Concordance index (85.41 vs. 84.41), indicating superior performance in terms of both prediction accuracy and ranking, demonstrating superior accuracy and robustness for candidate screening and establishing a new reference point for cheminformatics tasks. To illustrate practical impact, we performed a case study on COVID-19 drug repurposing: the top-ranked drugs have been also found potential drugs including Imunovir and Remdesivir from existing antiviral drugs.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 13:18
Network Pharmacology Prediction and Experimental Validation of Trichosanthes-Fritillaria thunbergii Action Mechanism Against Lung Adenocarcinoma
Published on: March 3, 2023
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:25
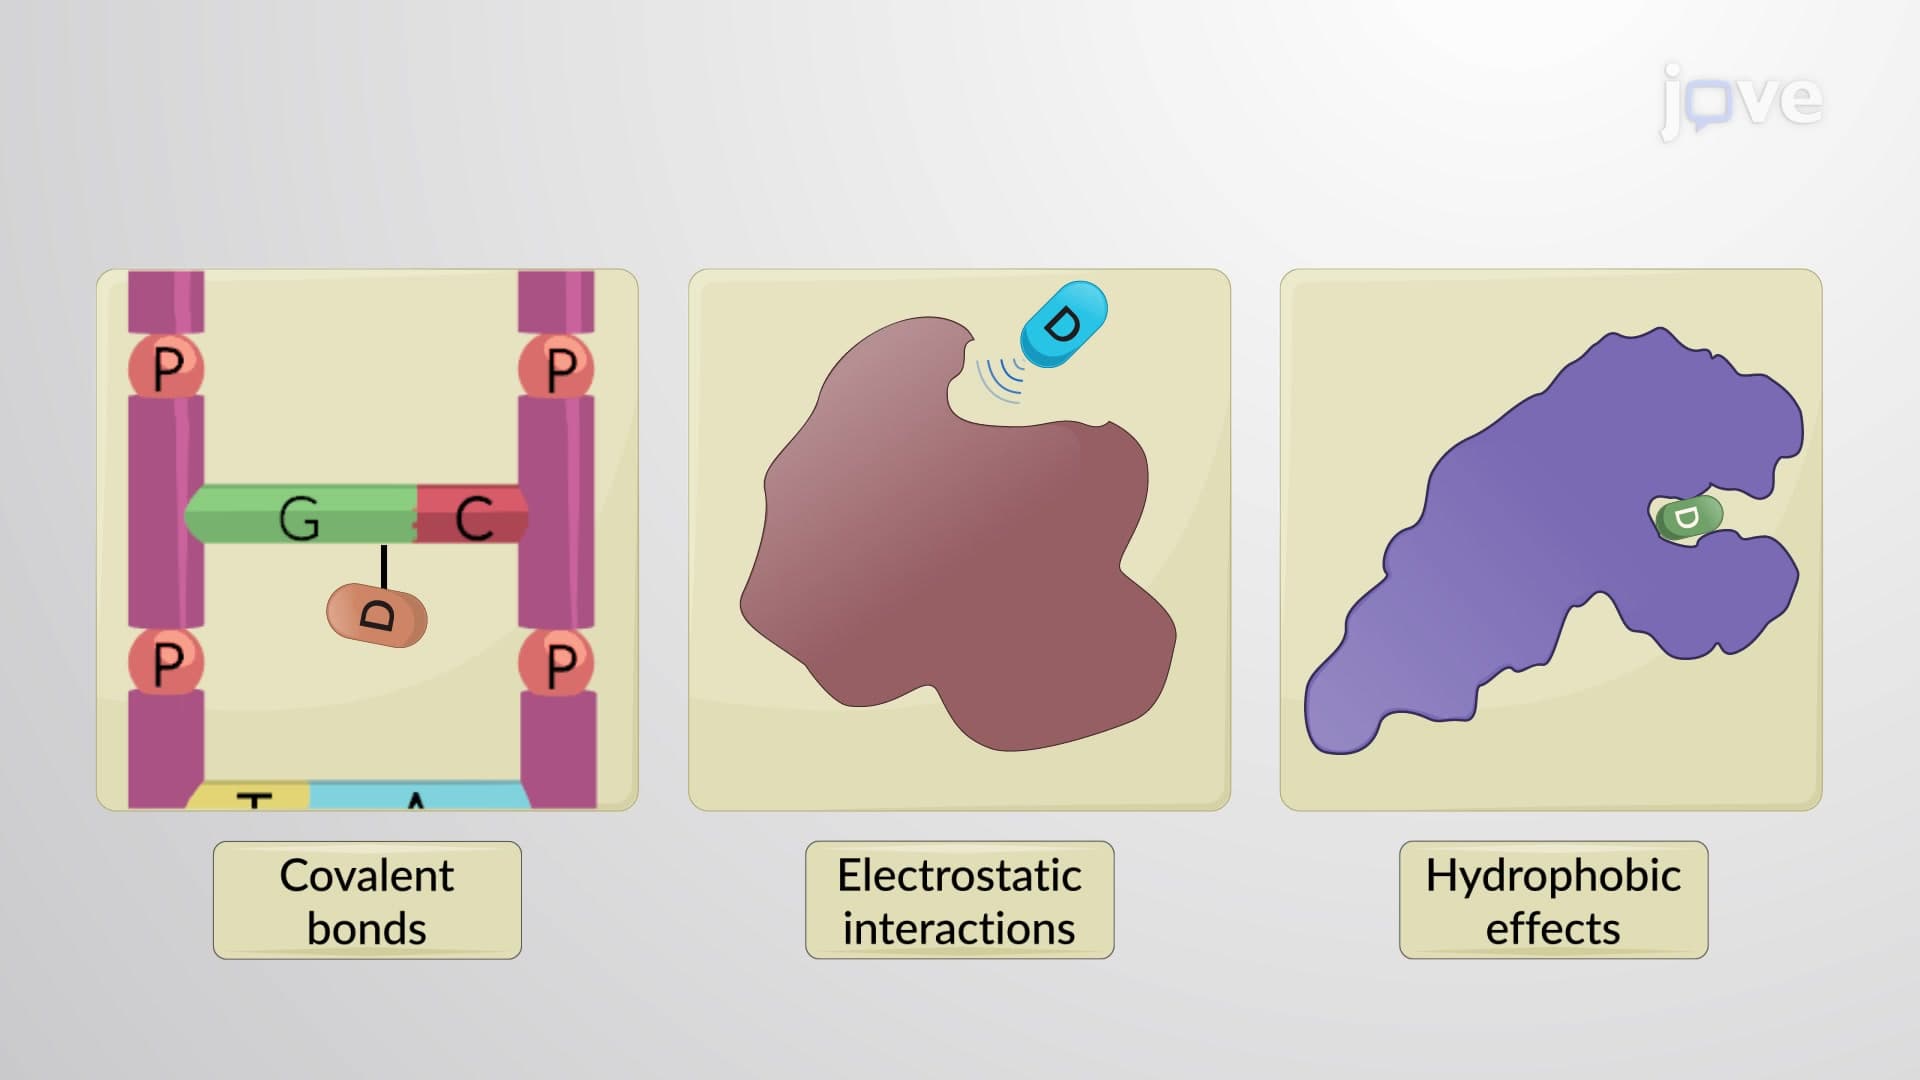
Drug-Receptor Bonds
In...
 01:29
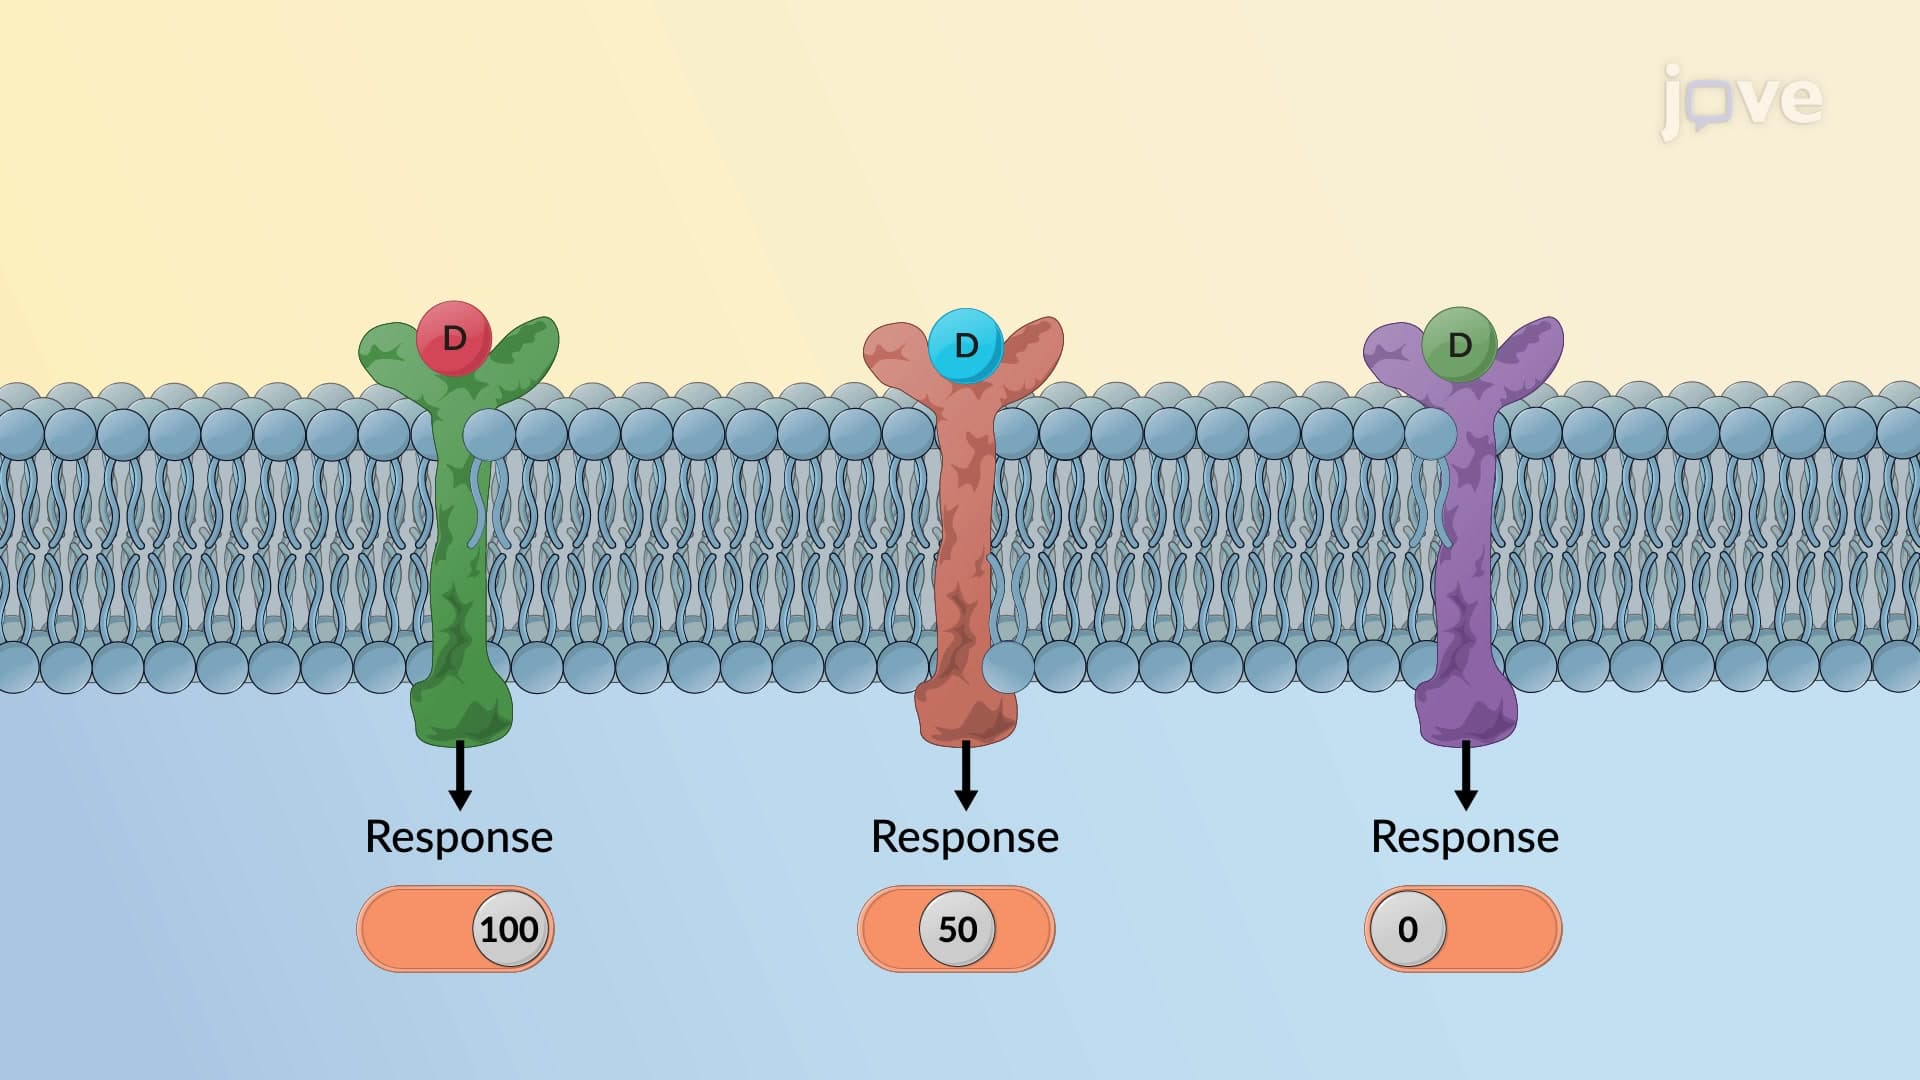
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
 01:26
Targets for Drug Action: Overview
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...
 01:16
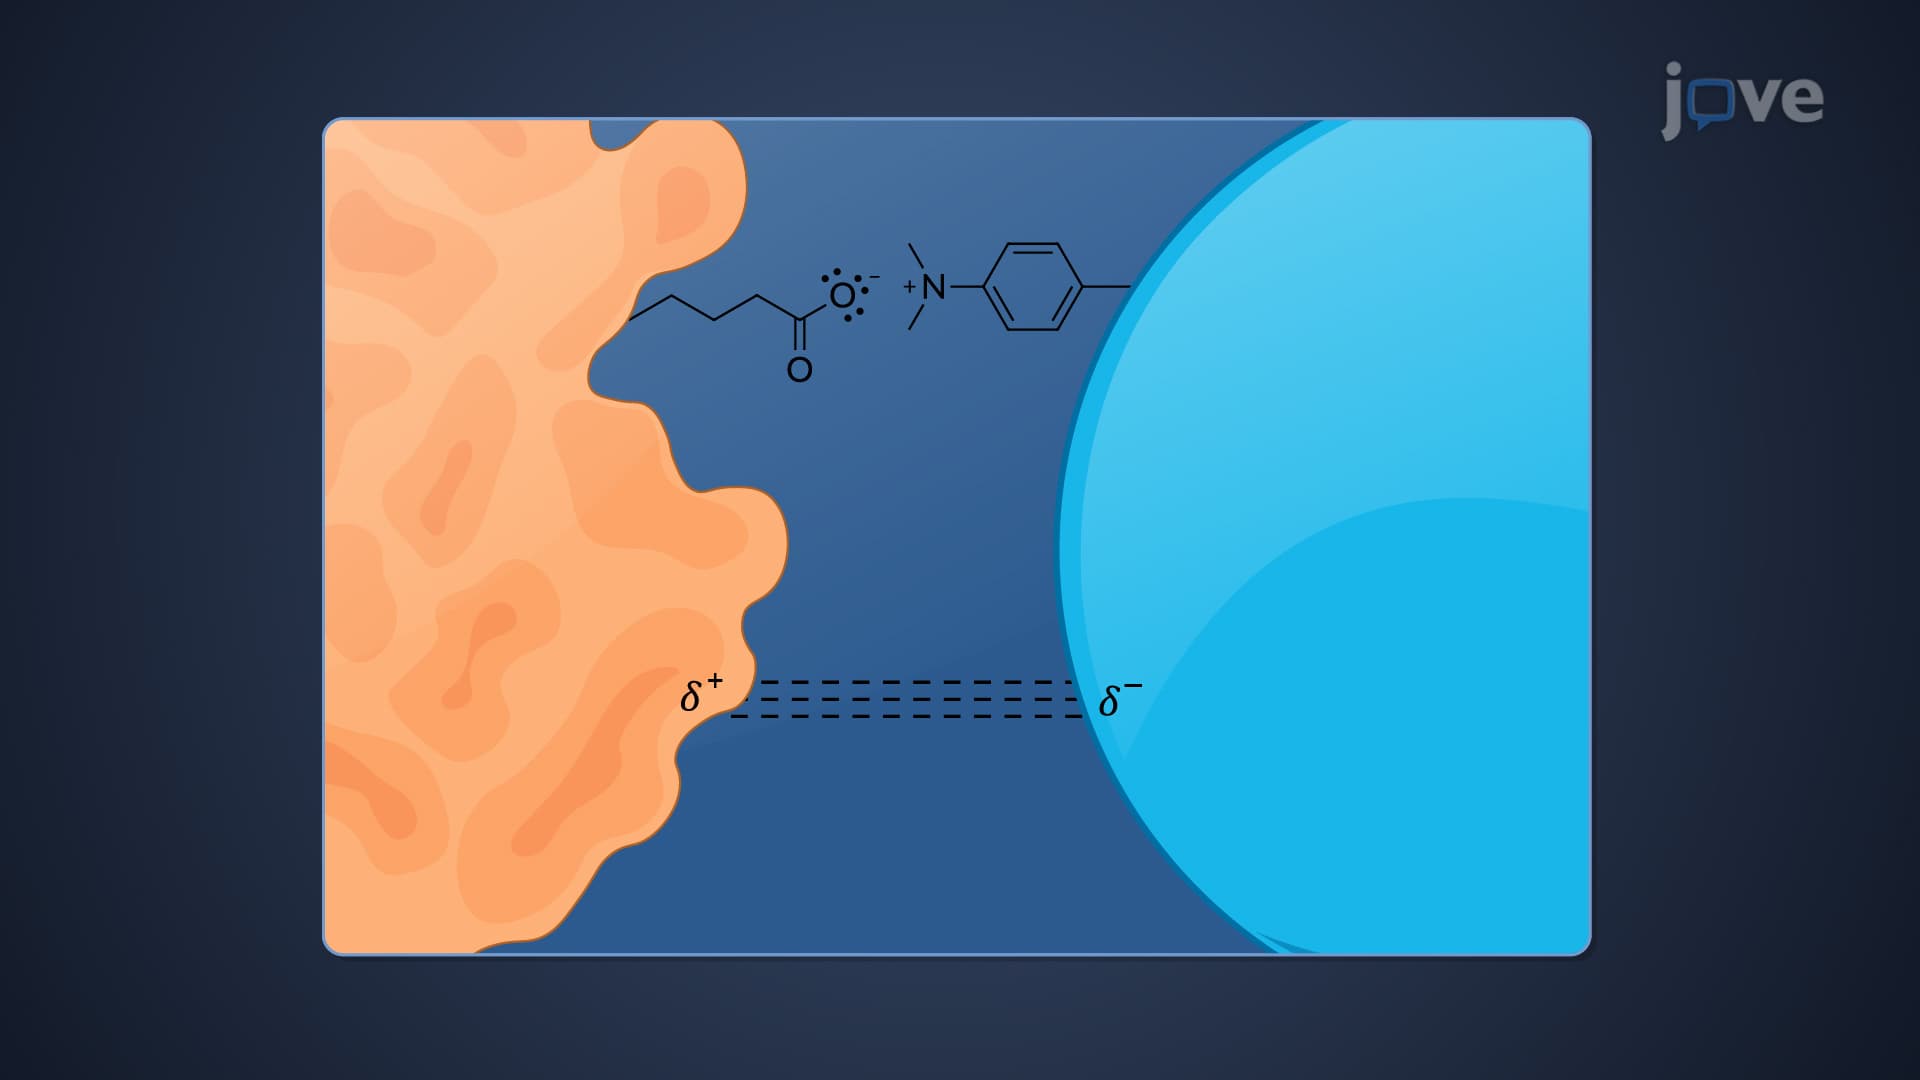
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...