Related Experiment Video
Updated: Jun 29, 2026

Biosynthesis of a Flavonol from a Flavanone by Establishing a One-pot Bienzymatic Cascade
Published on: August 14, 2019
Computational Insights into Flavonoids for ADAMTS-5 Exosite Inhibition in Knee Osteoarthritis: Docking, MD
Mayurakkhi Bhatia1, Nithyadevi Duraisamy1, Mohammed Cherkaoui1
1Department of Digital Engineering and Artificial Intelligence, College of Science, Long Island University, Brooklyn, NY 11201, USA.

Abstract:
The limited selectivity of most catalytic-site ADAMTS-5 inhibitors and the necessity to preserve aggrecan integrity in early-grade knee osteoarthritis require the development of selective aggrecanase inhibitors. The present study conducted rational in silico screening of flavonoids as potential ADAMTS-5 inhibitors by integrating high-throughput virtual screening, molecular docking, and molecular dynamic simulations targeting the exosite domains of ADAMTS-5 (the Disintegrin-like and spacer domain). The objective was to identify plant-derived flavonoids with favorable drug-like properties and specific interactions towards the ADAMTS-5 exosite as a more targeted alternative to catalytic-site inhibition. In this study, 847 flavonoids were screened using drug-likeness and ADME criteria to identify promising leads. The top 16 selected flavonoids were further subjected to molecular docking and SAR analysis. Of these compounds, Homoeriodictyol satisfied key drug-likeness criteria and exhibited the highest binding affinity to the Disintegrin-like domain, with a binding energy of -23.1 kcal/mol and favorable interactions. Molecular dynamics simulations of the Homoeriodictyol-ADAMTS-5 complex over 100 ns demonstrated stable binding throughout the trajectory. DCCM analysis and PCA further supported the proposed exosite-mediated modulation. To extend exosite mapping beyond the Disintegrin-like domain, this study also examined the spacer domain using a machine-learning-predicted structural model and identified key residues that contribute to ligand binding.
Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:16
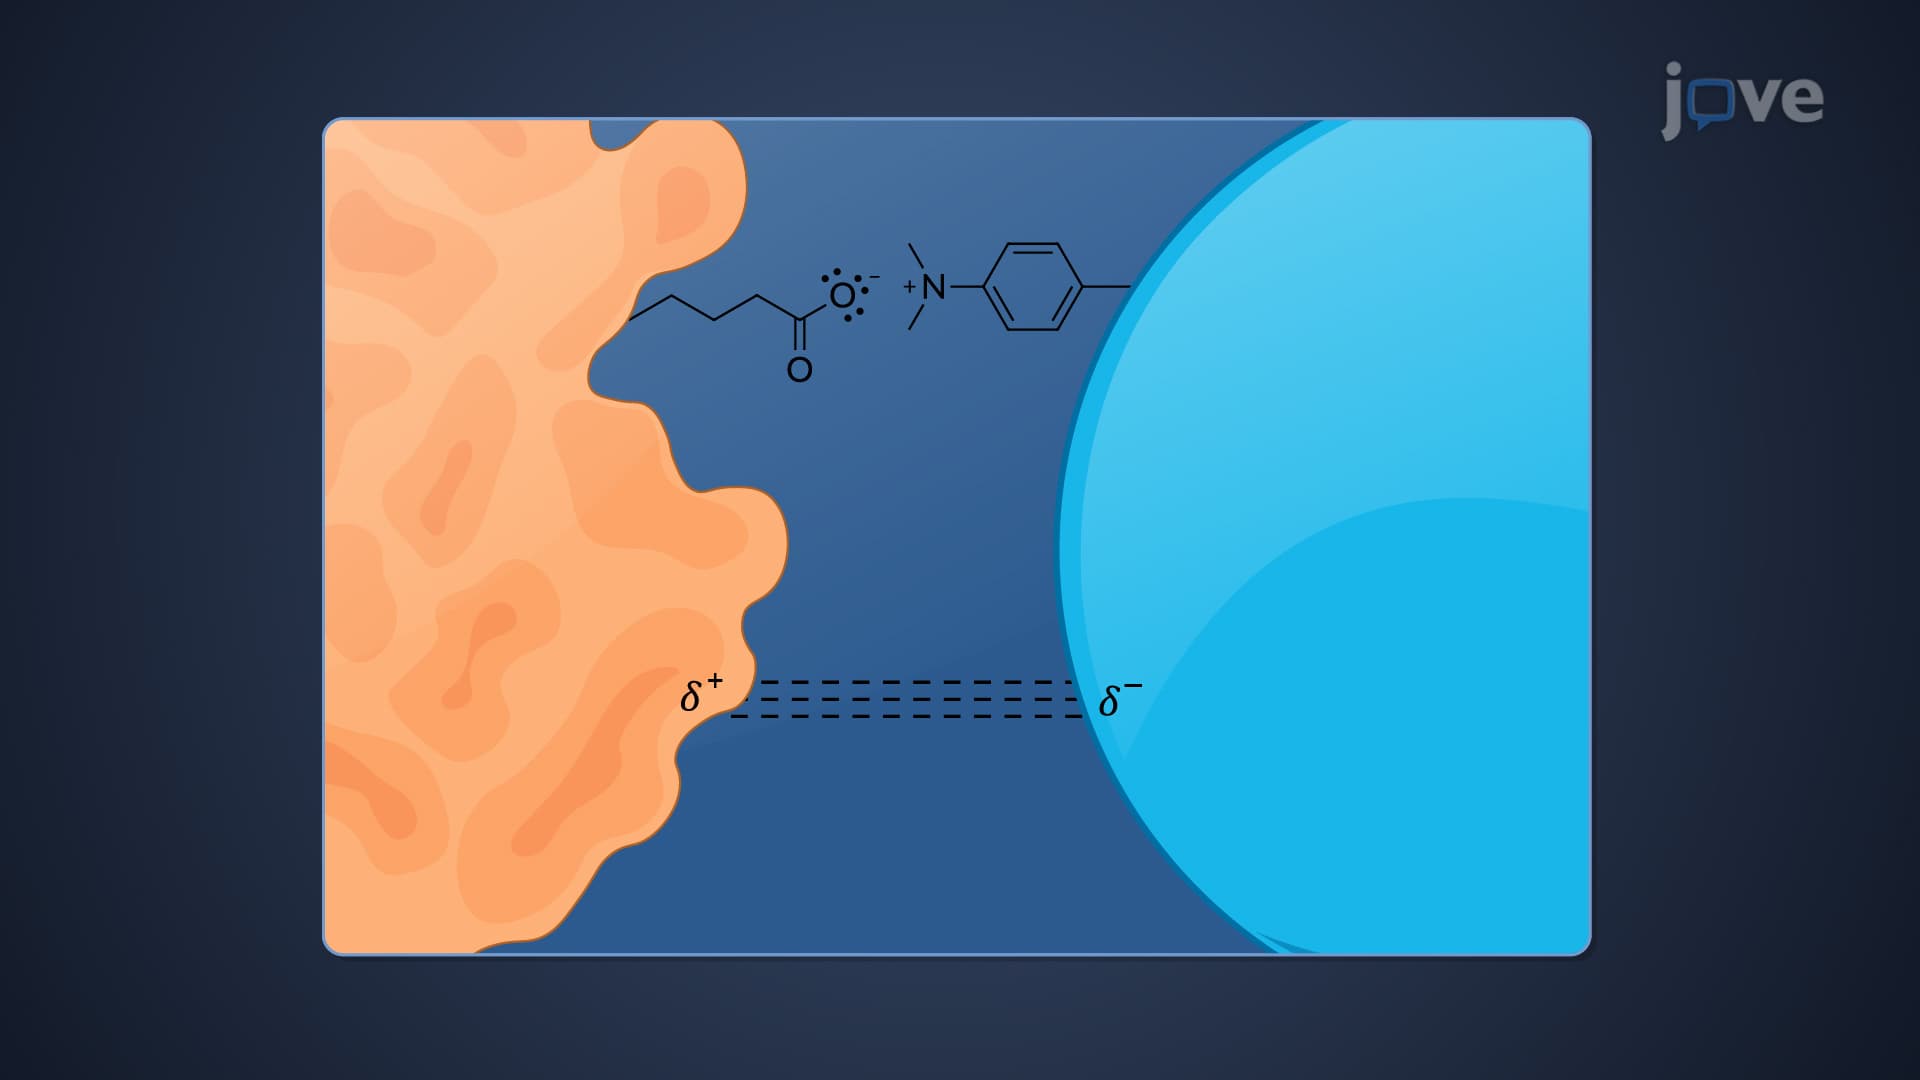
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...