Related Experiment Video
Updated: Apr 12, 2026

A Pipeline to Investigate the Structures and Signaling Pathways of Sphingosine 1-Phosphate Receptors
Published on: June 8, 2022
Structural insights into subtype-specific agonist recognition by sphingosine-1-phosphate receptors
Leiye Yu1,2, Haizhan Jiao3, Bin Pang1
1Shanghai Key Laboratory of Metabolic Remodeling and Health, Institute of Metabolism and Integrative Biology, Shanghai Xuhui Central Hospital, Zhongshan-Xuhui Hospital, Fudan University, Shanghai, China.

Abstract:
Sphingosine-1-phosphate (S1P), a key metabolite of sphingolipids, plays crucial roles in a wide range of physiological and pathological processes. S1P primarily exerts its functions by binding to G protein-coupled sphingosine-1-phosphate receptors (S1PRs), which comprise five subtypes (S1PR1-5) in humans, thereby activating these receptors and their downstream signaling pathways. Understanding the molecular determinants that govern agonist selectivity among different S1PR subtypes is vital for the rational and precise development of targeted therapeutic agents. Here, four cryo-electron microscopy structures of agonist-bound S1PR1-Gi1 complexes are reported. Through an integrated approach combining structural analysis, molecular dynamics simulations, and pharmacological assays, the molecular basis for the selectivity of CYM5442, HY-X-1011, Ponesimod, and SAR247799 toward S1PR1 over S1PR2-S1PR5 is uncovered. Nonconserved residues within the ligand-binding pocket and at the Gi1-protein interface contribute to S1PR1 selectivity by these agonists. A distinct agonist binding orientation toward transmembrane helices 5-7, combined with branched substituents that increase the agonist's molecular width, results in steric clashes with residues in S1PR3. Additionally, branched moieties located at the tail portions of the agonist restrict its deep insertion into the binding pocket of both S1PR3 and S1PR5. These structural features collectively enhance its selectivity for S1PR1 over S1PR3 and S1PR5. Furthermore, polar interactions with conserved polar residues in the top region of the binding pocket also influence agonist selectivity. Besides, the relatively broad molecular width of the agonist sterically hinders its binding into S1PR2 and S1PR4 pocket by nonconserved residue pairs bearing bulky side chains. These findings establish a structural framework for the rational design of next-generation S1PR1 highly selective agonists with improved therapeutic potential.
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:25
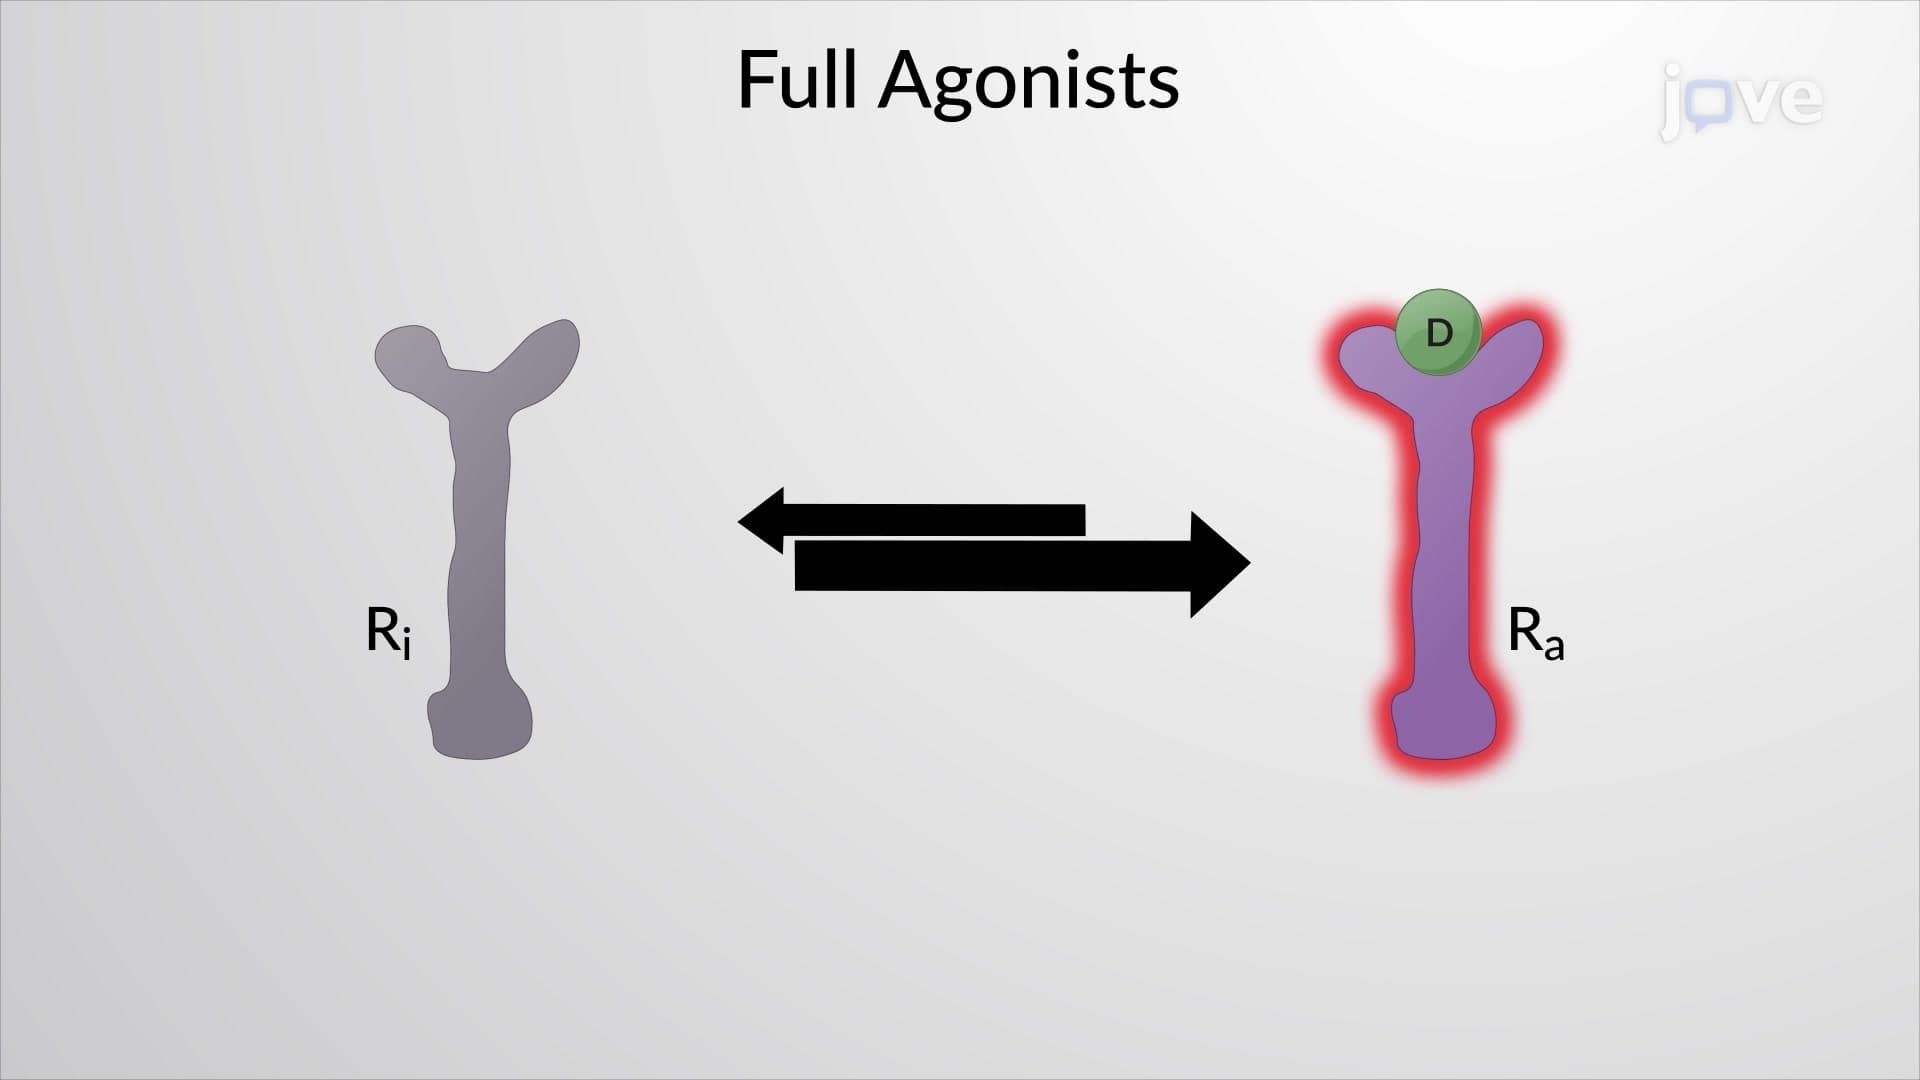
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous...
 01:31
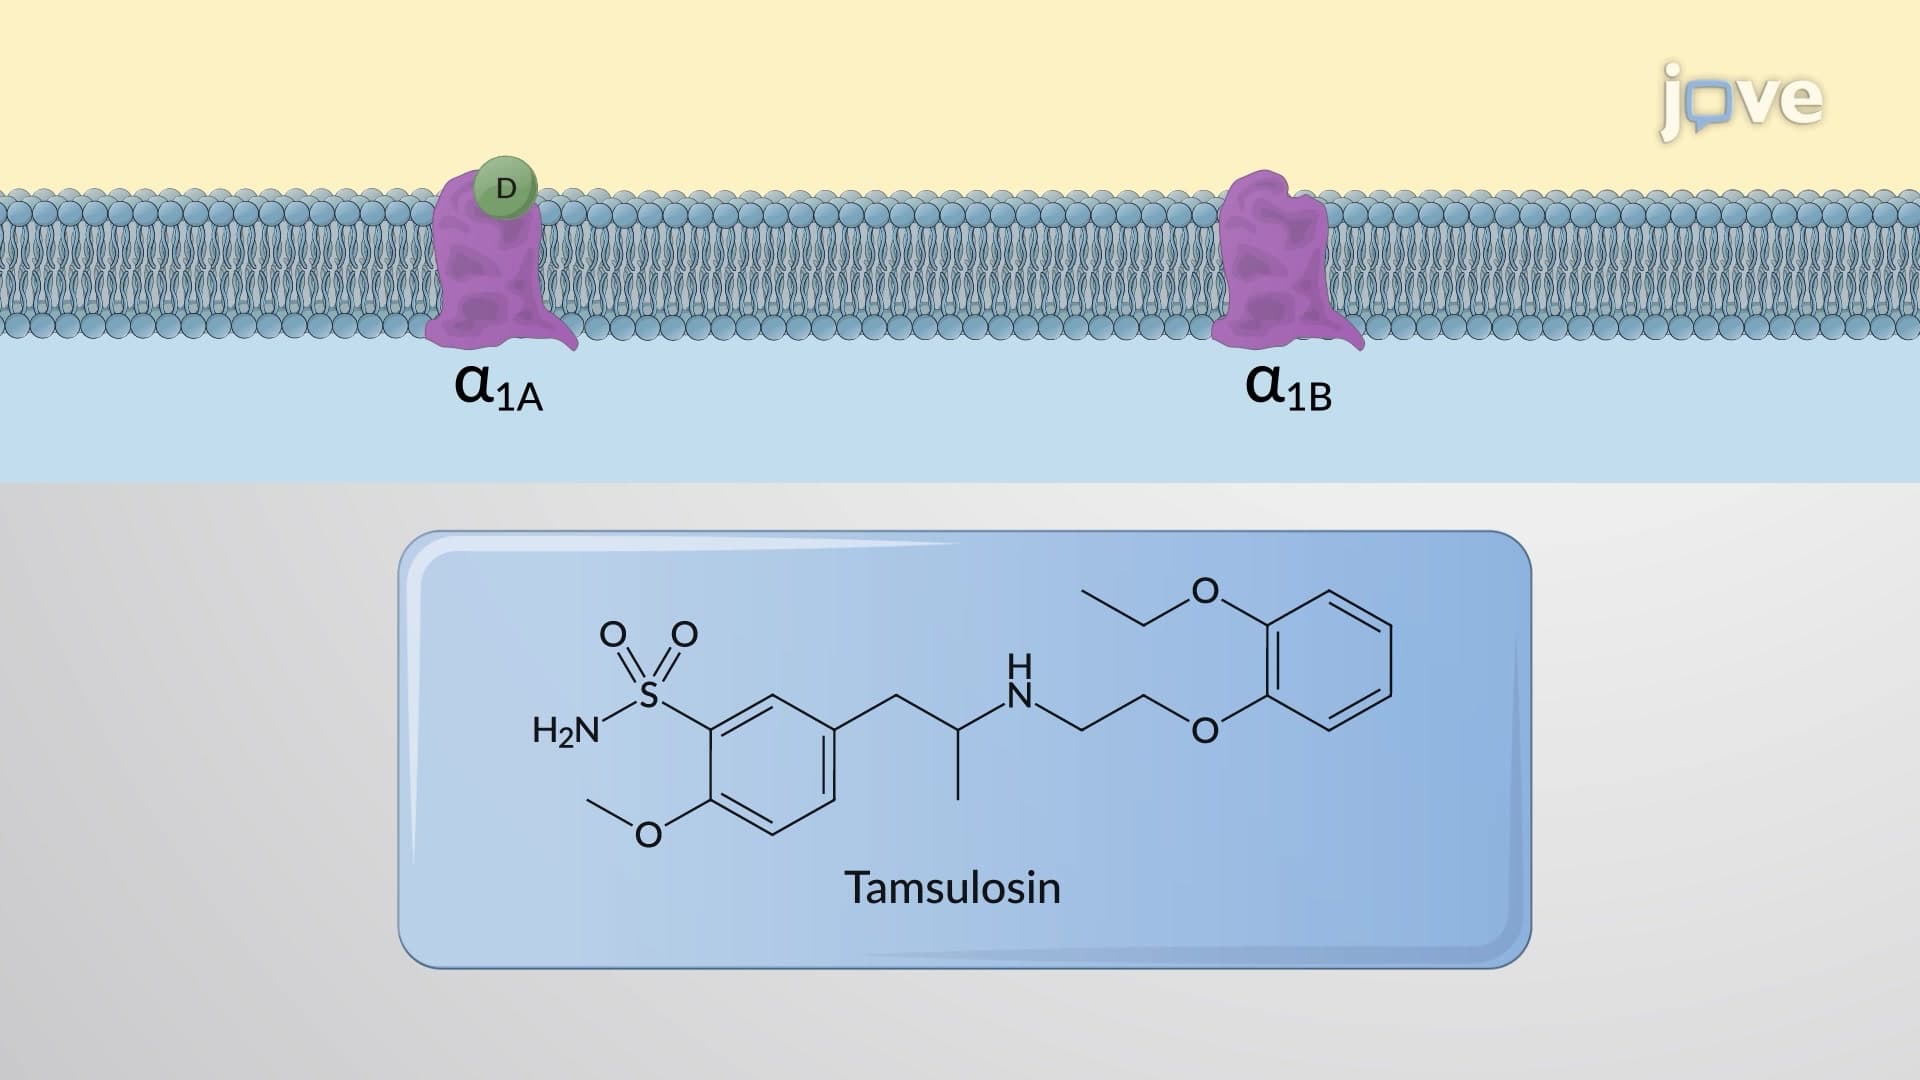
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase...
 01:22
Opioid Receptors: Overview
 01:15
G Protein-coupled Receptors
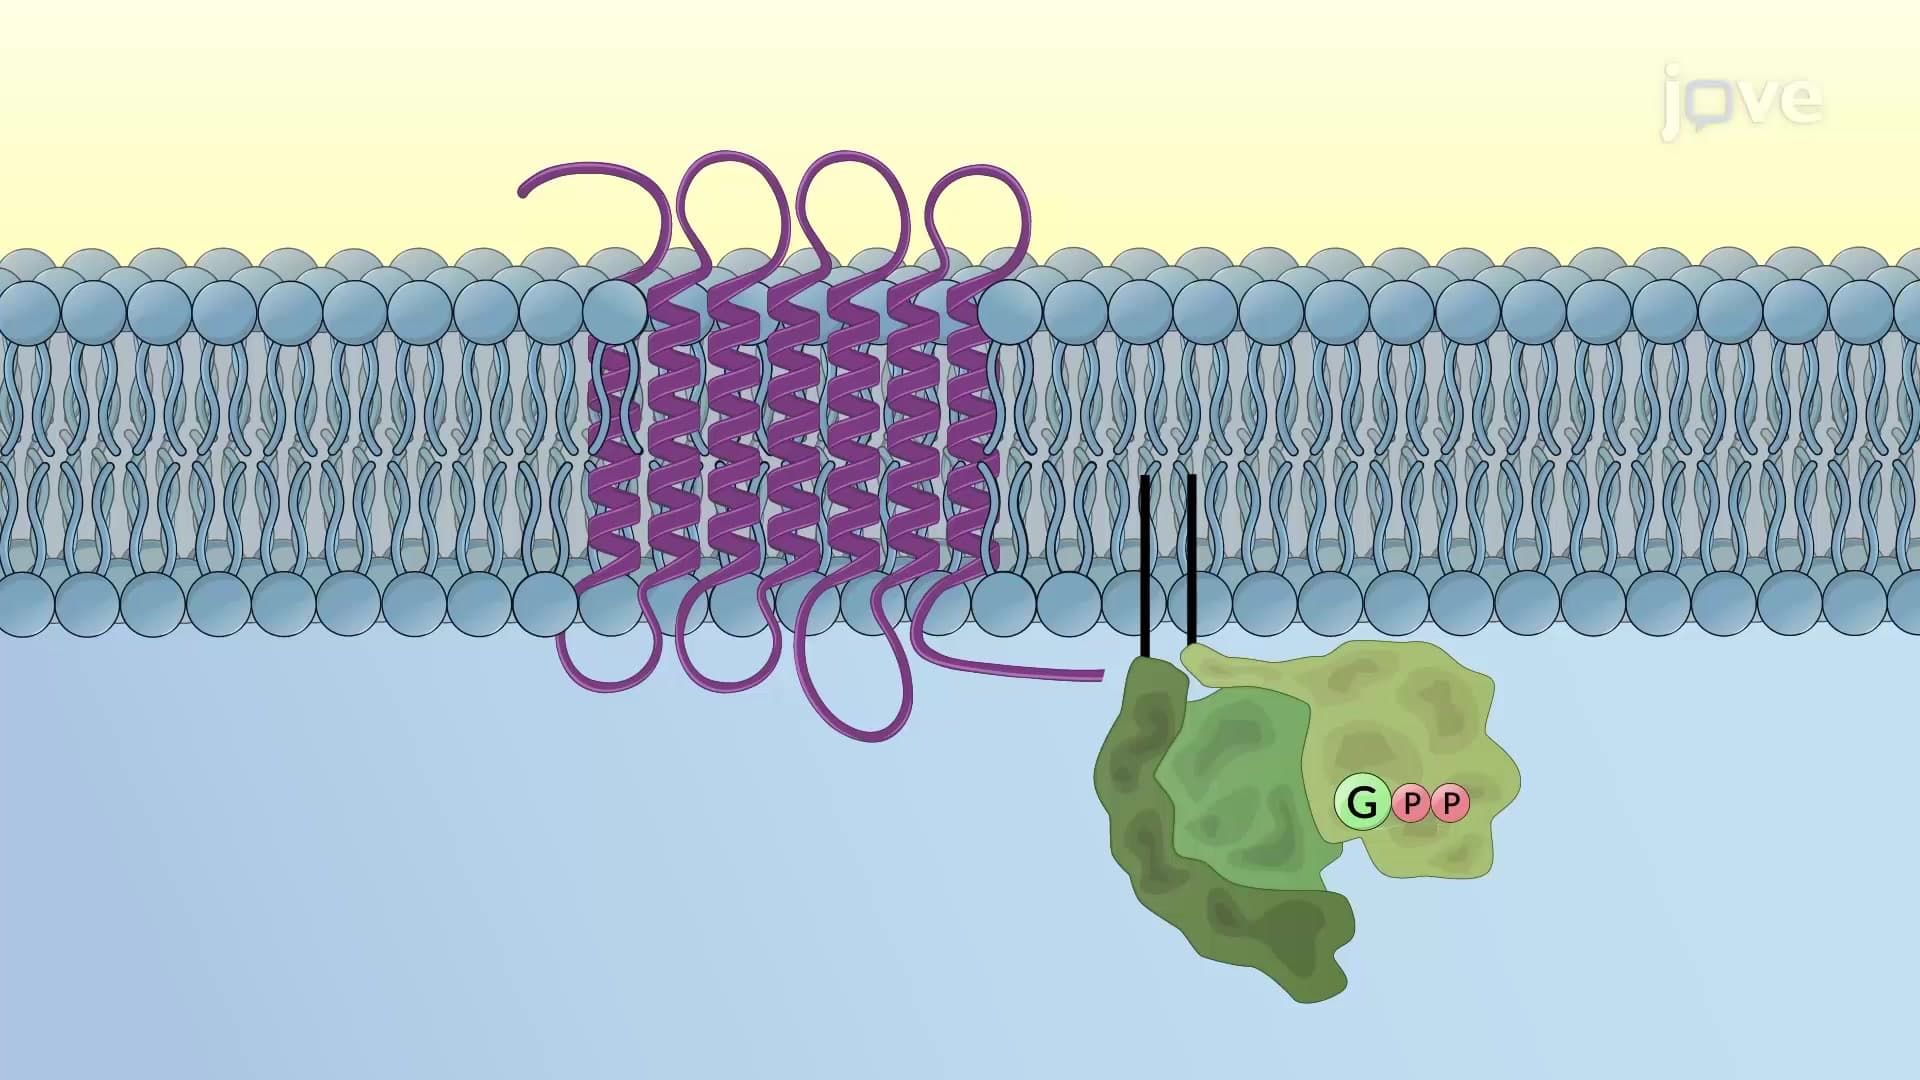
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...