Related Experiment Video
Updated: Apr 13, 2026
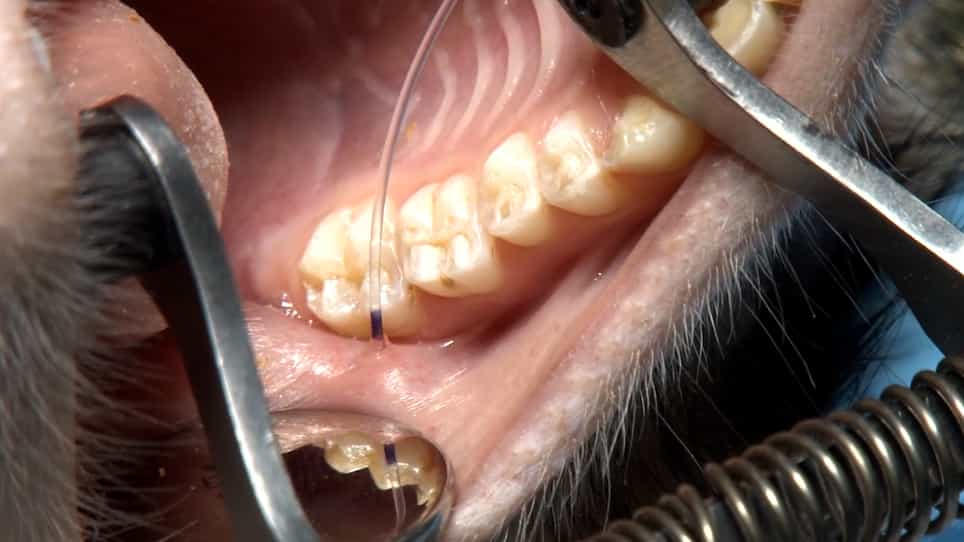

Retrograde Parotid Gland Infusion through Stensen's Duct in a Non-Human Primate for Vectored Gene Delivery
Published on: August 12, 2021
First-in-human intracisternal dosing of RGX-111 in severe MPS I is well tolerated and generates sustained
Raymond Y Wang1, Nina Movsesyan2, Shih-Hsin Kan2
1Division of Metabolic Disorders, Children's Hospital of Orange County Specialists, Orange, CA 92868, USA; Department of Pediatrics, University of California, Irvine, Orange, CA 92868, USA.

Mucopolysaccharidosis type I (MPS I), caused by α-L-iduronidase enzyme (IDUA) deficiency, manifests multisystemic symptoms from tissue accumulation of undegraded glycosaminoglycans. Severe MPS I (Hurler syndrome) is associated with developmental delay and loss of neurocognition. Although IDUA enzyme replacement therapy and allogeneic hematopoietic stem cell transplantation (HSCT) are standard therapies for Hurler syndrome, gene therapy represents potential alternative treatment of neurodevelopmental disease. We report more than 5 years of clinical, biochemical, and neurocognitive outcomes following a first-in-human, open-label, single-patient, single administration of intra-cisterna magna (ICM) RGX-111 (AAV9.CB7.hIDUA, 1E-10 vector genomes/g brain mass) in a 20-month-old boy with Hurler syndrome with two older affected siblings deceased from HSCT-related complications. The ICM procedure was safe and well tolerated; treatment-emergent adverse events were mild and self-limiting. Crucially, serial brain imaging has been normal. He has never received an HSCT and continues with weekly IDUA infusions. Neurocognitive testing demonstrates ongoing acquisition of developmental abilities, with cognitive, speech, and motor age equivalents measuring one to two standard deviations below the normative mean. His neurodevelopment is significantly above the natural history of untransplanted Hurler syndrome patients. This single-patient experience of central nervous system-directed gene therapy demonstrates therapeutic potential for severe MPS I and other genetic neurodegenerative diseases.
Mucopolysaccharidosis type I (MPS I), caused by α-L-iduronidase enzyme (IDUA) deficiency, manifests multisystemic symptoms from tissue accumulation of undegraded glycosaminoglycans. Severe MPS I (Hurler syndrome) is associated with developmental delay and loss of neurocognition. Although IDUA enzyme replacement therapy and allogeneic hematopoietic stem cell transplantation (HSCT) are standard therapies for Hurler syndrome, gene therapy represents potential alternative treatment of neurodevelopmental disease. We report more than 5 years of clinical, biochemical, and neurocognitive outcomes following a first-in-human, open-label, single-patient, single administration of intra-cisterna magna (ICM) RGX-111 (AAV9.CB7.hIDUA, 1E-10 vector genomes/g brain mass) in a 20-month-old boy with Hurler syndrome with two older affected siblings deceased from HSCT-related complications. The ICM procedure was safe and well tolerated; treatment-emergent adverse events were mild and self-limiting. Crucially, serial brain imaging has been normal. He has never received an HSCT and continues with weekly IDUA infusions. Neurocognitive testing demonstrates ongoing acquisition of developmental abilities, with cognitive, speech, and motor age equivalents measuring one to two standard deviations below the normative mean. His neurodevelopment is significantly above the natural history of untransplanted Hurler syndrome patients. This single-patient experience of central nervous system-directed gene therapy demonstrates therapeutic potential for severe MPS I and other genetic neurodegenerative diseases.