Related Experiment Video
Updated: Apr 16, 2026
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Multireference perturbation theory reveals the intricate excited-state electronic structure of the 2-phenylpyridine
Mario Taddei1, Davide Accomasso1, Giovanni Li Manni2
1Dipartimento di Chimica Industriale "Toso Montanari," University of Bologna, via Gobetti 85, 40129 Bologna, Italy.

Abstract:
2-Phenylpyridine (2-ppy) is a widely used cyclometalating ligand in transition-metal complexes (TMCs), yet its intrinsic excited-state electronic structure remains poorly characterized. Here, we investigate its singlet excited states of ππ* nature accessible in the ultraviolet spectral window, both as an isolated molecule and when coordinated in the TMC Ir(2-ppy)3, using extended multi-state restricted active-space second-order perturbation theory (XMS-RASPT2). Our study reveals multiple low-lying excited states, whose relative energy ordering and oscillator strengths are strongly dependent on the choice of the orbital active space. The XMS-RASPT2 calculations accurately reproduce the experimental absorption spectrum of the isolated 2-ppy and show that the characteristic two-band profile arises from mixing between the same bright configuration and dark transitions. Coordination to the iridium in Ir(2-ppy)3 induces an overall redshift of the ligand-centered transitions accompanied by a change in their energy ordering. In contrast, inter-ligand couplings, evaluated using a Frenkel exciton model, are found to have an almost negligible impact on the electronic structure of the 2-ppy manifold.
Related Concept Videos
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:17
Electron Paramagnetic Resonance (EPR) Spectroscopy: Organic Radicals
 01:37
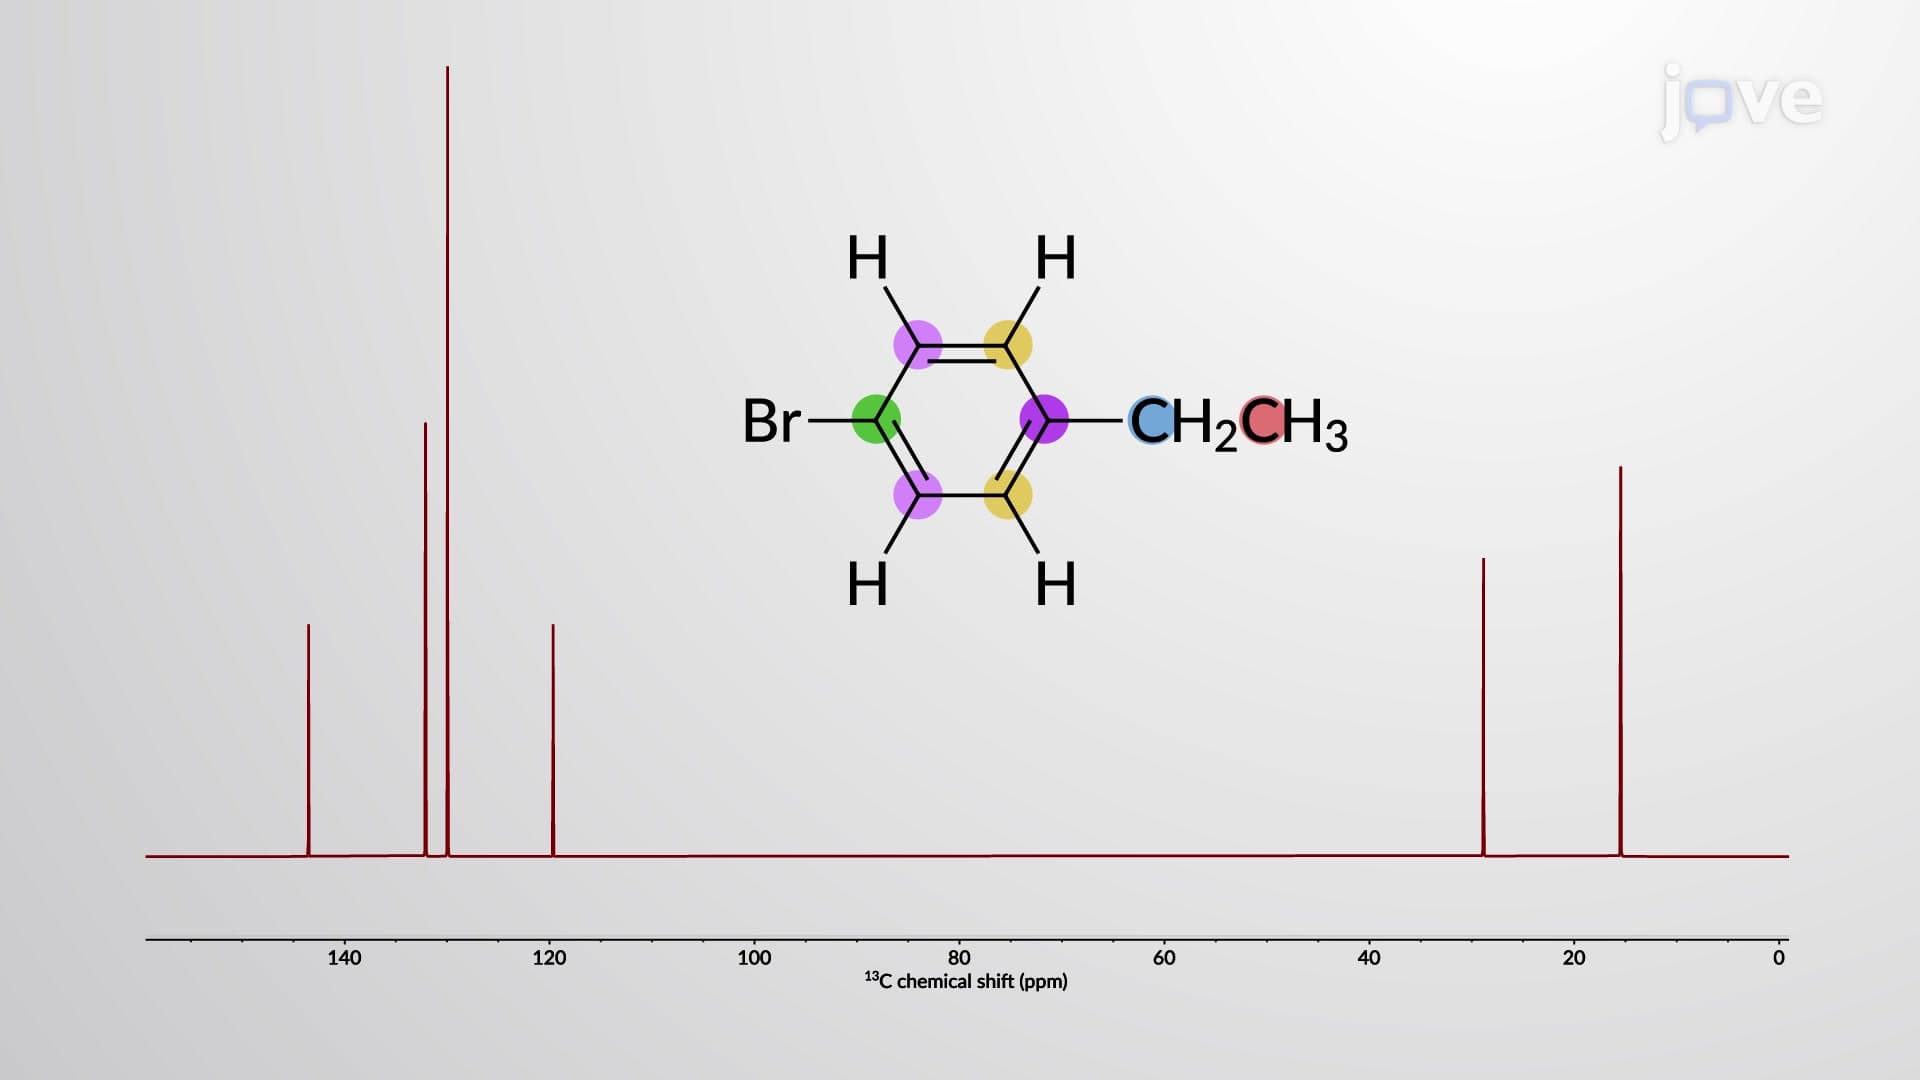
NMR Spectroscopy of Benzene Derivatives
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 01:18
Structure of Benzene: Molecular Orbital Model
 01:25
Basicity of Heterocyclic Aromatic Amines