Related Experiment Video
Updated: Apr 18, 2026


In Vivo Functional Study of Disease-associated Rare Human Variants Using Drosophila
Published on: August 20, 2019
Functional inactivation of MDR3 caused by a homozygous ABCB4 missense variant leading to liver failure
Sophia Heinrich1,2, Annika Behrendt3, Malte Sgodda1,4
1Department of Gastroenterology, Hepatology, Infectious Diseases and Endocrinology, Hannover Medical School, Hannover, Germany.

Abstract:
Progressive familial intrahepatic cholestasis (PFIC) is a rare hereditary liver disorder that is caused by defective hepatobiliary transport. Variants in ATP binding cassette 4 ( ), encoding phosphatidylcholine floppase MDR3, are a frequent cause; however, many remain classified as variants of uncertain significance (VUS), limiting molecular diagnosis. Here, we functionally characterized a previously reported homozygous ABCB4 missense variant (c.431G>A, p.(Arg144Gln)) without experimental evidence of pathogenicity. An in silico analysis using the ABCB4-specific prediction tool Vasor indicated a high probability of pathogenicity (0.88). Structural modeling suggested that Arg144Gln disrupted key electrostatic interactions essential for MDR3 membrane anchoring. Immunofluorescence analyses demonstrated markedly reduced membrane localization with residual cytoplasmic retention, consistent with complete loss of protein function. In conclusion, the ABCB4 p.(Arg144Gln) variant causes functional inactivation of MDR3 and represents a novel pathogenic mutation. Combined genetic, structural, and functional analyses are valuable tools for characterizing variants of uncertain significance in ABCB4-associated cholestatic liver disease.
Insights
The ABCB4 p.(Arg144Gln) variant causes progressive familial intrahepatic cholestasis (PFIC) by inactivating the MDR3 protein. This study clarifies a variant of uncertain significance, aiding molecular diagnosis of cholestatic liver disease.
Area of Science:
- Hepatology
- Genetics
- Molecular Biology
Background:
- Progressive familial intrahepatic cholestasis (PFIC) is a rare hereditary liver disorder.
- Defective hepatobiliary transport, often due to ATP binding cassette 4 (ABCB4) variants, causes PFIC.
- Many ABCB4 variants are of uncertain significance (VUS), hindering diagnosis.
Purpose of the Study:
- To functionally characterize the homozygous ABCB4 missense variant c.431G>A (p.(Arg144Gln)).
- To determine the pathogenicity of the p.(Arg144Gln) variant in ABCB4-associated cholestatic liver disease.
Main Methods:
- In silico analysis using the Vasor prediction tool.
- Structural modeling of the MDR3 protein.
- Immunofluorescence analyses to assess protein localization and function.
Main Results:
- In silico analysis predicted a high probability of pathogenicity (0.88).
- Structural modeling indicated disruption of electrostatic interactions essential for MDR3 membrane anchoring.
- Immunofluorescence showed reduced membrane localization and cytoplasmic retention, indicating loss of function.
Conclusions:
- The ABCB4 p.(Arg144Gln) variant causes functional inactivation of MDR3, representing a novel pathogenic mutation.
- Combined genetic, structural, and functional analyses are crucial for characterizing VUS in ABCB4-associated cholestatic liver disease.
Related Concept Videos
 01:16
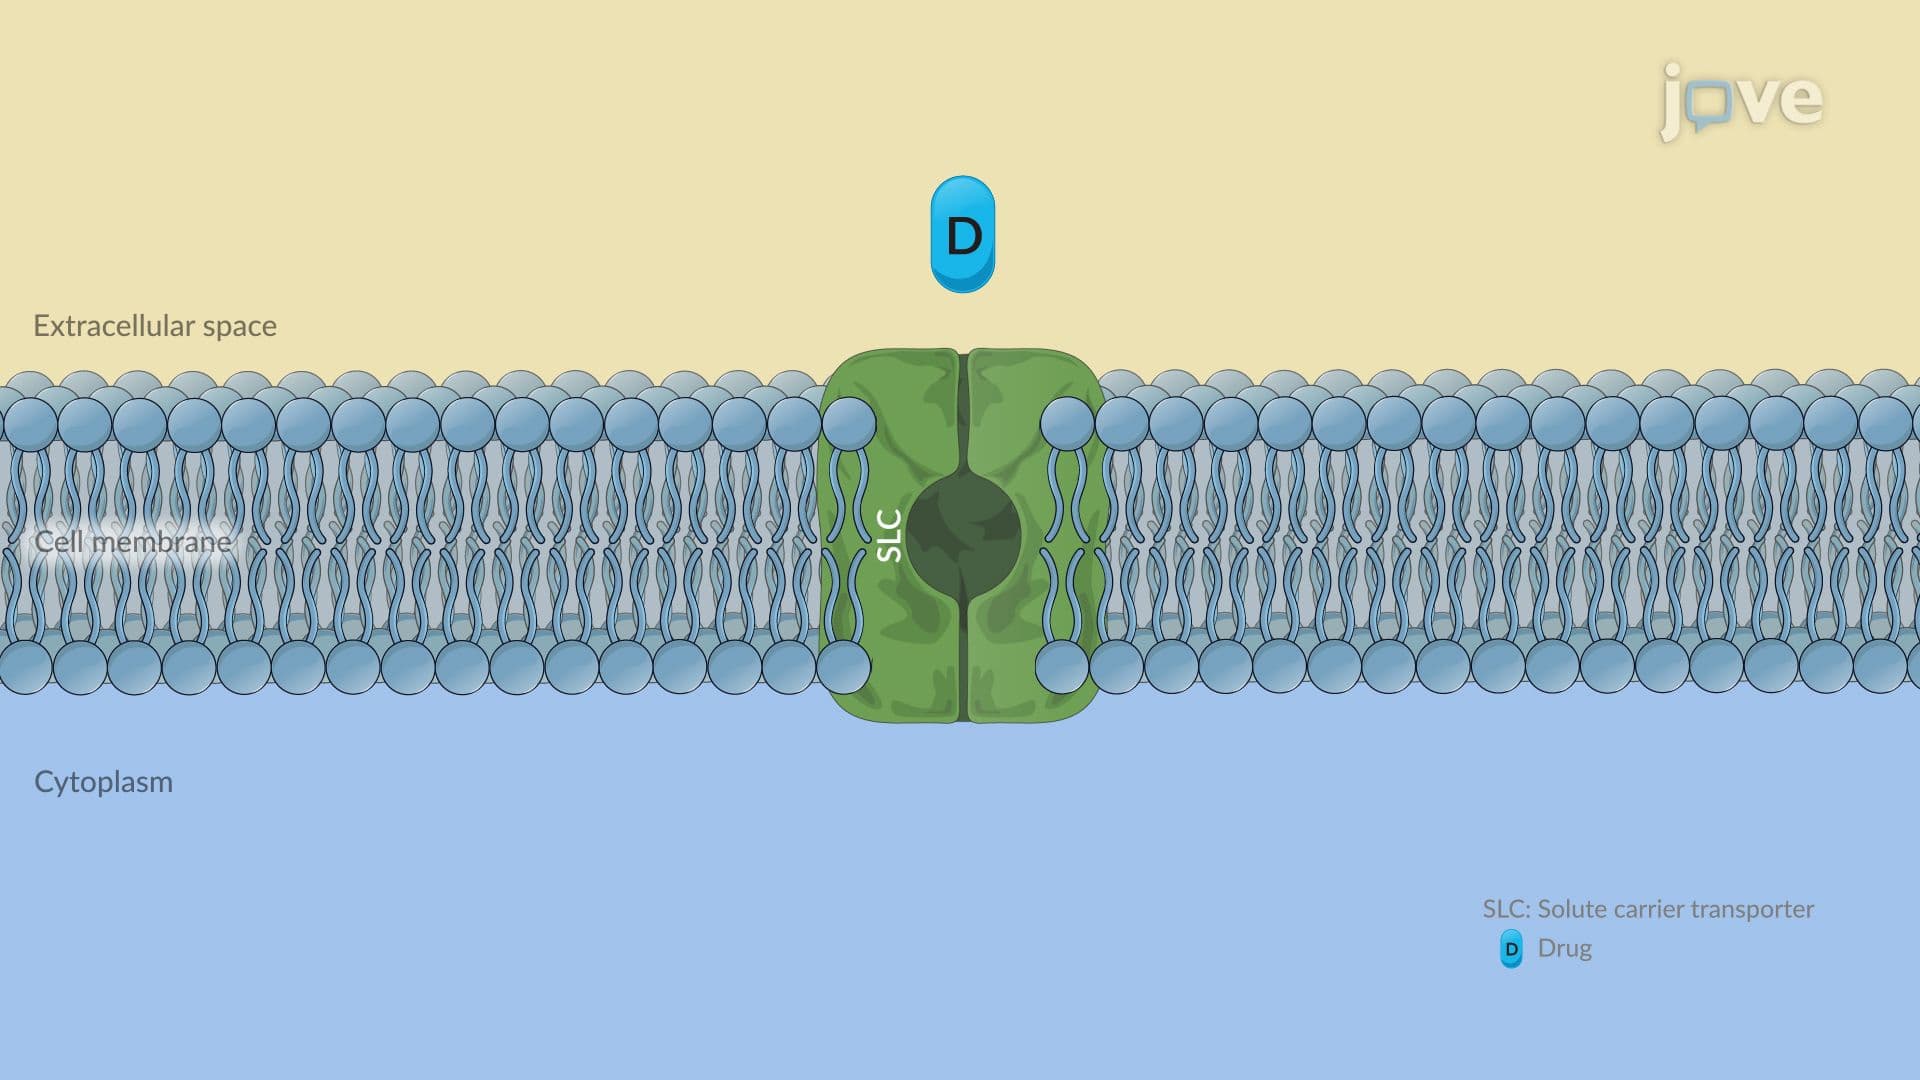
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:31
ABC Transporters: Exporter
 01:26
Effect of Hepatic Disease on Pharmacokinetics: Drug Dosing and Hepatic Blood Flow
 01:22
Effect of Hepatic Disease on Pharmacokinetics: Pathophysiologic Assessment and Liver Function Test
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:29
Pharmacogenetic Phenotypes: Alterations in Pharmacokinetics, Drug Targets and Biologic Milieu