Related Experiment Video
Updated: May 1, 2026
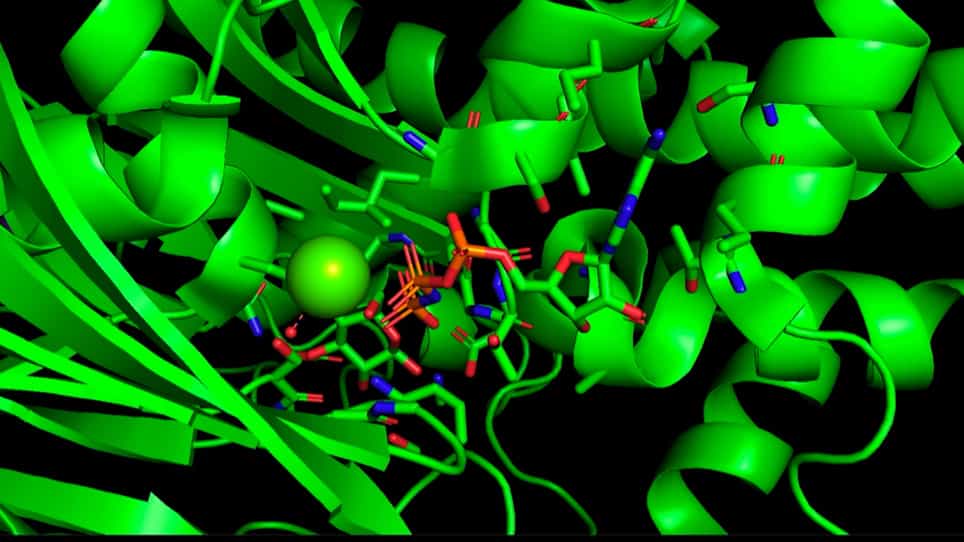

Modeling an Enzyme Active Site using Molecular Visualization Freeware
Published on: December 25, 2021
Structure-Based Discovery of Glyoxalase-I Inhibitors: Evaluating the Role of Active Site Water Molecules
Reema Anaam1, Israa H Isawi1, Qosay Al-Balas1
1Department of Medicinal Chemistry and Pharmacognosy, Faculty of Pharmacy, Jordan University of Science and Technology, Irbid, Jordan.

Background:
Glyoxalase-I (GLO-I) is a zinc-dependent metalloenzyme and a promising target for anticancer drug discovery. It catalyzes the detoxification of methylglyoxal, a cytotoxic byproduct of glycolysis, a metabolic shift commonly observed in cancer cells. GLO-I overexpression in tumor cells promotes multidrug resistance and tumor progression. However, the role of conserved water molecules within the GLO-I active site remains insufficiently explored, and understanding their influence on ligand binding may improve structure-based inhibitor design.
Purpose:
This study aimed to identify potential GLO-I inhibitors by examining the effect of conserved active site water molecules on ligand binding and activity predictions using a structure-based drug design approach.
Materials And Methods:
Three human GLO-I crystal structures were used to generate structure-based pharmacophore models under two conditions: with and without crystallographic active site water molecules. The models were applied to virtually screen the OTAVA Lead-Like library (commercial lead-like compound library). Molecular docking of the resulting hits was then performed under both hydration conditions to evaluate effects on ligand binding affinity and pose orientation. Top-ranked compounds were purchased and evaluated in vitro for GLO-I inhibition. The most active hit was further evaluated by 1000 ns molecular dynamics (MD) simulations (± crystallographic waters), including analysis of pose stability and binding-site water behavior.
Results:
Among the 22 compounds tested in vitro, five showed inhibitory activity, with IC50 values ranging from 12.07 to 25.36 μM. The most potent compound (hit 19) demonstrated an IC50 of 12.07 ± 0.31 μM and 85.63% inhibition at 50 μM. Docking analysis indicated that including crystallographic water molecules often increased docking scores but could distort binding orientations, whereas docking without conserved active site water molecules more consistently produced plausible poses and better agreement with experimental activity trends.
Conclusion:
For GLO-I, docking without conserved active site water molecules provided more accurate results and may represent a more reliable approach for studying ligand binding and guiding inhibitor design.
More Related Videos
Related Concept Videos
 01:30
Enzyme Inhibition
 01:20
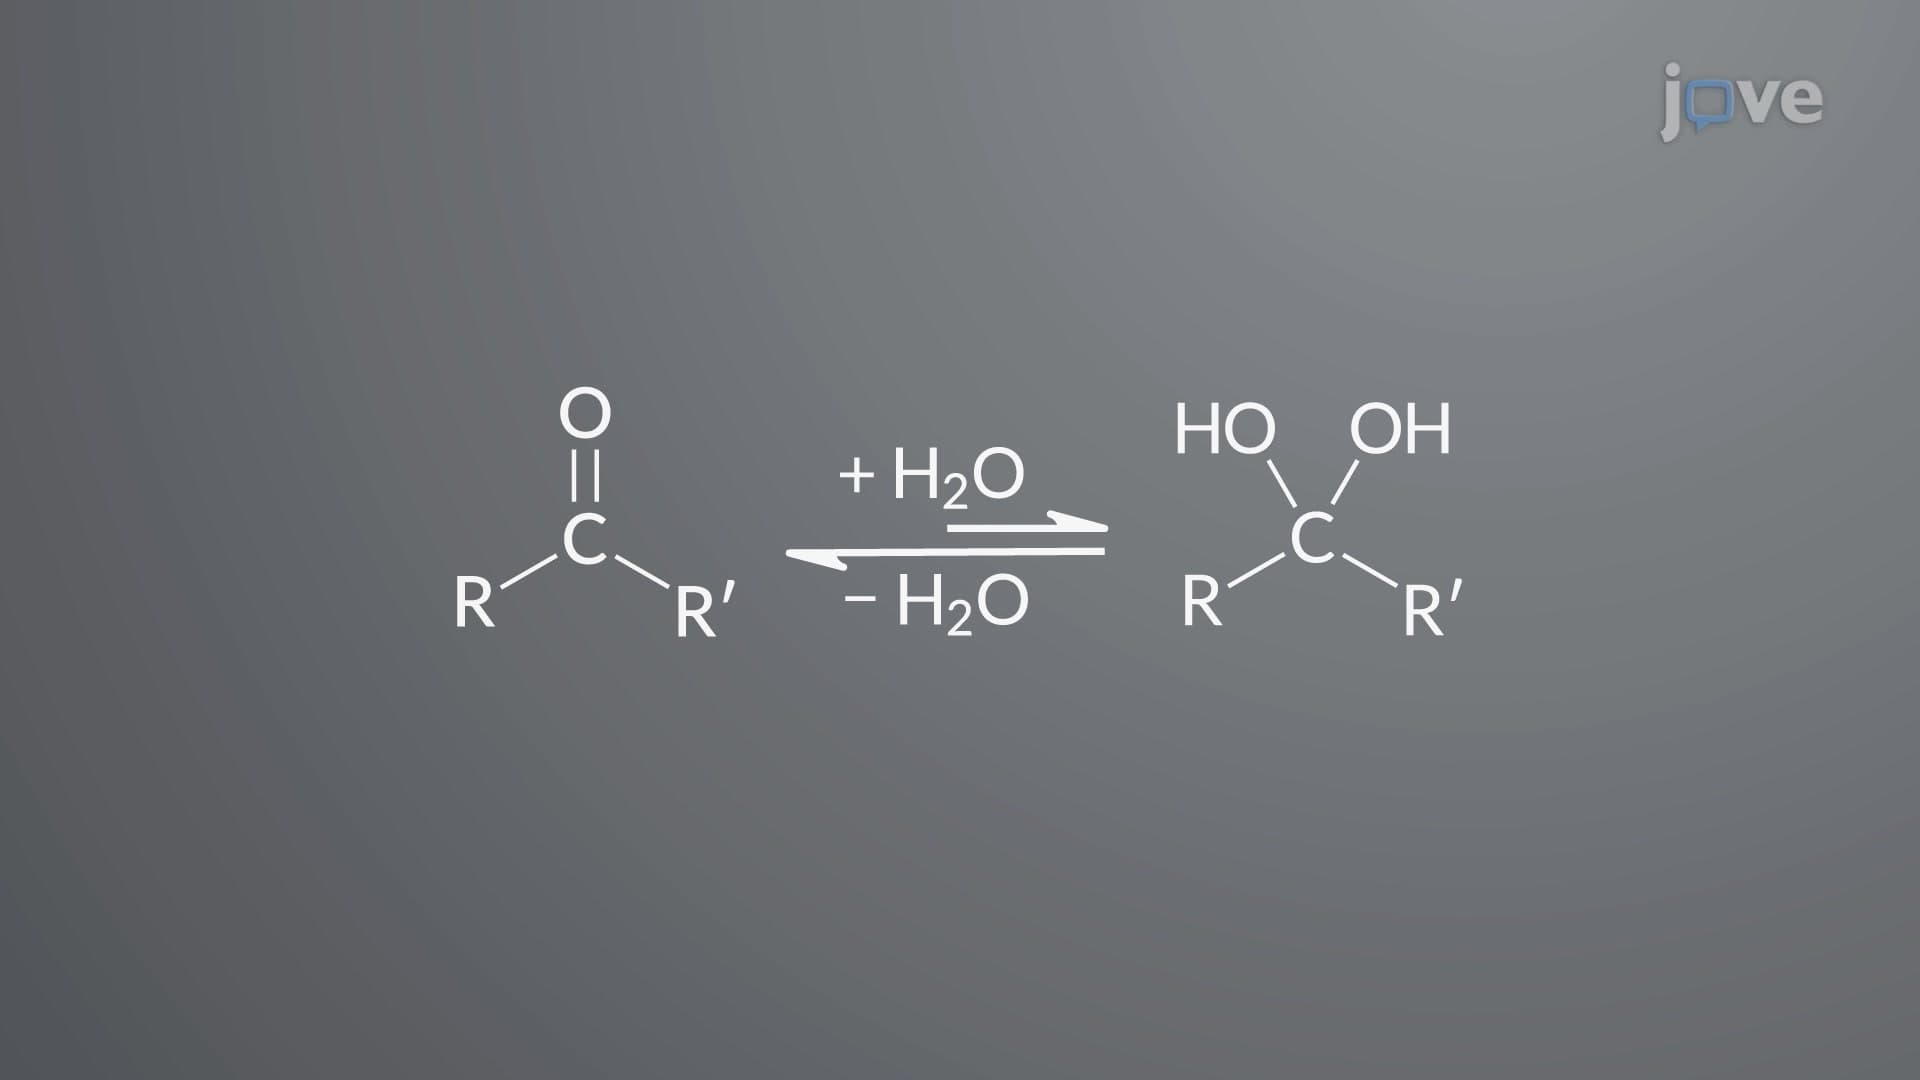
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 02:34
Enzymes
Enzyme deficiencies can often translate into life-threatening diseases. For example, a genetic abnormality resulting in the deficiency of the enzyme G6PD...
 00:49
Ligand Binding and Linkage
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...