Related Experiment Video
Updated: May 6, 2026


Fingerprinting Cardiolipin in Leukocytes by Mass Spectrometry for a Rapid Diagnosis of Barth Syndrome
Published on: March 23, 2022
β-ureidopropionase deficiency mimicking Leigh syndrome associated with methylmalonic aciduria
Giulia Ferrera1,2, Sara Boenzi3, Eleonora Lamantea4
1Child Neurology Unit, Fondazione IRCCS Istituto Neurologico Carlo Besta, Milan, Lombardy, Italy giulia.ferrera@unimi.it.

Abstract:
β-ureidopropionase (βUP) deficiency, caused by pathogenic variants in UPB1, is a rare autosomal recessive disorder with fewer than 60 reported cases to date. Clinical presentation is highly variable, ranging from asymptomatic individuals to severe neurodevelopmental disorders. Diagnosis relies on the detection of pyrimidine degradation metabolites, such as N-carbamyl-β-alanine (NCβA) and N-carbamyl-β-aminoisobutyrate (NCβAIBA), which are not widely available in routine clinical practice.We report a 7-year-old boy with early-onset developmental delay and regression, Leigh-like basal ganglia lesions on brain MRI, and persistent elevation of urinary methylmalonic acid (MMA). Muscle biopsy revealed an isolated respiratory chain complex I deficiency. Genetic testing identified two novel biallelic UPB1 variants, and urinary analysis confirmed marked elevation of pyrimidine degradation metabolites, establishing the diagnosis of βUP deficiency.This case expands the clinical and neuroradiological spectrum of βUP deficiency and identifies elevated urinary MMA as a previously unreported finding in this disorder. Given the widespread availability of urinary organic acid analysis, MMA may represent a possibly useful and accessible biomarker to support diagnosis. βUP deficiency should be considered in the differential diagnosis of children with Leigh-like encephalopathy and unexplained MMA elevation.
More Related Videos
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:22
Lysosomal Hydrolases
 01:18
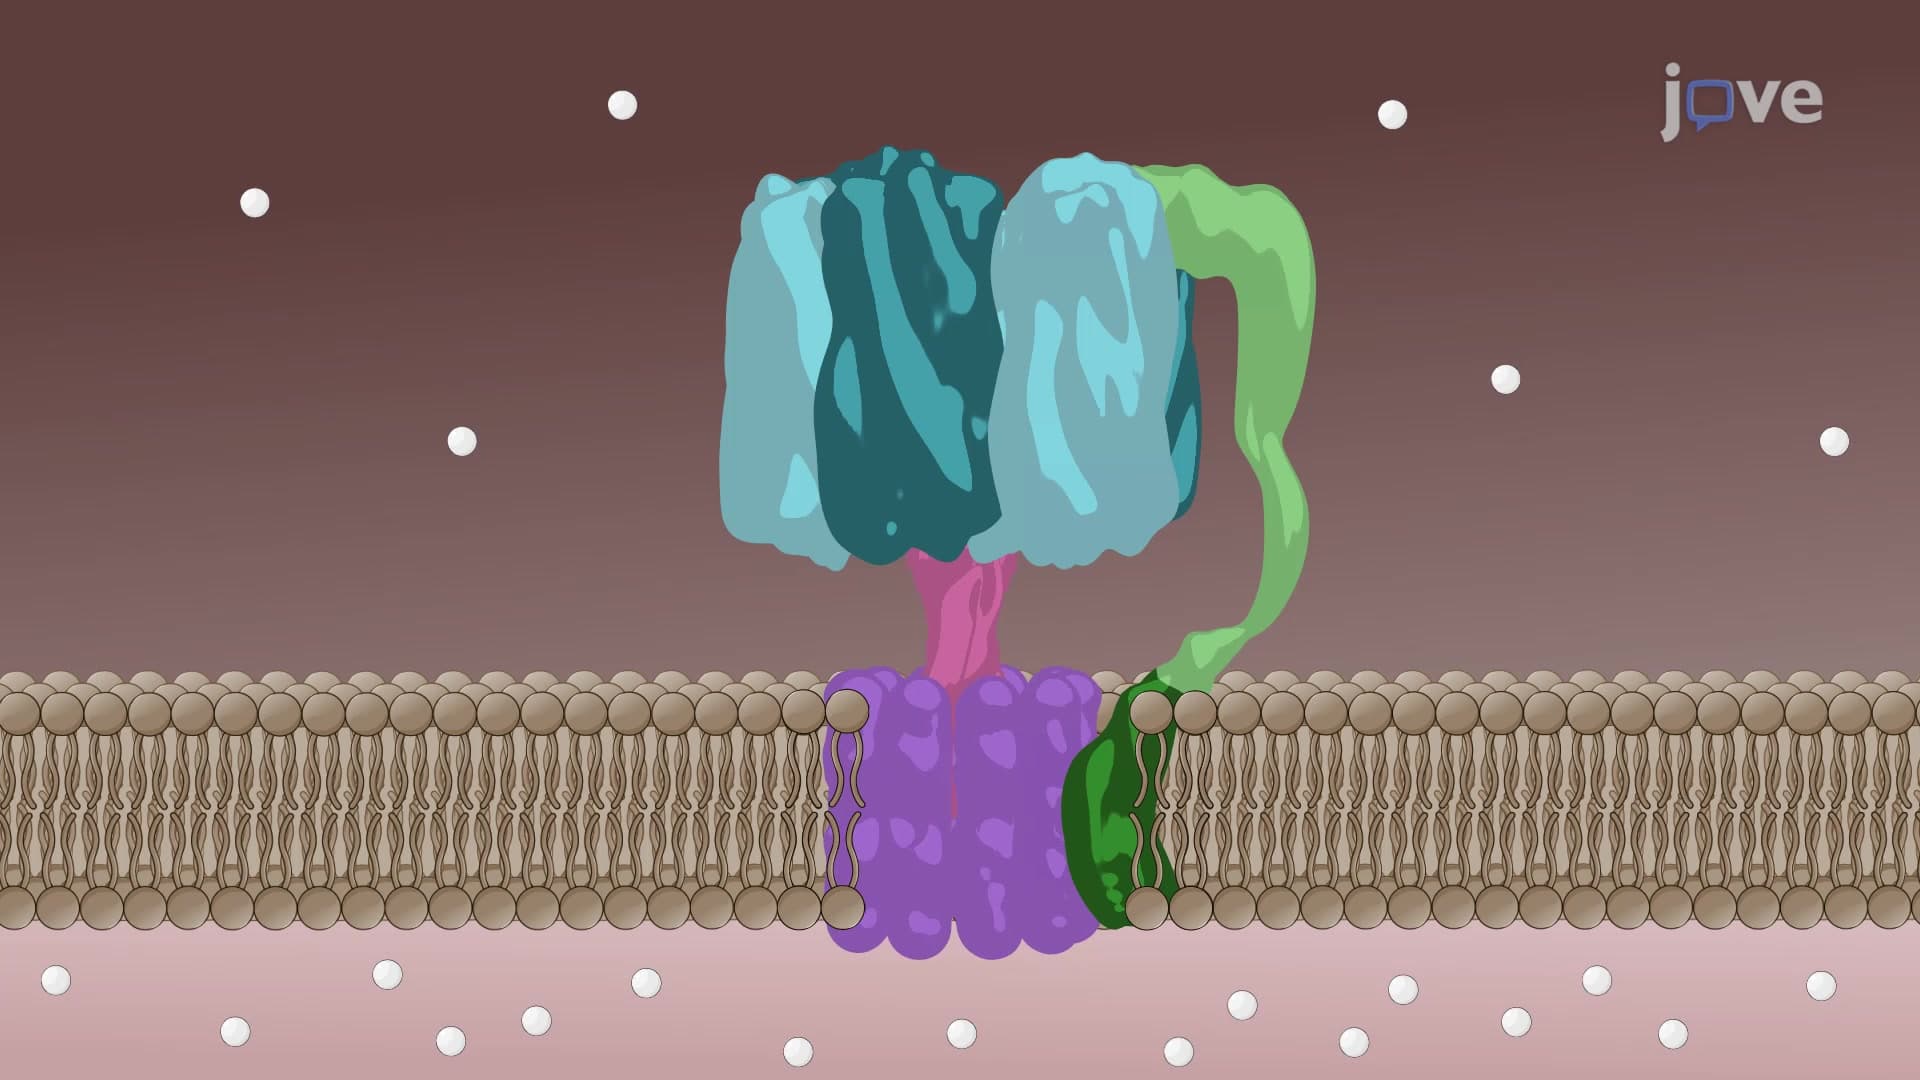
ATP Synthase: Structure
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:23
Urea Cycle
 01:28
Biosynthesis of Nucleic Acids