Related Experiment Video
Updated: May 9, 2026
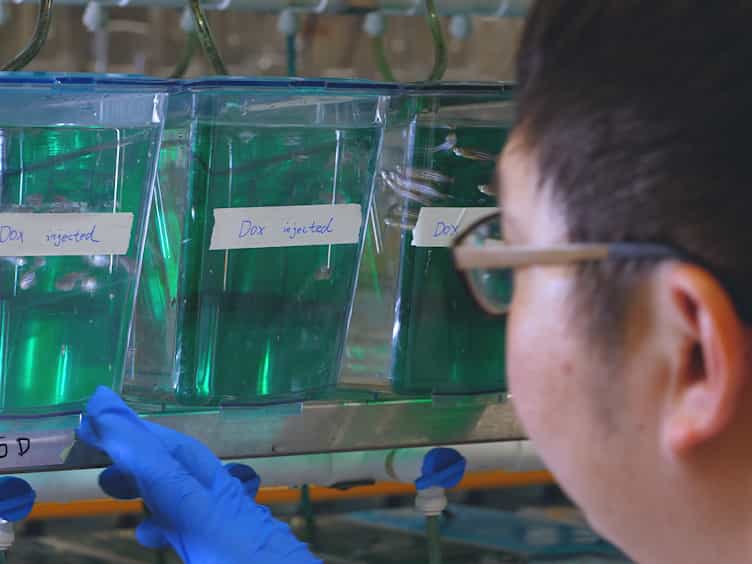
A Doxorubicin-induced Cardiomyopathy Model in Adult Zebrafish
Published on: June 7, 2018
BAG3V468M impairs proteasomal protein clearance and induces dilated cardiomyopathy in vivo
Federica Diofano1, Ionela Emilia Giorgiana Madac1, Dennis Koeble1
1Molecular Cardiology, Department of Internal Medicine II, University of Ulm, Ulm, Germany.

Insights
Genetic variants in BAG3 protein can cause dilated cardiomyopathy (DCM). This study shows a specific BAG3 mutation (V468M) impairs protein quality control, leading to DCM-like heart problems in zebrafish by disrupting protein clearance.
Area of Science:
- Cardiovascular Biology
- Molecular Genetics
- Proteostasis Research
Background:
- Idiopathic dilated cardiomyopathy (DCM) is a primary cause of heart failure, linked to genetic factors.
- BAG3 gene variants are implicated in DCM, suggesting a role in protein quality control (PQC) and proteinopathy.
Purpose of the Study:
- Investigate the in vivo molecular effects of a specific BAG3 missense variant (V468M) associated with familial DCM.
- Determine the impact of BAG3 V468M on cardiac function, myocardial structure, and proteostasis.
Main Methods:
- Ectopic expression of human BAG3 V468M in wild-type zebrafish embryos.
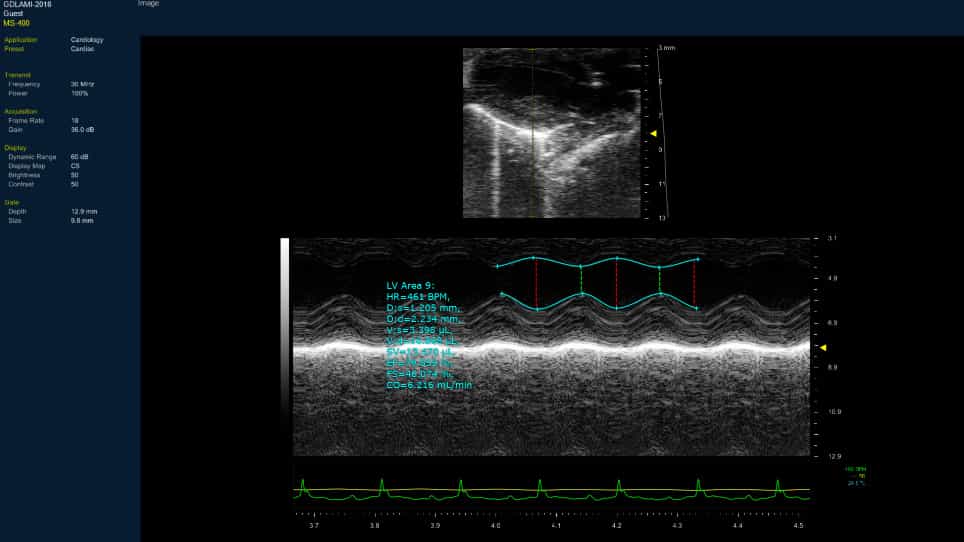
- Phenotypic analysis including echocardiography, heart rate, and contractility measurements.
- Transmission electron microscopy for ultrastructural analysis.
- Assessment of autophagy and ubiquitin-proteasome system markers.
Main Results:
- BAG3 V468M expression induced a DCM-like phenotype in zebrafish, with ventricular dilation, reduced heart rate, and impaired contractility.
- Ultrastructural analysis revealed disrupted myocardial architecture and sarcomeric organization.
- While autophagy markers were unchanged, ubiquitinated protein accumulation indicated impaired ubiquitin-proteasome system function.
Conclusions:
- The BAG3 V468M variant triggers a DCM-like phenotype in vivo.
- Disrupted myocardial architecture and altered proteostasis, specifically impaired ubiquitin-mediated protein turnover, contribute to BAG3-associated cardiomyopathy.
Abstract:
Idiopathic dilated cardiomyopathy (DCM) is one of the major causes of heart failure, characterized by left ventricular dilation and systolic dysfunction in the absence of an identifiable cause, and is associated with reduced life expectancy. Genetic studies, including genome-wide association studies, have identified variants in BAG3, a key regulator of protein quality control (PQC), as contributors to both familial and sporadic forms of DCM. Impaired PQC and the accumulation of misfolded proteins (proteinopathy) have emerged as potential pathogenic mechanisms. Here, we investigated the molecular consequences of a recently identified BAG3 missense variant (V468 M) associated with familial DCM. To assess the in vivo effects of the variant, human BAG3V468M was ectopically expressed in wild-type zebrafish embryos. Overexpression of BAG3V468M resulted in a DCM-like phenotype characterized by ventricular dilation, reduced heart rate, and impaired contractility. Transmission electron microscopy revealed marked disruption of myocardial ultrastructure and sarcomeric organization. To explore the impact on proteostasis, markers of autophagy (LC3-I/II and p62) were analyzed and showed no significant differences between BAG3V468M and control embryos under basal conditions. In contrast, analysis of the ubiquitin-proteasome system demonstrated a significant accumulation of ubiquitinated proteins in BAG3V468M-expressing embryos, suggesting impaired proteasomal protein clearance or increased proteotoxic stress. Expression of BAG3V468M induces a DCM-like phenotype in vivo associated with disrupted myocardial architecture and altered proteostasis. While canonical autophagy markers remain unchanged, the accumulation of ubiquitinated proteins points toward a disturbance in ubiquitin-mediated protein turnover. These findings implicate mutation-specific alterations in proteostasis as a potential mechanism contributing to BAG3-associated cardiomyopathy.
Related Concept Videos
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:30
Cardiomyopathy II: Dilated Cardiomyopathy
 01:29
Cardiomyopathy IV: Restrictive Cardiomyopathy