Related Experiment Video
Updated: May 9, 2026

Combining Analysis of DNA in a Crude Virion Extraction with the Analysis of RNA from Infected Leaves to Discover New Virus Genomes
Published on: July 27, 2018
Benchmarks for taxonomic classification of jingmenviruses and closely related viruses using newly identified genomic
Agathe M G Colmant1, Rhys H Parry2, Remi Charrel1
1Unite des Virus Emergents, Universita di Corsica, Marseille, France.

Abstract:
Jingmenviruses are a group of viruses related to orthoflaviviruses characterized by a segmented genome and multipartite organization that have been detected worldwide in a wide range of hosts. With the growing number of new jingmenvirus sequences identified in metagenomics data, it can be difficult to assess whether a new sequence is associated with a new virus species or with a strain of an existing species. The ICTV is about to ratify the reclassification of the Flaviviridae family, recognizing segmented viruses previously designated jingmenviruses as part of that family and proposing two genera to classify them: Jingmenvirus and Guaicovirus. These proposals do not include clear criteria to classify jingmenviruses and related sequences into species or genera. In order to determine such criteria, we generated a large sequence database from published and newly assembled sequences. Indeed, we screened public raw sequencing data from studies that did not search for or report jingmenvirus or related sequences, looking for new strains of previously described viruses. We then performed multiple sequence alignments and used the inferred percentage identity values to determine demarcation criteria based on the distribution of evolutionary distances upon pairwise comparisons. We report the identification of almost 60 libraries containing jingmenvirus and related sequences, in a wide range of sample types and geographical locations. Using these data and published sequences, we have determined that to be classified as a virus species, at least four segments are required, on which eight cut-off values in percentage identity (nucleotide and amino acid) are used for demarcation. The use of these criteria would enhance consistency in jingmenvirus taxonomy and provide a standardized framework for comparative genomics studies of these viruses, as they are still under-characterized.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:29
Modern Molecular Taxonomy
 01:20
Applications of Molecular Taxonomy
 01:26
Size and Structure of Viral Genomes
 01:24
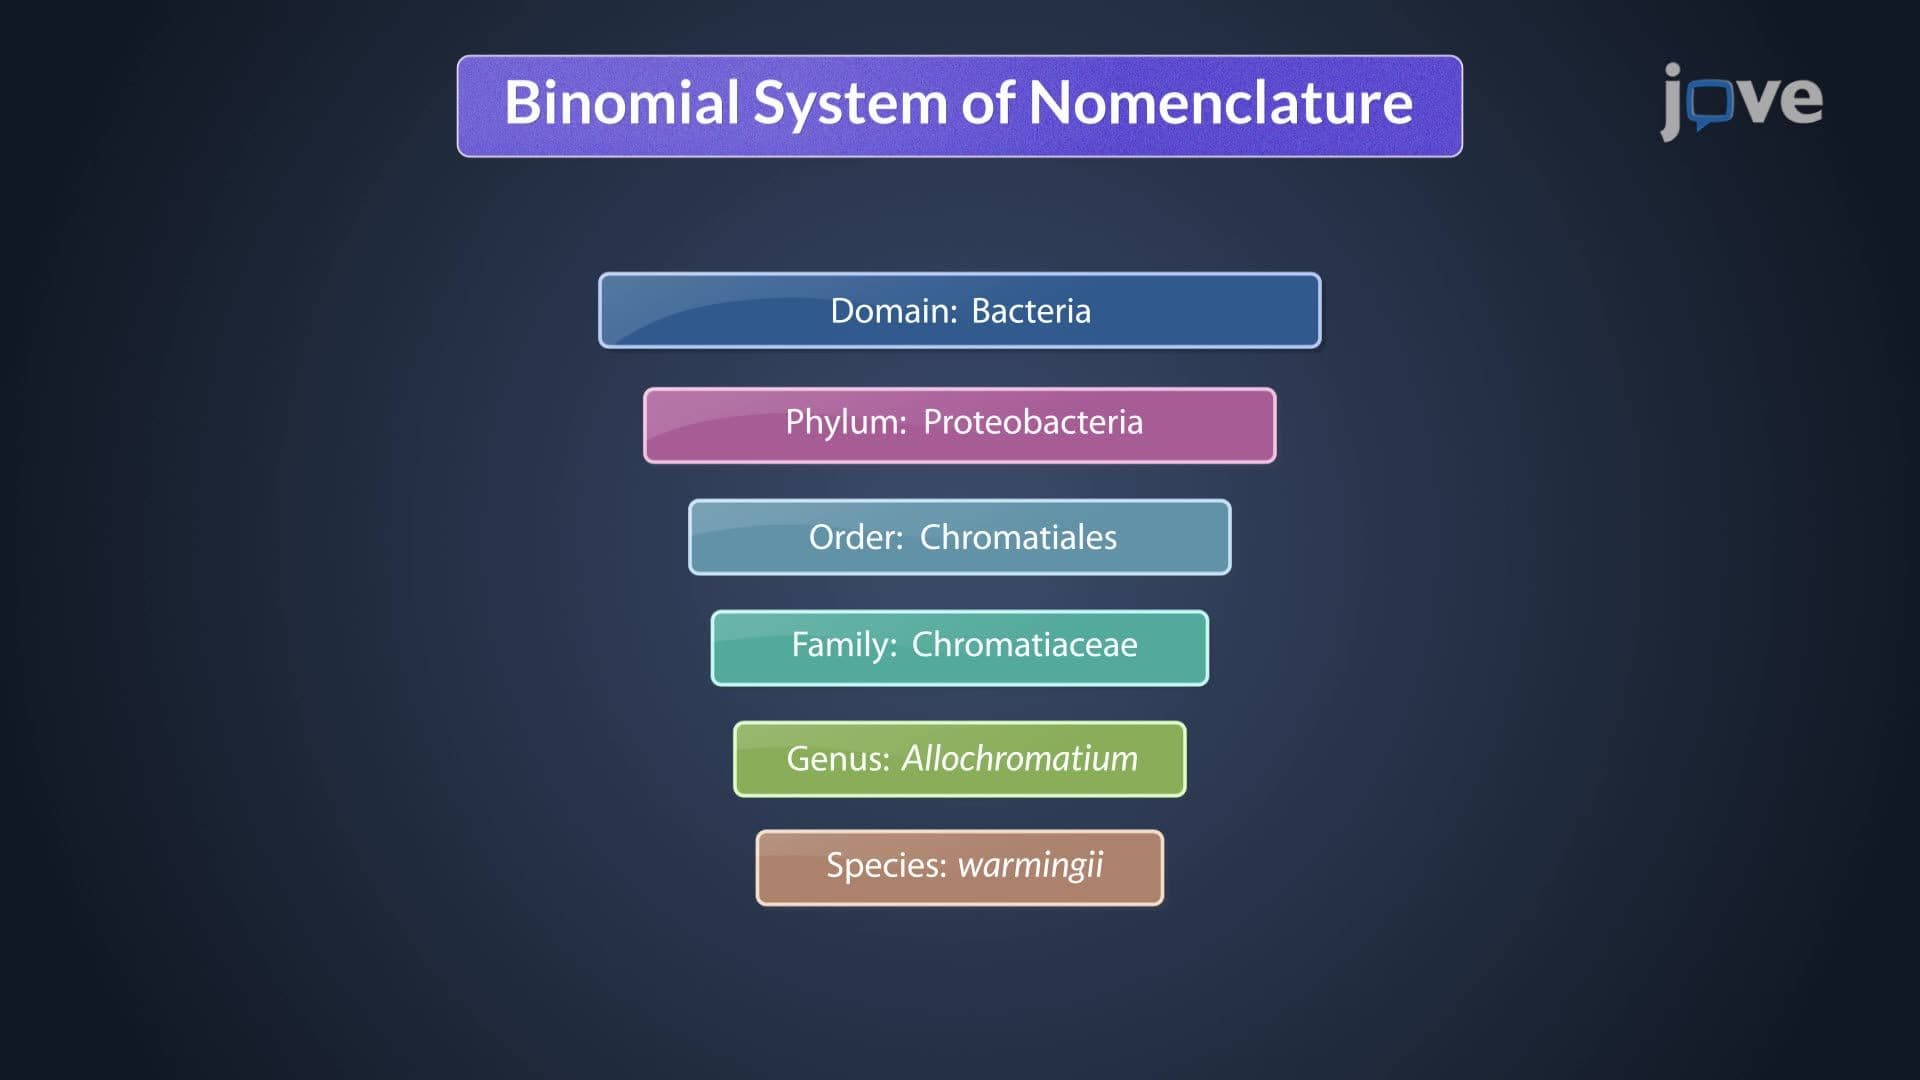
Microbial Classification System
 01:29
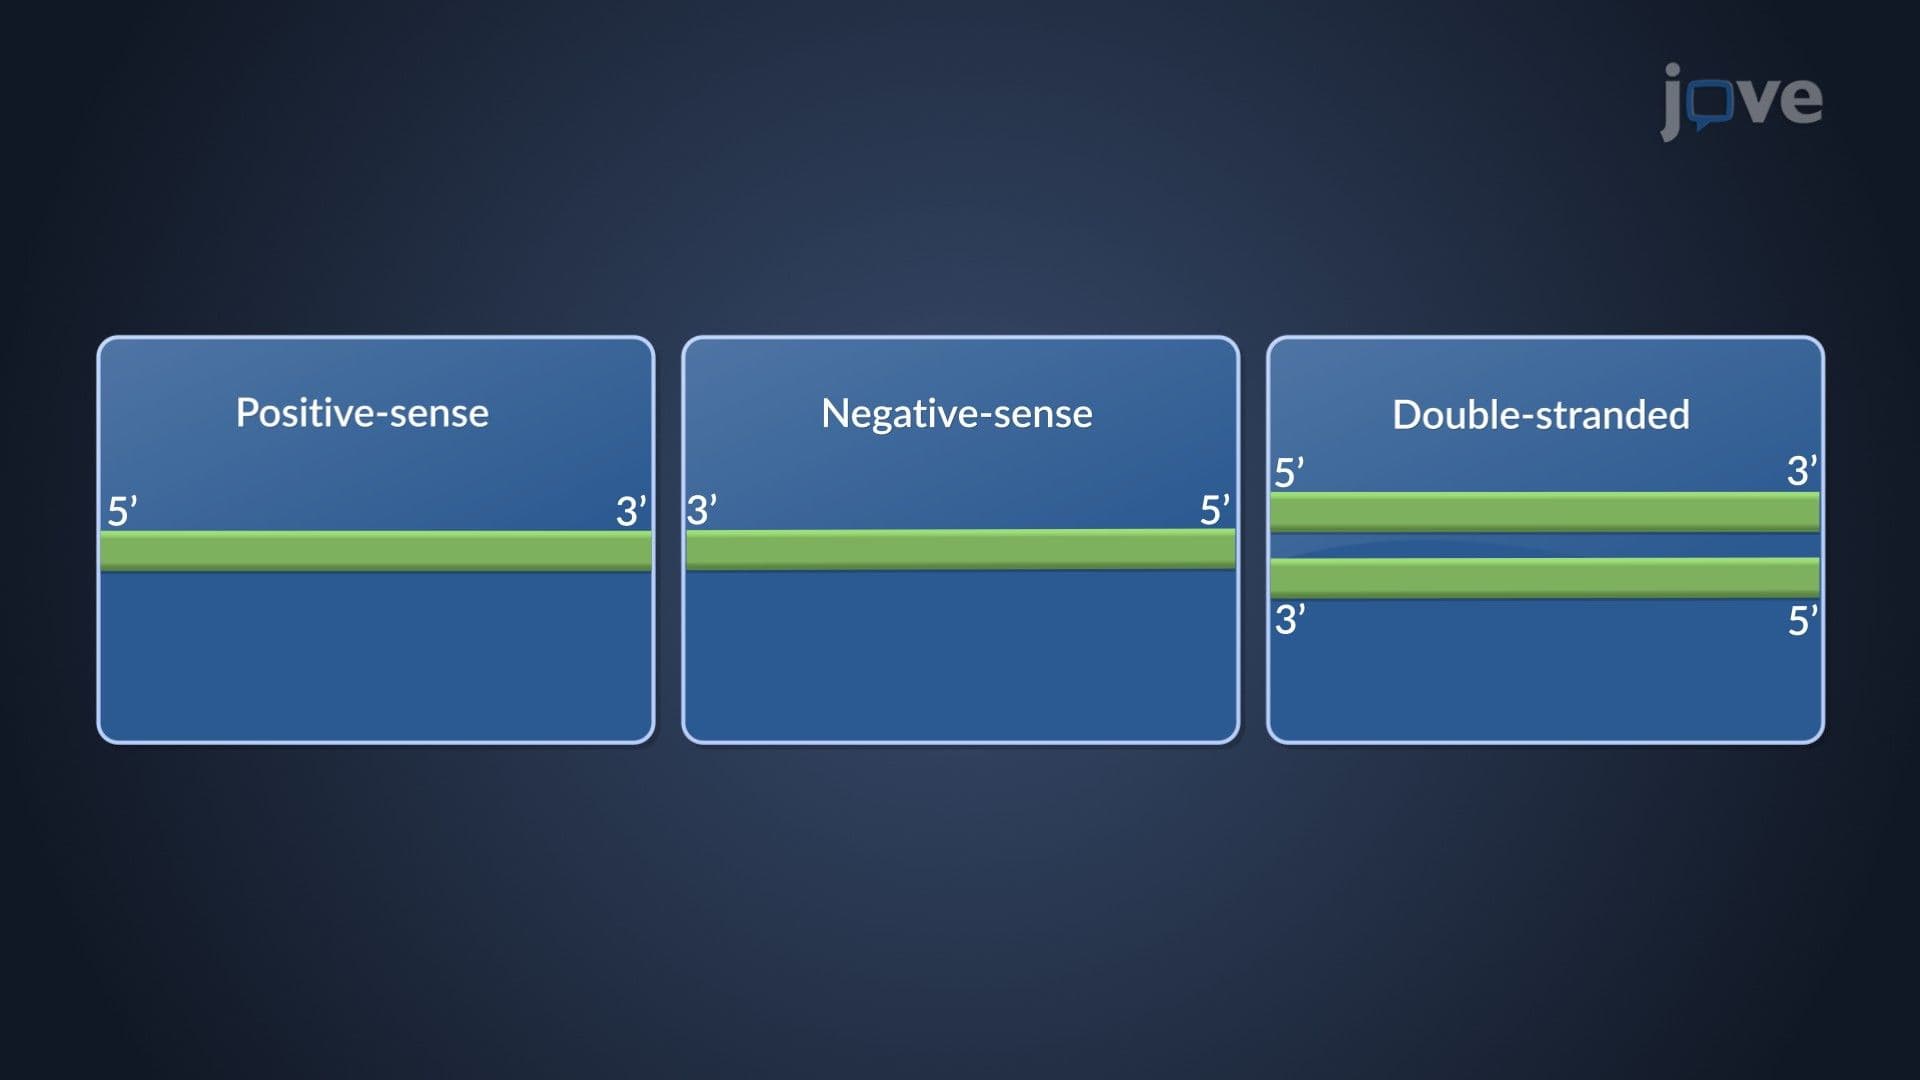
Viruses with RNA Genomes