Related Experiment Video
Updated: May 15, 2026

Influence of Hybrid Perovskite Fabrication Methods on Film Formation, Electronic Structure, and Solar Cell Performance
Published on: February 27, 2017
Dynamic lattice disorder overrides energetics for barrierless interfacial charge transfer in 2D hybrid perovskites
Tianjing Li1, Haoran Lu2, Ming Xia3
1Zhejiang Key Laboratory of Excited-State Energy Conversion and Energy Storage, Department of Chemistry, Institute of Fundamental and Transdisciplinary Research, Zhejiang University, Hangzhou, Zhejiang 310027, China.

Abstract:
Two-dimensional (2D) hybrid perovskites show promise in charge transfer (CT) applications, yet how their inherent dynamic lattice disorder redefines the CT dynamics remains unresolved. Using tetrathiophene-based 2D perovskites 4Tm2MAn-1PbnI3n+1 (n = 1 to 4), we reveal a general two-step CT process: ultrafast hole transfer followed by slower electron transfer from inorganic layer to form triplets in organic ligands. Strikingly, despite a broad driving force range (0.16 to 0.75 electron volts), CT rates increase monotonically with decreasing inorganic layer number n, exhibiting no Marcus inverted region behavior and negligible temperature dependence (80 to 295 kelvin). Such anomalous barrierless CT behavior cannot be described by classical Marcus CT or quantum tunneling models. We propose a dynamic lattice CT model where lattice fluctuation enables sampling a wide spectrum of transient lattice configurations, allowing CT to always proceed via barrierless pathways. This model, validated by ab initio calculations, establishes dynamic lattice disorder as a fundamental design principle for efficient CT in anharmonic materials, transcending conventional energetic constraints.
Related Concept Videos
 01:30
The Electrical Double Layer
 02:54
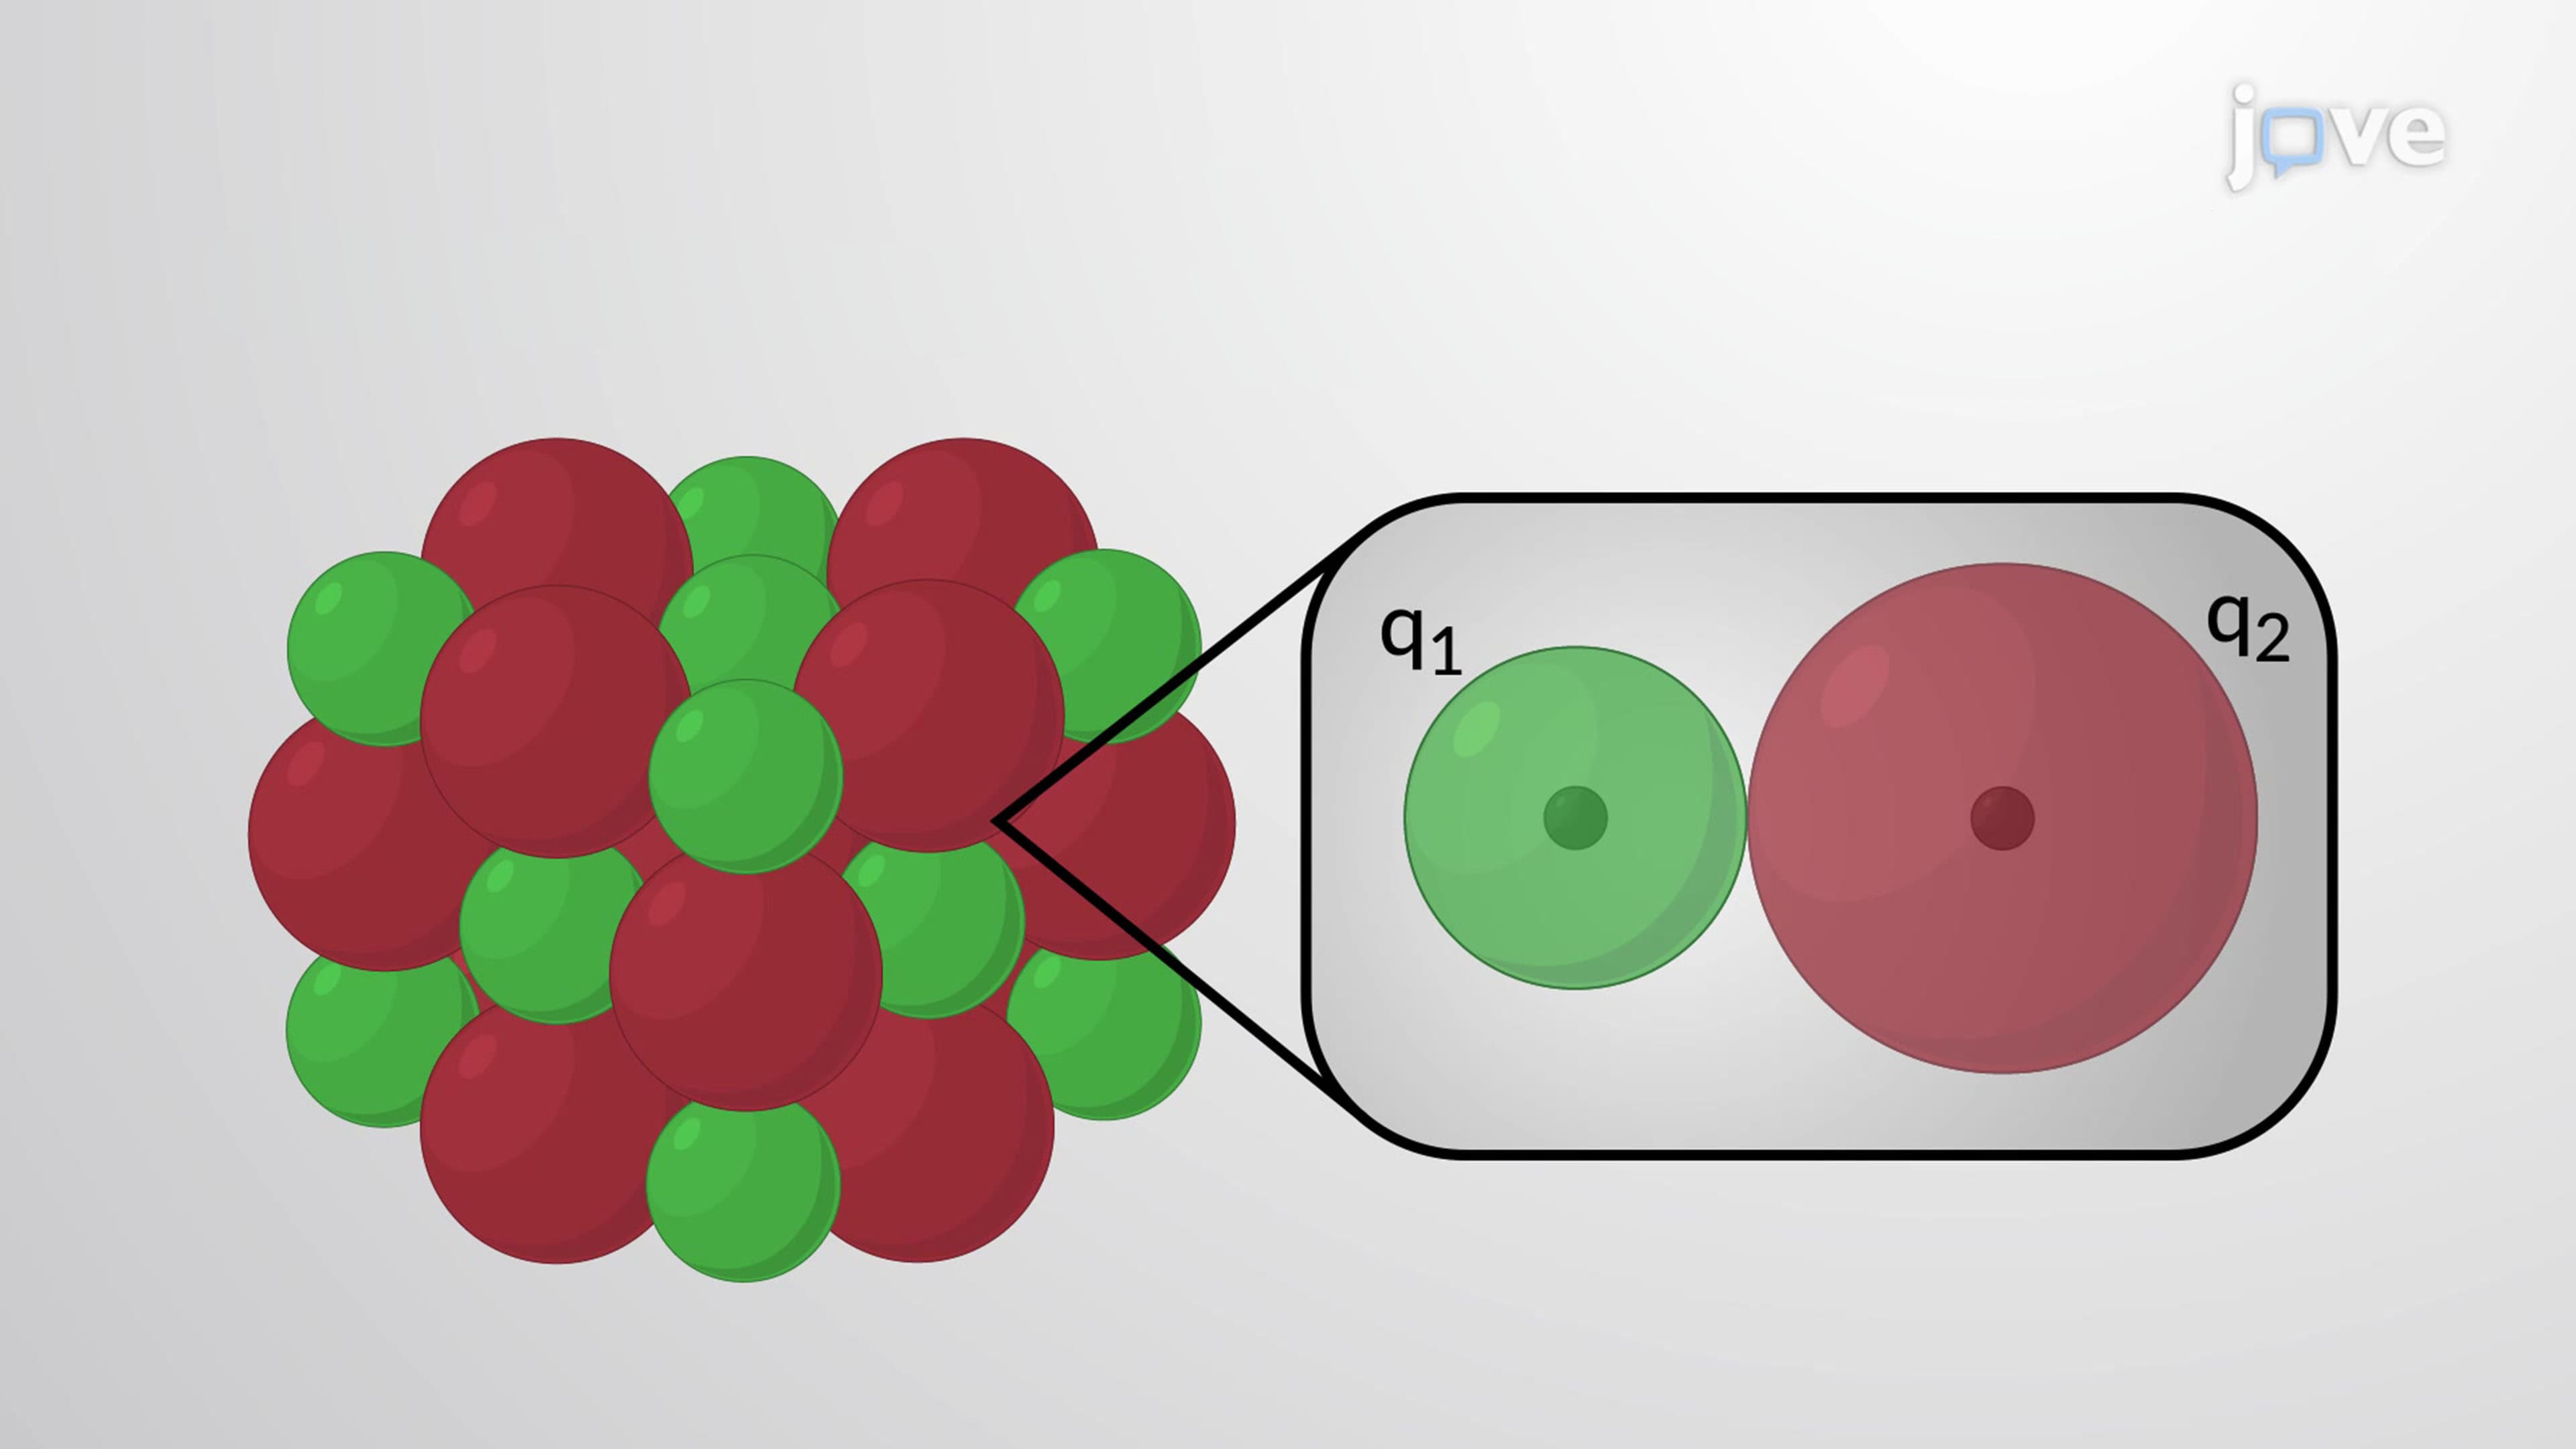
Trends in Lattice Energy: Ion Size and Charge
 01:27
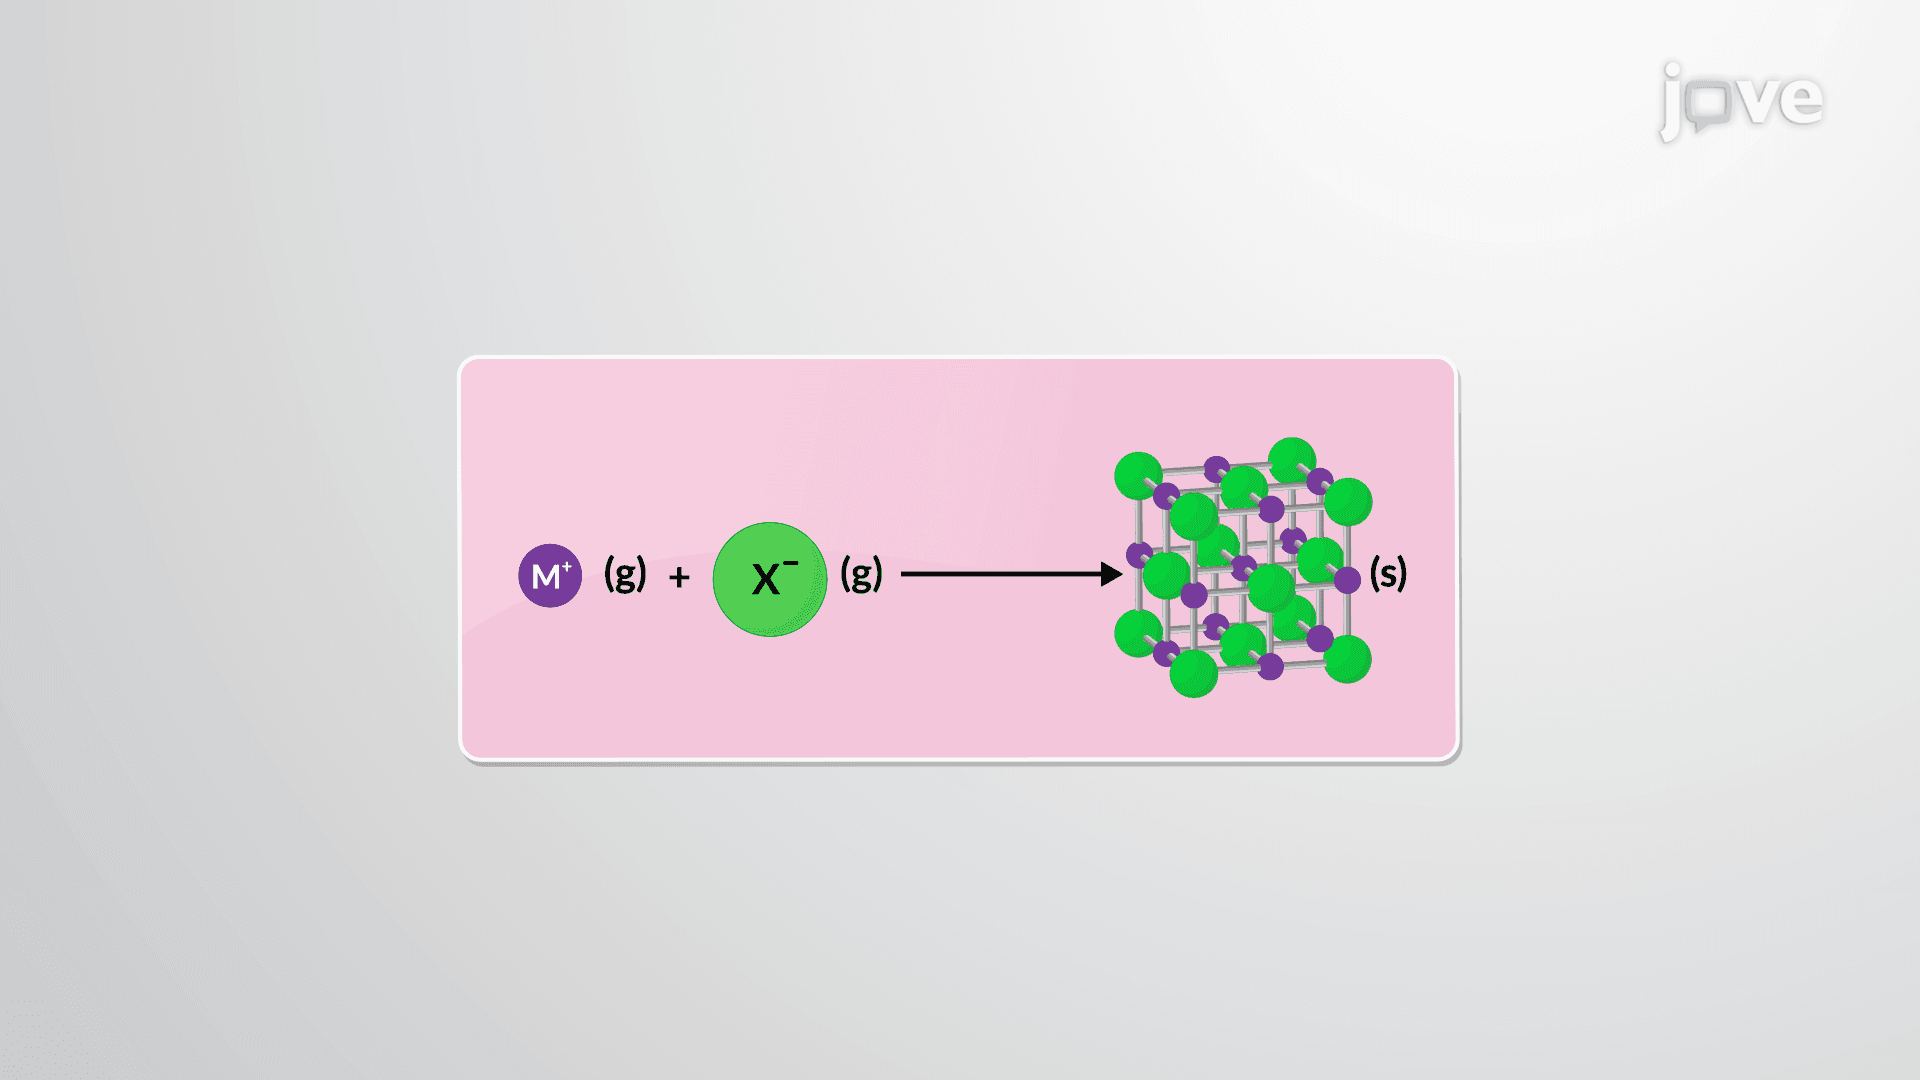
Lattice Energies of Ionic Crystals
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...