Related Experiment Video
Updated: May 24, 2026
Using Live Cell STED Imaging to Visualize Mitochondrial Inner Membrane Ultrastructure in Neuronal Cell Models
Published on: June 30, 2023
Genetic ablation of neuronal mitochondrial calcium uptake impedes Alzheimer's disease progression
Pooja Jadiya1,2, Elena Berezhnaya1, Devin W Kolmetzky1
1Aging + Cardiovascular Discovery Center, Department of Cardiovascular Sciences, Lewis Katz School of Medicine at Temple University, Philadelphia, PA, USA.

Abstract:
Loss of mCa2+ efflux capacity contributes to the pathogenesis and progression of Alzheimer's disease (AD) by promoting mitochondrial Ca2+ (mCa2+) overload. Here, we utilized loss-of-function genetic mouse models to causally evaluate the role of mCa2+ uptake by conditionally deleting the mitochondrial calcium uniporter channel (mtCU) in a robust mouse model of AD. Loss of neuronal mCa2+ uptake reduced Aβ and tau-pathology, synaptic dysfunction, and cognitive decline in 3xTg-AD mice. Knockdown of Mcu in an in vitro model of AD significantly reduced matrix Ca2+ content, redox imbalance, and mitochondrial dysfunction. The preservation of mitochondrial function rescued the AD-dependent decline in autophagic capacity and protected neurons against amyloidosis and cell death. This was corroborated by in vivo data showing improved mitochondrial structure and apposition in AD mice with loss of neuronal Mcu. These results suggest that inhibition of neuronal mCa2+ uptake represents a powerful therapeutic target to impede AD progression.
Insights
Inhibiting mitochondrial calcium uptake in Alzheimer's disease (AD) neurons reduces key pathologies like amyloid-beta and tau tangles. This approach improves mitochondrial function and cognitive decline, offering a potential therapeutic strategy for AD.
Area of Science:
- Neuroscience
- Mitochondrial Biology
- Neurodegenerative Diseases
Background:
- Mitochondrial calcium overload is implicated in Alzheimer's disease (AD) pathogenesis.
- Dysfunctional calcium handling contributes to neuronal damage and cognitive decline in AD.
Purpose of the Study:
- To investigate the causal role of mitochondrial calcium (mCa2+) uptake in AD progression.
- To evaluate the therapeutic potential of inhibiting the mitochondrial calcium uniporter (MCU) in AD models.
Main Methods:
- Utilized loss-of-function genetic mouse models (3xTg-AD) with conditional deletion of the MCU.
- Employed in vitro AD models with MCU knockdown.
- Assessed Aβ and tau pathology, synaptic function, cognitive performance, mitochondrial Ca2+ content, redox balance, and autophagy.
Main Results:
- Loss of neuronal MCU function reduced Aβ and tau pathology, synaptic dysfunction, and cognitive deficits in AD mice.
- MCU knockdown in vitro decreased matrix Ca2+, redox imbalance, and mitochondrial dysfunction.
- Preserved mitochondrial function rescued autophagic capacity and protected neurons from amyloidosis and cell death.
- In vivo studies showed improved mitochondrial structure and function in AD mice lacking neuronal MCU.
Conclusions:
- Inhibiting neuronal mitochondrial calcium uptake is a promising therapeutic strategy for Alzheimer's disease.
- Targeting MCU can ameliorate core AD pathologies and neurodegeneration.
- Restoring mitochondrial calcium homeostasis may reverse AD-associated functional deficits.
More Related Videos
 06:41
Quantitative Analysis of Mitochondria-Associated Endoplasmic Reticulum Membrane (MAM) Stabilization in a Neural Model of Alzheimer's Disease (AD)
Published on: January 10, 2025
 06:23
Visualizing Axonal Growth Cone Collapse and Early Amyloid β Effects in Cultured Mouse Neurons
Published on: October 30, 2018
Related Concept Videos
 01:23
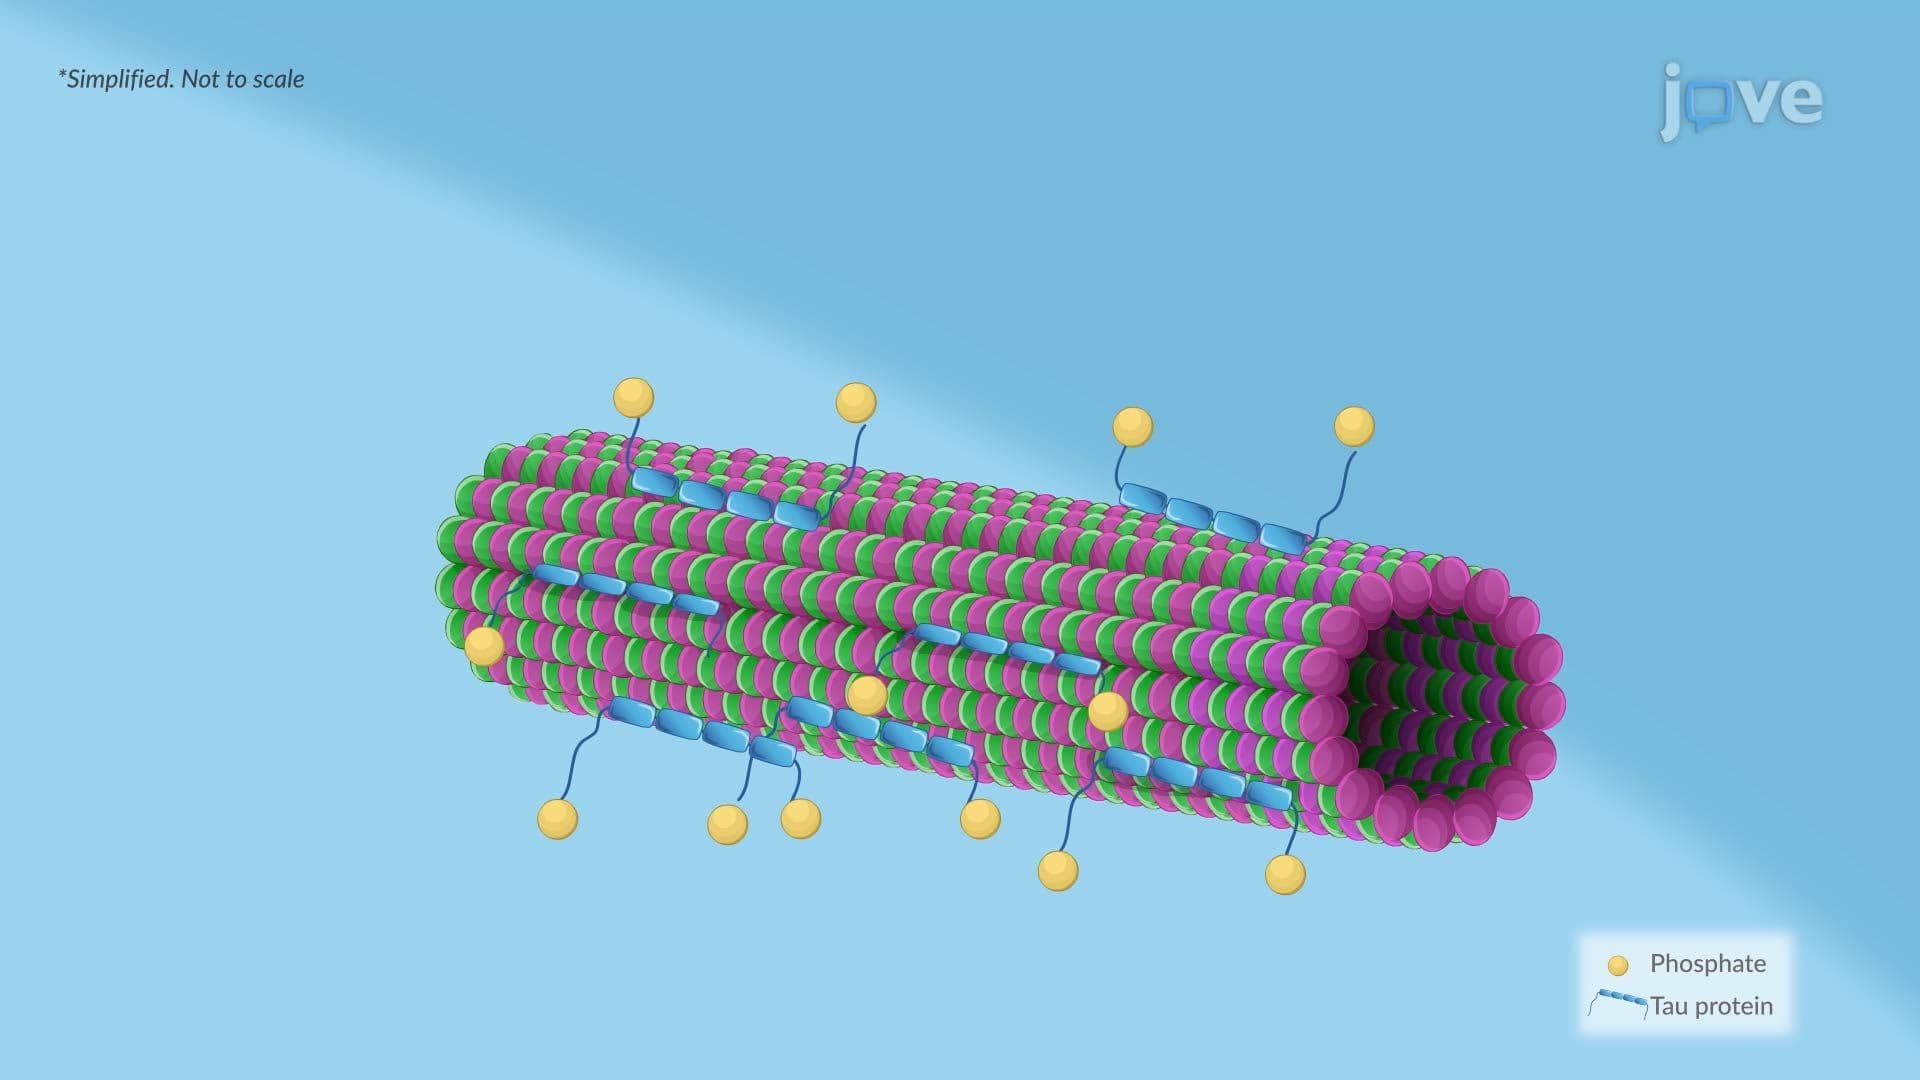
Alzheimer Disease ll: Pathophysiology
 01:26
Alzheimer's Disease: Overview
The clinical diagnosis of AD hinges on the presence of memory and other cognitive impairments. Biomarkers, such as changes in Aβ and tau...
 01:22
Alzheimer's Disease: Treatment
 01:29
Alzheimer Disease l: Introduction