Related Experiment Video
Updated: May 28, 2026

Analysis of Brain Mitochondria Using Serial Block-Face Scanning Electron Microscopy
Published on: July 9, 2016
A Case of Multiple Mitochondrial Dysfunctions Syndrome 1 and Review of the Literature
Charles R DiFalco1,2, Aaron Williams1,2, Claudia Soler-Alfonso3
1Department of Molecular and Human Genetics, Baylor College of Medicine, Houston, Texas, USA.

Abstract:
Multiple mitochondrial dysfunctions syndrome 1 (MMDS1, MIM #605711) due to NFU1 gene defects is an ultra-rare autosomal recessive inborn error of metabolism associated with reduced function of NFU1 iron-sulfur cluster (ISC) scaffold protein. This disorder demonstrates variable clinical features including hypotonia, epileptic encephalopathy, respiratory failure, lactic acidosis, hyperglycinemia, and hemodynamic instability. One unique feature of this mitochondrial disorder is the common manifestation of pulmonary arterial hypertension (PAH), for which the pathophysiologic mechanism is not well defined to date. Here, we report a case of infantile-onset lactic acidosis, refractory epilepsy with encephalopathy, hyperglycinemia, and severe PAH in a patient who underwent rapid genome sequencing (GS) which identified compound heterozygous variants in NFU1 consistent with a diagnosis of MMDS1. We additionally review the literature on published cases of MMDS1 including developments in the understanding of the intracellular impact of defective ISC proteins. Recent studies delineate a multifactorial pathogenesis including globally dysregulated energy metabolism, specific deficiencies in oxidative phosphorylation, and excess reactive oxygen species (ROS) production. A key difference between established animal models of this disorder and human cases published to date is that the previously described sex-based phenotype discrepancy in rats was not recapitulated in this review. Identification of the clinical combination of lactic acidosis, PAH, and hyperglycinemia during diagnostic evaluation should raise suspicion for a defect of the ISC pathway, and further investigations of the cellular pathophysiology of this disorder may provide insight into the phenotypic spectrum of human disease and potential therapeutic targets.
Insights
Multiple mitochondrial dysfunctions syndrome 1 (MMDS1) is caused by NFU1 gene defects. This case highlights lactic acidosis, epilepsy, hyperglycinemia, and pulmonary arterial hypertension (PAH) in an infant, emphasizing the need for early diagnosis of iron-sulfur cluster pathway defects.
Area of Science:
- Biochemistry
- Genetics
- Mitochondrial Biology
Background:
- Multiple mitochondrial dysfunctions syndrome 1 (MMDS1) is an ultra-rare metabolic disorder caused by NFU1 gene defects, affecting iron-sulfur cluster (ISC) protein function.
- Clinical features include hypotonia, epileptic encephalopathy, respiratory failure, lactic acidosis, hyperglycinemia, and hemodynamic instability, with pulmonary arterial hypertension (PAH) being a notable manifestation.
- The precise pathophysiologic mechanism underlying PAH in MMDS1 remains poorly understood.
Purpose of the Study:
- To report a case of infantile-onset MMDS1 diagnosed via rapid genome sequencing.
- To investigate the clinical presentation of MMDS1, focusing on the combination of lactic acidosis, refractory epilepsy, hyperglycinemia, and severe PAH.
- To review and synthesize current understanding of MMDS1 pathophysiology, including intracellular impacts of defective ISC proteins.
Main Methods:
- Rapid genome sequencing (GS) for genetic diagnosis.
- Literature review of published MMDS1 cases.
- Analysis of intracellular impacts of defective ISC proteins.
Main Results:
- A patient with infantile-onset lactic acidosis, refractory epilepsy, hyperglycinemia, and severe PAH was diagnosed with MMDS1 due to compound heterozygous NFU1 variants.
- Literature review revealed multifactorial pathogenesis involving dysregulated energy metabolism, impaired oxidative phosphorylation, and excess reactive oxygen species (ROS) production.
- A key finding was the lack of sex-based phenotype discrepancy in human cases compared to previously reported animal models.
Conclusions:
- The clinical triad of lactic acidosis, PAH, and hyperglycinemia should prompt suspicion for ISC pathway defects.
- Further research into MMDS1 pathophysiology may elucidate the broader phenotypic spectrum and identify potential therapeutic targets.
- Understanding the role of NFU1 and ISC proteins is crucial for managing this rare mitochondrial disorder.
Related Concept Videos
 01:45
Mitochondrial Membranes
 02:59
Animal Mitochondrial Genetics
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:37
Mitochondria
 01:28
The Inner Mitochondrial Membrane
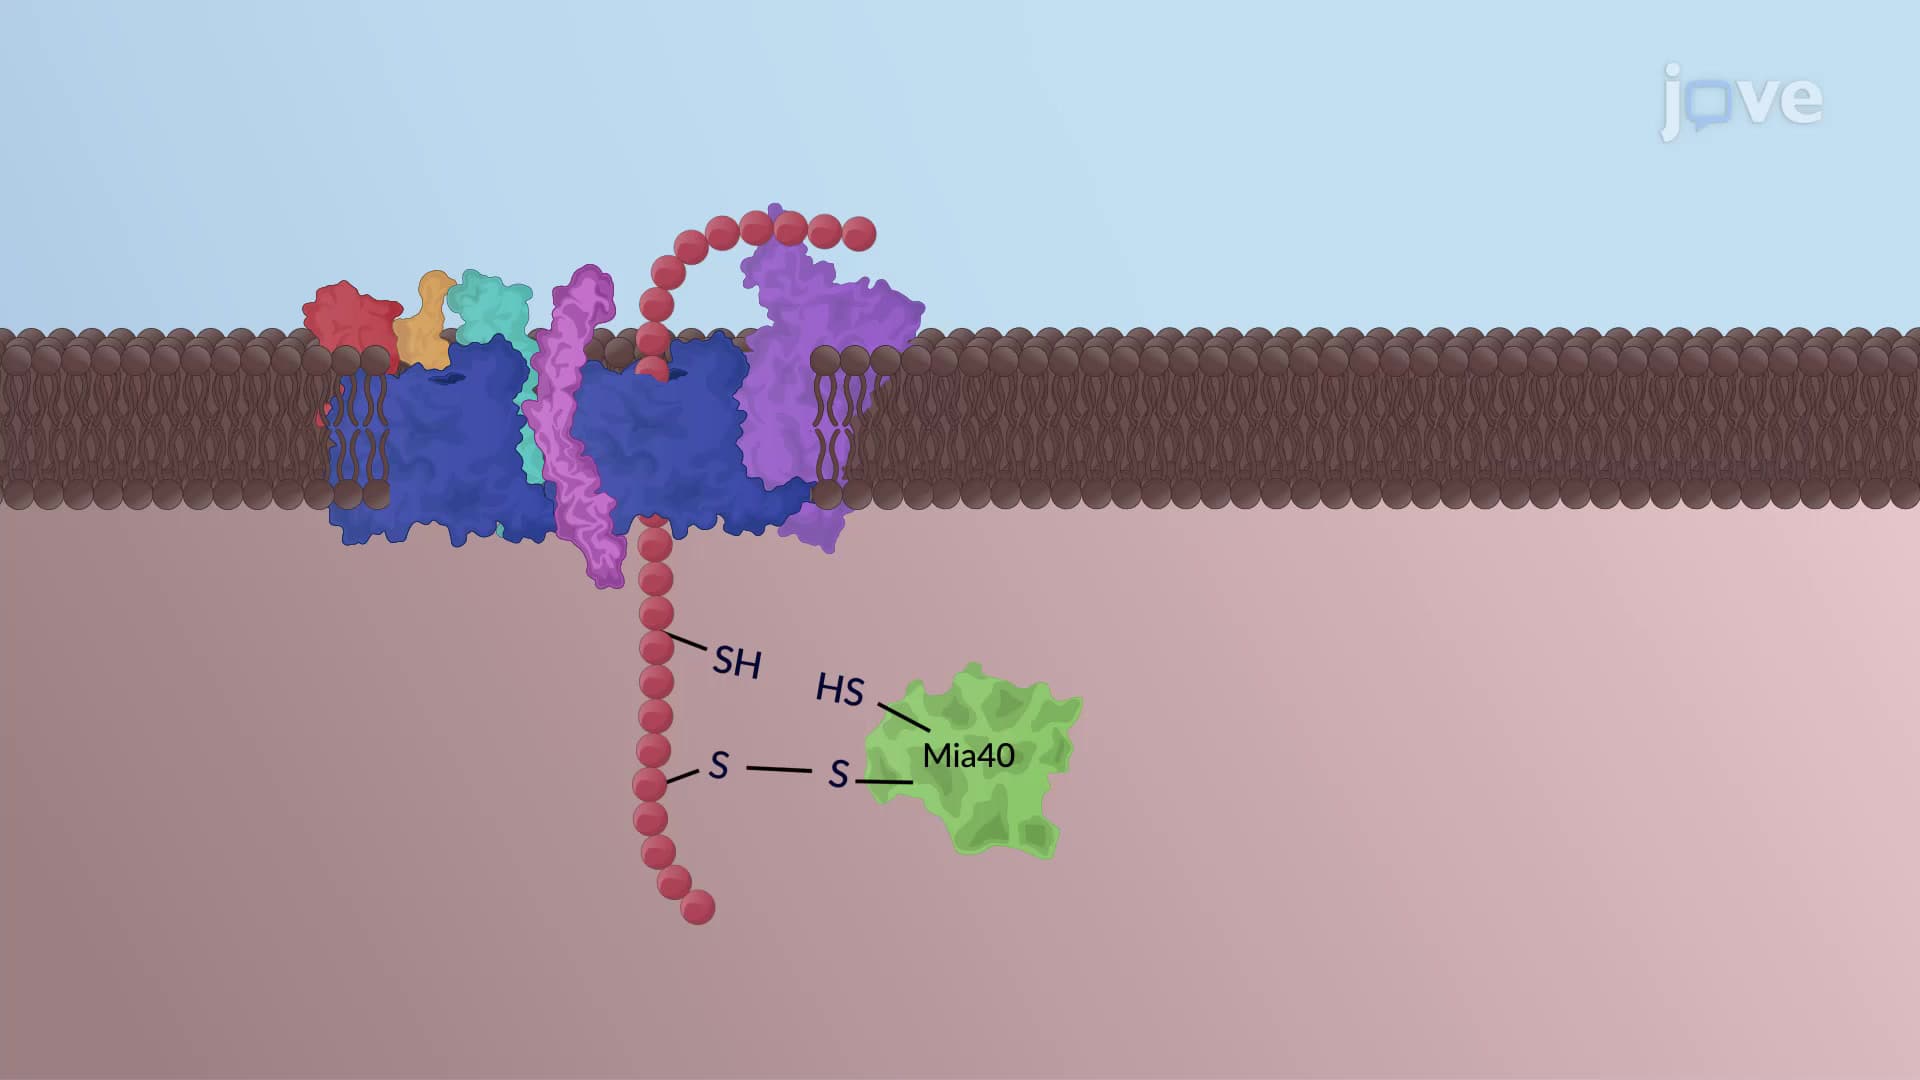 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...