Related Experiment Video
Updated: May 28, 2026

A Protocol for Electrochemical Evaluations and State of Charge Diagnostics of a Symmetric Organic Redox Flow Battery
Published on: February 13, 2017
A Local Charge State Fixation Approach for Resolving Charge-Flow Problems in Reorganization Energy Calculations of
Hyeok Yun1, Jiyoung Bang1, Hayun Kim1
1Department of Chemistry, Chonnam National University, Gwangju 61186, Republic of Korea.

Abstract:
Marcus theory provides a fundamental framework for describing hopping-type charge transport, and donor-bridge-acceptor (D-B-A) dimer models are widely used to evaluate reorganization energies and electron transfer rates. In practice, constrained density functional theory (CDFT) has been extensively employed to construct charge-localized states because the direct evaluation of diabatic states using post-Hartree-Fock (post-HF) methods is computationally demanding. However, in systems containing π-conjugated bridges, maintaining charge-localized states within CDFT can become more difficult, which may affect the evaluated reorganization energies. Standard remedies, including partial geometry fixation and ONIOM-based approaches, do not sufficiently resolve this limitation. In this study, we propose a local charge state fixation (LCSF) method for calculating reorganization energies in D-B-A dimers without imposing explicit constraints on the electronic density. Charged donor and acceptor monomers are optimized independently and subsequently translated and rotated to satisfy the Eckart condition with respect to the corresponding fragments of the neutral dimer. These monomers are then grafted to construct charge-localized dimer geometries, enabling reorganization energy evaluation without charge flow. The method was applied to 1,4-diphenylbutane (PS), trans,trans-1,4-diphenyl-1,3-butadiene (PD), and 1,4-diphenylbutadiyne (PT). For the PS, reorganization energies obtained from LCSF and CDFT are comparable, whereas for PD and PT, LCSF yields consistently larger values, indicating reduced electron delocalization effects. The applicability of the method was further demonstrated using Hartree-Fock and MP2 calculations. In addition, LCSF was applied to zinc oxo cluster dimers, allowing electron transfer rates to be evaluated. Although the method introduces structural constraints, it offers a computationally efficient strategy for reorganization energy calculations in dimer systems, particularly within post-HF frameworks.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 06:53
Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Related Concept Videos
 01:30
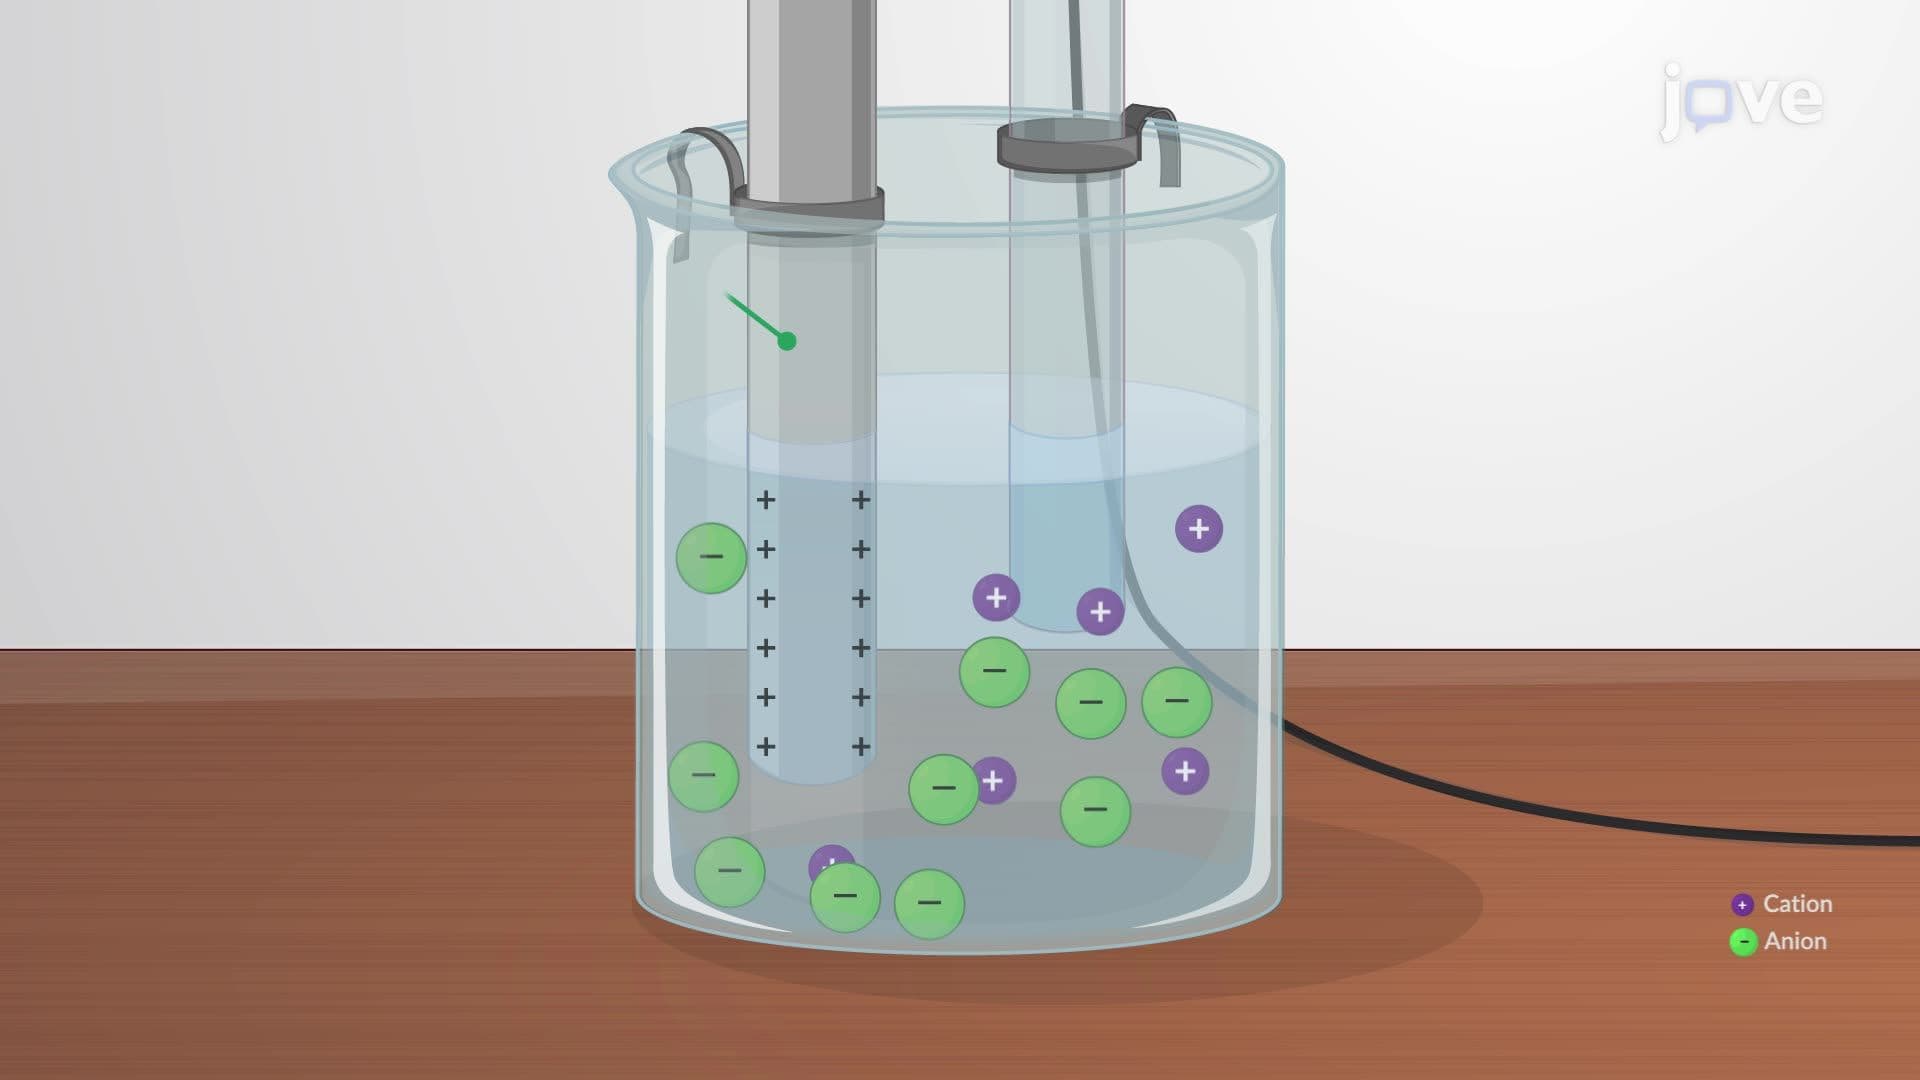
The Electrical Double Layer
 02:42
Formal Charges
 01:24
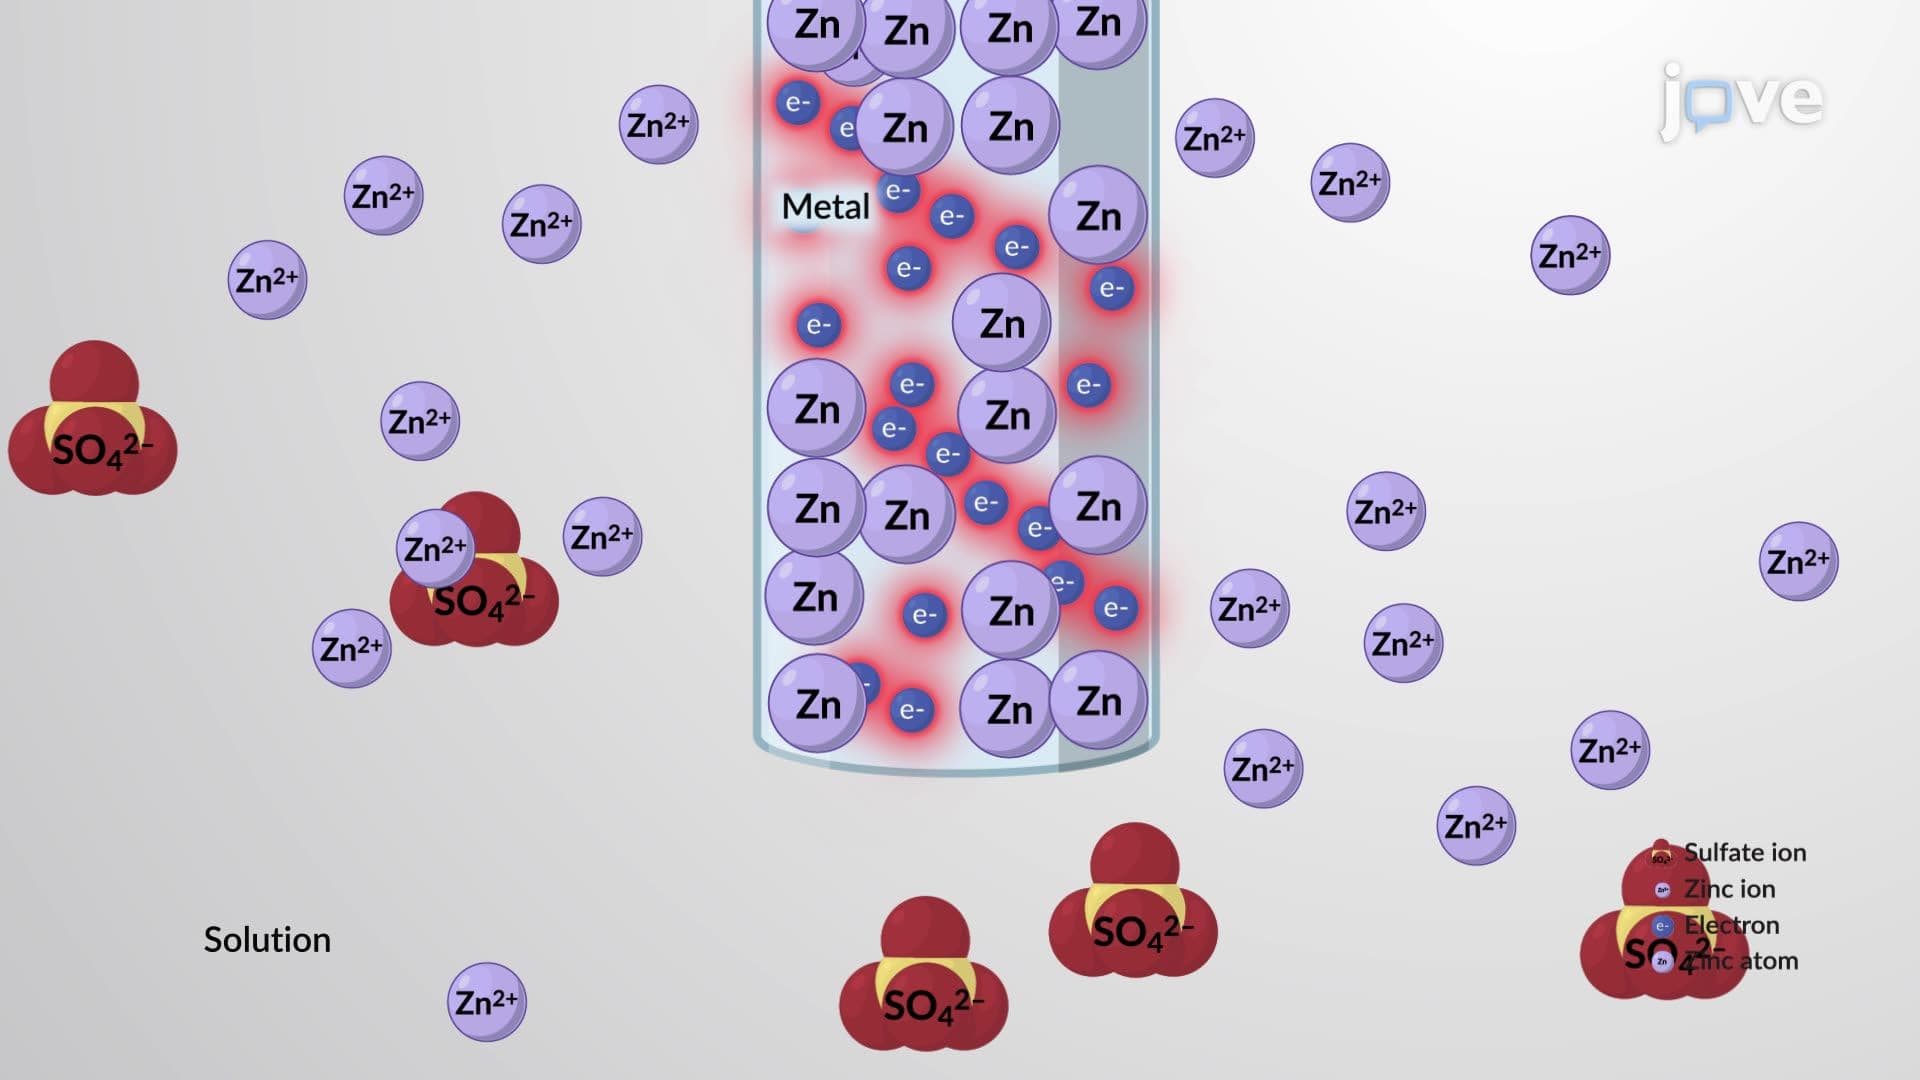
Electrochemical Systems
 01:30
Processes at Electrodes
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...