Related Experiment Video
Updated: May 29, 2026

Drug Repurposing Hypothesis Generation Using the "RE:fine Drugs" System
Published on: December 11, 2016
Machine learning-driven drug repurposing for GPR17: activity prediction via graph neural networks and multistage
Noushin Agha Babaie1, Morteza Farnia2, Jahan B Ghasemi2
1Department of Chemistry, Kish International Campus, University of Tehran, Tehran, Iran.

Abstract:
To identify novel GPR17-targeting ligands with potential relevance to multiple sclerosis (MS) therapy, we developed an integrated computational workflow combining graph neural network (GNN)-based prediction with multistage structure-based validation. GPR17 is a class A G protein-coupled receptor implicated in oligodendrocyte differentiation, neuroinflammation, and remyelination, making it a remarkable target for myelin repair. A curated dataset of 323 chemically diverse ligands with experimentally reported pIC50 values was assembled from the literature and used to train an NNConv-based GNN model. The model demonstrated robust predictive performance across training, validation, and independent test sets (R2 = 0.868, 0.727, and 0.790, respectively). To further assess model generalizability, scaffold-based validation was additionally performed to evaluate prediction across structurally distinct chemotypes. Analysis of the learned chemical space revealed enrichment of antihistamine- and leukotriene-related scaffolds among highly ranked compounds, motivating a drug-repurposing strategy. Candidate molecules, including barmastine, astemizole, sulukast, and iralukast, were evaluated alongside reference ligands pranlukast and cangrelor. Predicted hits were subsequently refined through pharmacophore-guided virtual screening, molecular docking, ADMET profiling, molecular dynamics simulations, and MM-PBSA/MM-GBSA free-energy calculations. Several ligands exhibited stable binding modes supported by persistent hydrophobic interactions and hydrogen-bond occupancies reaching 99.8%. Importantly, this study proposes computationally prioritized candidates rather than experimentally validated inhibitors. Overall, the proposed hybrid workflow provides a practical strategy for early-stage GPR17 ligand discovery and future CNS-oriented drug development.
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:26
Drug Discovery: Overview
 01:29
Pharmacogenomics: Identification of New Drug Targets
 01:15
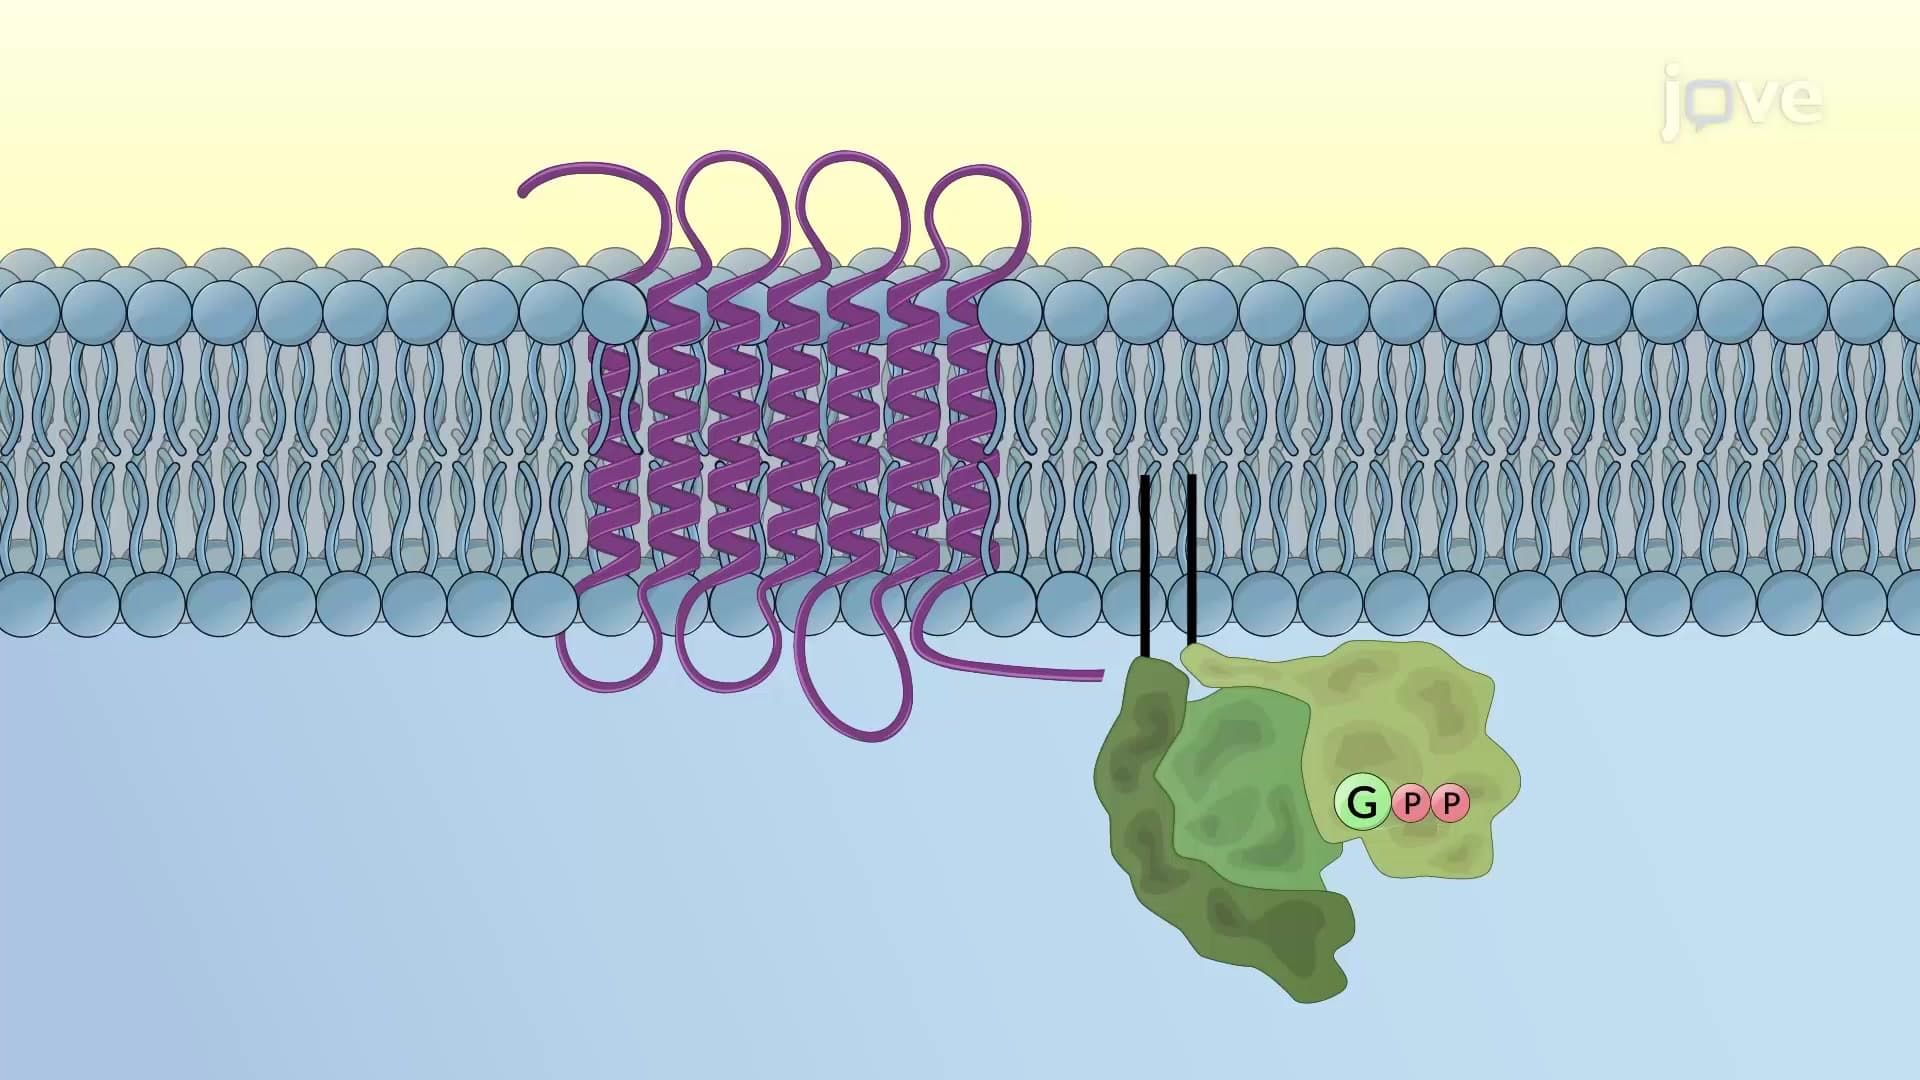
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...