Related Experiment Video
Updated: Jun 1, 2026
Assessment of Open Probability of the Mitochondrial Permeability Transition Pore in the Setting of Coenzyme Q Excess
Published on: June 1, 2022
COXFA4L2 upregulation preserves residual cytochrome c oxidase activity in COXFA4-related Leigh-like encephalopathy
Micol Falabella1, Sandra Lopez Calcerrada2, Jana Aref3
1Department of Neuromuscular Diseases, University College London Queen Square Institute of Neurology, London, UK. m.falabella@ucl.ac.uk.

Abstract:
Primary mitochondrial diseases (PMDs) affect approximately 1 in 4300 individuals and cause early-onset neuromuscular and multisystem dysfunction with reduced lifespan. They result from pathogenic variants in mitochondrial or nuclear DNA that impair oxidative phosphorylation. Cytochrome c oxidase (COX; complex IV) deficiency is a well-established cause of PMD, leading to a broad spectrum of phenotypes. COXFA4 (cytochrome c oxidase subunit FA4), formerly NDUFA4, is a nuclear-encoded COX subunit, but its role in disease remains poorly defined. We report the largest genetically confirmed cohort of COXFA4-related PMD to date, comprising 13 individuals from 12 families with biallelic pathogenic COXFA4 variants. All present with Leigh-like encephalopathy and complete loss of COXFA4 protein; however, patient-derived fibroblasts retain residual COX activity, with upregulation of COXFA4L2 (cytochrome c oxidase subunit FA4-like 2), a poorly characterised paralog. Here, we show that COXFA4 is a late-stage COX assembly subunit and identify a paralog-mediated compensatory mechanism with translational potential.
Insights
Primary mitochondrial diseases (PMDs) are linked to COXFA4 gene variants. A compensatory mechanism involving COXFA4L2 may offer therapeutic potential for Leigh-like encephalopathy.
Area of Science:
- Genetics
- Biochemistry
- Neurology
Background:
- Primary mitochondrial diseases (PMDs) are genetic disorders affecting ~1 in 4300 individuals, causing early-onset neuromuscular and multisystem dysfunction.
- Oxidative phosphorylation defects, particularly cytochrome c oxidase (COX; complex IV) deficiency, are common causes of PMDs with diverse clinical presentations.
- The role of COXFA4 (cytochrome c oxidase subunit FA4), a nuclear-encoded COX subunit, in disease pathogenesis is not well understood.
Purpose of the Study:
- To investigate the clinical and genetic spectrum of PMD caused by pathogenic variants in the COXFA4 gene.
- To elucidate the molecular mechanisms underlying COXFA4 deficiency and identify potential compensatory pathways.
- To explore the functional relationship between COXFA4 and its paralog, COXFA4L2.
Main Methods:
- Genetic analysis of 13 individuals from 12 families with suspected PMD.
- Assessment of COXFA4 protein levels and COX activity in patient-derived fibroblasts.
- Analysis of COXFA4L2 expression and its role in COX assembly and activity.
Main Results:
- Identified 13 individuals with biallelic pathogenic COXFA4 variants, presenting with Leigh-like encephalopathy and complete loss of COXFA4 protein.
- Patient-derived fibroblasts showed residual COX activity despite absent COXFA4.
- Upregulation of the poorly characterized paralog COXFA4L2 was observed, suggesting a compensatory mechanism.
Conclusions:
- COXFA4 is a crucial late-stage assembly subunit of cytochrome c oxidase.
- A paralog-mediated compensatory mechanism involving COXFA4L2 can partially restore COX activity in COXFA4-deficient cells.
- This compensatory pathway holds translational potential for therapeutic strategies in COXFA4-related PMD.
More Related Videos
 05:27
Inner Mitochondrial Membrane Sensitivity to Na+ Reveals Partially Segmented Functional CoQ Pools
Published on: July 20, 2022
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase (COX/SDH) Double-labeling Histochemistry
Published on: November 23, 2011
Related Concept Videos
 01:29
Hepatic Encephalopathy
 01:24
Cofactors and Coenzymes
Cofactors can be metallic ions or organic molecules called coenzymes. These types of helper...
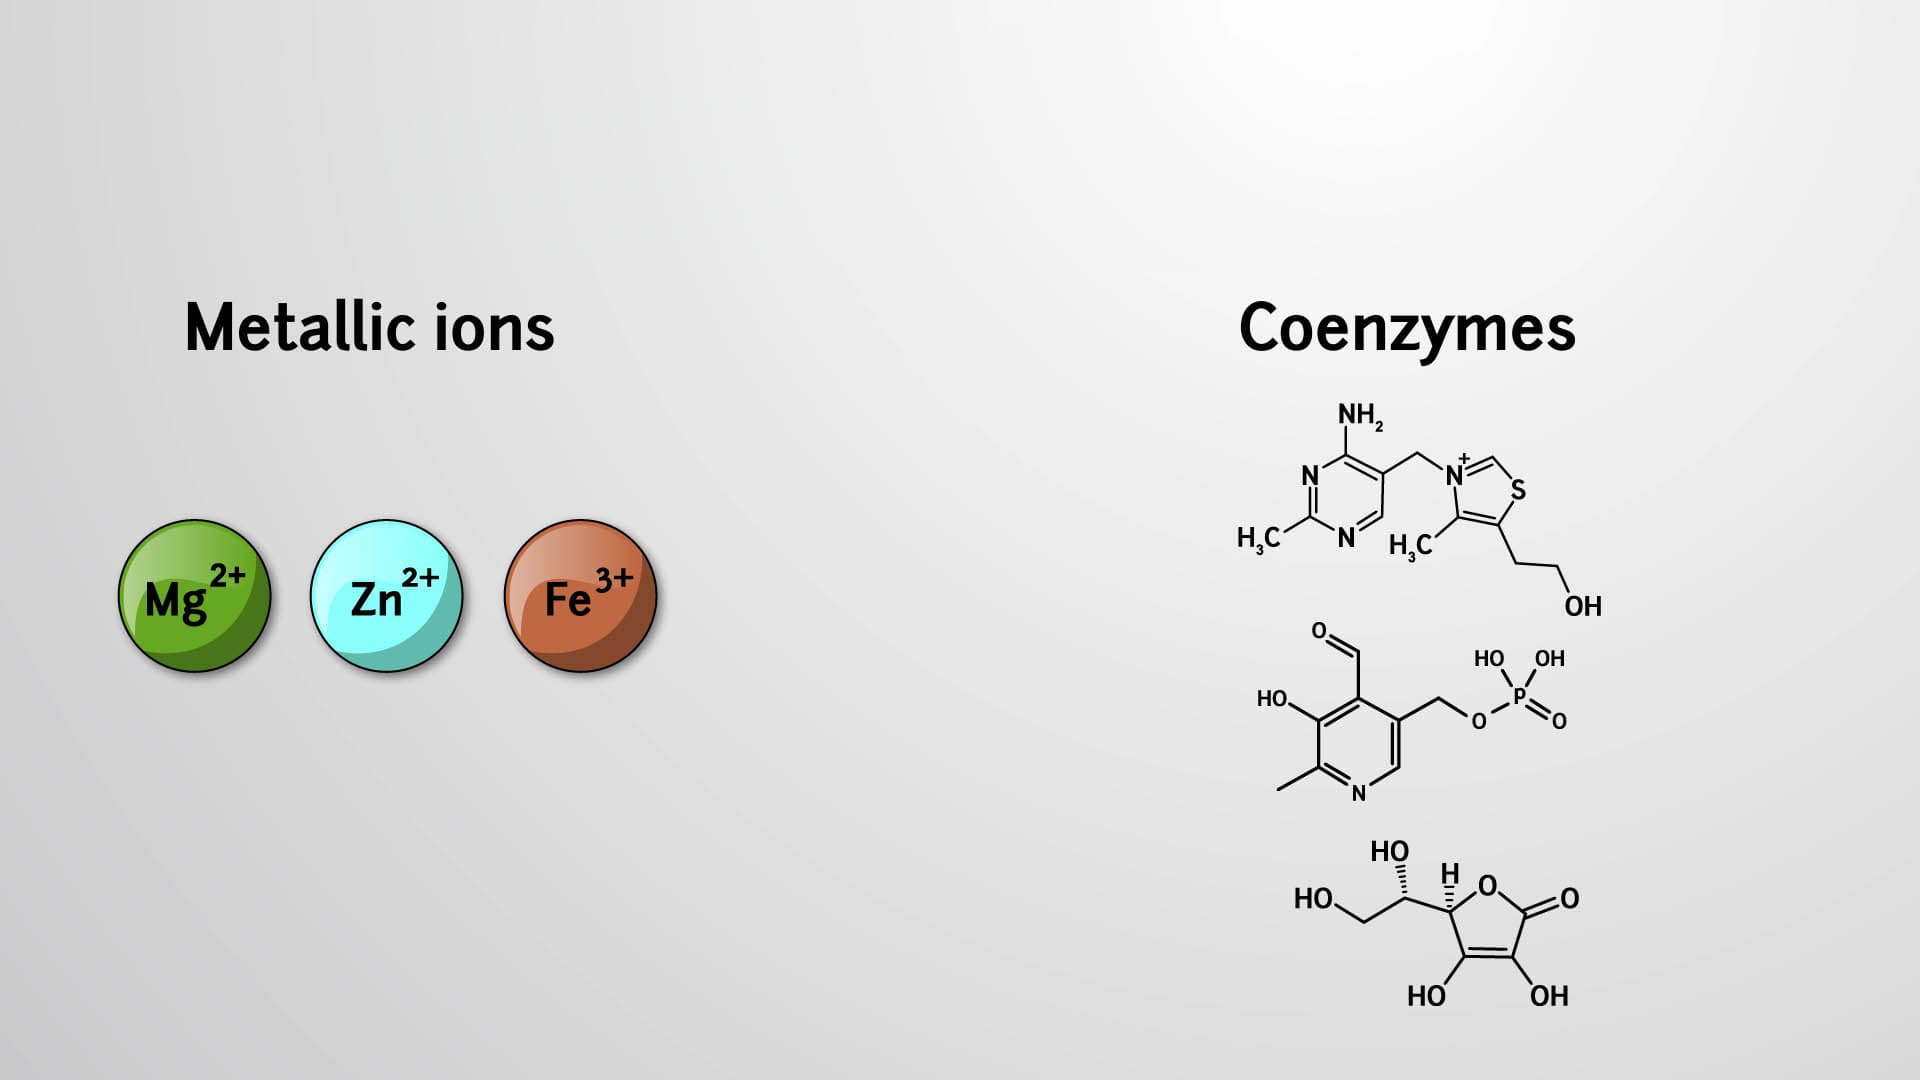 01:27
Cofactors and Coenzymes
01:27Cofactors and Coenzymes
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:43
Electron Transport Chain: Complex III and IV