Related Experiment Video
Updated: Jun 8, 2026
Purifying the Impure: Sequencing Metagenomes and Metatranscriptomes from Complex Animal-associated Samples
Published on: December 22, 2014
Metatranscriptomic analysis of upper and lower respiratory tract microbiomes in patients with tuberculosis and
Liangyu Li1,2, Meng Xu3,4, Mengling Liu1
1Department of Pulmonary and Critical Care Medicine, Renmin Hospital of Wuhan University, Wuhan, Hubei, 430060, China.

Background:
Although changes in the lung microbiome have been observed in many respiratory diseases, the lung microbiome of patients with tuberculosis (TB) remains largely undefined. The aim of this study was to determine and compare the composition of upper and lower respiratory microbial communities, changes in host gene expression, and functional pathway activation in patients with TB and community-acquired pneumonia using a Metatranscriptomic approach.
Methods:
From November 2020 to November 2021, 42 bronchoalveolar lavage fluid samples, 10 oropharyngeal swabs, and 10 nasopharyngeal swabs were collected from patients hospitalized with TB or community-acquired pneumonia for RNA-sequencing, within 72 h of admission. Data from 28 healthy controls were downloaded from the National Center for Biotechnology Information database.
Results:
The most common microorganisms in the samples from patients with TB were Prevotella, Escherichia, Mycobacterium, and Verrucoccus. Notable differences in microorganism diversity were observed between the TB and community-acquired pneumonia groups as well as between the upper and lower respiratory tracts compared to that in healthy controls. Altered microbial interactions were also observed. Rothia and Mycobacterium were identified as marker microorganisms in the TB group, which exhibited increased expression of SNAP25, MGAM2, CNTN4, MMP12, and GPR174, in parallel to IL36A, RHCG, and CYP2B6 downregulation relative to the community-acquired pneumonia group. The pathways enriched for differential genes were similar among all patient groups, particularly involving neuroactive ligand receptor interaction and Ca2+ signaling. Patients with TB differed from those with community-acquired pneumonia, particularly with regard to the cytokine-cytokine receptor interaction pathway. The most commonly detected antibiotic resistance genes conferred resistance against β-lactams and macrolides. Antibiotic resistance genes were more abundant in oropharyngeal than in nasopharyngeal samples.
Conclusions:
The composition of microbial communities and host expression landscapes differs between TB and community-acquired pneumonia as well as between different respiratory tract sites. These changes in lung microbiota may impact lung disease development and prognosis.
Related Concept Videos
 01:29
Microbiota of the Respiratory Tract
 01:29
Pneumonia I: Introduction
 01:29
Pulmonary Tuberculosis I
Causative Organism
The primary infectious agent causing tuberculosis is Mycobacterium tuberculosis, a slow-growing, acid-fast, aerobic rod that exhibits sensitivity to heat and ultraviolet light. Instances of Mycobacterium bovis and Mycobacterium avium contributing to the development of TB infection are rare.
Mode of...
 01:26
Pulmonary Tuberculosis IV
Several diagnostic approaches are used to detect TB. The conventional method is the Tuberculin Skin Test (TST), also known as the Mantoux test. However, this method has...
 01:28
Pulmonary Tuberculosis II
Here is a detailed explanation of its pathophysiology:
Transmission: The process begins when a person inhales droplet nuclei containing M. tuberculosis. These are typically released into the air when an individual with pulmonary or...
 01:23
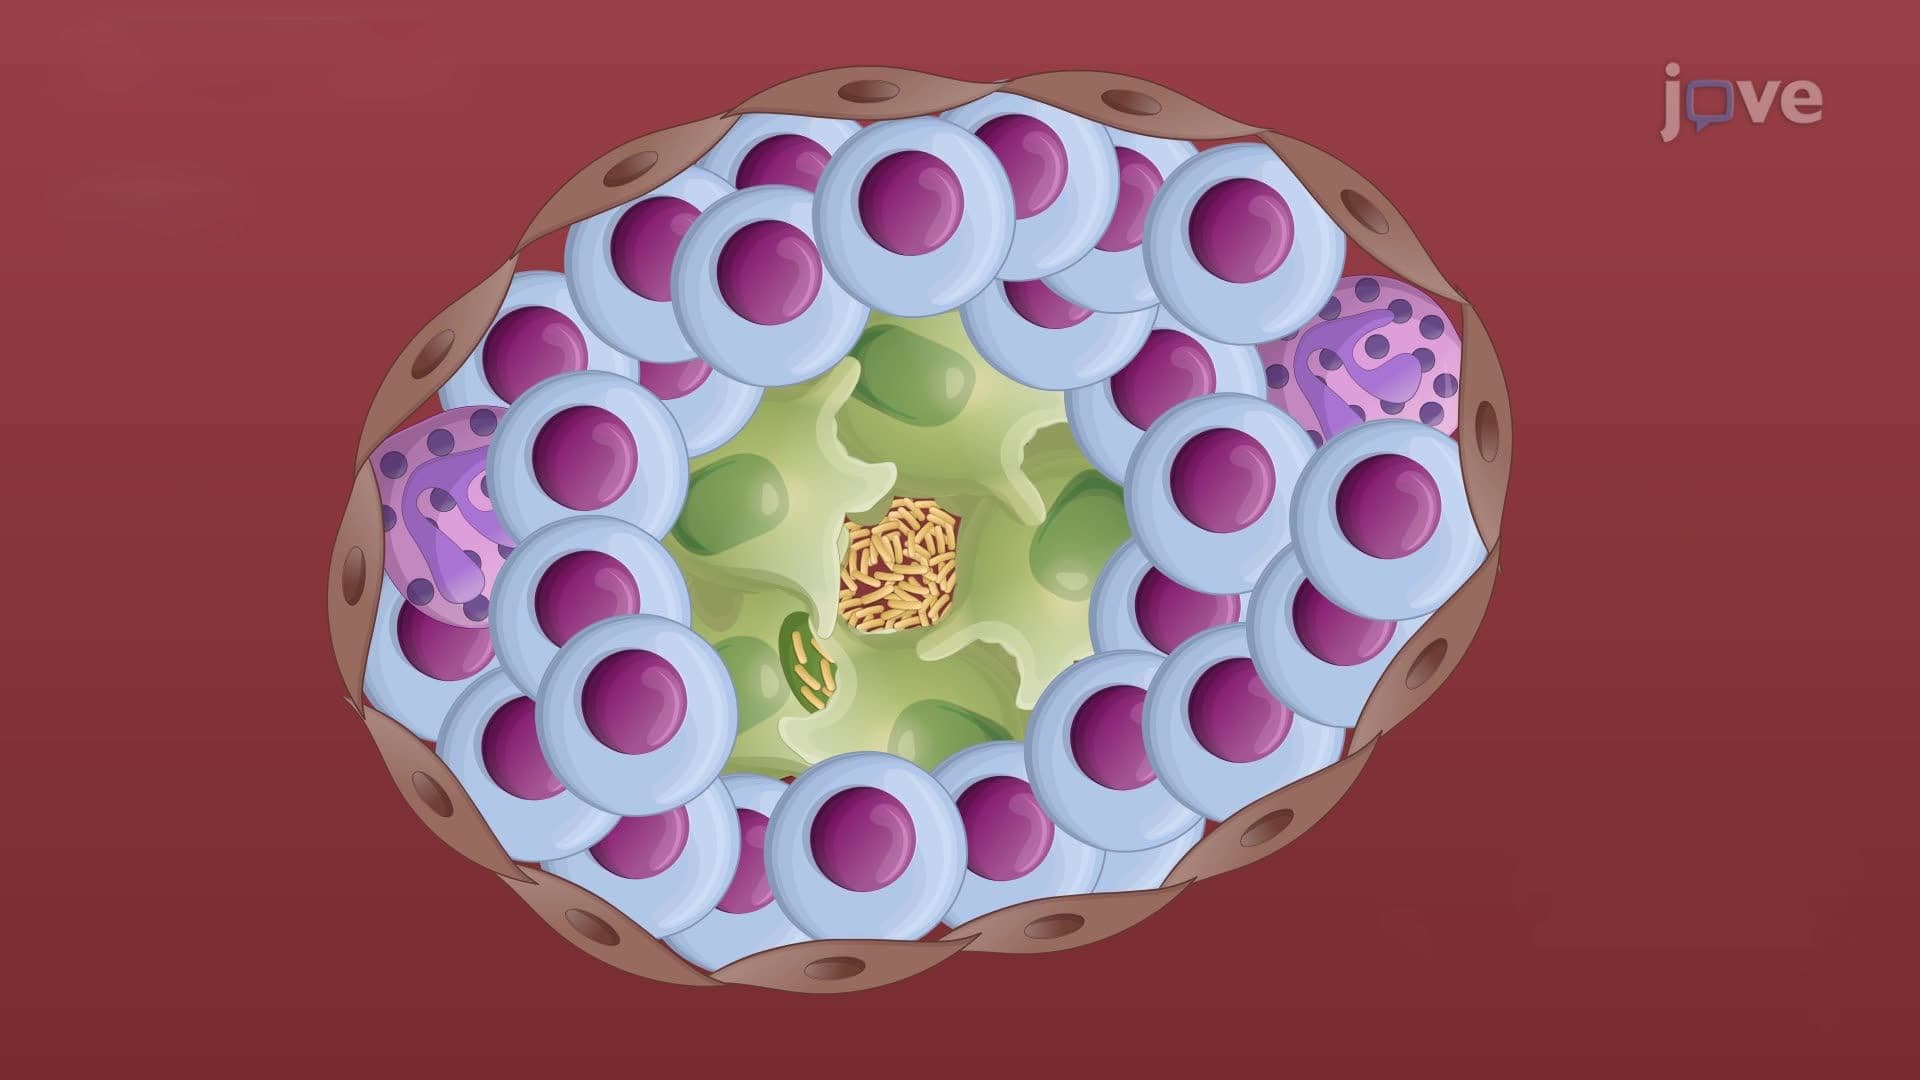
Tuberculosis