Related Experiment Video
Updated: Jun 11, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Adiabatic Connection Correlation Functionals in Metallic Solids from Hartree-Fock Gaussian Basis Set Ground State
Fabio Della Sala1,2, Fulvio Sarcinella1,2, Lucian A Constantin1
1Institute for Microelectronics and Microsystems (CNR-IMM), Via Monteroni, Campus Unisalento, 73100 Lecce, Italy.

Abstract:
Adiabatic Connection Integrand Interpolation (ACII) methods, which mix the Görling-Levy second-order perturbation theory (GL2) correlation with density functionals from the strong interaction regime, have been successfully applied to molecular systems, strongly correlated systems, as well as the uniform electron gas (UEG). Here, we consider real metallic solids, including transition metals, where the GL2 term diverges. We tested different ACII approaches, with increased accuracy in reproducing the UEG correlation, and with different strong interaction functionals within the Point-charge-plus-Continuum (PC) model. Calculations are based on the Hartree-Fock (HF) ground state, which has been obtained with a Gaussian Type Orbital basis set, properly reoptimized toward the HF global minimum, with a derivative-free approach introduced here. We compare different metallic properties such as lattice constants, cohesive energies, and bulk total correlation energies with accurate reference results and state-of-the-art Density Functional Theory (DFT) approaches. We found that only the ACII approach with high accuracy for the UEG, namely, the Density Parameter Interpolation (DPI), and with the strong interaction functional reproducing the exact PC model for the Wigner crystal yields a correct second-order gradient-expansion correlation coefficient. This allows reproducing lattice constants and bulk correlation energies with an accuracy comparable to the best state-of-the-art approaches. When DPI is combined with GL2 correlation of atoms, accurate cohesive energies are obtained without error cancellation, which is common in DFT methods.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:28
Debye–Huckel–Onsager Conductance Equation
 01:17
Theory of Metallic Conduction
In this theory, Newton's second law of motion is used to determine the acceleration of an electron in the presence of an applied electric field. Then, its velocity is expressed via this acceleration.
An electron moves through the crystal, containing positive ions,...
 02:32
Bonding in Metals
 01:10
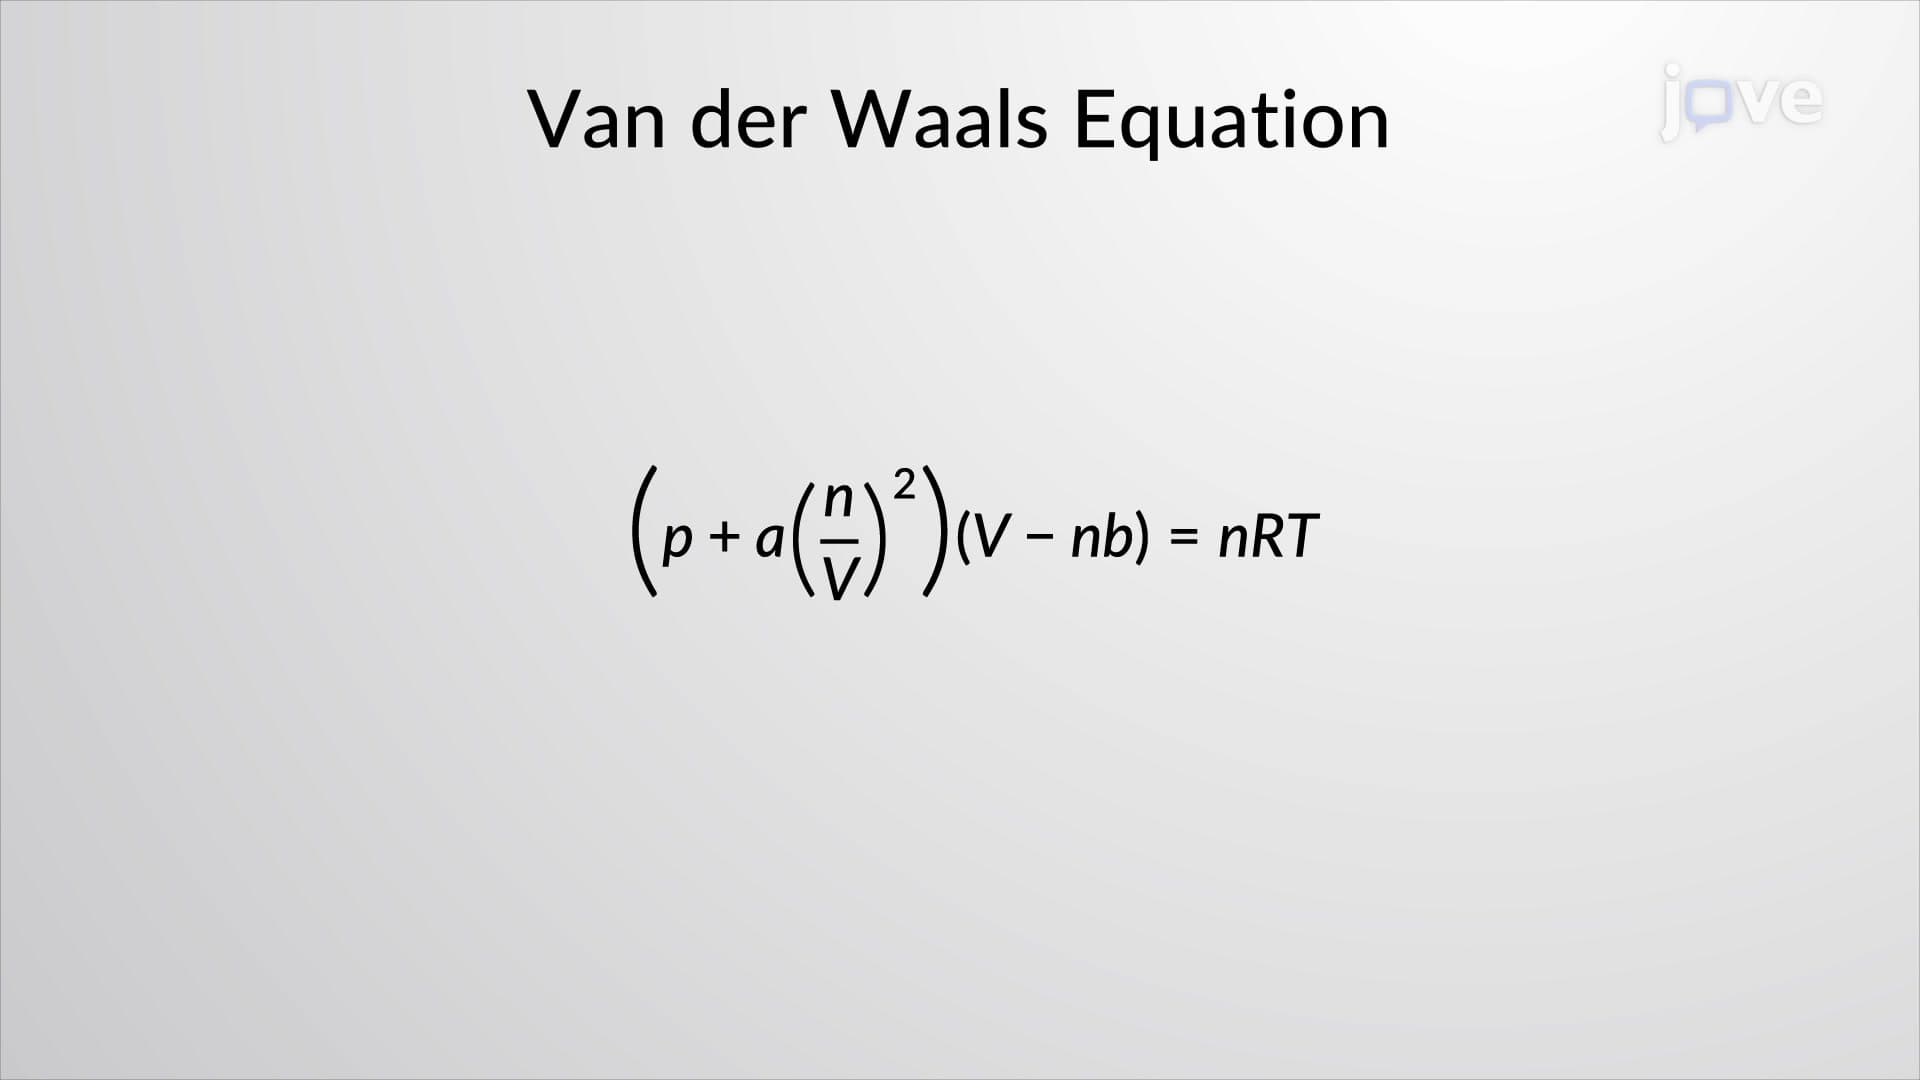
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory