Related Experiment Video
Updated: Jun 17, 2026

Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Electric field-induced bond length and bandgap tunability for enhanced reactivity of nitride nanorings
Riddhi Sainda1, Prafulla K Jha1
1Department of Physics, Faculty of Science, The Maharaja Sayajirao University of Baroda, Vadodara, Gujarat 390002, India. prafullaj@yahoo.com.

Abstract:
Electric-field engineering offers an efficient and reversible route to modulate the properties of low-dimensional nanomaterials. In this work, we present a comprehensive density functional theory (DFT) investigation of the two nitride nanorings, i.e., Al9N9 and B9N9, under an external electric field applied perpendicular to the ring plane. The influence of the electric field on the structural, electronic, and reactivity-related properties is systematically analyzed. Structural parameters reveal gradual bond elongation, angular distortion, and ring contraction with increasing field strength, indicating enhanced polarization while maintaining overall structural stability. The electronic structure exhibits a clear Stark effect, characterized by a pronounced downward shift of the LUMO and a smooth reduction of the HOMO-LUMO bandgap, confirming tunable semiconducting behavior. A steady increase in dipole moment and a significant shift in the Fermi level demonstrate strong field-induced charge redistribution. Global conceptual DFT descriptors, including chemical potential, hardness, softness, electrophilicity, nucleophilicity, and work function, reveal a controlled decrease in electronic hardness and a gradual enhancement of reactivity under the applied field. The linear and systematic evolution of these descriptors suggests that Al9N9 operates within a stable Stark regime without electronic instability. Overall, this study establishes the nitride nanoring as a robust electric-field-tunable inorganic semiconductor, highlighting its potential for applications in nanoelectronics, sensing, and field-controlled molecular devices.
Related Concept Videos
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states that no two...
 01:22
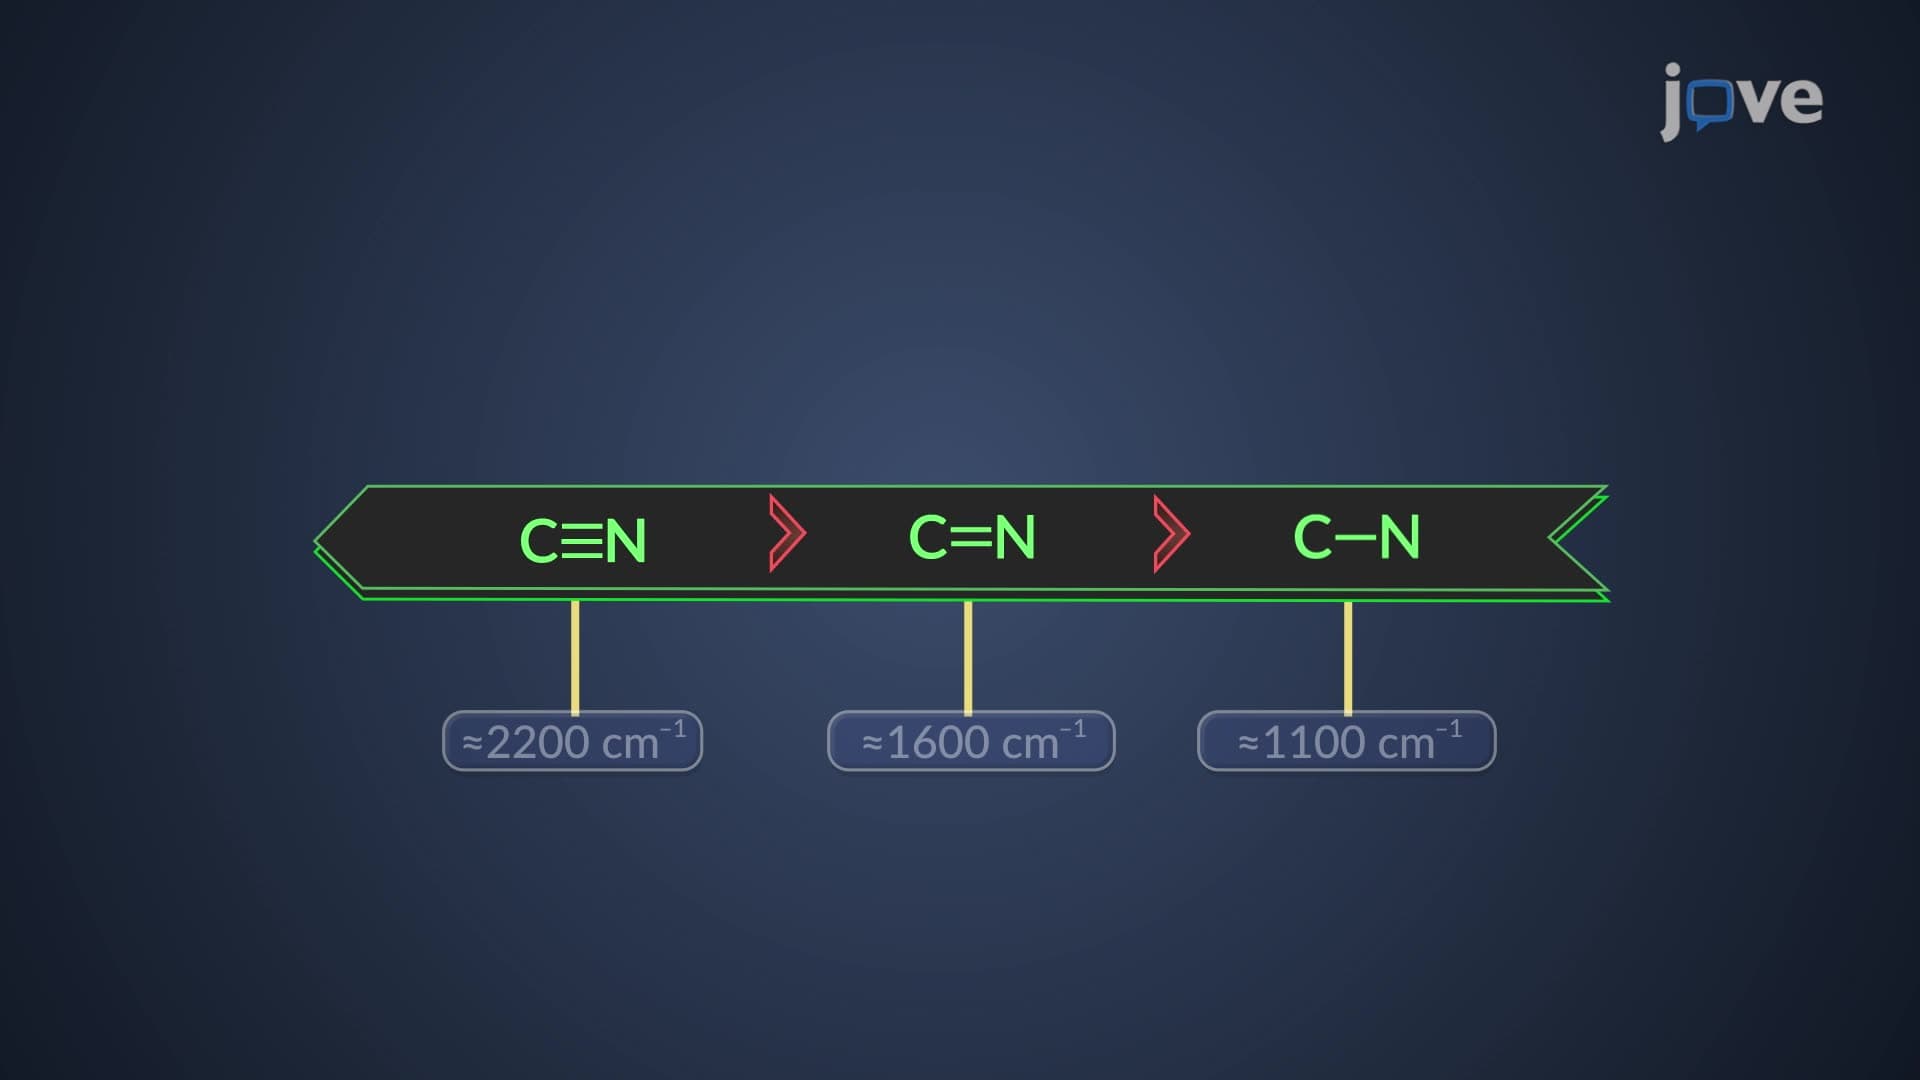
IR Frequency Region: Alkyne and Nitrile Stretching
Comparing the stretching vibrational frequency of C≡C triple bonds with that of double and single bonds, it is evident that C≡C triple bonds exhibit a higher stretching frequency than C=C double and C–C single bonds. Similarly, the C≡N triple bond exhibits higher stretching absorption than the C=N...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:27
π Electron Effects on Chemical Shift: Overview
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...