Related Experiment Video
Updated: Jun 17, 2026
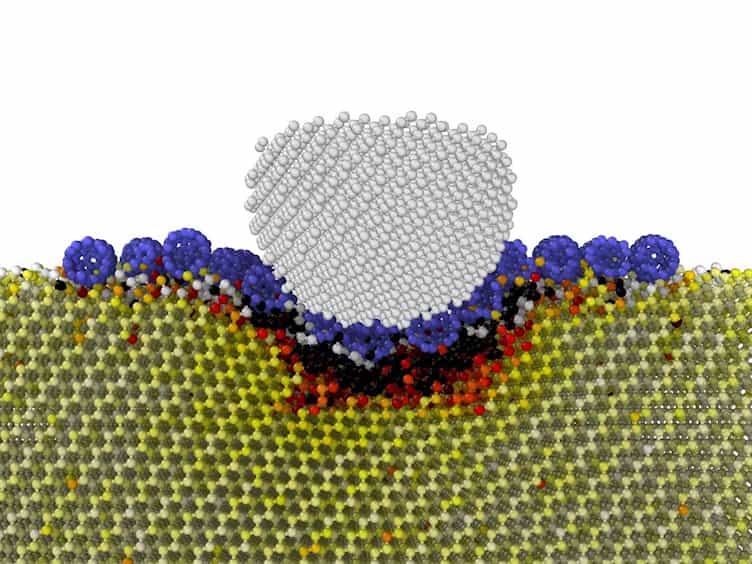
Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
A supramolecular C60S8 adduct: dynamic and interaction energy considerations of an orbiting S8 motif from MD + EDA
Margot Paco-Chipana1, Alvaro Muñoz-Castro2
1Doctorado en Biología Computacional, Facultad de Ingeniería, Universidad San Sebastián, Bellavista 7, Santiago, 8420524, Chile.

Abstract:
Dynamics of supramolecular complexes serve to rationalize the constant motion inherent to the formation of non-covalent adducts. Here, the C60S8 adduct is evaluated through the combined molecular dynamics energy decomposition analysis approach (MD + EDA), unraveling the constant orbit-like motion of S8 around the C60 fullerene globe, showing comparable supramolecular interaction energies for the entire dynamic process, arising from a balance between stabilizing (electrostatic, London dispersion, orbital) and destabilizing (Pauli repulsion) forces. Thus, molecular dynamics simulations reveal the dynamic flexibility, with the S8 ring freely reorienting in relation to the C60 surface, remaining at a similar distance with similar interaction energy. The charge transfer analysis indicates negligible net electron transfer, but interestingly, the calculated electronic coupling and charge hopping rates suggest efficient charge transport. These findings highlight the delicate interplay of weak non-covalent interactions governing the formation of the C60-S8 system and its potential for supramolecular charge transport applications, combining dynamic adaptability with specific binding properties for applications in supramolecular chemistry and related fields.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:44
Measurements of Long-range Electronic Correlations During Femtosecond Diffraction Experiments Performed on Nanocrystals of Buckminsterfullerene
Published on: August 22, 2017
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:35
Molecular Orbital Theory I
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:18
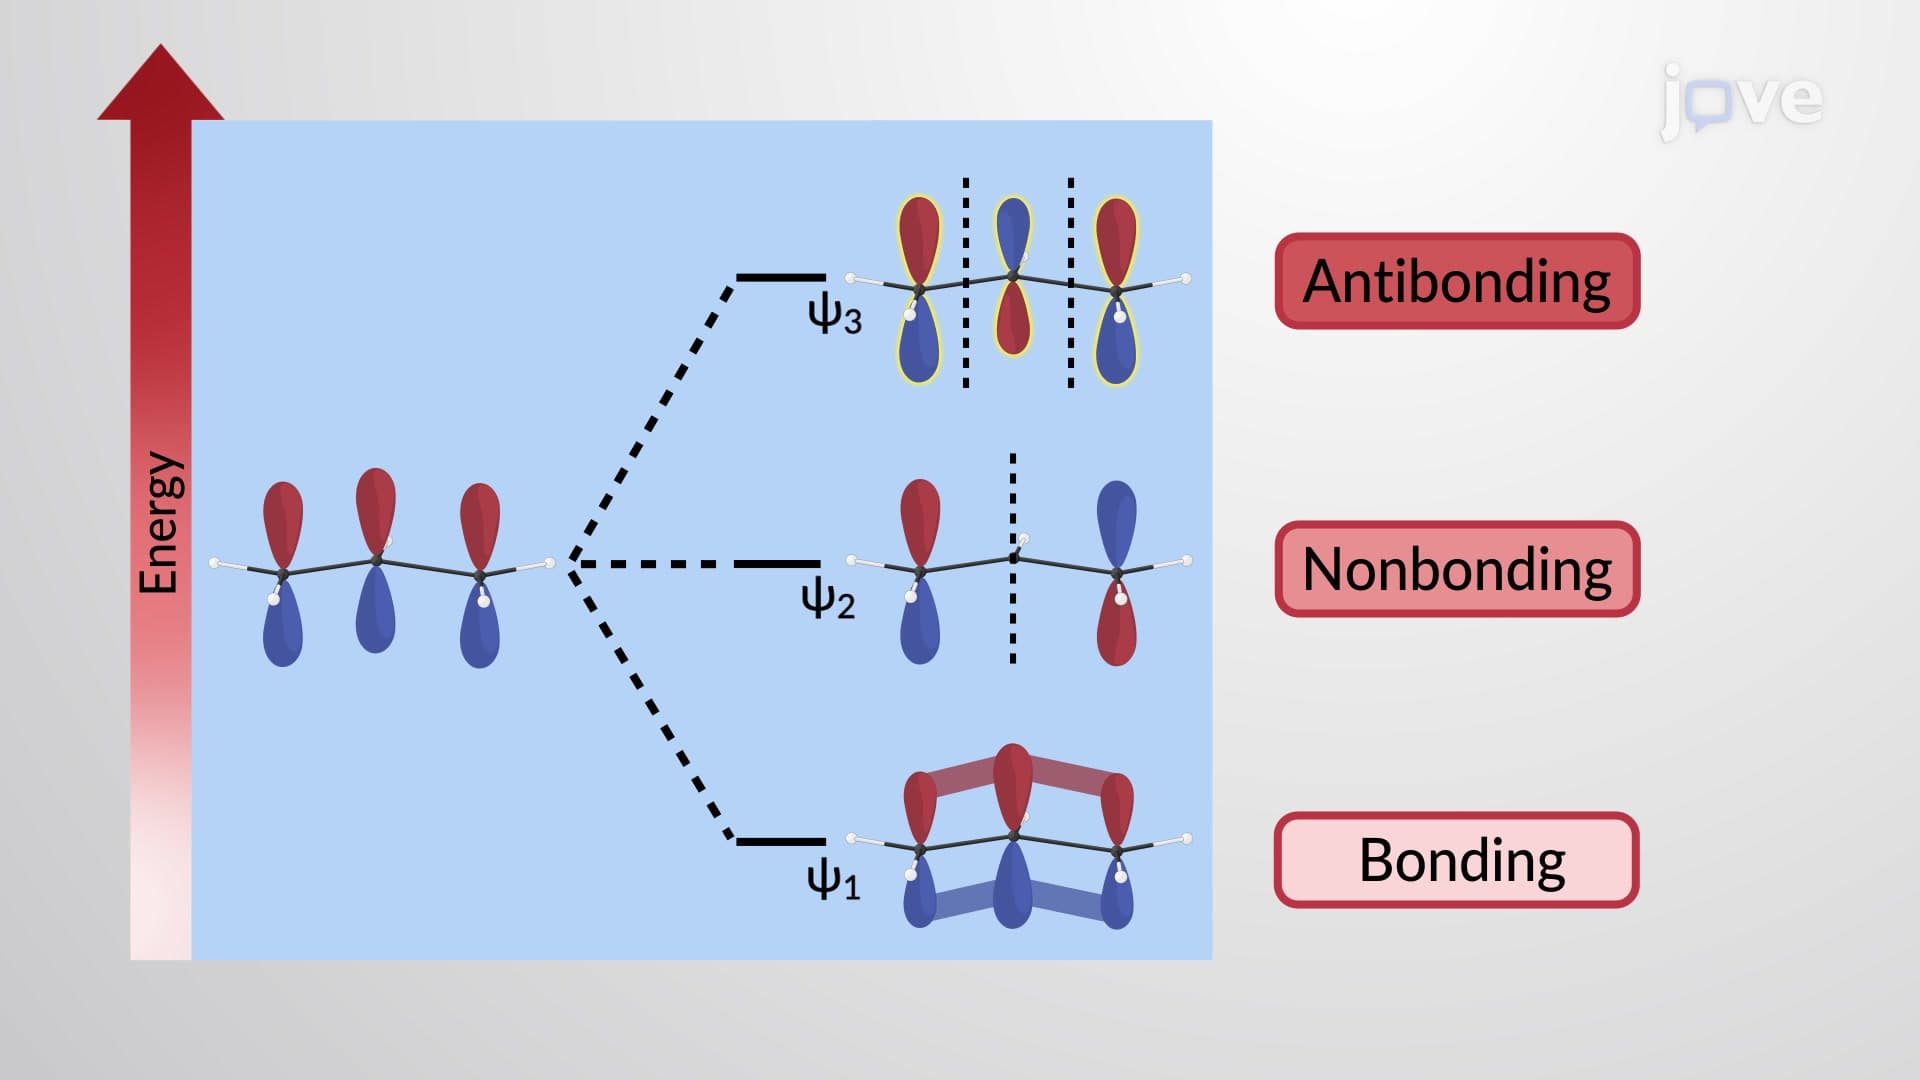
π Molecular Orbitals of the Allyl Cation and Anion
 02:40
MO Theory and Covalent Bonding