Related Experiment Video For O-GlcNAcylation
Updated: Jun 23, 2026
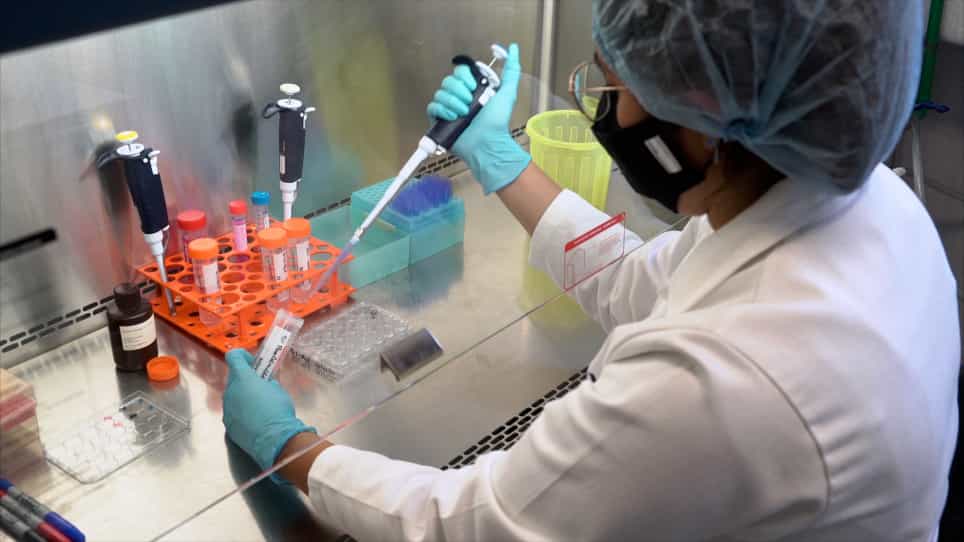

A Model of Experimental Steatosis In Vitro: Hepatocyte Cell Culture in Lipid Overload-Conditioned Medium
Published on: May 18, 2021
Cholesterol Overload Drives Hepatic Steatosis by Inhibiting OGT-dependent PPARα O-GlcNAcylation and Transactivation
Rui Guo1,2, Yanhui Li1, Qinchao Ding2
1Department of Kinesiology and Nutrition, University of Illinois Chicago, Chicago, IL, USA.

Abstract:
Although dietary cholesterol is known to exacerbate liver disease progression, whether and how it contributes to hepatic steatosis, the hallmark early pathological feature of both MASLD and ALD, remains poorly understood. Here, we investigated how cholesterol disrupts hepatic triacylglycerol metabolism using both dietary and cellular cholesterol-loading models. Integrated transcriptomic, metabolomic, and biochemical analyses were performed, and causality was examined through genetic and pharmacologic modulation in multiple hepatocyte systems and mice. Our results demonstrate that cholesterol overload induces hepatocellular fat accumulation in a dose-dependent, cell-autonomous manner, primarily by suppressing fatty acid β-oxidation. Mechanistically, we identified PPARα inhibition as a key event underlying this effect. Cholesterol overload suppressed PPARα transactivation, thereby impairing fatty acid β-oxidation and promoting hepatocellular fat accumulation. This inhibition was mechanistically linked to reduced O-GlcNAcylation. Specifically, cholesterol overload downregulated OGT, leading to reduced protein O-GlcNAcylation and consequent PPARα inhibition; similarly, liver-specific OGT knockout mice exhibited suppressed PPARα activity and increased hepatic fat accumulation. RNA-sequencing and co-immunoprecipitation analyses identified PPARα as an O-GlcNAc-modified protein, and loss of this modification impaired its transactivity. Functionally, restoration of O-GlcNAcylation via genetic OGA knockdown or pharmacological activation of PPARα with WY14643 alleviated cholesterol-induced hepatic steatosis in mice without altering hepatic cholesterol levels. Lastly, we identified SREBP2 as the upstream transcriptional regulator linking cholesterol overload to OGT suppression. In conclusion, our findings in this study uncover a previously unrecognized cholesterol-OGT-PPARα axis that suppresses hepatic fatty acid β-oxidation and drives steatosis. Targeting O-GlcNAc cycling or activating PPARα represents a promising therapeutic strategy for MASLD.
Related Concept Videos
 01:29
Cholesterol: Significance and Regulation
Considering cholesterol and...
 01:20
Lipid-Lowering Drugs: Statins and Miscellaneous Agents
 01:21
Chronic Pancreatitis II: Pathophysiology
 01:09
GPCRs Regulate Adenylyl Cylase Activity
Two...
 01:20
Cholecystitis
 01:16
Hepatic Drug Excretion: Influencing Factors