Related Experiment Videos
Parameter Efficient Deep Learning Models for Multi-Target Binding Affinity and hERG Cardiotoxicity Prediction

Abstract:
Accurately predicting binding affinity and toxicity is critical to drug discovery, offering the potential to reduce development costs and enhance safety profiles. Traditional computational methods, such as molecular docking and quantitative structure-activity relationship (QSAR) models, rely heavily on handcrafted features and domain expertise, limiting their scalability and generalizability to novel compounds. Recent advances in chemical language models (CLMs), which leverage Simplified Molecular Input Line Entry System (SMILES) representations, offer a scalable alternative by capturing complex molecular properties through learned representations. In this study, we propose a dual-paradigm framework for binding affinity prediction of 3 protein targets and cardiotoxicity for human ether-á-go-go related gene (hERG). The first component utilizes a specialized graph neural network (GNN) that operates directly on molecular graphs, treating atoms as nodes and bonds as edges. The second component employs a CLM fine-tuned via Low-Rank Adaptation (LoRA), enabling efficient adaptation of large pre-trained models to specific toxicological tasks with minimal parameter overhead. Our hybrid framework demonstrates superior performance, achieving an average AUROC of 0.92 across three protein targets, while the LoRA-adapted CLM attains an AUROC of 0.93 for cardiotoxicity prediction, offering a 98% reduction in trainable parameters, outperforming the performance of existing models.
Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:25
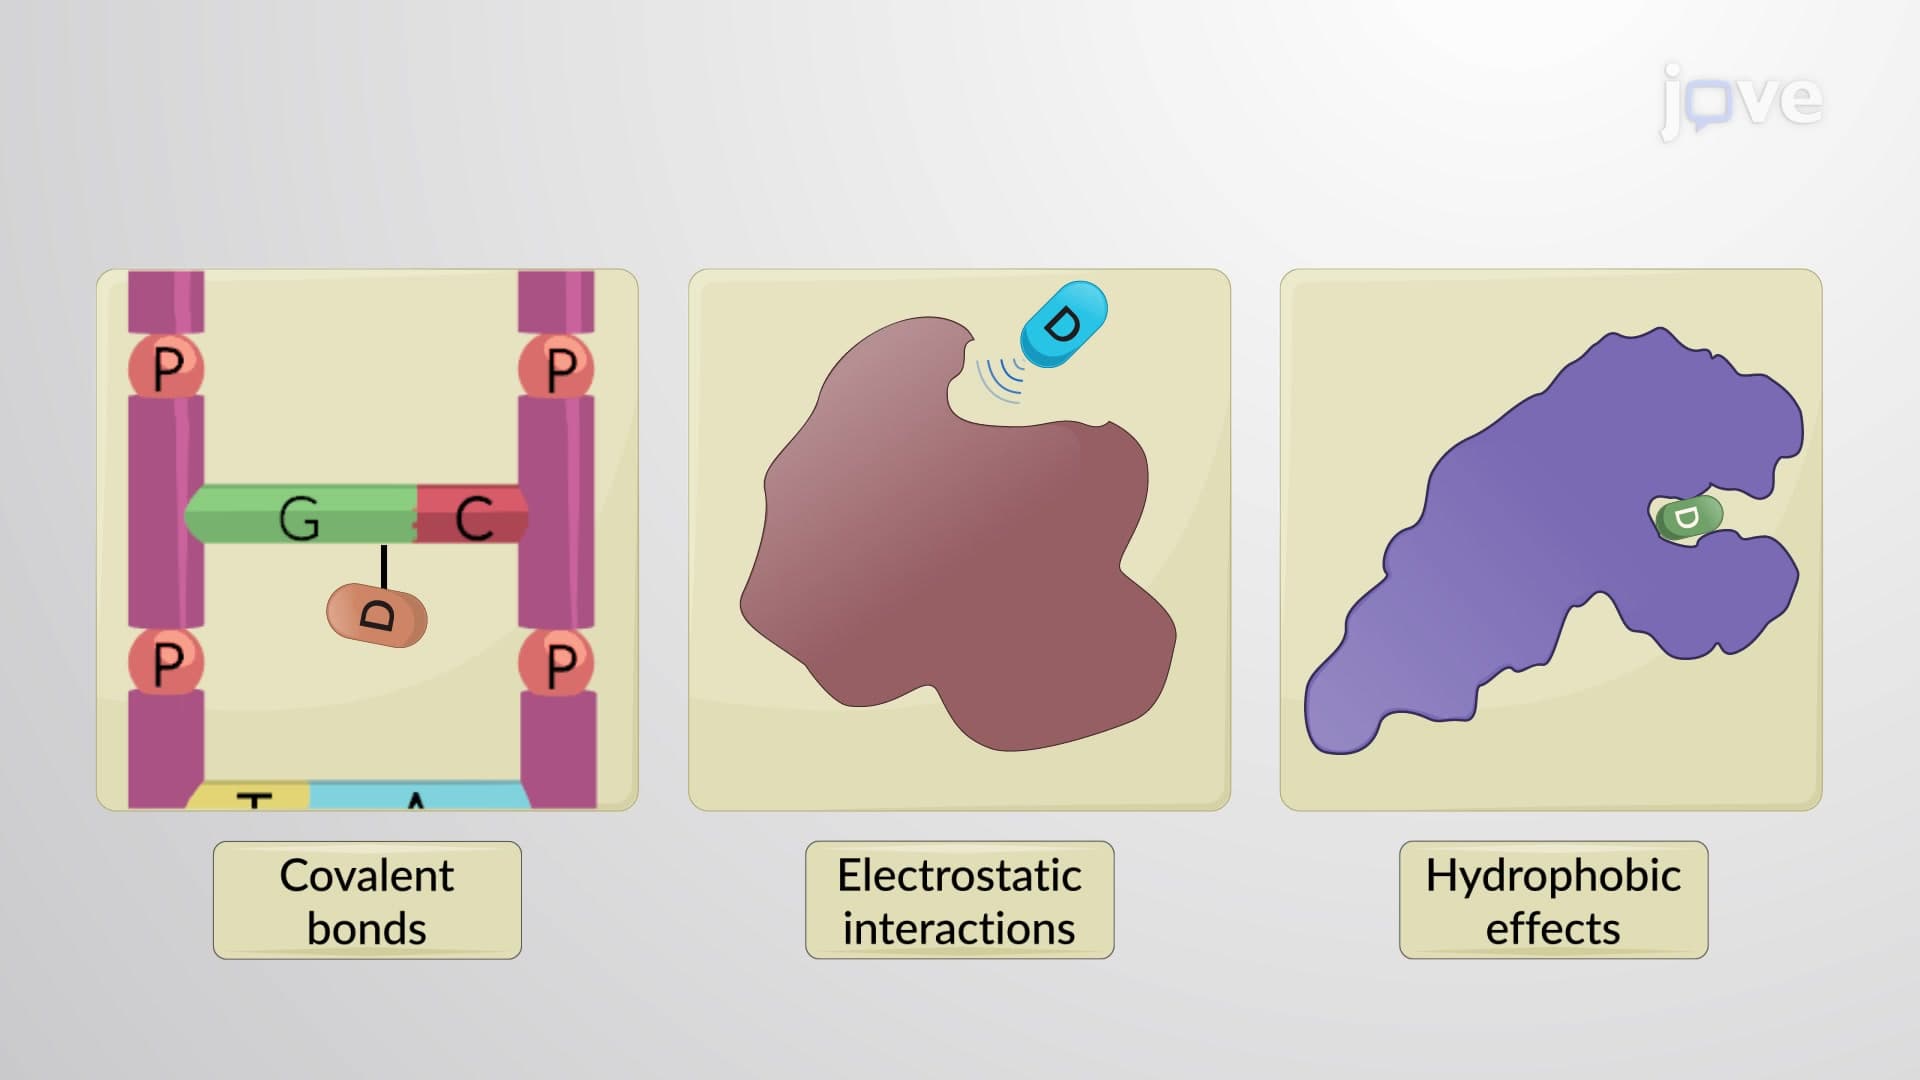
Drug-Receptor Bonds
In...
 01:15
Impact of Pharmacokinetic–Pharmacodynamic Models: Regulatory Decisions
 00:49
Ligand Binding and Linkage