Related Experiment Video
Updated: Jun 26, 2026
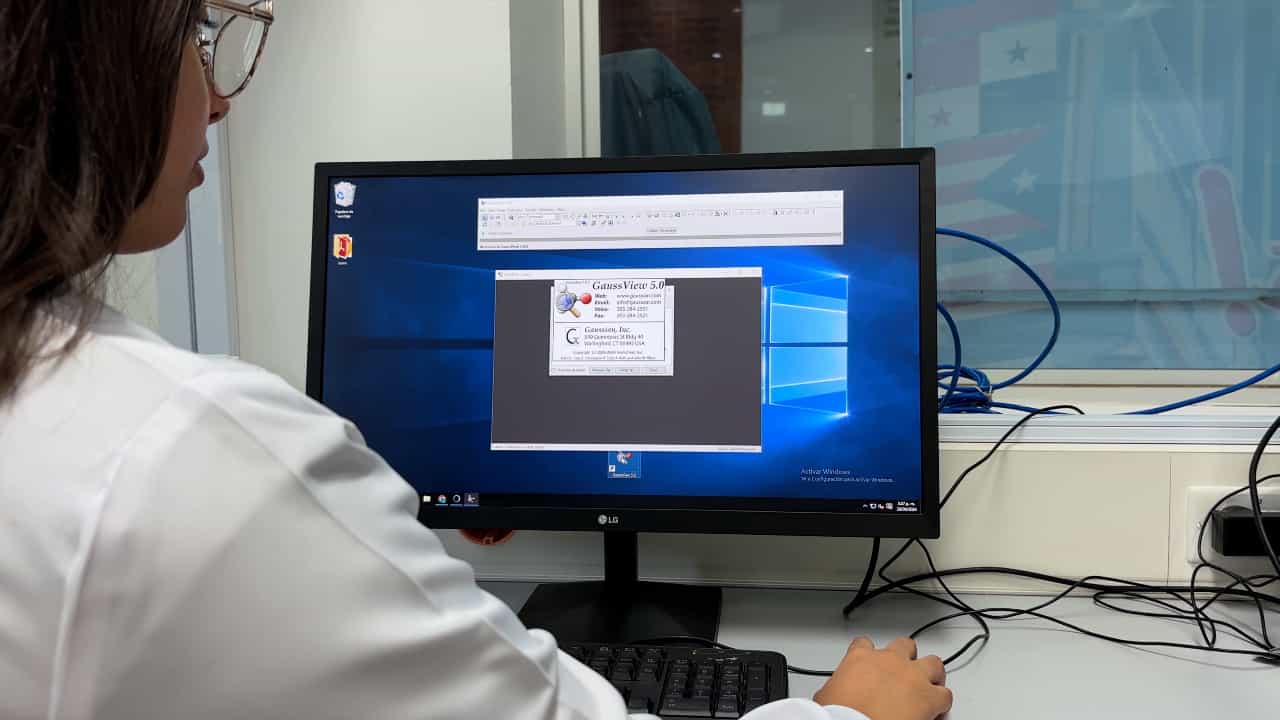

Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
GPU Accelerated Minimal Auxiliary Basis Approach TDDFT for Large Organic Molecules
Zehao Zhou1, Xiaojie Wu2, Yanheng Li3,4
1Zhongguancun Academy, Beijing 100094, China.

Abstract:
We introduce a GPU-accelerated implementation of time-dependent density functional theory with the minimal auxiliary basis approach (TDDFT-risp) in GPU4PySCF, together with large system demonstrations carried out using the Tamm-Dancoff approximation (TDA-risp). The method combines GPU-accelerated three-center integral evaluation, tensor contractions, exchange-space truncation, omission of hydrogen atoms from the auxiliary basis, and a host memory assisted Davidson solver. On a benchmark test set of 42 molecules (13-99 atoms), a conservative 40 eV exchange cutoff yields excitation-energy errors relative to standard TDA of about 0.03-0.05 eV for low-lying states. For systems of 300 to 3000 atoms, we demonstrate that TDA-risp calculations of 15 low-lying excited states with ωB97XD/def2-SVP complete on a single A100 GPU with wall times ranging from minutes to hours. Relative to RIJCOSX TDDFT in ORCA (32 MPI processes), GPU TDA-risp yields wall-time speedups of roughly 140-340× on systems with 72-480 atoms, using ωB97X-D3BJ/def2-TZVP. These results position GPU-TDDFT-risp as a practical route toward excited-state calculations for large organic and biomolecular systems with thousands of atoms.
More Related Videos


Related Concept Videos
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:29
Reaction Mechanisms: Rate-limiting Step Approximation
 01:14
Crossed Aldol Reaction Using Weak Bases
 03:35
Hybridization of Atomic Orbitals II
 02:27
Predicting Molecular Geometry
 01:26
Spectroscopy of Carboxylic Acid Derivatives
In the...