Related Experiment Video
Updated: Jun 26, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
The deformation energy gap in computational drug design: why interaction energy alone cannot rank drug candidates and
Iqra Malik1, Muhammad Javid Iqbal2
1Department of Pharmaceutical Chemistry, Faculty of Pharmacy, Bahauddin Zakariya University, Multan, Pakistan.

Context:
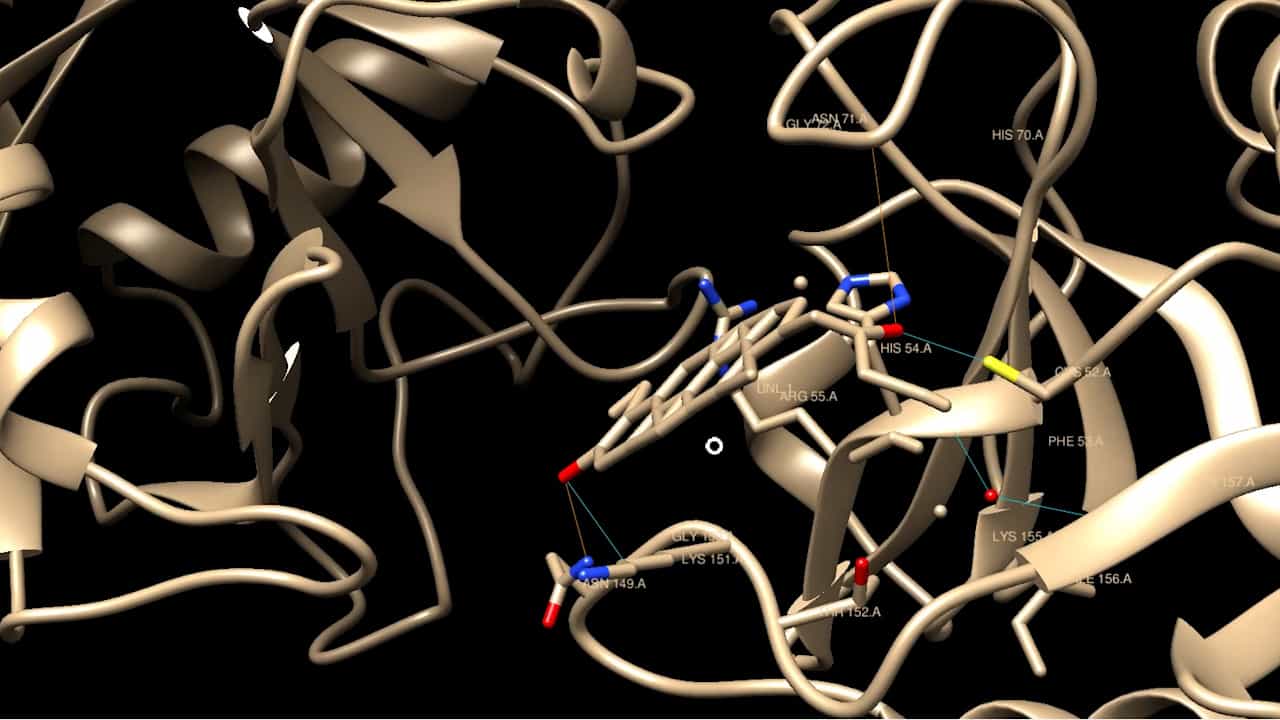
DFT calculations are increasingly combined with molecular docking to rank drug candidates, yet most studies report the interaction energy (ΔEint), computed at the complex geometry, as a surrogate for binding affinity. This quantity omits the deformation energy (ΔEdef): the thermodynamic penalty of distorting both partners from their free-state geometries into their bound conformations. Because ΔEdef is always positive (typically 2-20 kcal mol-1) and molecule-dependent, its omission systematically overestimates binding strength and can reverse predicted rank-orderings. We present the energetic decomposition ΔEbind = ΔEint + ΔEdef, demonstrate using published crystallographic strain data from over 3,000 protein-ligand complexes that deformation energies do not cancel between structurally distinct ligands, and propose a minimal five-step correction protocol applicable to any DFT-based drug design study. The protocol requires only two additional geometry optimizations beyond the standard workflow, adding only modest additional computational cost. This work does not introduce new computational data; it highlights an energetic inconsistency in common computational practice and provides a straightforward correction to enable more consistent electronic binding energy evaluation and improved candidate comparison. METHODS: The analysis is based on the supramolecular energy decomposition framework and the activation strain model (ASM), in which binding energy is partitioned into interaction and deformation (strain) components using standard variational principles. No new DFT calculations are reported. The argument draws on published conformational strain datasets obtained at various DFT levels and molecular mechanics force fields from crystallographic analyses of the PDBBind database. The proposed correction protocol is general and can be applied with any DFT functional, basis set, and quantum chemistry software package (e.g., Gaussian, ORCA, or equivalent).
Related Concept Videos
 01:25
Drug-Receptor Bonds
In...
 01:13
Force and Potential Energy in One Dimension
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:20
Strain-Energy Density
In the elastic region of a material, the relationship between the stress and the strain is linear and follows Hooke's Law. The strain energy density in this region...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 02:49
Calculating Standard Free Energy Changes