Related Experiment Video
Updated: Jun 30, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Theoretical Investigation of Optoelectronic and Photophysical Properties in the Carbazole-based Molecules via DFT and
Muhammad Zohaib Sabir1,2, Sania Ismaeel3, Yasir Ali4
1Department of chemistry, Faculty of Science King, Mongkut's Institute of Technology Ladkrabang, Bangkok, 10520, Thailand. 67056115@kmitl.ac.th.

Abstract:
The growing need for renewable energy resources has brought small-molecule organic solar cells into focus as one of the potential candidates for future generation photovoltaics owing to their easy manufacturing process, flexibility, and lightweight nature. This paper presents the theoretical investigation carried out through density functional theory (DFT) and time-dependent DFT (TDDFT) methods for designing and characterizing a series of five novel donor materials (CA1-CA5) based on carbazole core, thiophene spacer, and various functionalized terminal electron-withdrawing acceptors. Analysis of the results obtained from frontier molecular orbitals suggests that functionalization of the terminal acceptor can modulate the electronic properties, such as the energy level and band gap of these molecules. Remarkably, CA2 and CA1 possess lower band gaps (2.71 eV and 2.81 eV, respectively) than the reference material (3.48 eV) and are therefore expected to have higher charge transfer efficiency. Strong intramolecular charge transfer (ICT) is observed in all molecules using transition density matrix and molecular electrostatic potentials. Additionally, CA1 and CA2 are identified to have high oscillator strength and absorption at the longest wavelength, implying high light-harvesting ability.
More Related Videos
 11:44
Using Cyclic Voltammetry, UV-Vis-NIR, and EPR Spectroelectrochemistry to Analyze Organic Compounds
Published on: October 18, 2018
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:29
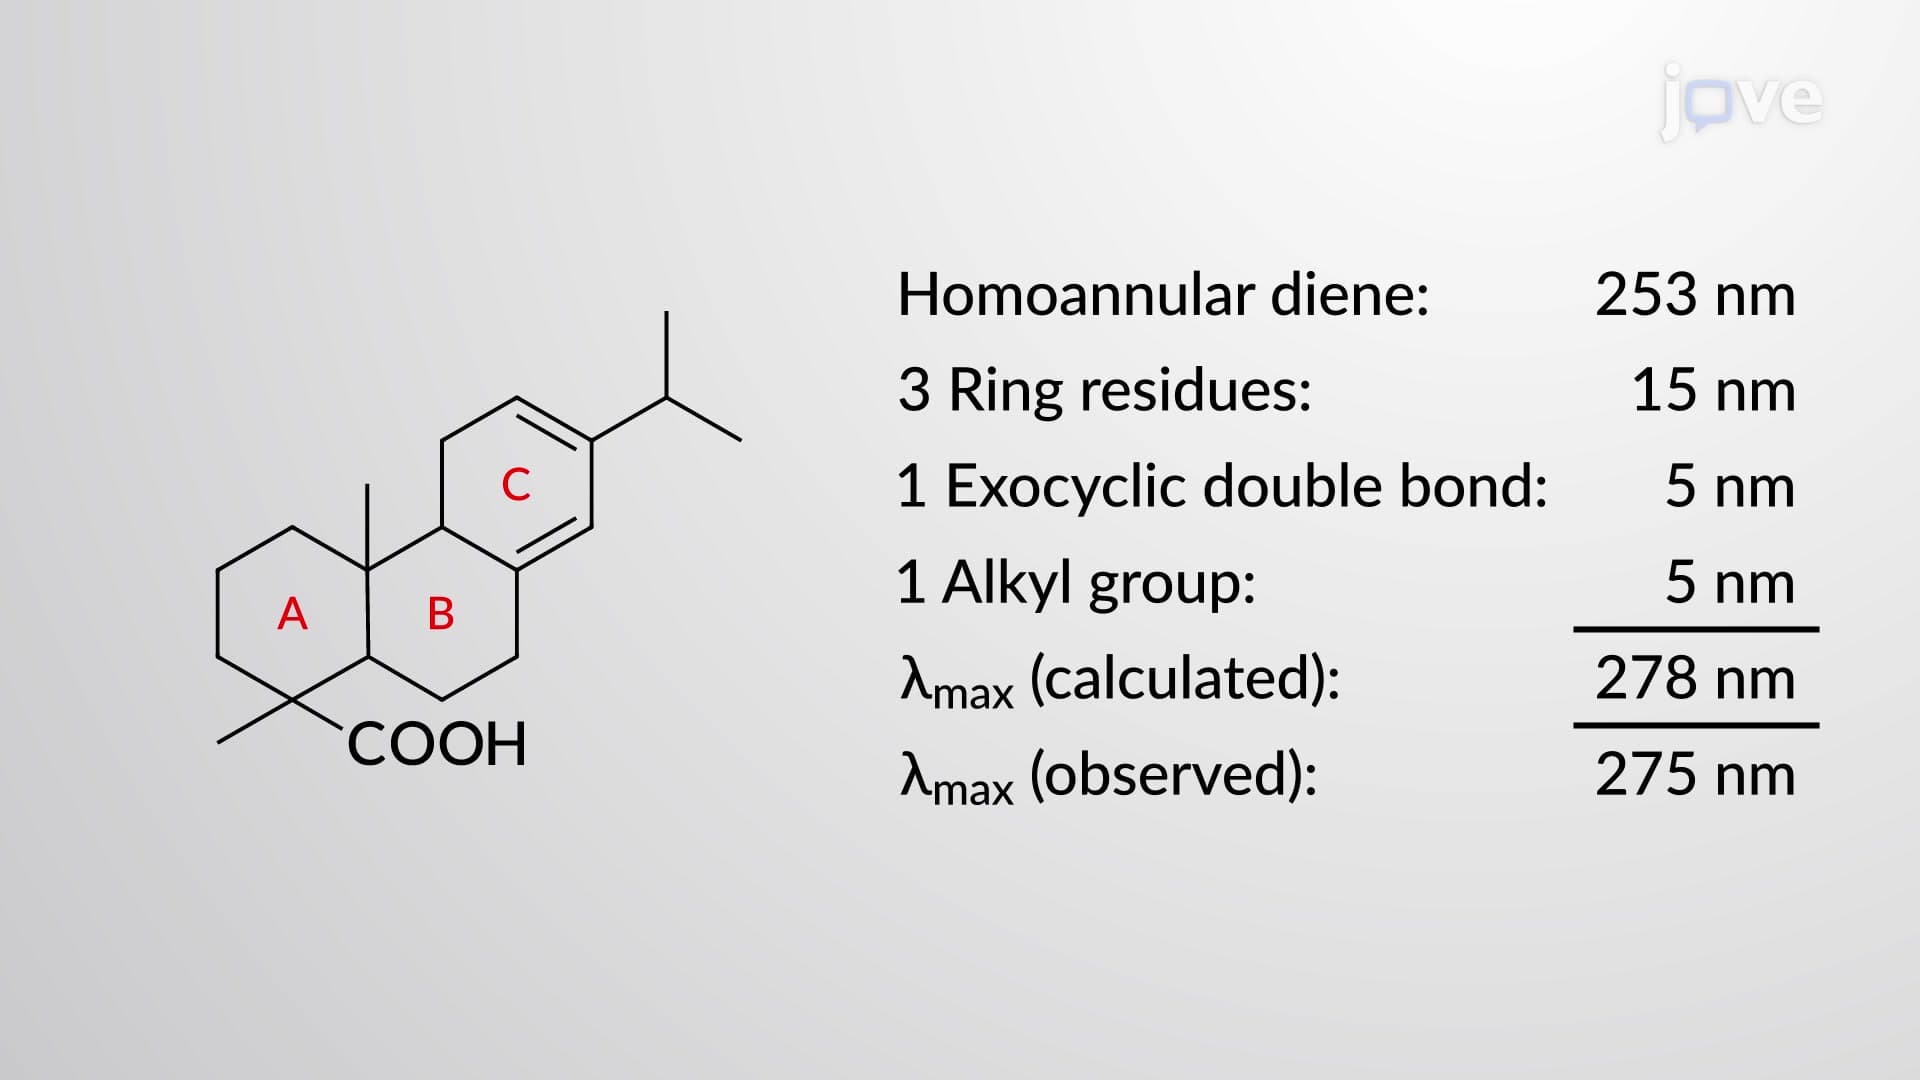
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:26
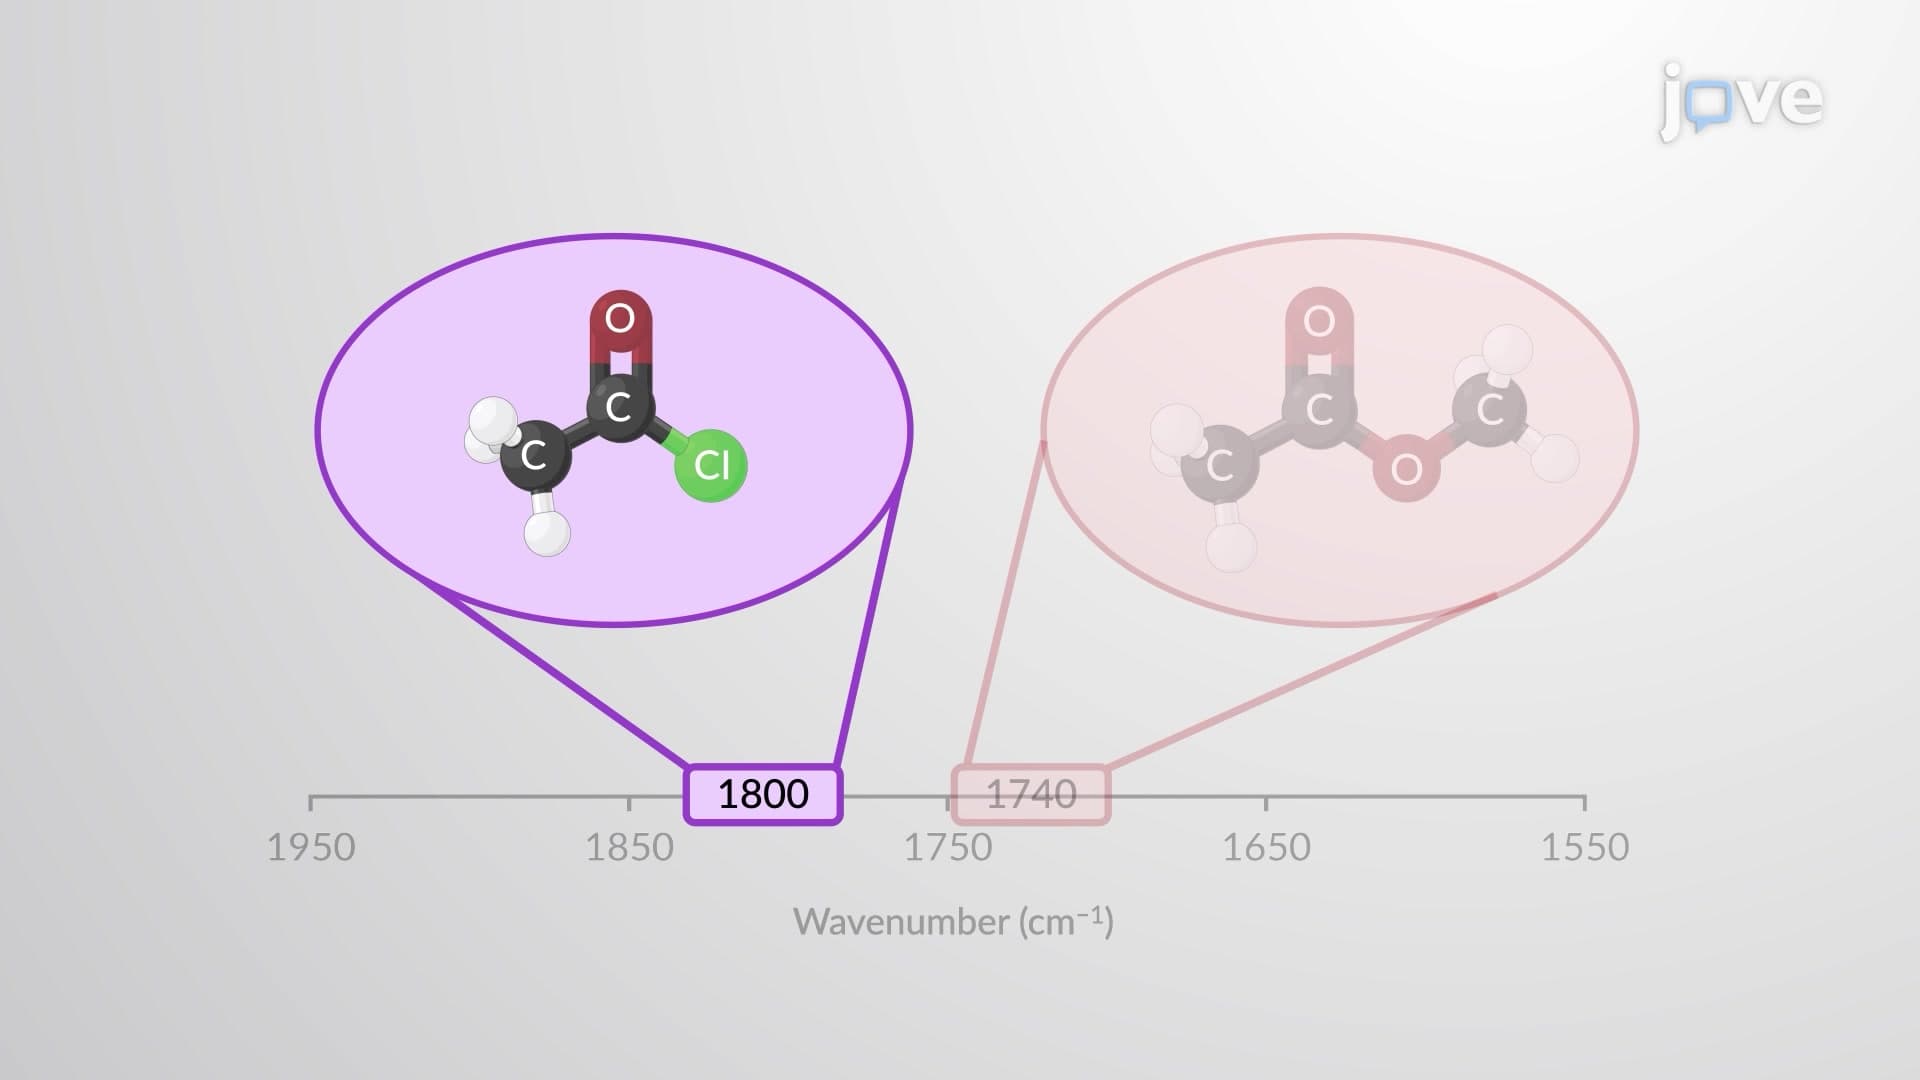
Spectroscopy of Carboxylic Acid Derivatives
In the...
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in the...
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:28
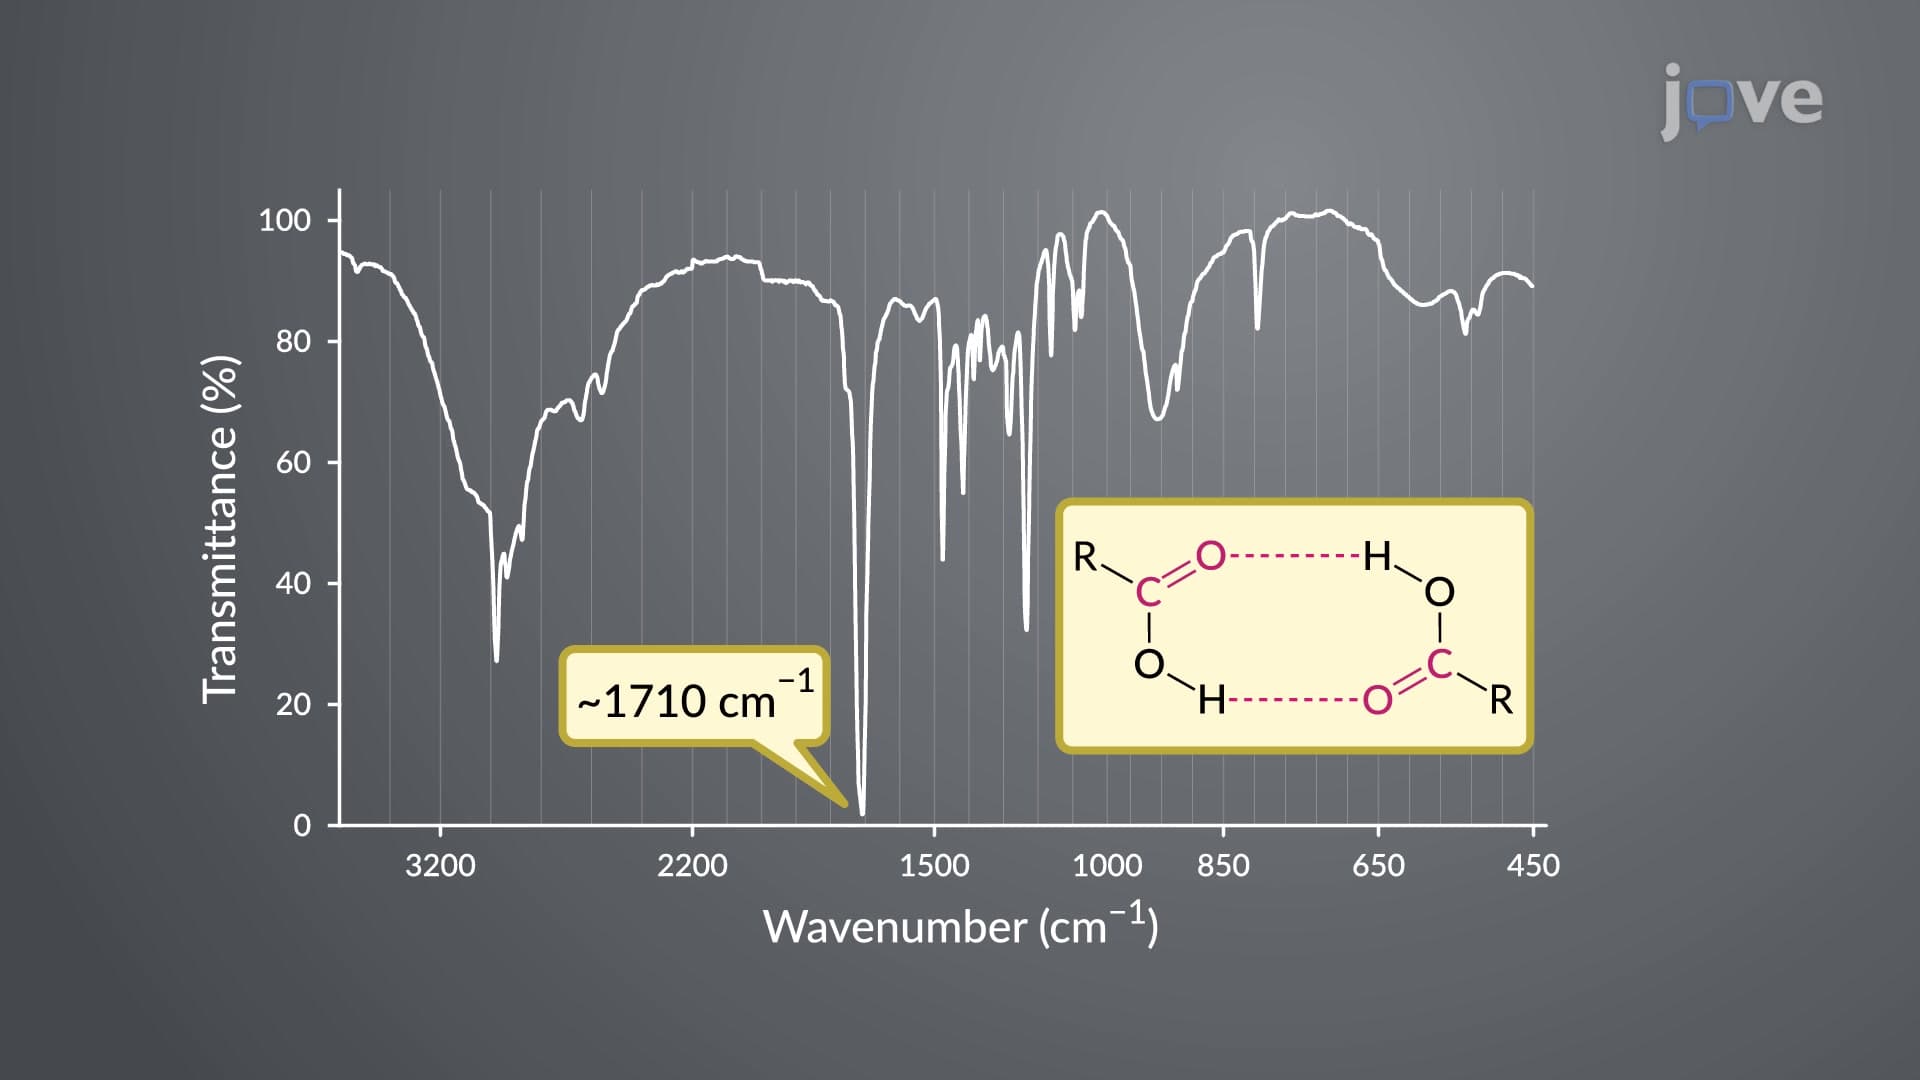
IR and UV–Vis Spectroscopy of Carboxylic Acids
However, the stretching absorptions for the C=O bond vary depending on the structure of carboxylic acids. The C=O bond of the free carboxylic acids shows a higher stretching frequency, 1760 cm−1, while H-bonded carboxylic acids (dimers) exhibit stretching absorptions at a lower frequency, 1710 cm−1. The C=O bond of the...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions