Related Experiment Video
Updated: Jul 5, 2026


Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Bridging local-global transmembrane protein contexts with contrastive pretraining for alignment-free pathogenicity
Yihang Bao1,2,3, Zhe Liu4, Fangyi Zhao1
1Shanghai Mental Health Center, Shanghai Jiao Tong University School of Medicine, School of Biomedical Engineering, Shanghai Jiao Tong University, 1954 Huashan Road, Xuhui District, Shanghai 200030, China.

Abstract:
Predicting the pathogenic consequences of protein mutations is a cornerstone of precision medicine, yet it remains a formidable challenge for transmembrane proteins (TMPs), a clinically vital class of drug targets. Existing computational methods are often hampered by their reliance on evolutionary data and fail to model TMP-specific biophysical constraints. Here, we introduce Memo-Patho, a deep learning framework for robust, alignment-free pathogenicity prediction of TMP variants. The core innovation is a within-protein, label-informed supervised contrastive pretraining strategy that learns sequence-encoded biophysical signatures distinguishing pathogenic and benign variants by directly comparing them within the same protein context. By fusing sequence-level representations from protein language models with local structural proxies derived from sequence, Memo-Patho achieves accurate predictions without multiple sequence alignments or experimental structures. Across diverse TMP benchmarks and under protein-level group splits, Memo-Patho consistently outperforms leading predictors, achieving up to 0.93 accuracy, and it transfers to an independent KCNQ1 ion-channel cohort without re-training. Its resource-efficient, alignment-free design enables routine large-scale screening when evolutionary or structural data are sparse. Conceptually, Memo-Patho addresses a key gap by directly learning discriminative, sequence-anchored signatures pertinent to TMP-specific constraints, offering a principled and generalizable foundation for research-use clinical variant triage and proteome-wide mutation-effect modeling.
More Related Videos

Related Concept Videos
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 02:07
Improving Translational Accuracy
02:07Improving Translational Accuracy
02:26Conservation of Protein Domains
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 01:09
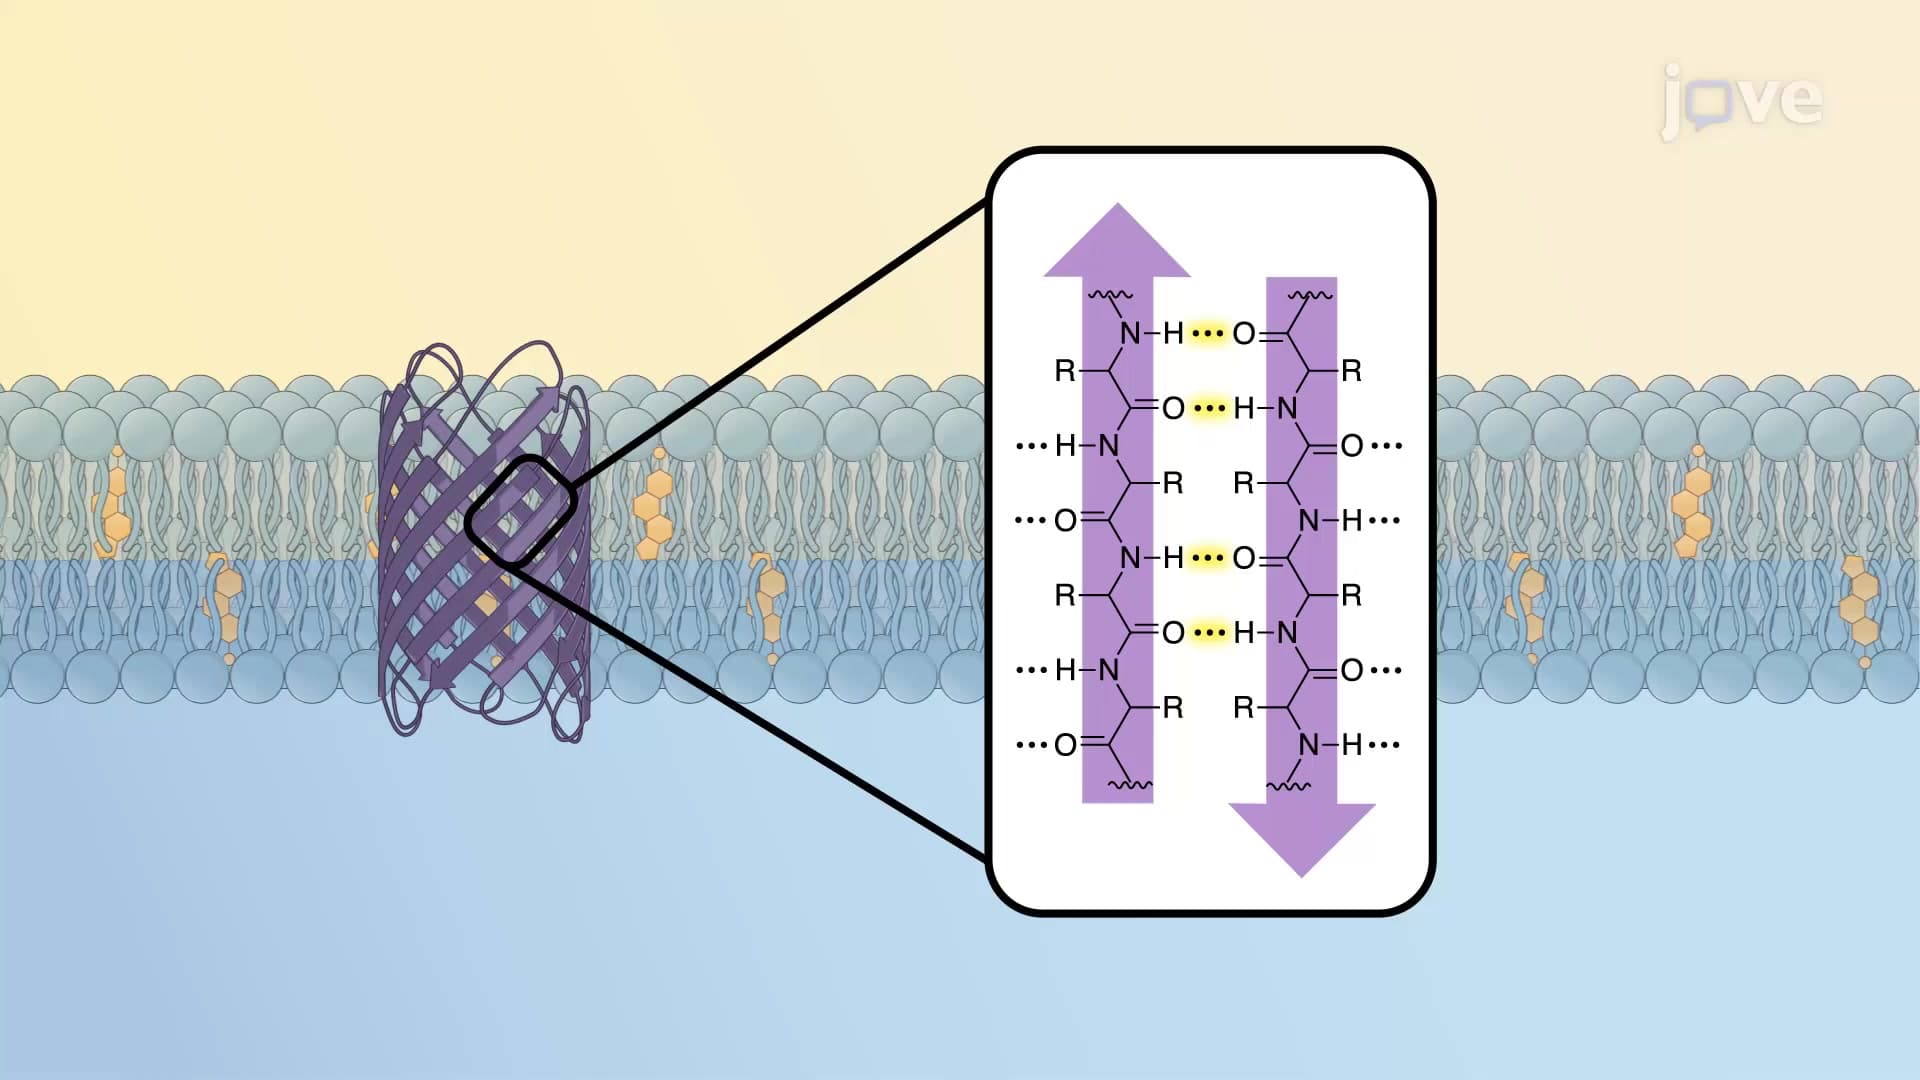
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as G-protein-linked receptors (GPCRs) and...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...