Related Experiment Video
Updated: Jul 8, 2026

Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
Resolving Conformational Preferences of Monosaccharides from 1H and 13C NMR Chemical Shifts Using an Integrated MD
Wojciech Plazinski1, Göran Widmalm2
1Jerzy Haber Institute of Catalysis and Surface Chemistry, Polish Academy of Sciences, 30-239Krakow, Poland.

Abstract:
Solution-state NMR spectroscopy is a powerful experimental technique that provides insight into the molecular structure and dynamics of saccharides in aqueous solution. Computational tools commonly used for modeling carbohydrate conformations, such as molecular dynamics (MD) simulations and quantum mechanical (QM) calculations, provide information on the inherent dynamics of conformational changes and structure-dependent NMR parameters, respectively; however, understanding how NMR parameters depend on dynamic conformational behavior remains challenging. Herein, we present an integrated MD and QM approach to accurately determine NMR parameters, in particular 1H and 13C NMR chemical shifts of saccharides. Classical MD simulations are used to sample conformational space, and representative structures are subsequently subjected to QM calculations to obtain NMR parameters, which are averaged according to populations in different conformational states. This approach reproduces experimental NMR chemical shifts with high accuracy (MAE = 0.96 ppm for 13C and 0.066 ppm for 1H relative to experimental data) across 11 monosaccharide entities. In parallel, we establish a quantitative relationship between conformational properties of monosaccharides and the chemical shift values. In this context, empirical relationships between chemical shifts and torsional angles enable mapping of structural descriptors onto NMR observables. Furthermore, we demonstrate that the use of conformation-dependent chemical shifts allows quantitative description of conformational equilibria within monosaccharide molecules. Average chemical shift values assigned to discrete conformers (e.g., ring conformers or hydroxymethyl rotamers) and to atoms in the vicinity of torsion angle transitions are analyzed; populations in distinct conformational states are optimized by minimizing deviations from experimental NMR chemical shifts. This approach enables determination of gt:gg:tg populations of hydroxymethyl group rotamers in β-d-Glcp-OMe, α-d-Manp-OMe and α-d-Galp-OMe, as well as the chair:inverted chair ratio for the β-d-Arap-OMe six-atom membered ring. Overall, this study establishes a framework in which NMR chemical shifts serve as quantitative probes of carbohydrate conformation, complementing traditional J coupling-based analyses.
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:20
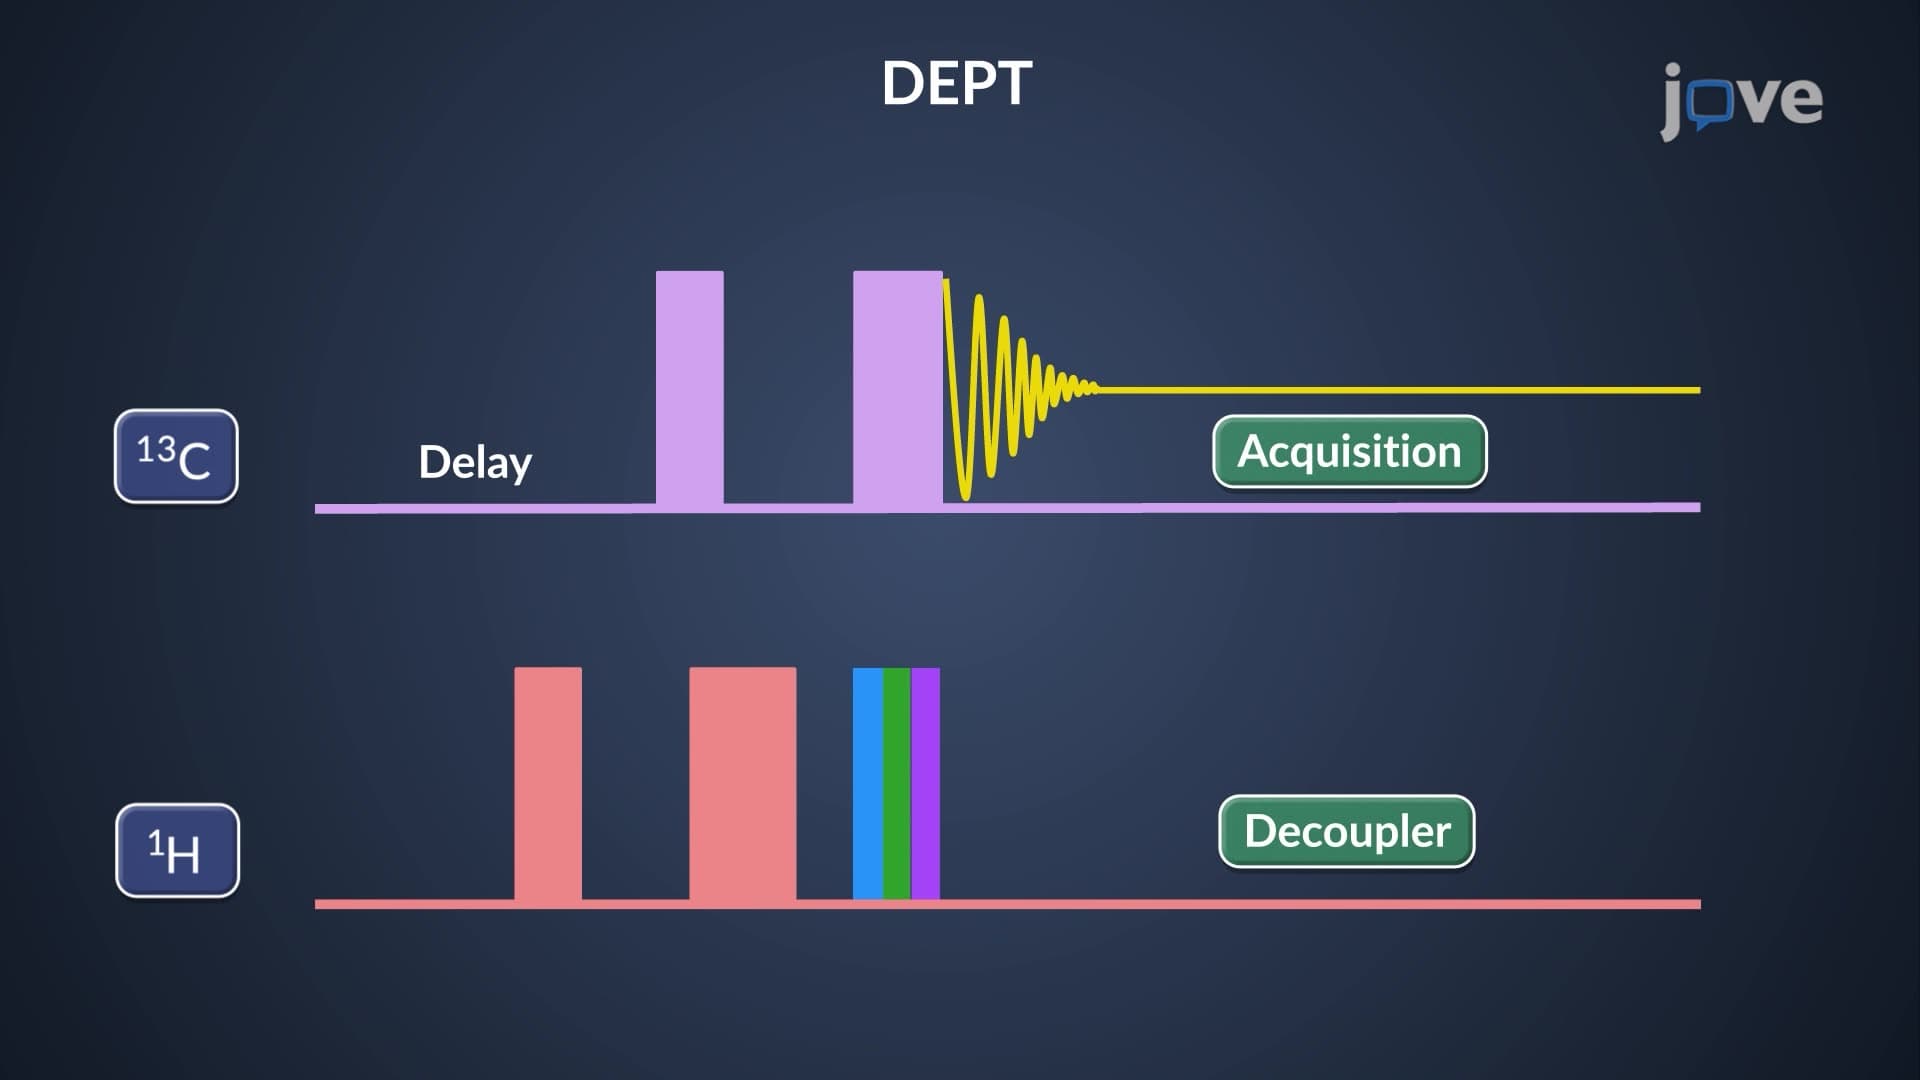
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:19
2D NMR: Heteronuclear Single-Quantum Correlation Spectroscopy (HSQC)
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:10
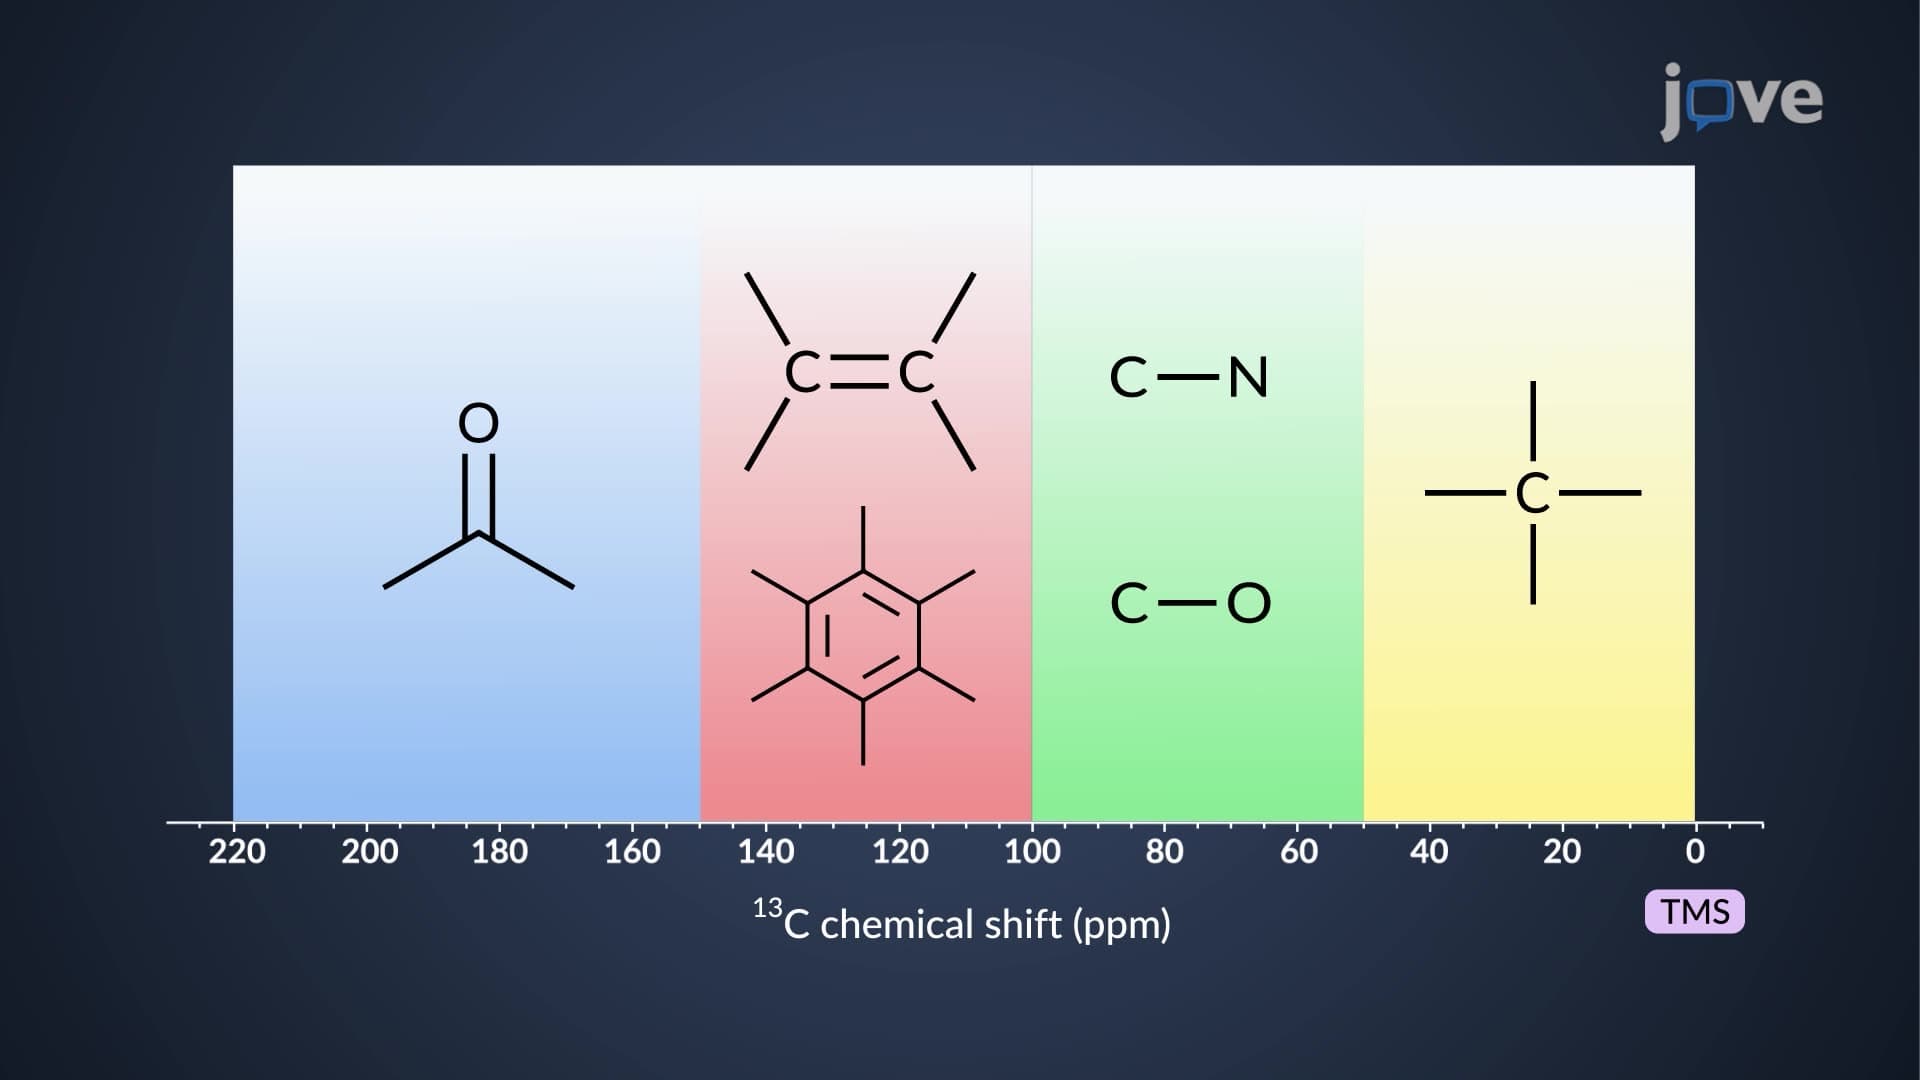
Carbon-13 (¹³C) NMR: Overview