Related Experiment Video
Updated: Jul 15, 2026

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico
Elham Omer Mahgoub1, Syed Asif Husain2
1College of Arts and Sciences, Qatar University; ilhamomer@yahoo.com.

Abstract:
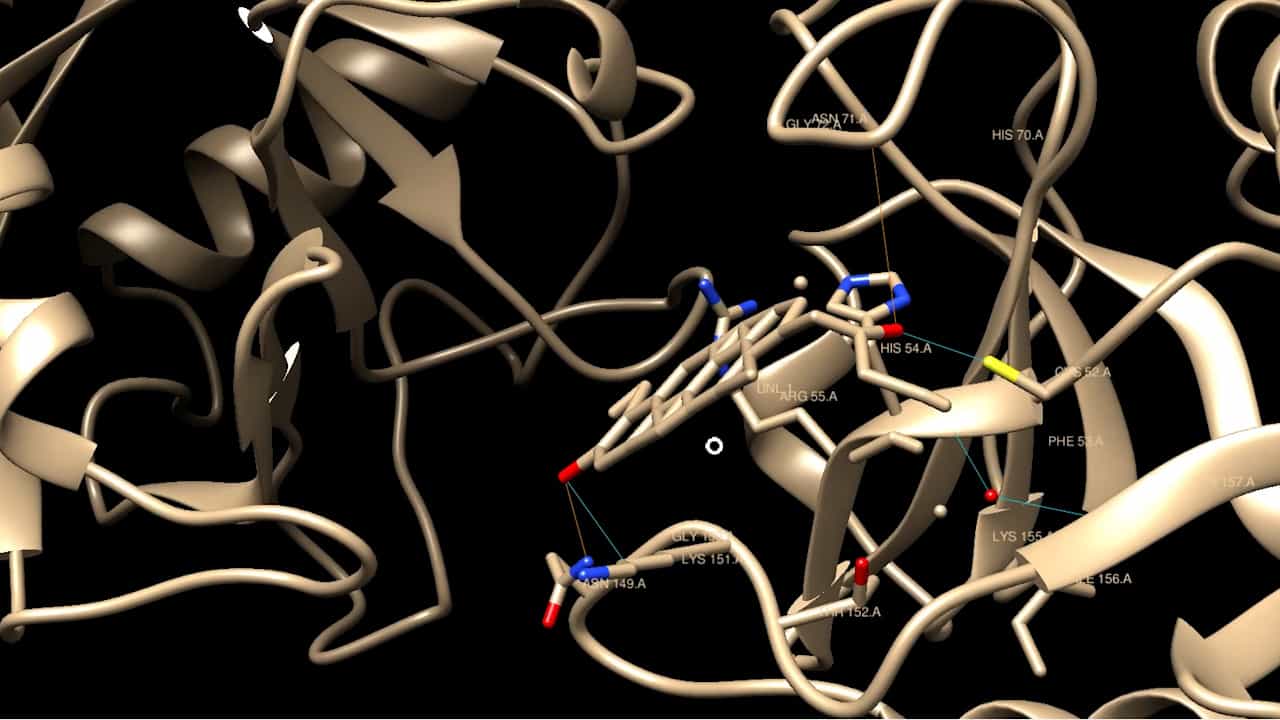
The cyclin Ds-CDKs axis (Cyclophilin A (Cyp /Ganoderiol F)) plays a critical role in cancer through various processes, such as controlling proliferation and inhibiting cancer cells. Ganoderiol F derivatives can serve as oncogenic targets for multiple cancer types. This study demonstrated that the structure-guided design and development of novel Cyp A inhibitors and Ganoderiol F derivatives can be employed to select calculated positions for increased effectiveness in chemical inhibitors and their interaction with the chosen receptors. For this approach, numerous software applications were utilized to achieve optimal molecular interactions. Overall, 117 novel cyclophilin inhibitor ligand molecules were constructed using Chemsketch software. The converted PDB files of the novel Cyp A inhibitor ligands were prepared for introduction to the Sanglifehrin A enzyme receptor. Ligand-protein docking interactions were performed using AutoDock 4.2.6 software to reveal the best geometric interactions. The docking confirmations were carried out using PyMOL 2.5 software. Finally, hydrogen bonds were detected using Chimera and Maestro software. The selected ligand-receptor docked complex was subjected to molecular dynamics simulation. The MD simulation process demonstrated the stability of the complex and indicated that the chosen ligand can be utilized as a drug inhibitor for cancer cells. In the results, the active amino acids in binding site were identified as Arginine 55 that in motion of 1.53Å, Asparagine (Asn), Glycine (Gly), Threonine (Thr), Lysine (Lys), Isoleucine (Iso), Histidine (Hid), Phenylalanine (Phe), and Cysteine (Cys) with the highest number of hydrogen bond ligand of 45 hydroxymangiferonic. That connected with 1nmk receptor in grid box coordinate of X 38.86, Y of 10.987, and Z of 40.734. The 1nmk receptor-ligand complex evaluation in molecular dynamics (MD) simulation in Gromacs version CHARMM 36 force field has optimum degrees.
Insights
Novel cyclophilin A inhibitors and Ganoderiol F derivatives were designed and tested as potential cancer drug candidates. Molecular docking and simulations confirmed a stable interaction, identifying key amino acids for drug development against cancer cells.
Area of Science:
- Medicinal Chemistry
- Computational Biology
- Oncology
Background:
- The cyclin Ds-CDKs axis, involving Cyclophilin A (Cyp A) and Ganoderiol F, is crucial in cancer progression.
- Ganoderiol F derivatives show promise as oncogenic targets for various cancer types.
Purpose of the Study:
- To design and develop novel Cyp A inhibitors and Ganoderiol F derivatives for cancer therapy.
- To optimize inhibitor effectiveness and receptor interaction through structure-guided design.
- To computationally evaluate potential drug candidates for cancer treatment.
Main Methods:
- Structure-guided design and molecular modeling using Chemsketch, AutoDock 4.2.6, PyMOL, Chimera, and Maestro.
- Construction of 117 novel cyclophilin inhibitor ligand molecules.
- Ligand-protein docking, docking confirmation, hydrogen bond detection, and molecular dynamics (MD) simulations using Gromacs with CHARMM 36 force field.
Main Results:
- Identified active amino acids in the binding site, including Arginine 55, Asparagine, Glycine, Threonine, Lysine, Isoleucine, Histidine, Phenylalanine, and Cysteine.
- Determined the optimal grid box coordinates (X: 38.86, Y: 10.987, Z: 40.734) for the 1nmk receptor-ligand complex.
- MD simulations confirmed the stability of the selected ligand-receptor complex, indicating its potential as a cancer drug inhibitor.
Conclusions:
- Novel Cyp A inhibitors and Ganoderiol F derivatives can be effectively designed using computational methods.
- The identified ligand-receptor complex demonstrates stability and potential for development as an anti-cancer therapeutic agent.
- This study provides a foundation for further investigation into these compounds for cancer treatment.
More Related Videos
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
 08:48
Development of a Backbone Cyclic Peptide Library as Potential Antiparasitic Therapeutics Using Microwave Irradiation
Published on: January 26, 2016
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:29
Pharmacogenomics: Identification of New Drug Targets