Related Experiment Video
Updated: Jul 16, 2026

Facile Preparation of (2Z,4E)-Dienamides by the Olefination of Electron-deficient Alkenes with Allyl Acetate
Published on: June 21, 2017
Reaction-Coordinate Analysis of DMAP-Mediated Allylation of Allylic Alcohols: Coupled Proton Transfer and C-C Bond
Emna Cherni1, Khaled Essalah2,3, Sameh Ayadi4
1NMR Research Unit, Faculty of Science, University of Oulu, P.O. Box 3000, OuluFI-90014, Finland.

Abstract:
Density functional theory calculations reveal that the DMAP-mediated allylation of cyclic Baylis-Hillman alcohols proceeds through a concerted substitution pathway in which proton transfer is directly coupled to bond reorganization, as an alternative to stepwise pathways involving discrete ionization. Although formation of an allylic cation has been invoked to rationalize this transformation, the energetic feasibility of ionization under neutral conditions has not been explicitly examined. Intrinsic reaction-coordinate (IRC) analysis provides direct insight into how proton transfer and bond reorganization evolve along the substitution coordinate. The computed reaction landscape shows that heterolytic C-O bond cleavage from the DMAP adduct is associated with a prohibitively high barrier (ΔG‡ ≈ 45 kcal·mol-1) and does not yield a discrete cationic intermediate, but instead collapses into a strongly associated ion pair, indicating that charge separation is incomplete and disfavors pathways requiring prior ionization under the reaction conditions. In contrast, rapid proton transfer between DMAP and the active methylene compound establishes a low-barrier acid-base pre-equilibrium (ΔG‡ ≈ 13 kcal·mol-1), generating a reactive carbanion-DMAPH+ ion pair. Subsequent C-C bond formation proceeds through a single proton-coupled transition state in which nucleophilic attack, proton transfer, and C-O bond cleavage occur in a concerted manner. This pathway avoids high-energy charge separation and provides a kinetically viable route consistent with prolonged heating under refluxing toluene, where slow but feasible turnover can be achieved over extended reaction times. Competing O-allylation pathways involve multiple high-energy intermediates and are therefore mechanistically less competitive. These results demonstrate that substitution can proceed through a concerted reaction coordinate in which proton transfer and bond cleavage are directly coupled, offering an alternative to pathways that rely on prior ionization and providing a general physical-chemistry perspective on metal-free allylic substitution.
Related Concept Videos
 01:18
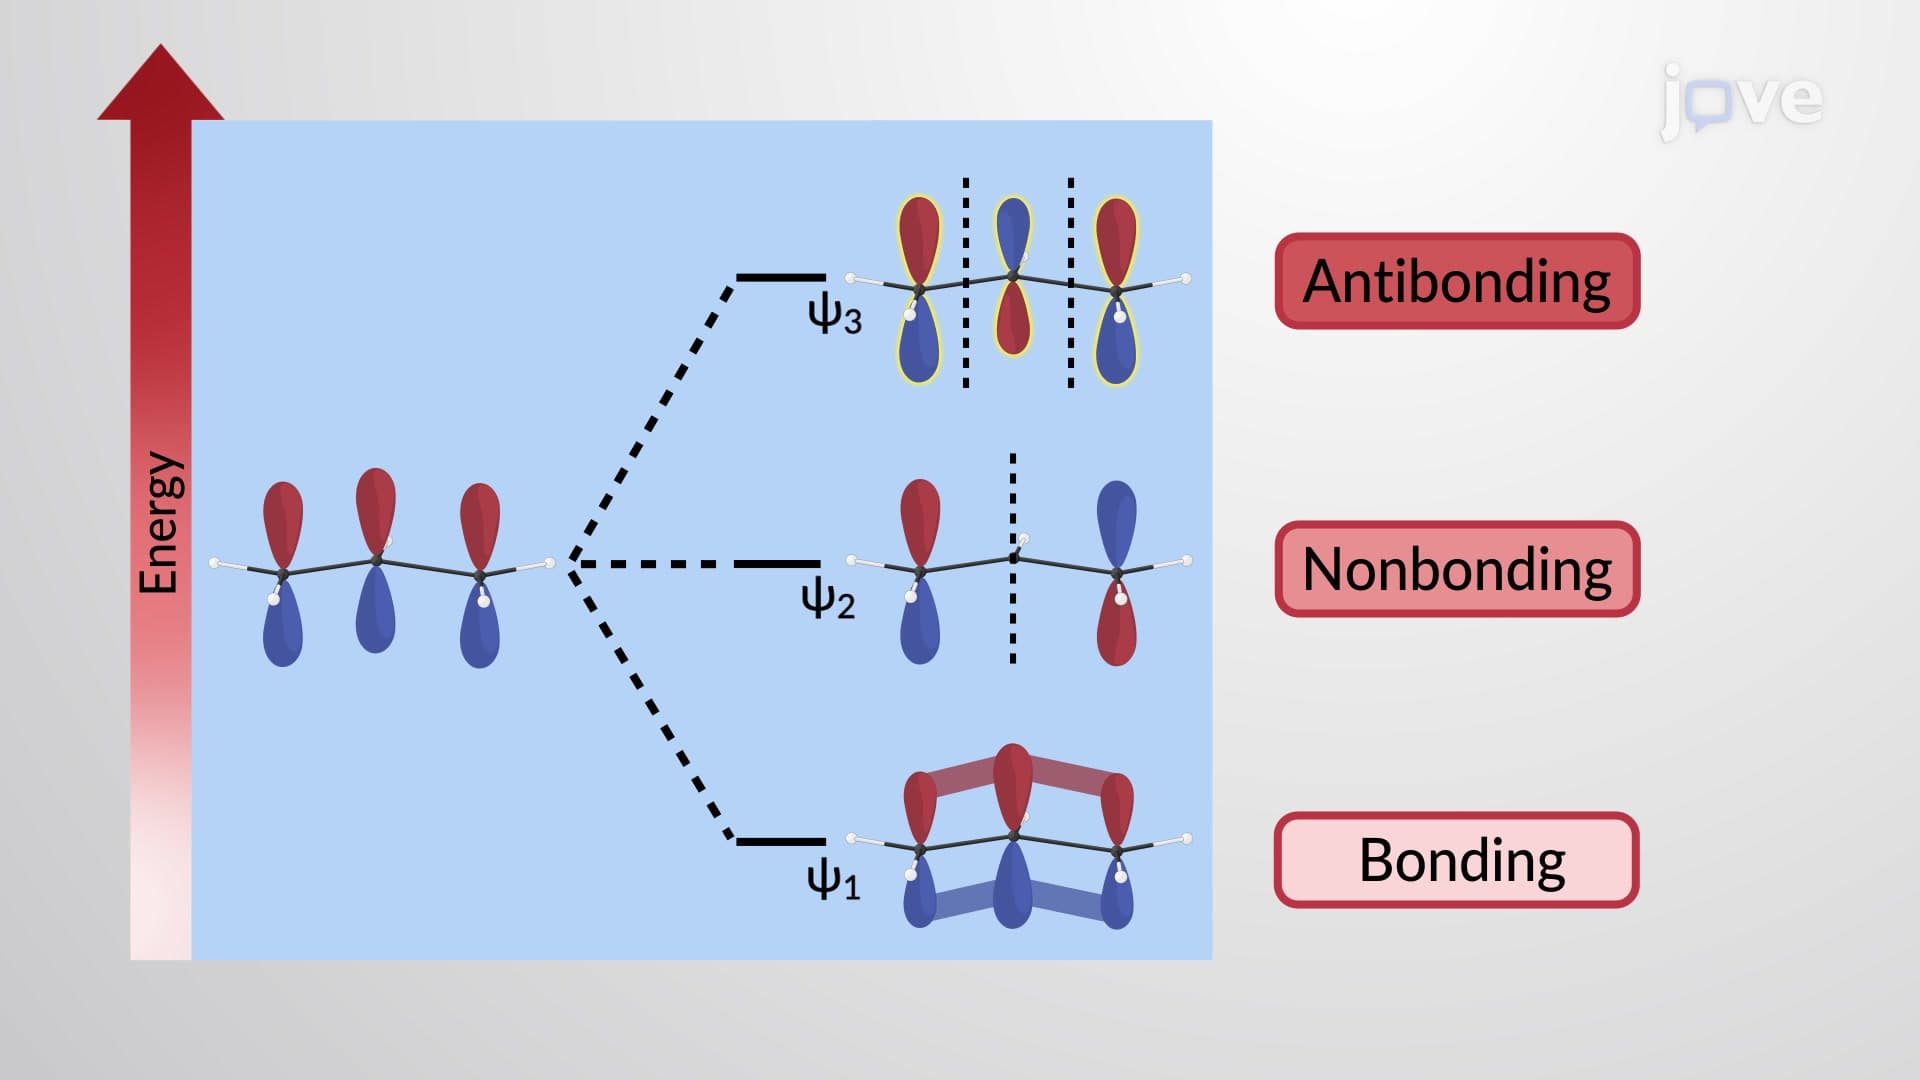
π Molecular Orbitals of the Allyl Cation and Anion
 01:16
[4+2] Cycloaddition of Conjugated Dienes: Diels–Alder Reaction
 01:24
[3,3] Sigmatropic Rearrangement of Allyl Vinyl Ethers: Claisen Rearrangement
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 01:22
Alkenes via Reductive Coupling of Aldehydes or Ketones: McMurry Reaction
 01:10
α-Alkylation of Ketones via Enolate Ions