Related Experiment Video
Updated: Jul 17, 2026
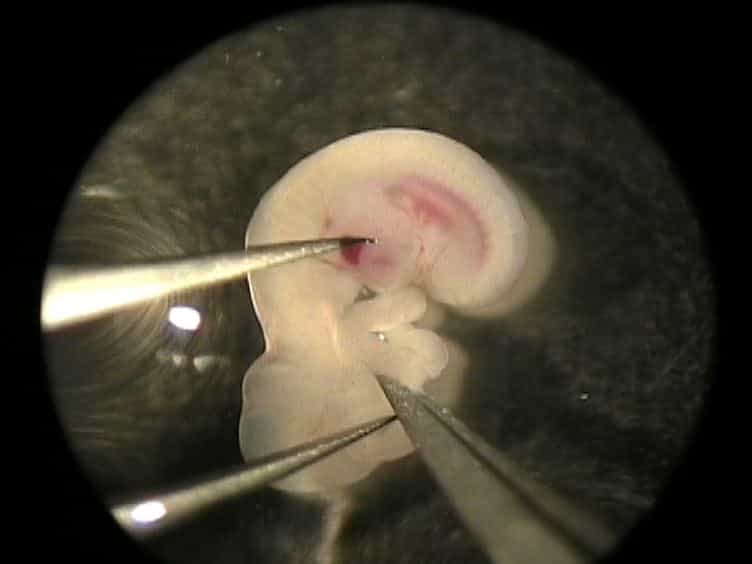

Separation of Mouse Embryonic Facial Ectoderm and Mesenchyme
Published on: April 12, 2013
SPEN deficiency contributes to the development of orofacial clefts in humans and mice
Bum Jun Kim1, Andrés Hernández-García1, David L Curtis1
1Department of Molecular & Human Genetics, Baylor College of Medicine, One Baylor Plaza, Houston, TX 77030, United States.

Abstract:
Haploinsufficiency of SKI, PRDM16, RERE, PAX7, and GRHL3 have been implicated in the development of orofacial clefting (OFC) associated with chromosome 1p36 deletions based on human and/or mouse data. Haploinsufficiency of SPEN, a 1p36 gene that encodes a transcriptional repressor, causes Radio-Tartaglia syndrome, a neurodevelopmental syndrome in which high/narrow palates are common, and OFC is occasionally observed. We show that Spen-null embryos have abnormal palatal shelf elevation and extension leading to the development of cleft palate. Mesenchymal cell proliferation in the medial halves of the palatal shelves of Spen-null embryos at E13.5 is significantly reduced. This contributes to the delay of palatal shelf elevation. Tissue specific ablation of Spen in the cranial neural crest cells results in delayed palatal development. This pattern of abnormal palatal development mimics the pattern described in RERE-deficient mice. We show that Rere and Spen are expressed in same cell types, that Rere and Spen interact genetically in the development of the palate, that Spen expression is reduced in the palates of RERE-deficient embryos at E14.5, and that the rate of OFC in individuals with 1p36 deletions involving both RERE and SPEN is higher than those of individuals with RERE or SPEN haploinsufficiency. Our results suggest that SPEN is required for normal mammalian palatal development, that RERE and SPEN interact in a common pathway during palatal development, and that haploinsufficiency of RERE and SPEN are likely to contribute to the development of OFC in individuals with 1p36 deletions.