Related Experiment Video
Updated: Jul 17, 2026

Exploring the Regulation of Lipid Droplet Catabolism through Lipophagy
Published on: January 31, 2025
Lipidation of the KV1.3 blocking peptide HsTX1[R14A] alters its pharmacokinetics and biodistribution
Lihuan Lin1, Thomas R Mitchell2, Karoline Sanches2
1Drug Delivery, Disposition and Dynamics, Monash Institute of Pharmaceutical Sciences, Monash University, Parkville 3052, Victoria, Australia.

Upregulation of the voltage-gated potassium channel KV1.3 in effector memory T cells has been implicated in several autoimmune diseases, making selective KV1.3 blockade an attractive therapeutic strategy. HsTX1[R14A], a 34-residue peptide with picomolar potency and high selectivity for KV1.3, is effective in a rodent model of rheumatoid arthritis. However, the in vivo half-life of HsTX1[R14A] can be improved to enhance its potential as a therapeutic candidate. Here we explore how conjugation to C14, C16 or C18 acyl chains affects HsTX1[R14A] binding to serum albumin and lipoproteins, potency at KV1.3, pharmacokinetics (PK), and biodistribution to key organs, tissues, and lymph nodes. The fluorophore Cy5 was conjugated to the HsTX1[R14A] analogs for biodistribution studies. LC-MS/MS assays were developed to quantify the lipidated peptides in mouse plasma. Conjugation of palmitic and stearic acids to HsTX1[R14A] caused over 20-fold loss in potency but significantly enhanced its binding to albumin, while showing minimal association with plasma lipoproteins. This modification extended the plasma exposure by 9-fold and prolonged the elimination half-life by 2-fold after subcutaneous (SC) injection to mice. Biodistribution studies demonstrated that Cy5-labelled lipidated analogs enhanced accumulation at the dosing site and draining lymph nodes at 4 and 12 h after SC injection compared to Cy5-HsTX1[R14A]. Cy5-labelled lipidated analogs of HsTX1[R14A] also accumulated in the gastrointestinal tract more than HsTX1[R14A], while having significantly less exposure in the kidneys at 4 and 12 h after SC administration. Thus, lipidation is a promising approach to optimize the PK, biodistribution and therapeutic potential of HsTX1[R14A].
Upregulation of the voltage-gated potassium channel KV1.3 in effector memory T cells has been implicated in several autoimmune diseases, making selective KV1.3 blockade an attractive therapeutic strategy. HsTX1[R14A], a 34-residue peptide with picomolar potency and high selectivity for KV1.3, is effective in a rodent model of rheumatoid arthritis. However, the in vivo half-life of HsTX1[R14A] can be improved to enhance its potential as a therapeutic candidate. Here we explore how conjugation to C14, C16 or C18 acyl chains affects HsTX1[R14A] binding to serum albumin and lipoproteins, potency at KV1.3, pharmacokinetics (PK), and biodistribution to key organs, tissues, and lymph nodes. The fluorophore Cy5 was conjugated to the HsTX1[R14A] analogs for biodistribution studies. LC-MS/MS assays were developed to quantify the lipidated peptides in mouse plasma. Conjugation of palmitic and stearic acids to HsTX1[R14A] caused over 20-fold loss in potency but significantly enhanced its binding to albumin, while showing minimal association with plasma lipoproteins. This modification extended the plasma exposure by 9-fold and prolonged the elimination half-life by 2-fold after subcutaneous (SC) injection to mice. Biodistribution studies demonstrated that Cy5-labelled lipidated analogs enhanced accumulation at the dosing site and draining lymph nodes at 4 and 12 h after SC injection compared to Cy5-HsTX1[R14A]. Cy5-labelled lipidated analogs of HsTX1[R14A] also accumulated in the gastrointestinal tract more than HsTX1[R14A], while having significantly less exposure in the kidneys at 4 and 12 h after SC administration. Thus, lipidation is a promising approach to optimize the PK, biodistribution and therapeutic potential of HsTX1[R14A].
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:14
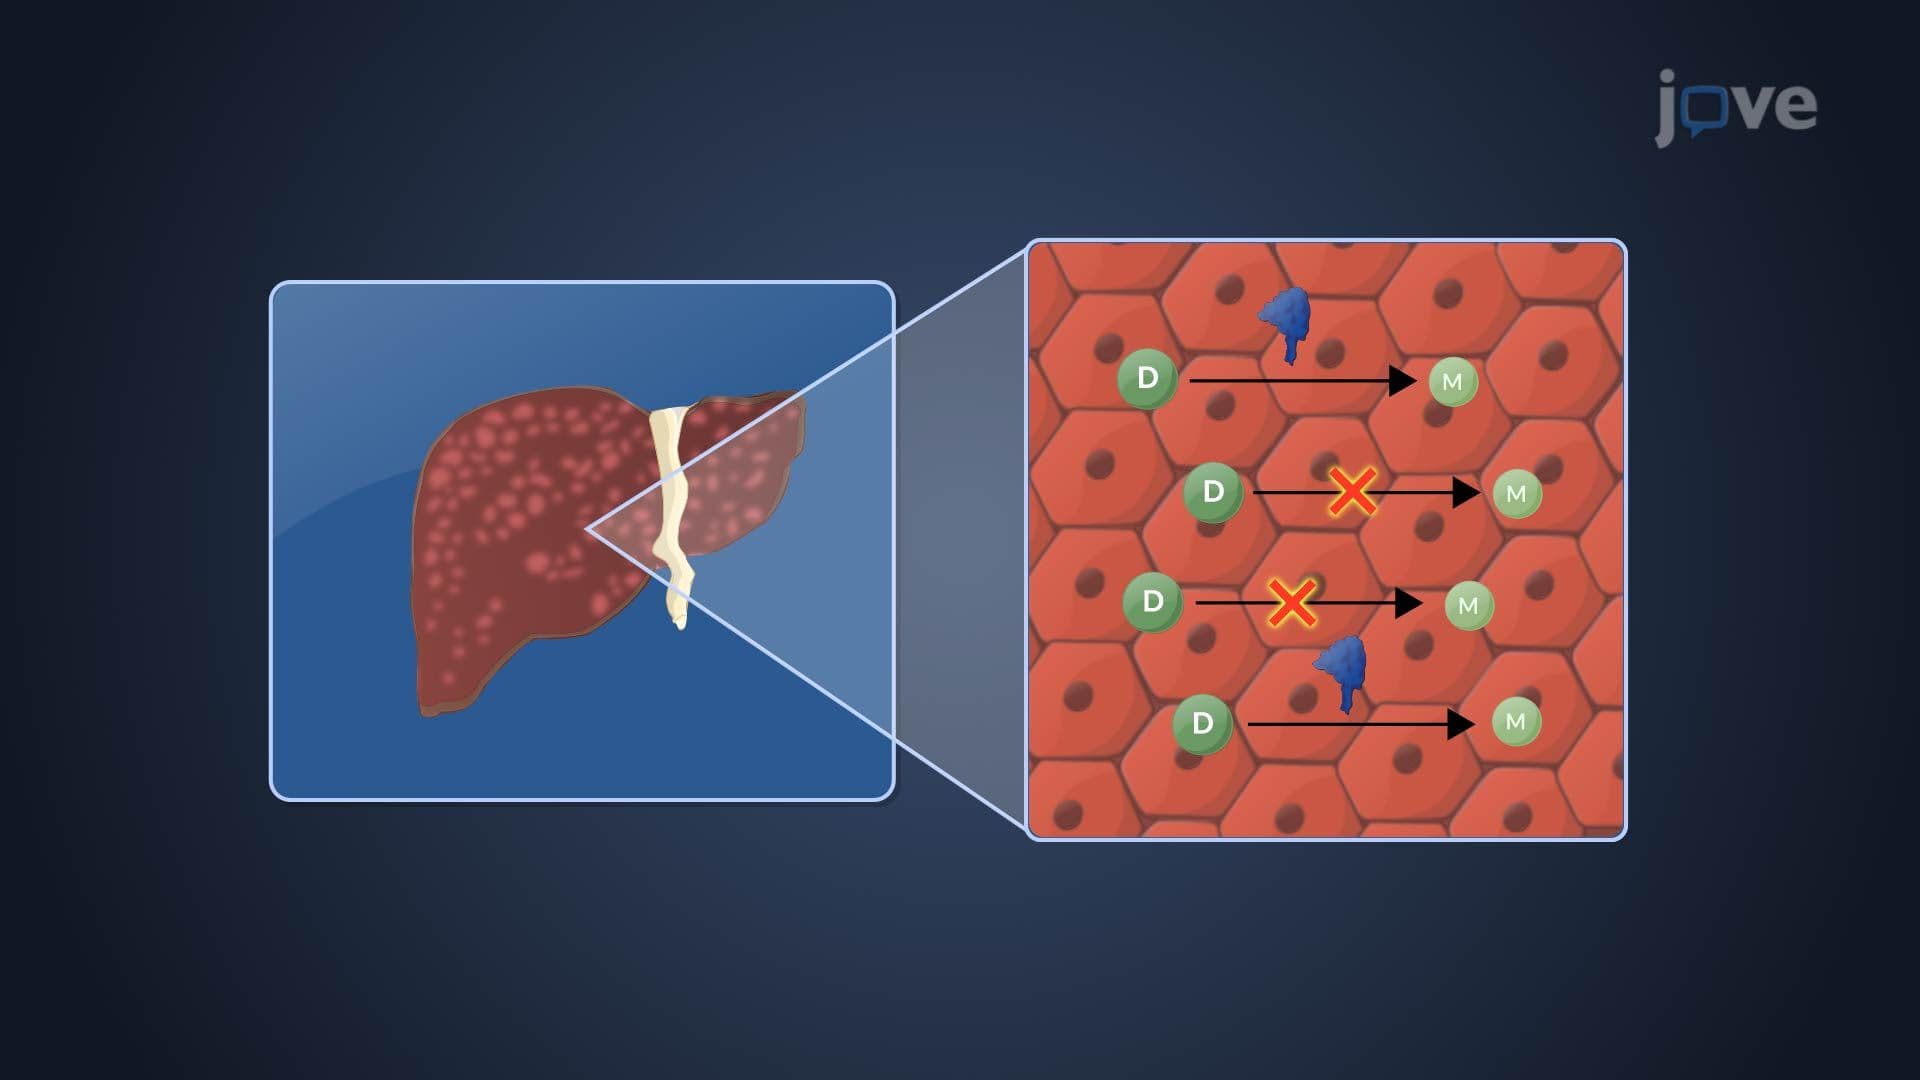
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug
 01:16
Drug Biotransformation: Overview
 01:27
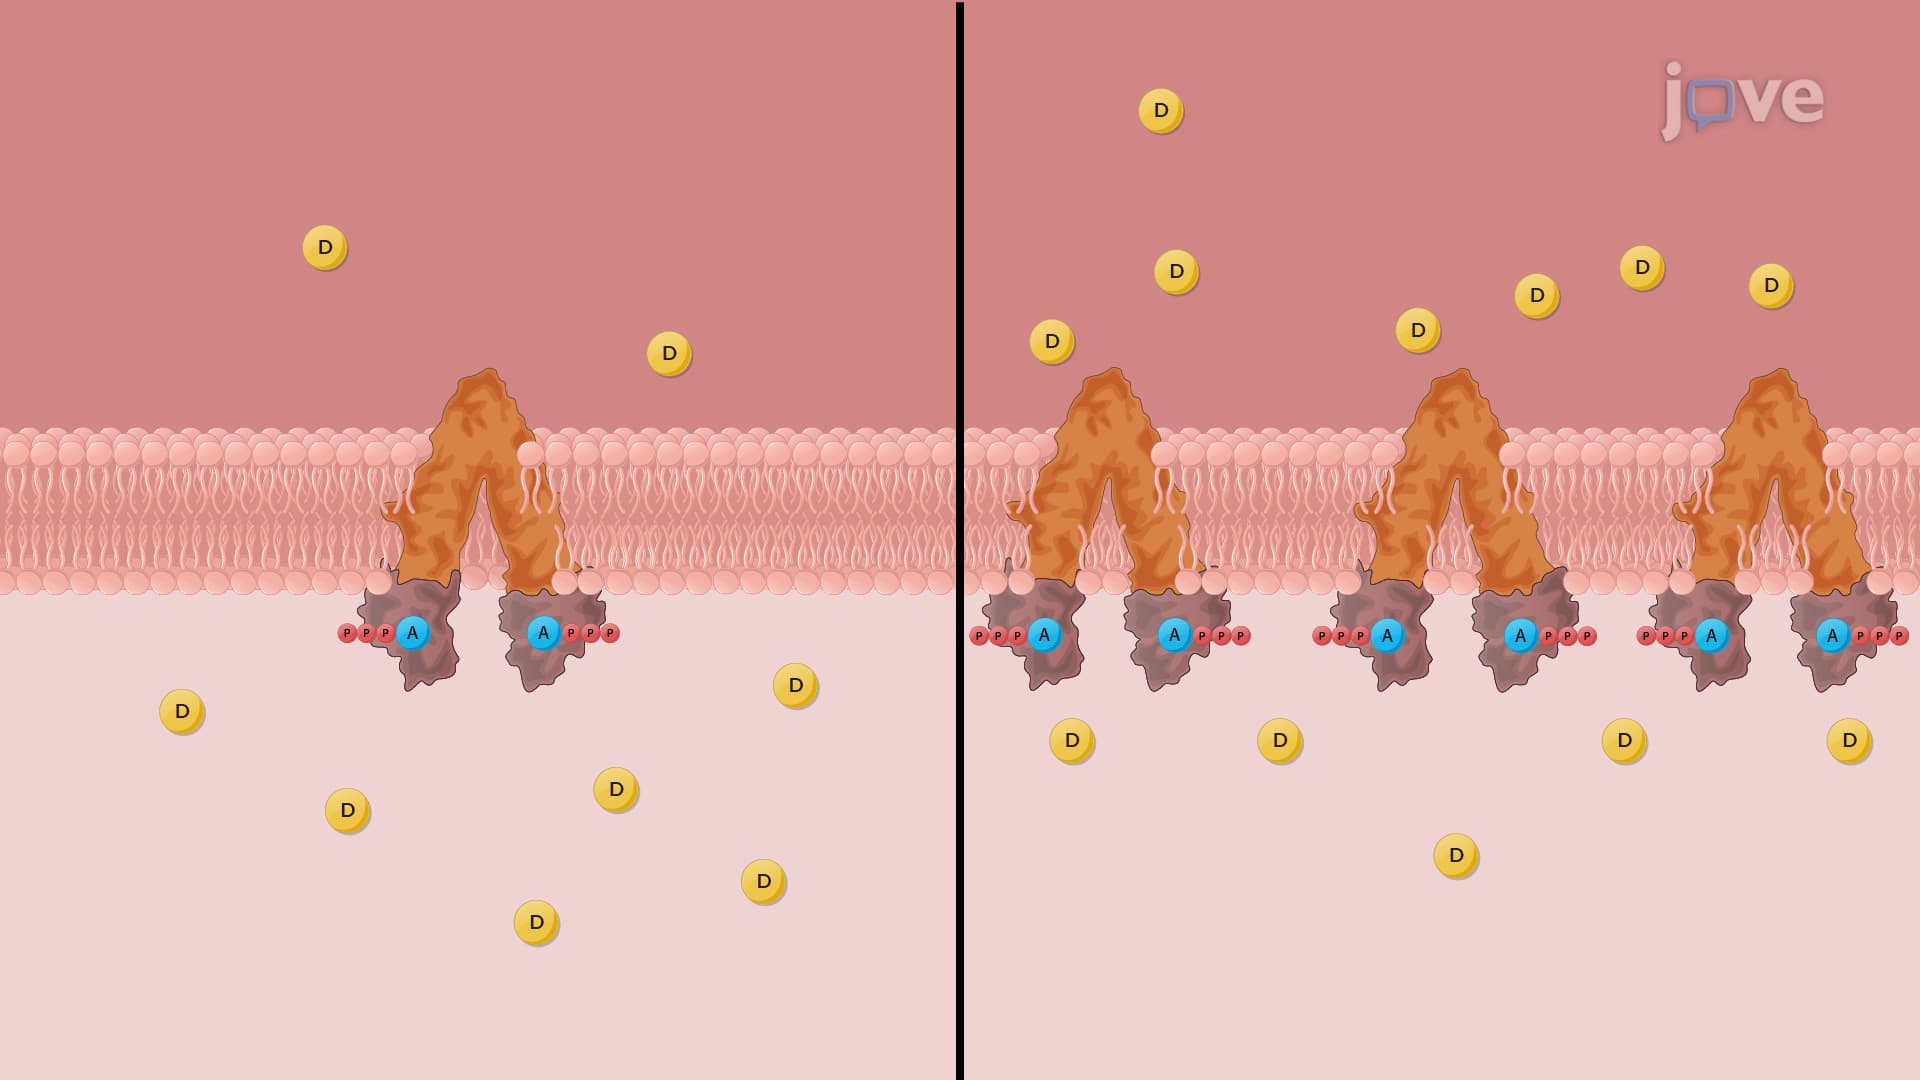
Nonlinear Pharmacokinetics: Role of Transporters
Polymorphisms occurring in drug transporters can alter...
 01:22
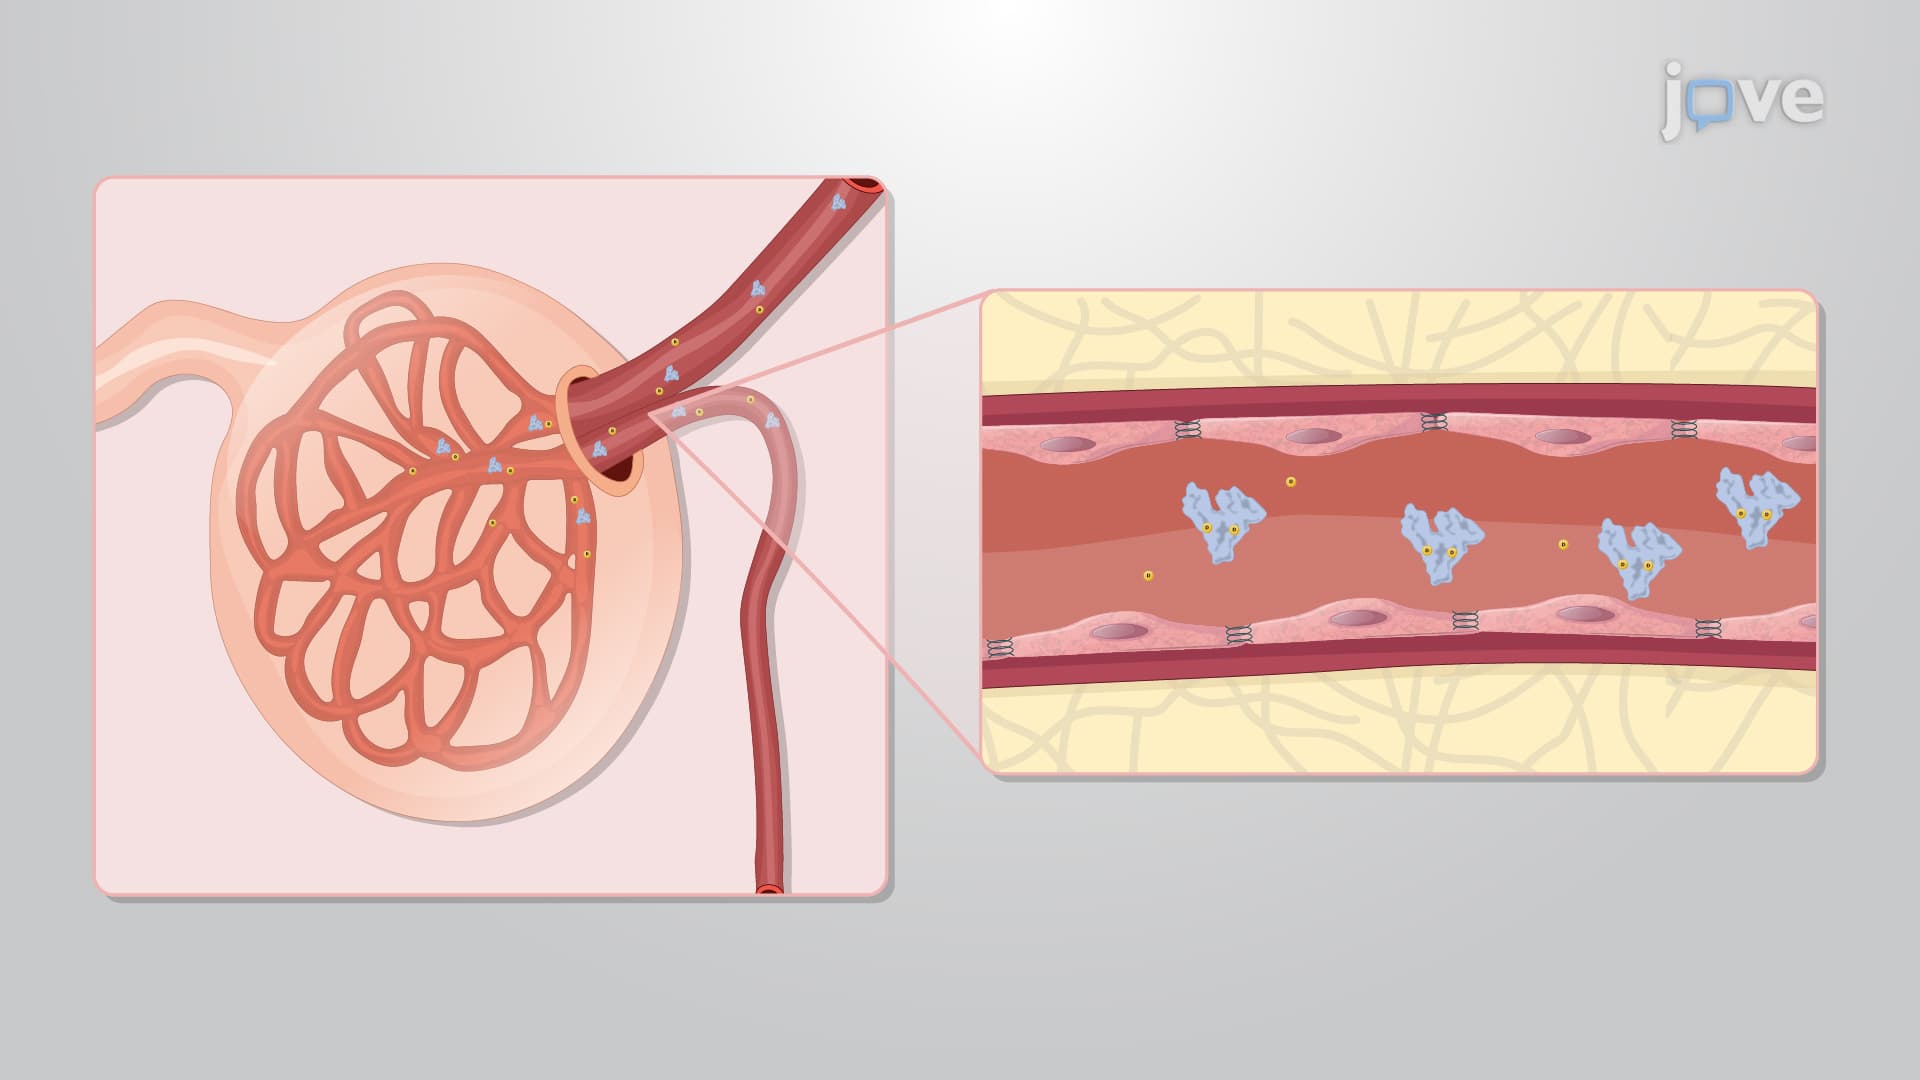
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:26
Effect of Hepatic Disease on Pharmacokinetics: Drug Dosing and Hepatic Blood Flow