Related Experiment Video
Updated: Aug 6, 2026
Refined Murine Model of Idiopathic Pulmonary Fibrosis
Published on: June 17, 2025
Targeting alveolar type II cell dysfunction in idiopathic pulmonary fibrosis: Molecular mechanisms and emerging
Tianhao Xie1, Kang Hu1, Shu Pan1
1Department of Thoracic Surgery, The First Affiliated Hospital of Soochow University, Suzhou, Jiangsu 215000, P.R. China.

Idiopathic pulmonary fibrosis (IPF) is a fatal and progressive form of interstitial lung pathology. It is characterized by the relentless replacement of functional alveolar epithelium by aberrant fibroblasts and excessive extracellular matrix deposition, resulting in respiratory failure. Currently, the treatment options for this disease are limited. Approved antifibrotic agents primarily slow disease progression by inhibiting fibroblast proliferation and downstream fibrotic pathways; however, they do not halt or reverse the underlying pathology. Recent studies have elucidated the proliferation and differentiation characteristics of type II alveolar epithelial (AT2) cells, identifying them as facultative stem cells of the distal lung with significant therapeutic potential in IPF. However, there are certain limitations in clinical translation. This review comprehensively summarizes the regulatory pathways governing AT2 proliferation and differentiation, and investigates key pathogenic drivers of IPF, including cellular senescence and mechanical tension. Furthermore, it evaluates current treatment strategies and methods to facilitate safer, more effective clinical delay or even reversal of pulmonary fibrosis by identifying or improving existing therapies.
Idiopathic pulmonary fibrosis (IPF) is a fatal and progressive form of interstitial lung pathology. It is characterized by the relentless replacement of functional alveolar epithelium by aberrant fibroblasts and excessive extracellular matrix deposition, resulting in respiratory failure. Currently, the treatment options for this disease are limited. Approved antifibrotic agents primarily slow disease progression by inhibiting fibroblast proliferation and downstream fibrotic pathways; however, they do not halt or reverse the underlying pathology. Recent studies have elucidated the proliferation and differentiation characteristics of type II alveolar epithelial (AT2) cells, identifying them as facultative stem cells of the distal lung with significant therapeutic potential in IPF. However, there are certain limitations in clinical translation. This review comprehensively summarizes the regulatory pathways governing AT2 proliferation and differentiation, and investigates key pathogenic drivers of IPF, including cellular senescence and mechanical tension. Furthermore, it evaluates current treatment strategies and methods to facilitate safer, more effective clinical delay or even reversal of pulmonary fibrosis by identifying or improving existing therapies.
Related Concept Videos
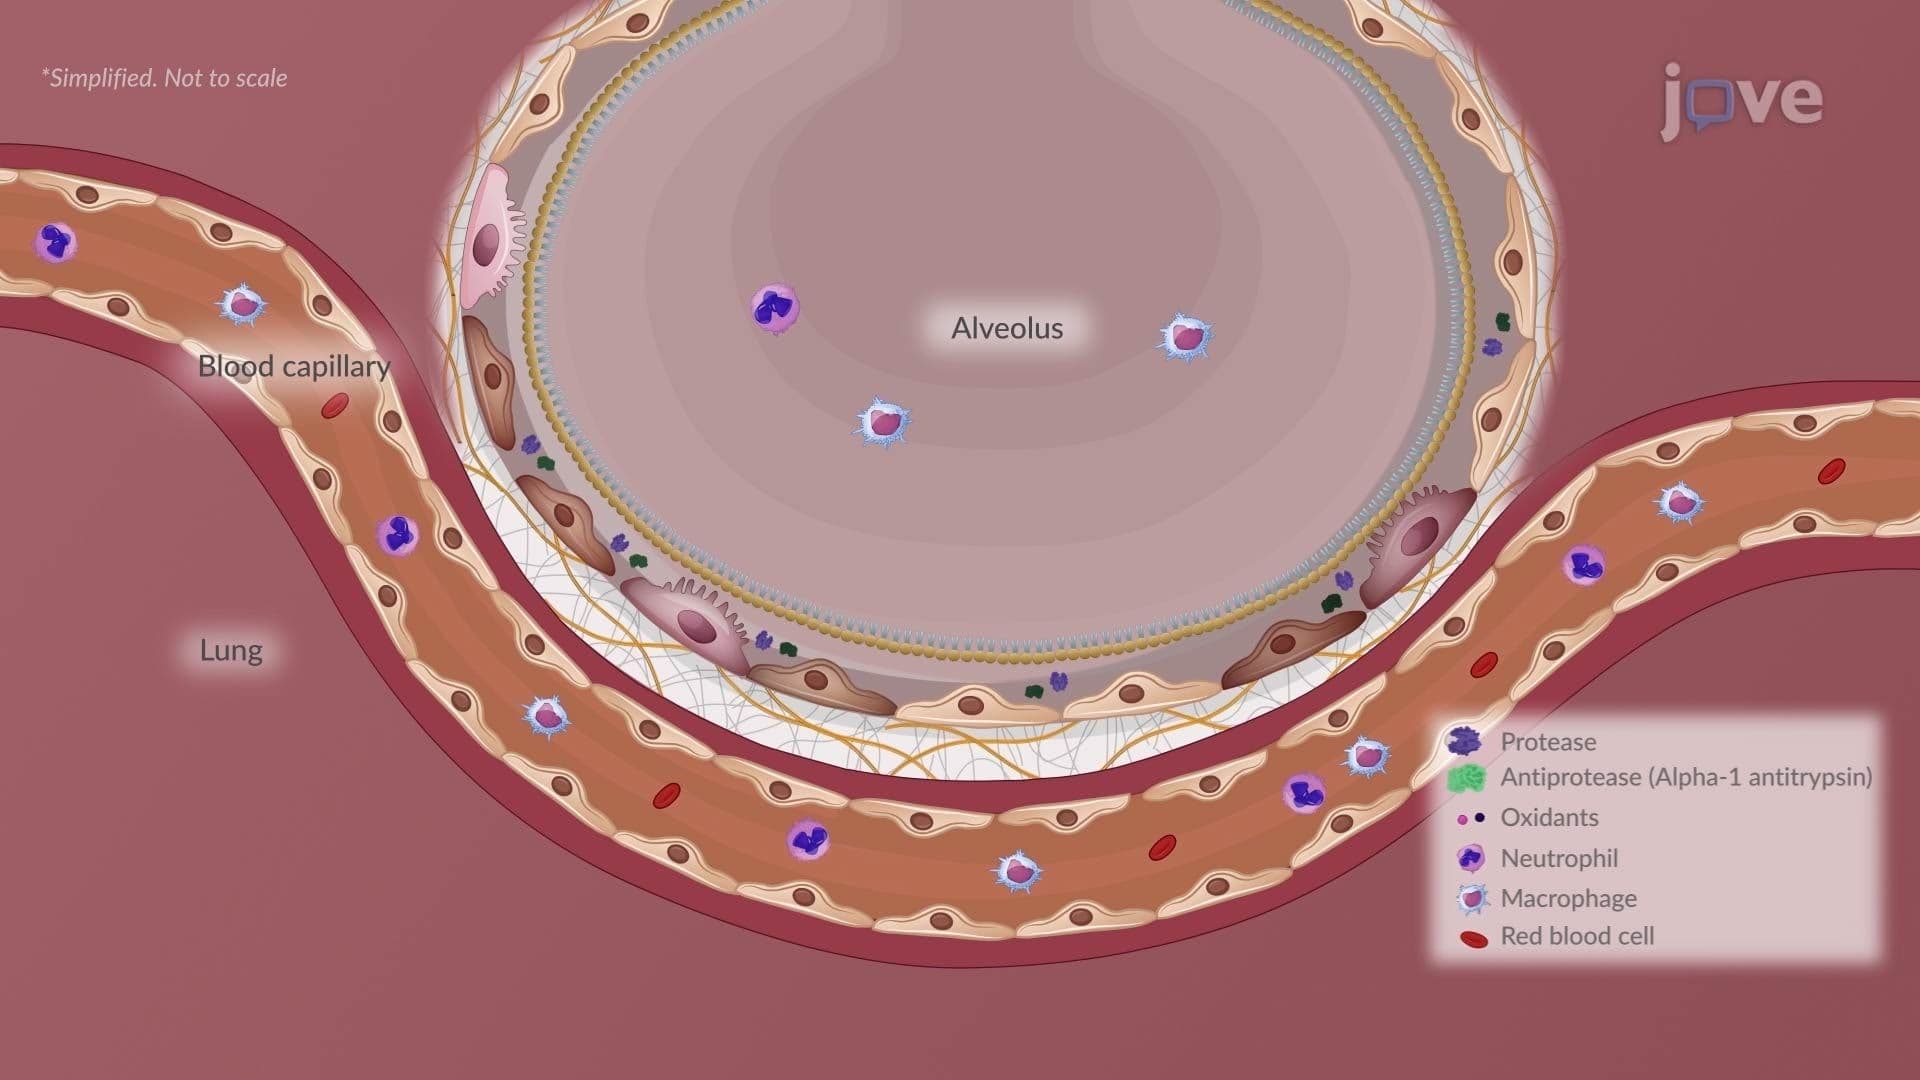 01:23
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:21
Acute Respiratory Failure-II
The underlying physiological abnormalities that contribute to hypoxemic respiratory failure include:
 01:10
Atelectasis II: Pathophysiology