Related Experiment Video
Updated: Aug 6, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Benchmarking semiempirical quantum chemical methods on liquid water
Xin Wu1,2, Hossam Elgabarty1, Vahideh Alizadeh3,4,5
1Dynamics of Condensed Matter and Center for Sustainable Systems Design, Chair of Theoretical Chemistry, Paderborn University, Warburger Str. 100, D-33098 Paderborn, Germany.

Abstract:
Stimulated by the renewed interest and recent developments in semiempirical quantum chemical (SQC) methods for noncovalent interactions, we examine the properties of liquid water under ambient conditions by means of molecular dynamics (MD) simulations, both with the conventional neglect of diatomic differential overlap-type methods, e.g., AM1 and PM6, and with DFTB-type (density-functional tight-binding) methods, e.g., DFTB2 and GFN-xTB (Geometry-Frequency-Noncovalent eXtended Tight-Binding). Besides the original parameter sets, some specifically reparameterized SQC methods (denoted as AM1-W, PM6-fm, and DFTB2-iBi) targeting various smaller water systems ranging from molecular clusters to bulk are considered as well. The quality of these different SQC methods for describing liquid water properties under ambient conditions is assessed by comparison with well-established experimental data and also with BLYP-D3 density functional theory-based ab initio MD simulations. Our analyses reveal that static and dynamic properties of bulk water are poorly described by all considered SQC methods with the original parameters, regardless of the underlying theoretical models, with most of the methods suffering from too weak hydrogen bonds and hence predicting a far too fluid water with highly distorted hydrogen bond kinetics. Meanwhile, the reparameterized force-matched PM6-fm method is shown to be able to quantitatively reproduce the static and dynamic features of liquid water and thus can be used as a computationally efficient alternative to electronic structure-based MD simulations for liquid water that requires extended length and time scales. DFTB2-iBi predicts a slightly overstructured water with reduced fluidity, whereas AM1-W gives an amorphous ice-like structure for water under ambient conditions.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:39
Enthalpy of Solution
 01:16
Titration in Nonaqueous Solvents
 01:16
Solubility Equilibria: Ionic Product of Water
The ionic product of water varies with temperature, and its value is 1.0 x 10⁻¹⁴ at standard experimental conditions. Per Le Chatelier's...
 02:05
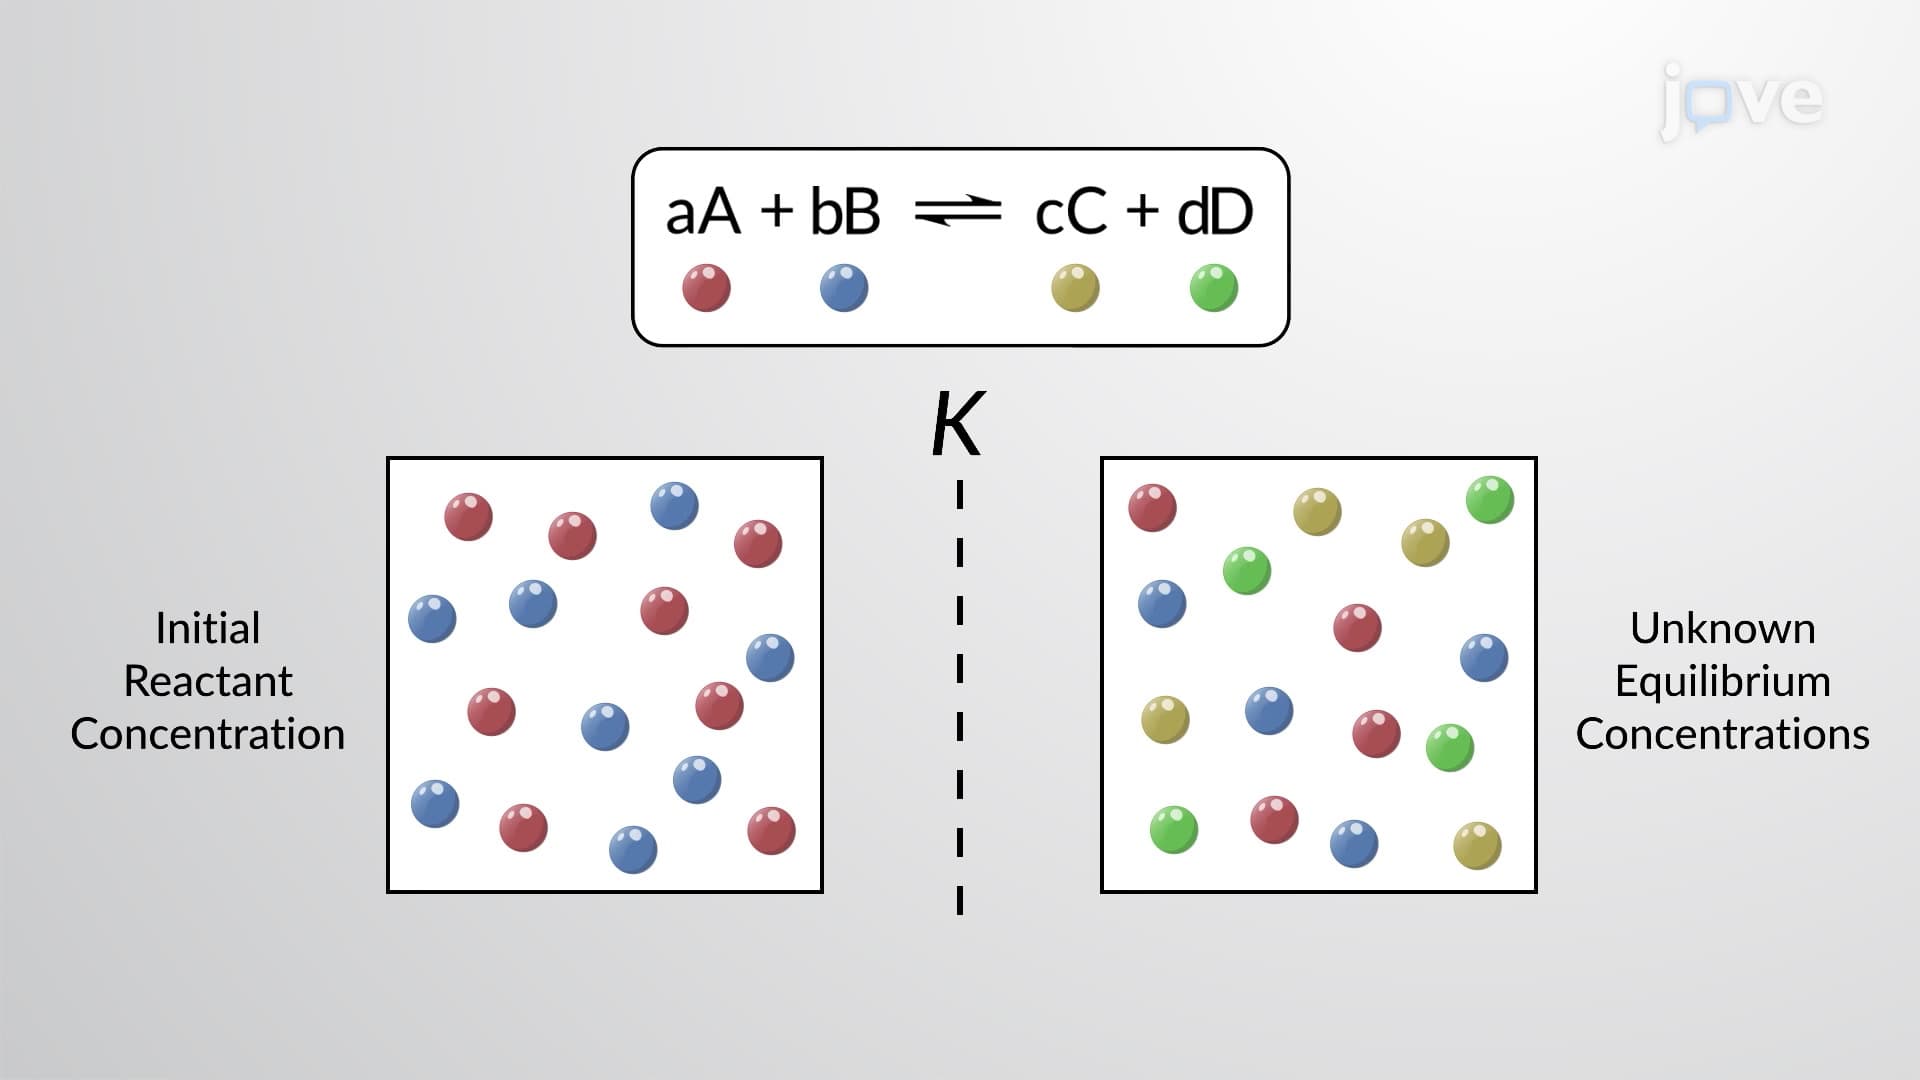
Calculating Equilibrium Concentrations
A more...