Related Experiment Video
Updated: Aug 6, 2026

Atom Probe Tomography Studies on the Cu(In,Ga)Se2 Grain Boundaries
Published on: April 22, 2013
First-principles study on the interaction between Gr/GO and C-S-H gel units
Jianlin He1,2, Chunwei Zhang3
1School of Materials Science and Engineering, Shenyang University of Technology, Shenyang, 110870, People's Republic of China.

Context:
Based on density functional theory (DFT), this study investigated the interfacial interactions between pristine graphene (Gr), Gr with single-vacancy (SV), double-vacancy (DV), and Stone-Wales (SW) defects, as well as graphene oxide (GO) modified with different oxygen-containing functional groups, and representative units of calcium silicate hydrate (C-S-H) gel. The results show that the adsorption strength of Si(OH)4 on pristine Gr/GO surfaces is mainly controlled by the polarity of the functional groups, following the order of GO-COOH > GO-OH > GO-O > Gr. The SW defect can enhance the adsorption on Gr and GO-OH, but weakens the short-range interaction between GO-COOH and Si(OH)4. For negatively charged SiO(OH)3-, the SV defect can significantly promote interfacial charge redistribution. The adsorption energy of GO-OH-SV reaches -1.907 eV, accompanied by proton transfer and local structural reconstruction. The introduction of Ca2+ and OH- can further improve adsorption stability through coordination bridging, charge compensation, and the synergistic effect between defects and functional groups. Among them, the adsorption energy of Si(OH)4 on Gr-DV decreases to -3.902 eV, while that of SiO(OH)3- on GO-O-SV decreases to -4.974 eV. The interaction between pristine Gr and the silicate chain is mainly characterized by weak van der Waals interactions, whereas hydroxyl and carboxyl groups can form specific bonding with the dangling oxygen atoms of the silicate chain through active hydrogen atoms and enhance charge transfer. In the Ca2+/OH- environment, oxygen-containing functional groups and defect sites jointly construct multiple adsorption centers, further strengthening the interfacial bonding between Gr/GO and C-S-H gel units.
Methods:
Density functional theory (DFT) calculations were carried out using the CASTEP module in Materials Studio software. The GGA-PBE functional, ultrasoft pseudopotential, DFT-D dispersion correction, and Monkhorst-Pack k-point sampling were used for geometry optimization and energy calculations. Adsorption energy, formation energy, and charge density difference were calculated to evaluate the interaction between Gr/GO and C-S-H gel units.
Related Concept Videos
 01:17
Spin–Spin Coupling: One-Bond Coupling
 02:21
Chemical and Solubility Equilibria
 01:13
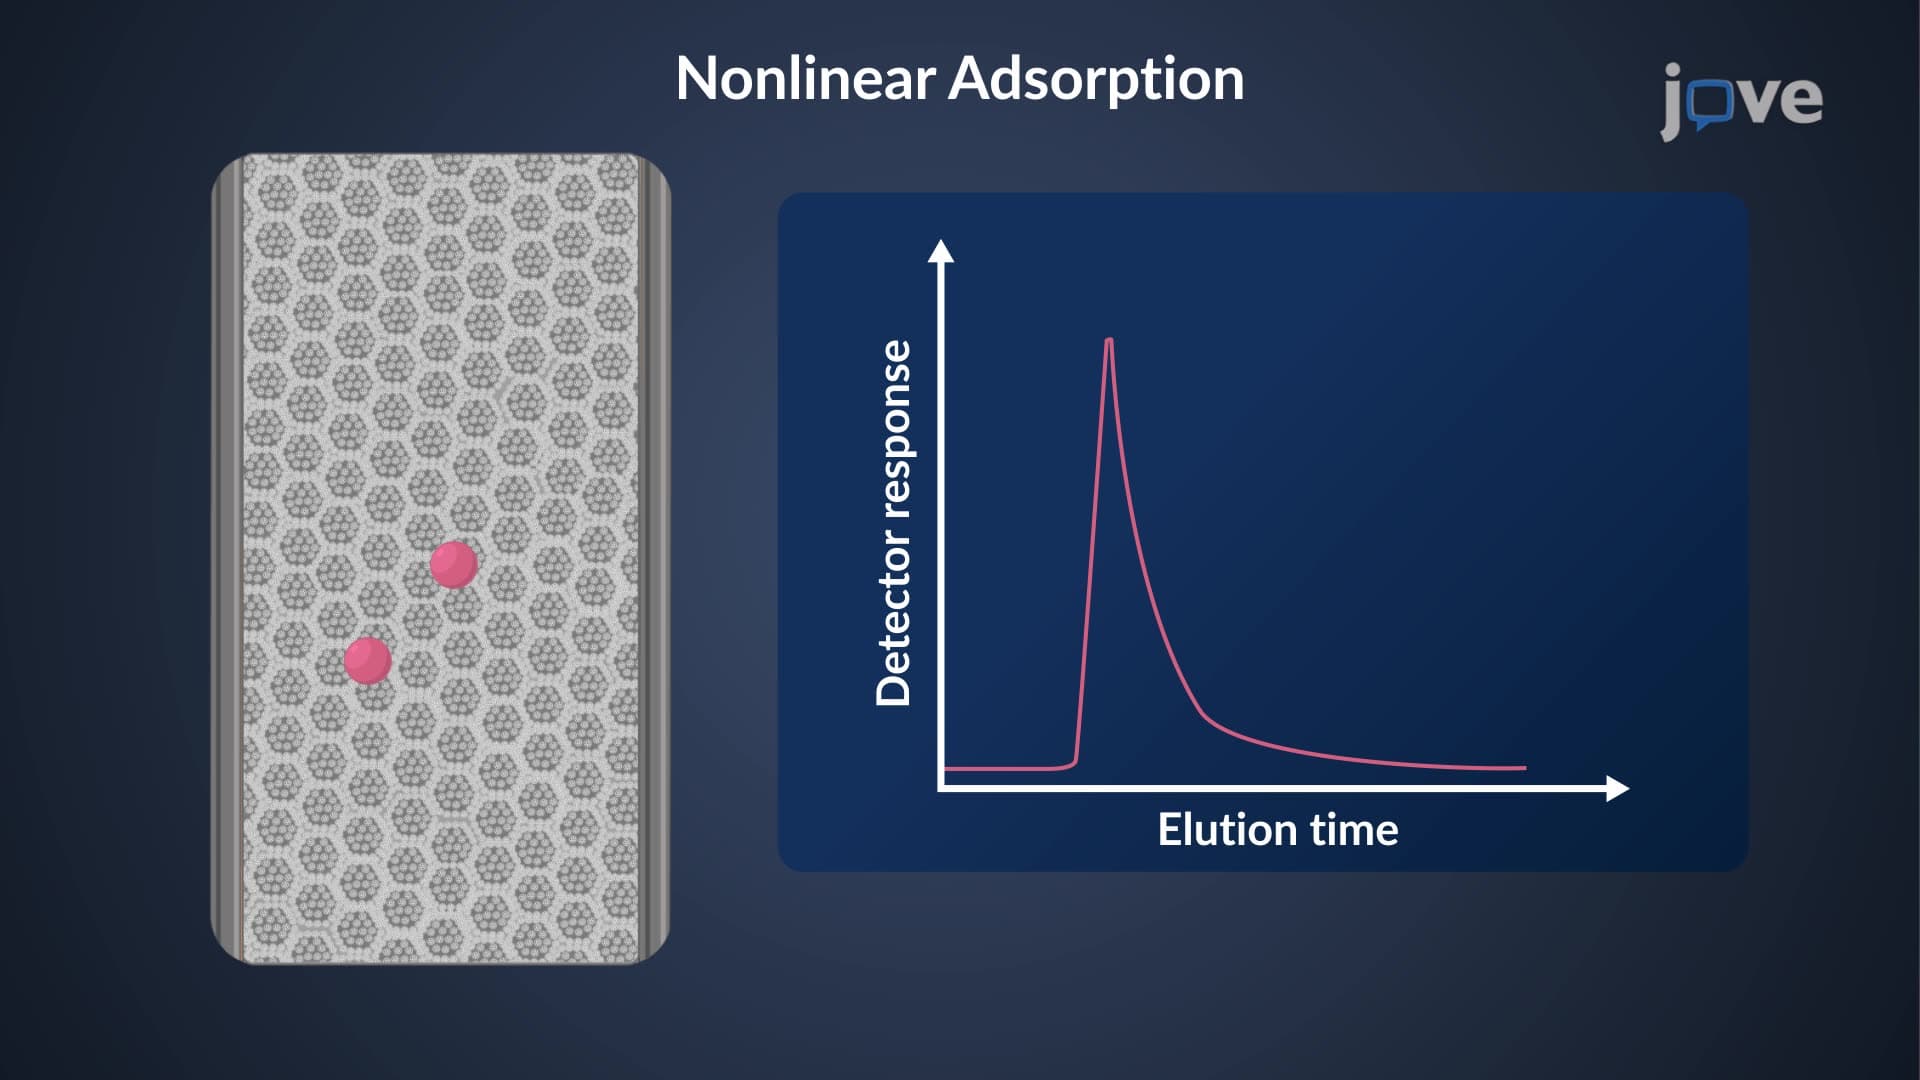
Gas Chromatography: Introduction
In GC, a sample is vaporized and mixed with an inert carrier gas (the mobile phase), which transports it through a column.
 01:22
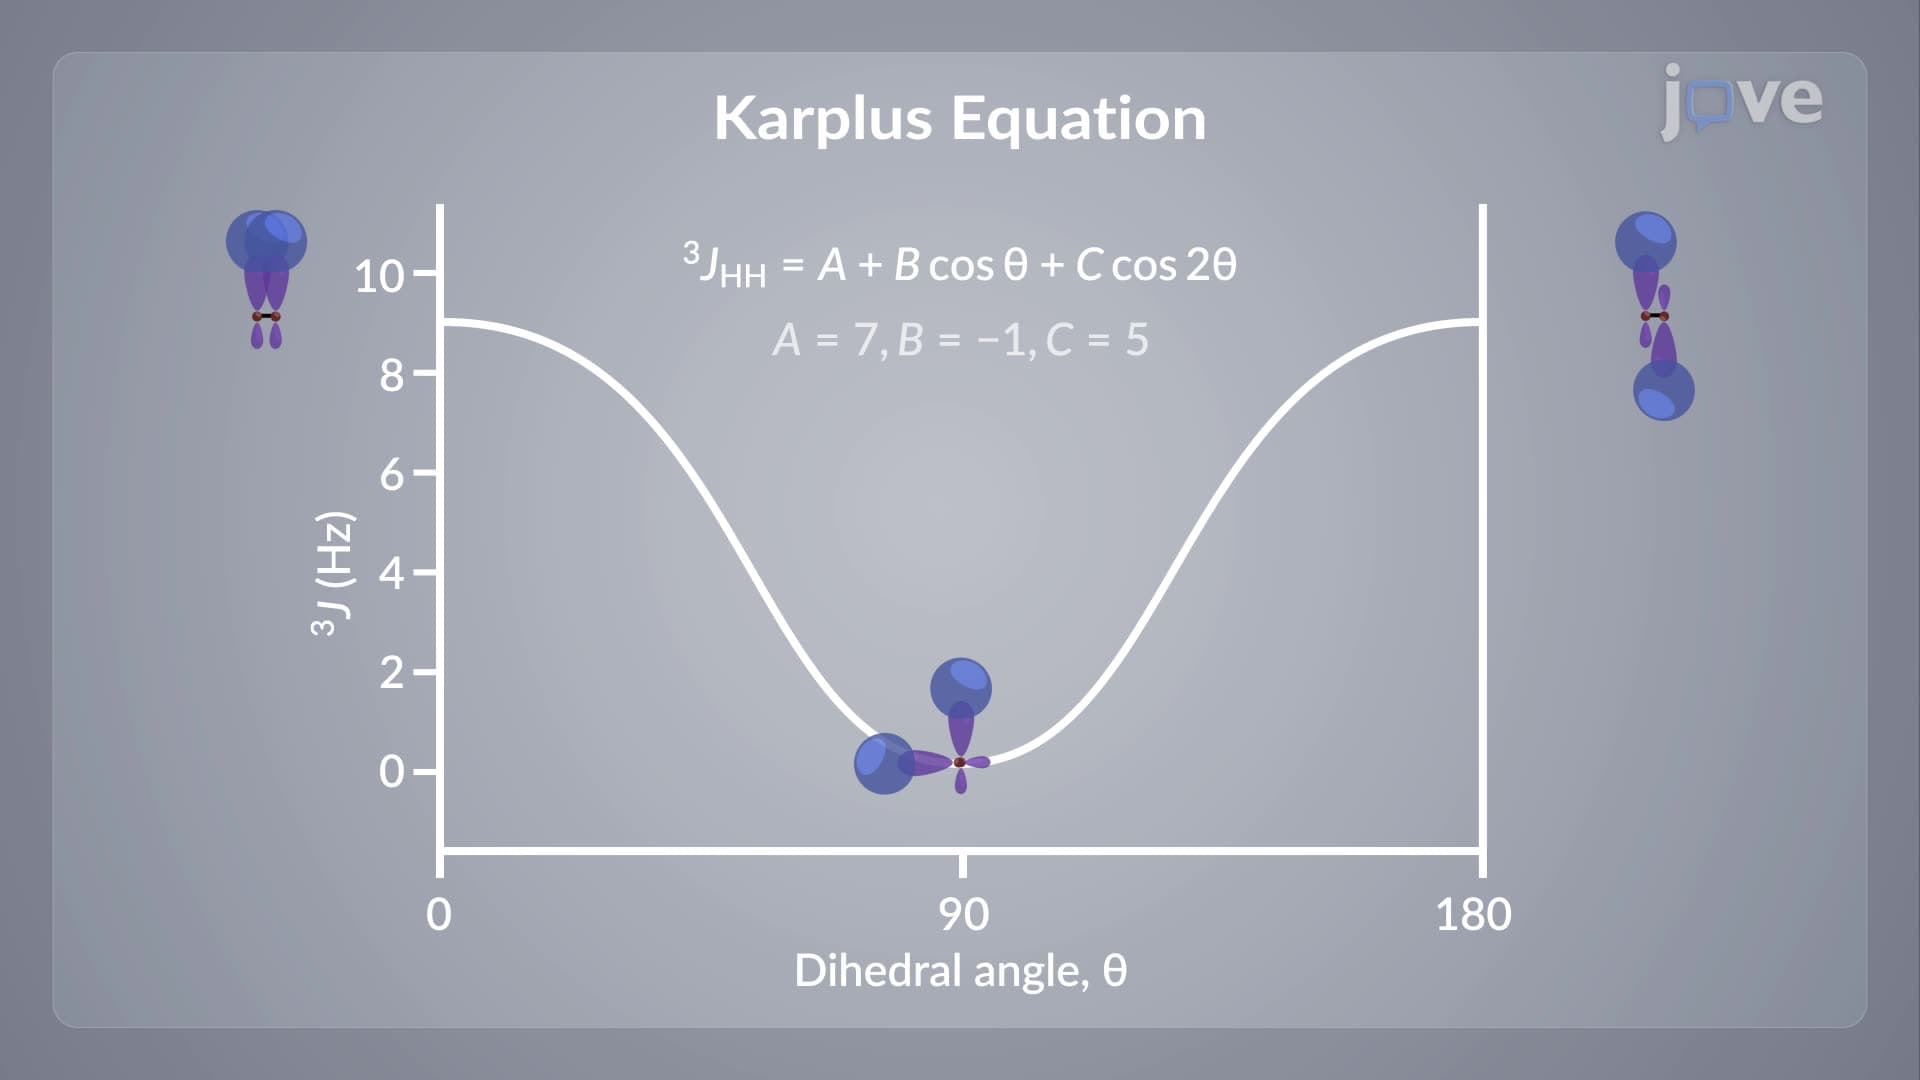
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 02:30
Chirality at Nitrogen, Phosphorus, and Sulfur
A consequence of chirality is the need for enantiomeric resolution. While this is theoretically possible for all...
 03:24
Hybridization of Atomic Orbitals I