Related Experiment Video
Updated: Aug 6, 2026


Realistic Membrane Modeling Using Complex Lipid Mixtures in Simulation Studies
Published on: September 1, 2023
Dual-Coordinate Relative Free Energy Simulations Using Machine-Learned Interatomic Potentials
Anna Katharina Picha1,2, Stefan Boresch1
1University of Vienna , Faculty of Chemistry, Institute of Computational Biological Chemistry, Vienna1090, Austria.

Abstract:
The integration of machine-learned interatomic potentials (MLIPs) into free energy simulations (FES) offers the promise of near-quantum mechanical accuracy at a reduced computational cost. However, employing MLIPs for alchemical relative free energy calculations remains challenging since MLIPs are typically not trained to handle the unphysical intermediate states required for such transformations. In previous work, we demonstrated how atoms and molecules can be gradually decoupled in systems fully described by MLIPs by manipulating the neighbor list and introducing an artificial offset to interatomic distances. Here, we extend this approach to general alchemical transformations between two states and apply the methodology to computing relative solvation free energies (RSFEs) between arbitrary solute pairs in systems fully described by an unmodified MLIP. We employ a dual-coordinate approach where both solutes are explicitly present but do not interact with one another. By introducing λ-dependent distance offsets to the neighbor list, we perform a simultaneous transformation: smoothly decoupling the first solute from the solvent while coupling the second solute to the same environment. To enforce spatial overlap without modifying the internal intramolecular dynamics of either solute, we utilize a harmonic ″anchor″ restraint to a central atom on each molecule. We demonstrate the robustness of this method using the MACE-OFF23(S) potential across a diverse set of small molecule pairs, including transformations between species with no chemical similarity (e.g., toluene to tetrahydrofuran). The method's accuracy is validated through thermodynamic cycle closure by combining direct RSFE calculations with absolute solvation free energy (ASFE) calculations. The methodology achieves excellent internal consistency, with cycle closure errors less than or equal to ±0.11 kcal/mol, well within the estimated statistical uncertainty. By requiring only a single energy evaluation per simulation step and avoiding architecture-specific modifications and retraining, this anchor-based dual-coordinate approach provides a highly adaptable and efficient route for relative alchemical FES with modern MLIPs.

Related Concept Videos
 01:26
Thermodynamic Potentials
 02:27
Predicting Molecular Geometry
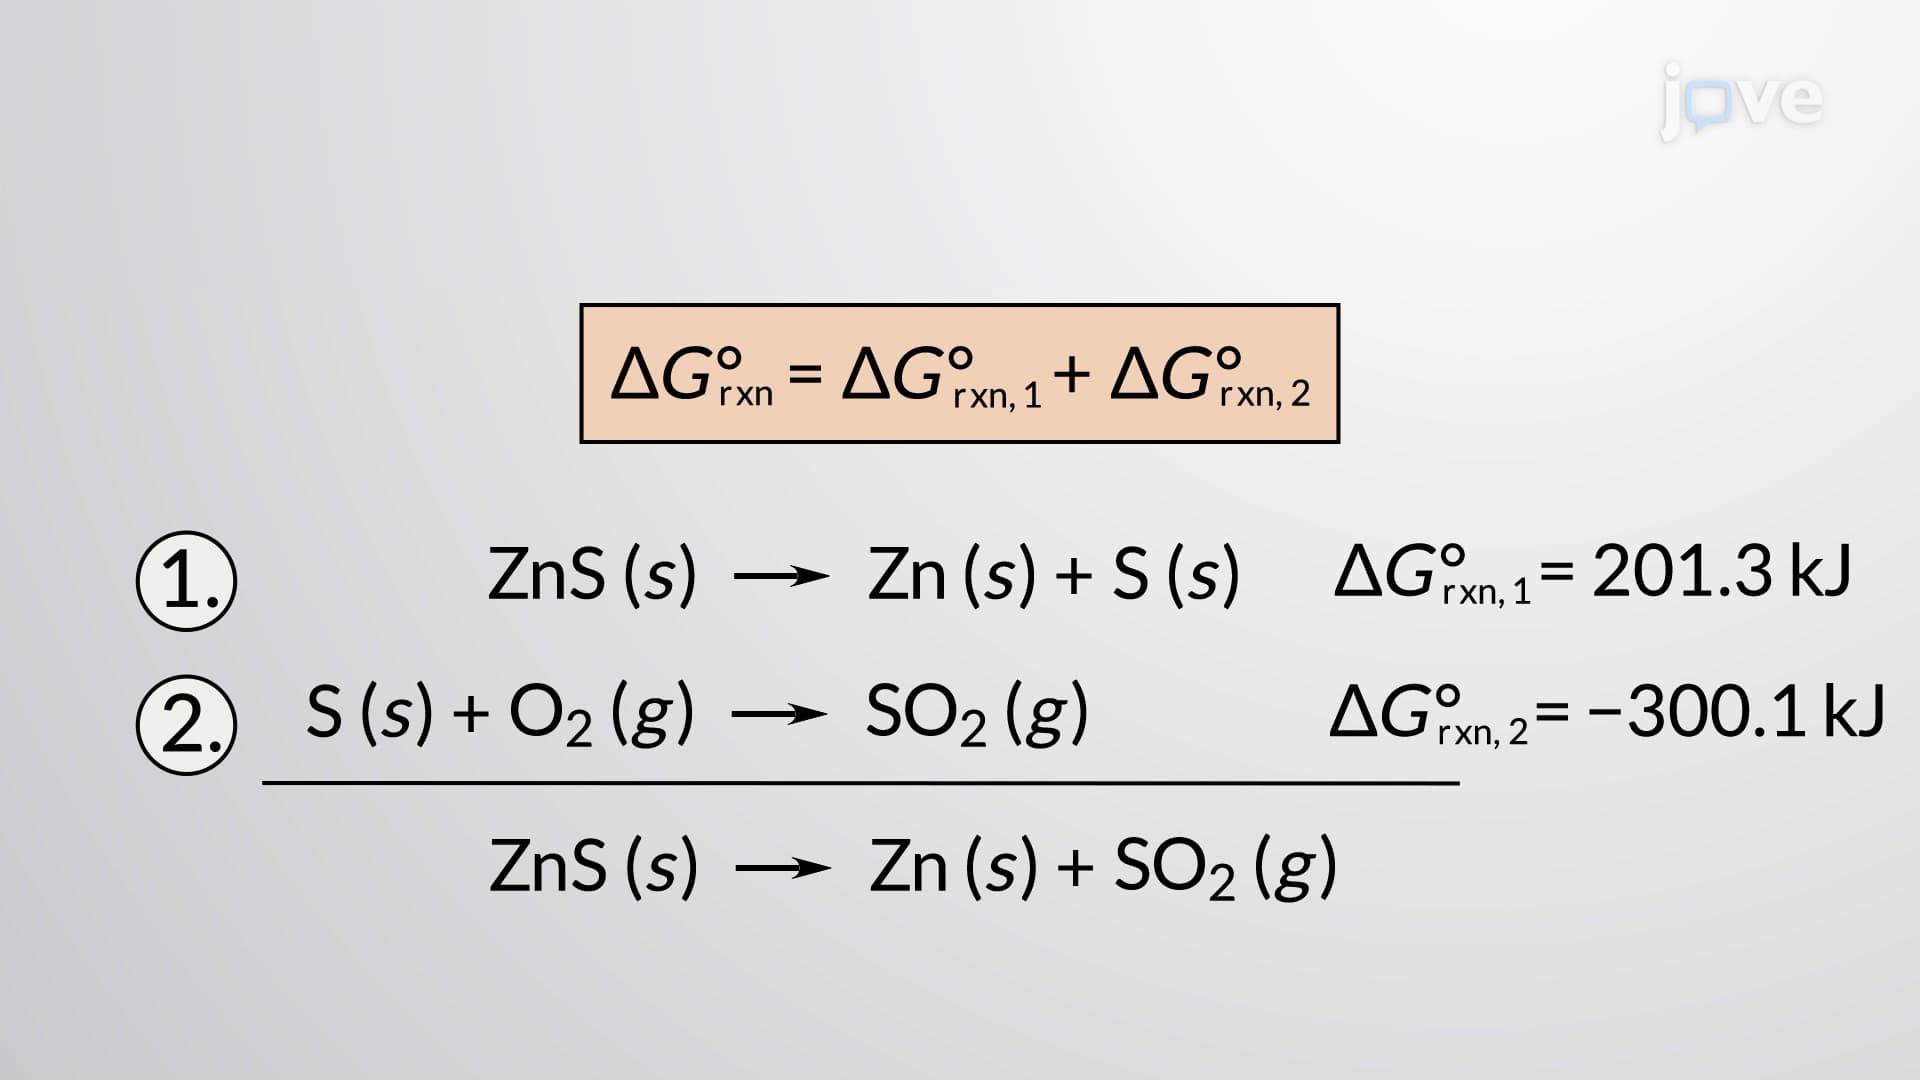 02:49
Calculating Standard Free Energy Changes
 01:16
Potential-Energy Criterion for Equilibrium
 03:25
Free Energy Changes for Nonstandard States
 01:20
Two-Dimensional Force System