Related Experiment Video
Updated: Aug 6, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Benchmarking mixed quantum-classical molecular dynamics for electronic strong coupling
Arun Kumar Kanakati1, Oriol Vendrell2, Gerrit Groenhof1
1Nanoscience Center and Department of Chemistry, University of Jyväskylä, P.O. Box 35, 40014 Jyväskylä, Finland.

Abstract:
Experiments indicate that collective coupling of molecular ensembles to confined optical modes can modify excited-state dynamics and photochemical reactivity. To describe such cavity-induced effects at atomic resolution, semi-classical molecular dynamics approaches have been developed that treat nuclear motion classically while describing the collective light-matter interaction within the Tavis-Cummings framework of quantum electrodynamics. Here, we benchmark mixed quantum-classical approaches, Ehrenfest dynamics, and Fewest-Switches Surface Hopping (FSSH) for simulating nonadiabatic dynamics of electronically strongly coupled carbon monoxide molecules. Their predictions are compared against numerically exact quantum dynamics simulations performed with the multi-configuration time-dependent Hartree method, which treats both electronic and nuclear degrees of freedom quantum mechanically. We find that the semi-classical approaches reproduce the qualitative features of the full quantum dynamics. Quantitative agreement is best achieved with FSSH when a decoherence correction is included. These results demonstrate that mixed quantum-classical methods provide a computationally efficient and quantitatively reliable alternative to fully quantum simulations for investigating nonadiabatic photochemistry under collective electronic strong coupling in systems beyond the reach of exact quantum treatments.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.
 03:35
Hybridization of Atomic Orbitals II
 01:02
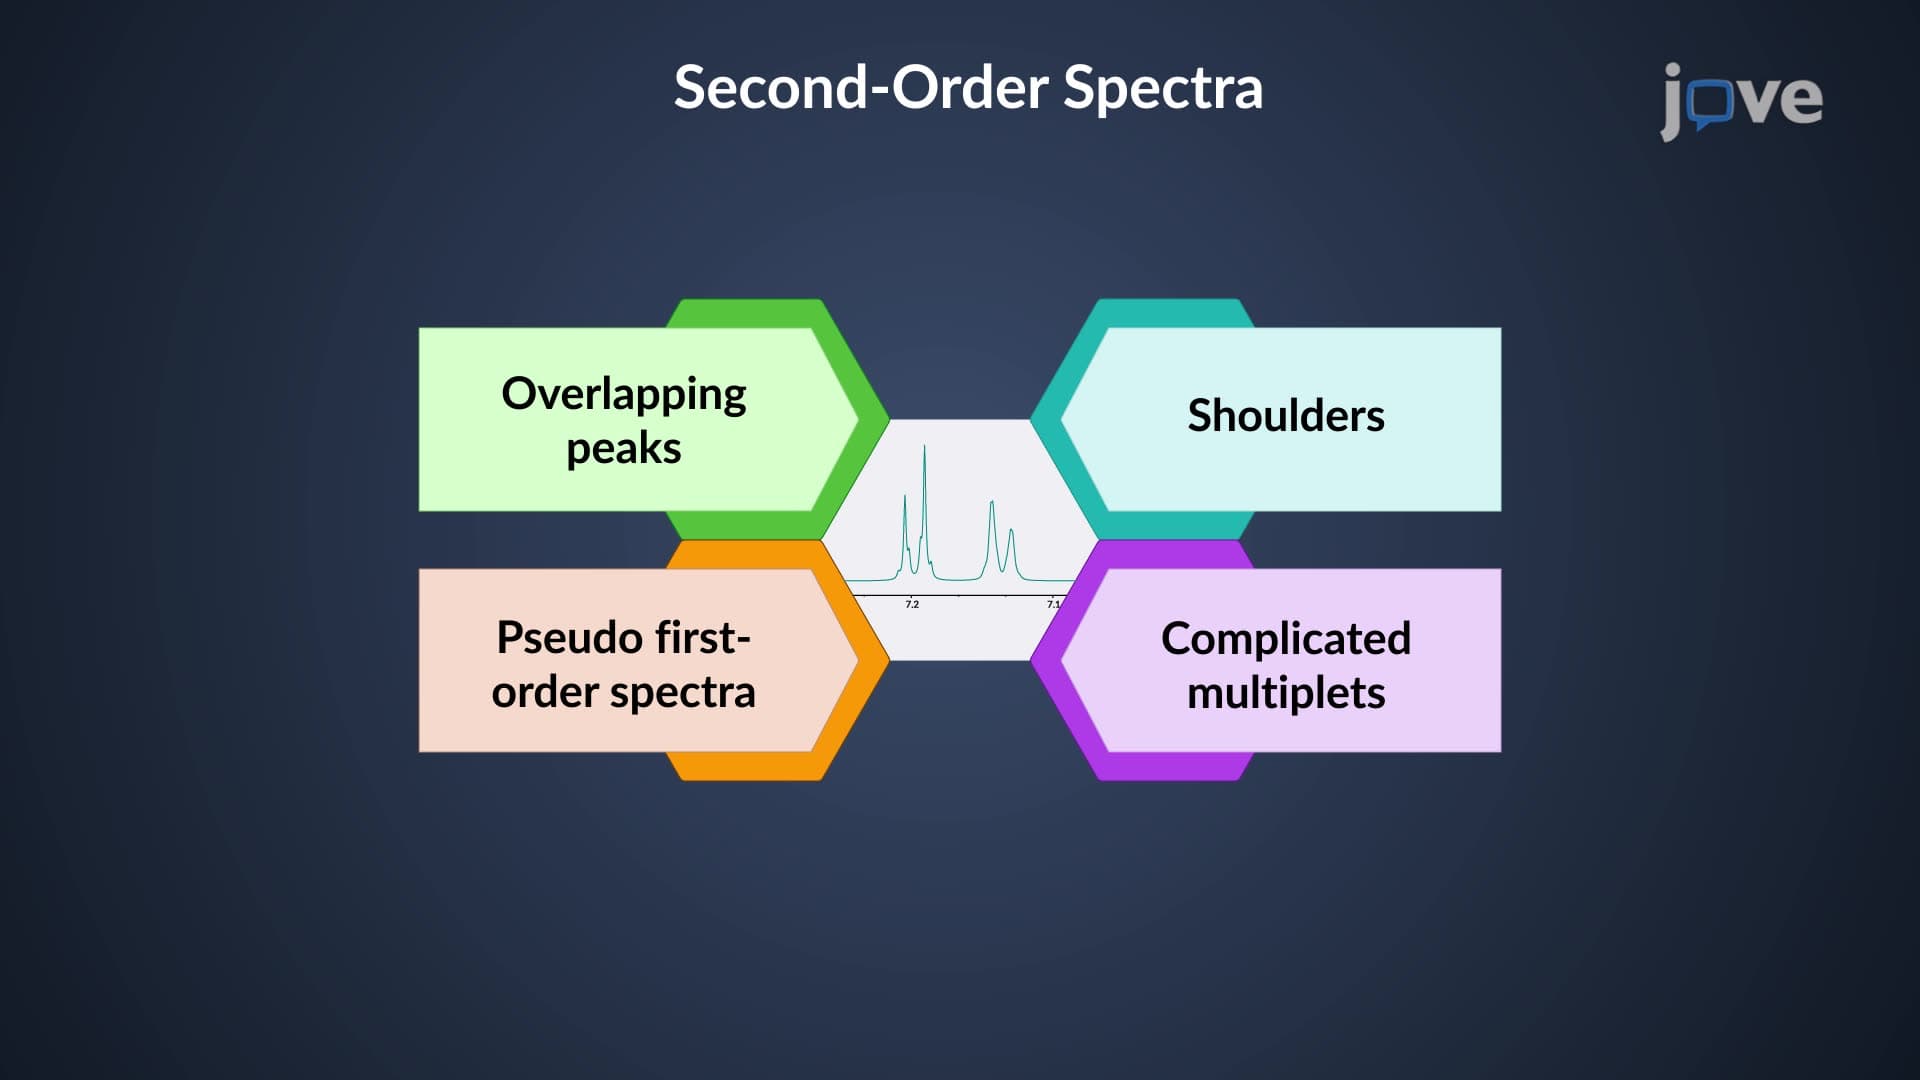
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 03:24
Hybridization of Atomic Orbitals I
 01:08
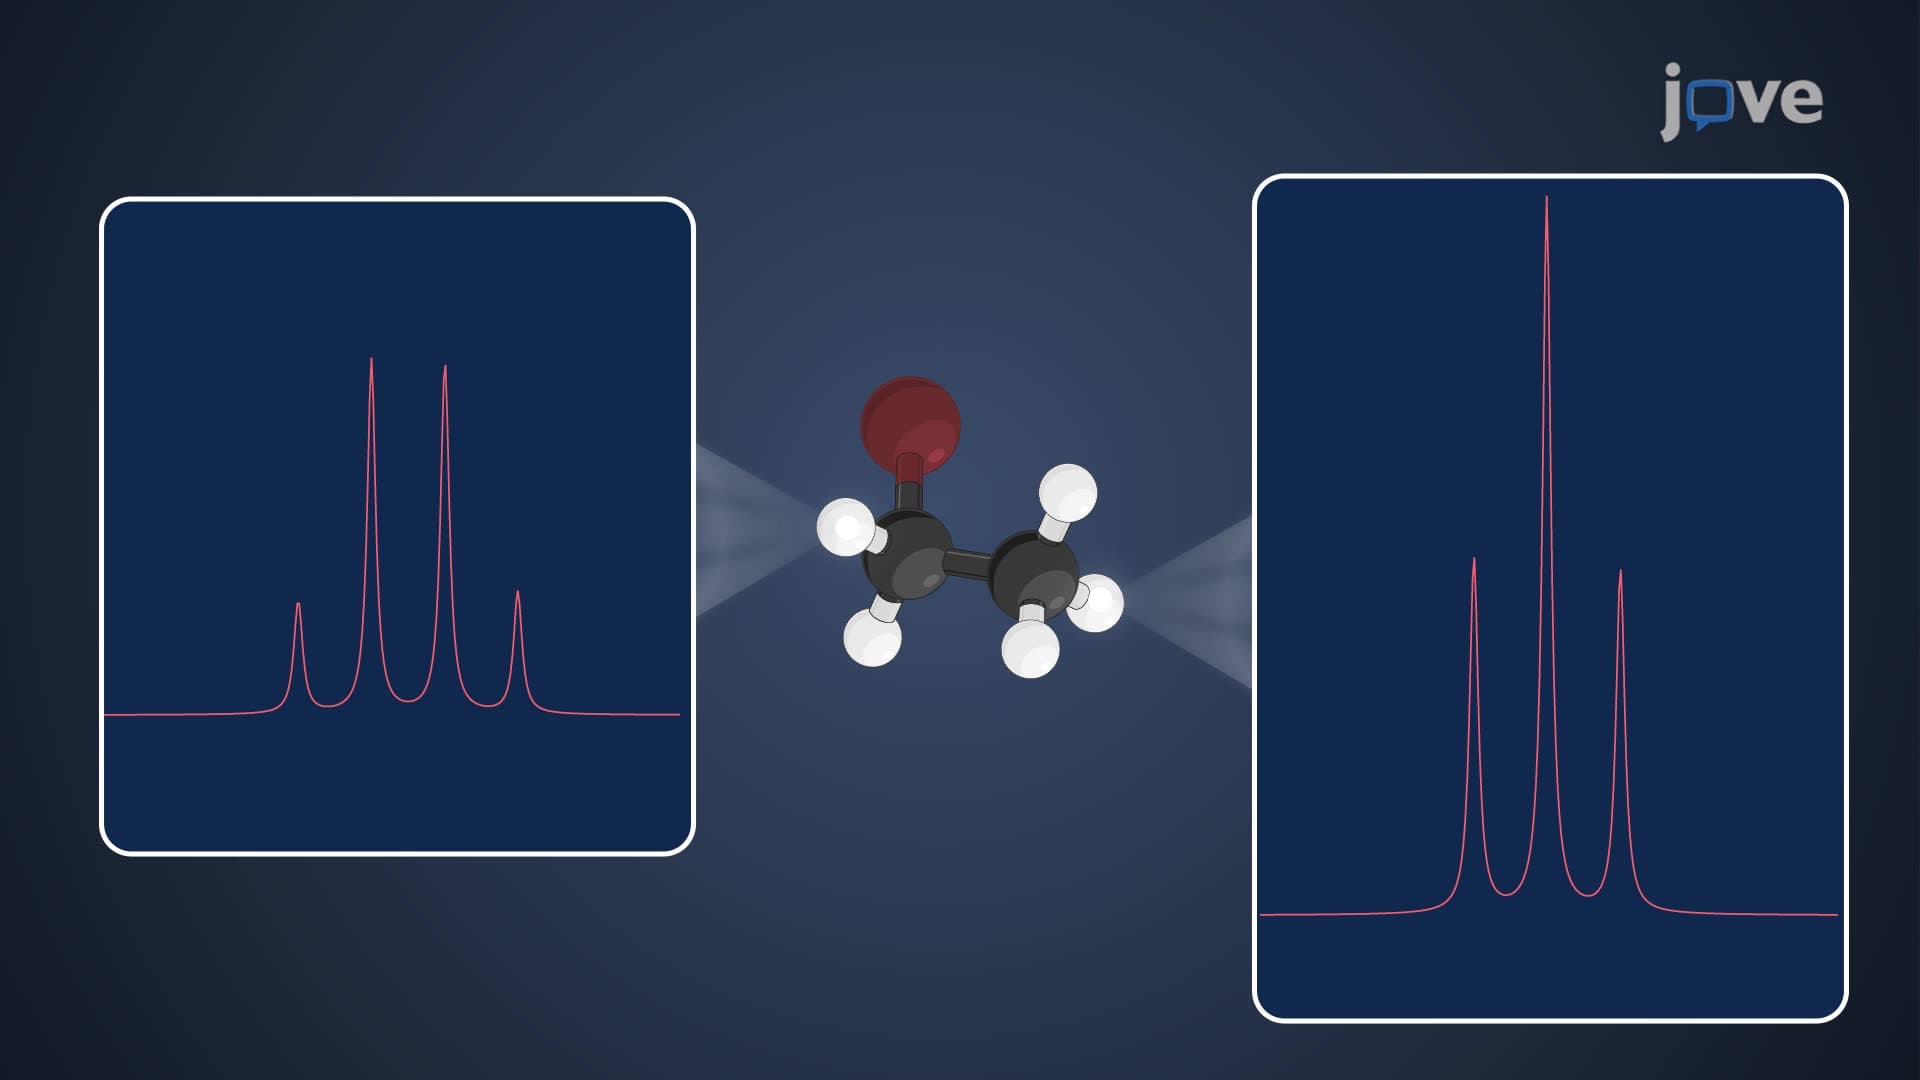
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must have a...
 01:17
Spin–Spin Coupling: One-Bond Coupling