Related Experiment Video
Updated: Aug 6, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Cooperative stability, many-body expansion, and σ-aromaticity of (LiH)n clusters (n = 1-6): A CCSD(T) study at the
Emmanouil Semidalas1, Filippos Drakopoulos1, E Alexandros Routsi1,2
1Laboratory of Physical Chemistry, Department of Chemistry, National and Kapodistrian University of Athens, Panepistimiopolis Zografou, Athens 15784, Greece.

Abstract:
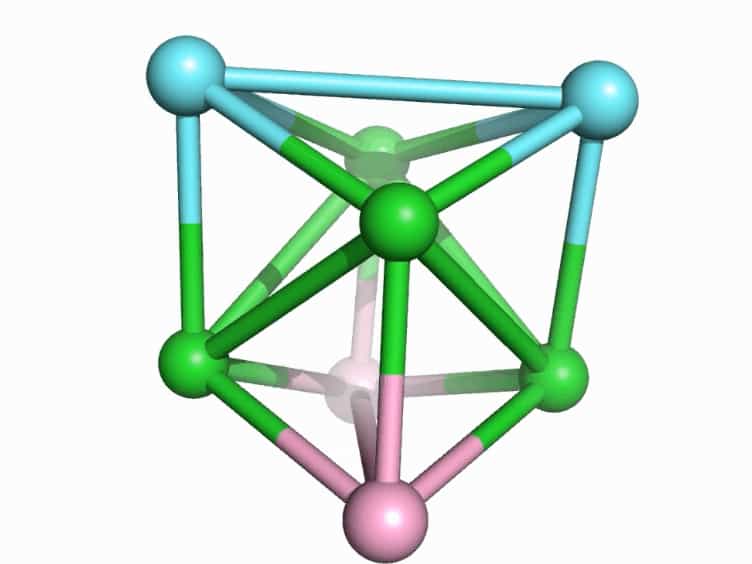
We investigated the energetics and bonding of lithium hydride clusters (LiH)n (n = 1-6) using a composite ab initio scheme inspired by W2 theory to achieve sub-kcal/mol accuracy. This approach combines CCSD(T) results extrapolated to the complete basis set limit with a rigorous treatment of core-valence correlation, scalar relativistic effects, and the diagonal Born-Oppenheimer correction. Our results show that while Hartree-Fock theory captures the primary electrostatic binding, correlation effects are crucial for determining the energetic preference of compact isomers over cyclic rings. A parallel density functional theory study shows that while standard hybrid functionals like B3LYP-D4 and M06-2X exhibit larger deviations, the double-hybrid revDSD-PBEP86-D4 functional closely matches our benchmarks, delivering sub-kcal/mol accuracy. Structural and chemical bonding analyses, including intrinsic bond orbital, nucleus-independent chemical shift, and many-body expansion (MBE) methods, reveal high ionic character and multi-center bonding (3c-2e and 4c-2e) within the (LiH)n clusters. MBE analysis of the interaction energy of the monocyclic clusters with respect to the LiH molecules reveals that the two- and three-body terms are consistently negative (stabilizing), while all higher-order terms are negligible. We find that σ-aromaticity in these systems is predominantly local and bond-centered. As the rings expand, the interior becomes magnetically decoupled from the σ-skeleton, precluding the formation of a global ring current. These results establish definitive benchmarks for the stability of prototypical electron-deficient clusters.
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 01:27
¹H NMR: Long-Range Coupling
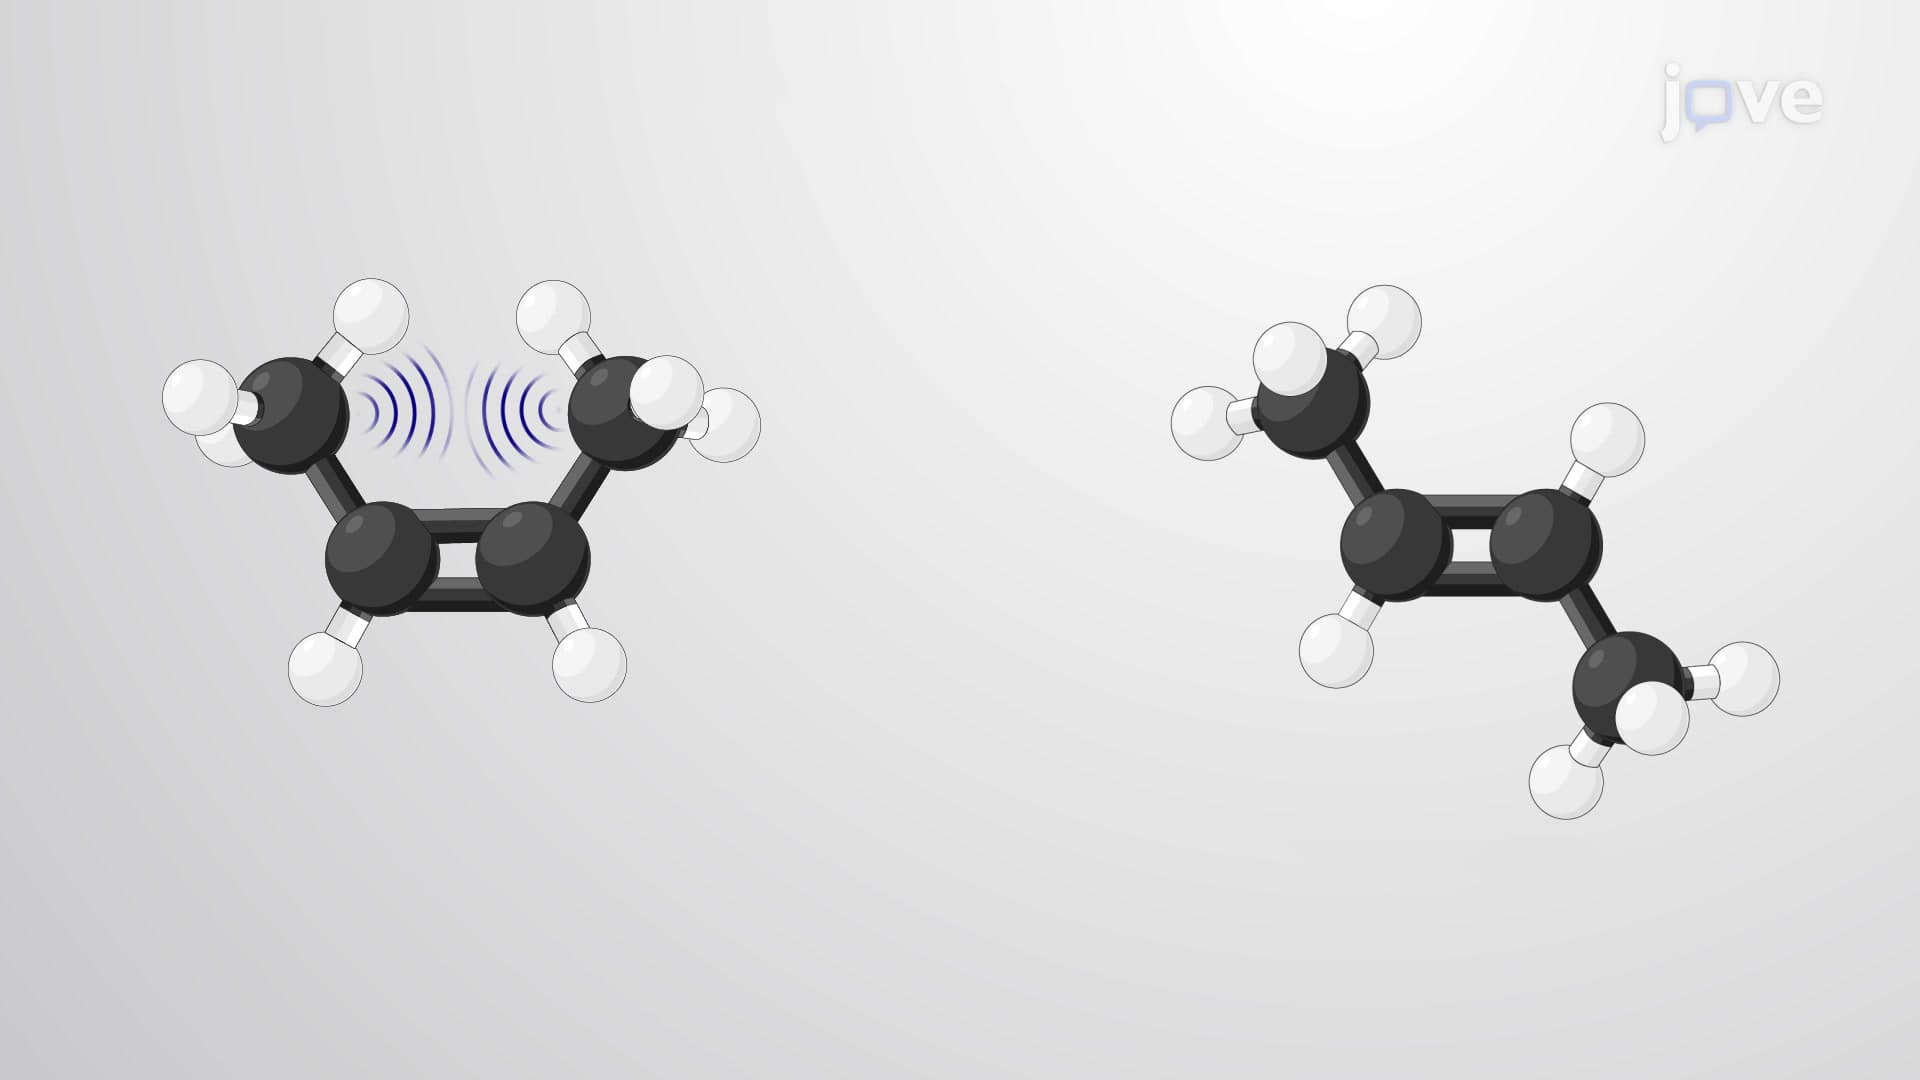
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.
 01:59
Relative Stabilities of Alkenes
 02:35
Molecular Orbital Theory I
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...