Related Experiment Video
Updated: Aug 5, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Improved gene tree inference from removing alignment errors both from focal genes and when training substitution
Andrew L Wheeler1, Chiragdeep Chatur2, Peter W Goodman3
1Genetics Graduate Interdisciplinary Program, University of Arizona, Tucson, AZ, USA.

Abstract:
Multiple sequence alignment (MSA) is a key step in phylogenetic analysis and is prone to error. Unfortunately, algorithms that remove likely alignment errors from MSAs sometimes also remove informative residues, making phylogenetic tree inference worse. Here we present a novel MSA cleaning algorithm based on consensus between MSAs using a range of Hidden Markov Models and guide trees, named CLOAK (CLeaning On the basis of Alignment C(K)onsensus). CLOAK is a gentle filter, with a low false positive rate for removal from MSAs according to the BALiBASE benchmarks, while still removing a significant fraction of likely alignment errors. Gentle vs. stringent MSA filtering methods are appropriate for different tasks. We assess methods based on their ability to bring the gene trees of single copy orthologs closer to the accepted species tree. Amino acid substitution models trained on filtered MSAs improve gene tree inference, with stricter filtering methods providing the biggest model improvements. In contrast, it is gentler filtering of single gene MSAs that provides additional improvements to gene tree inference, with CLOAK performing best.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 03:21
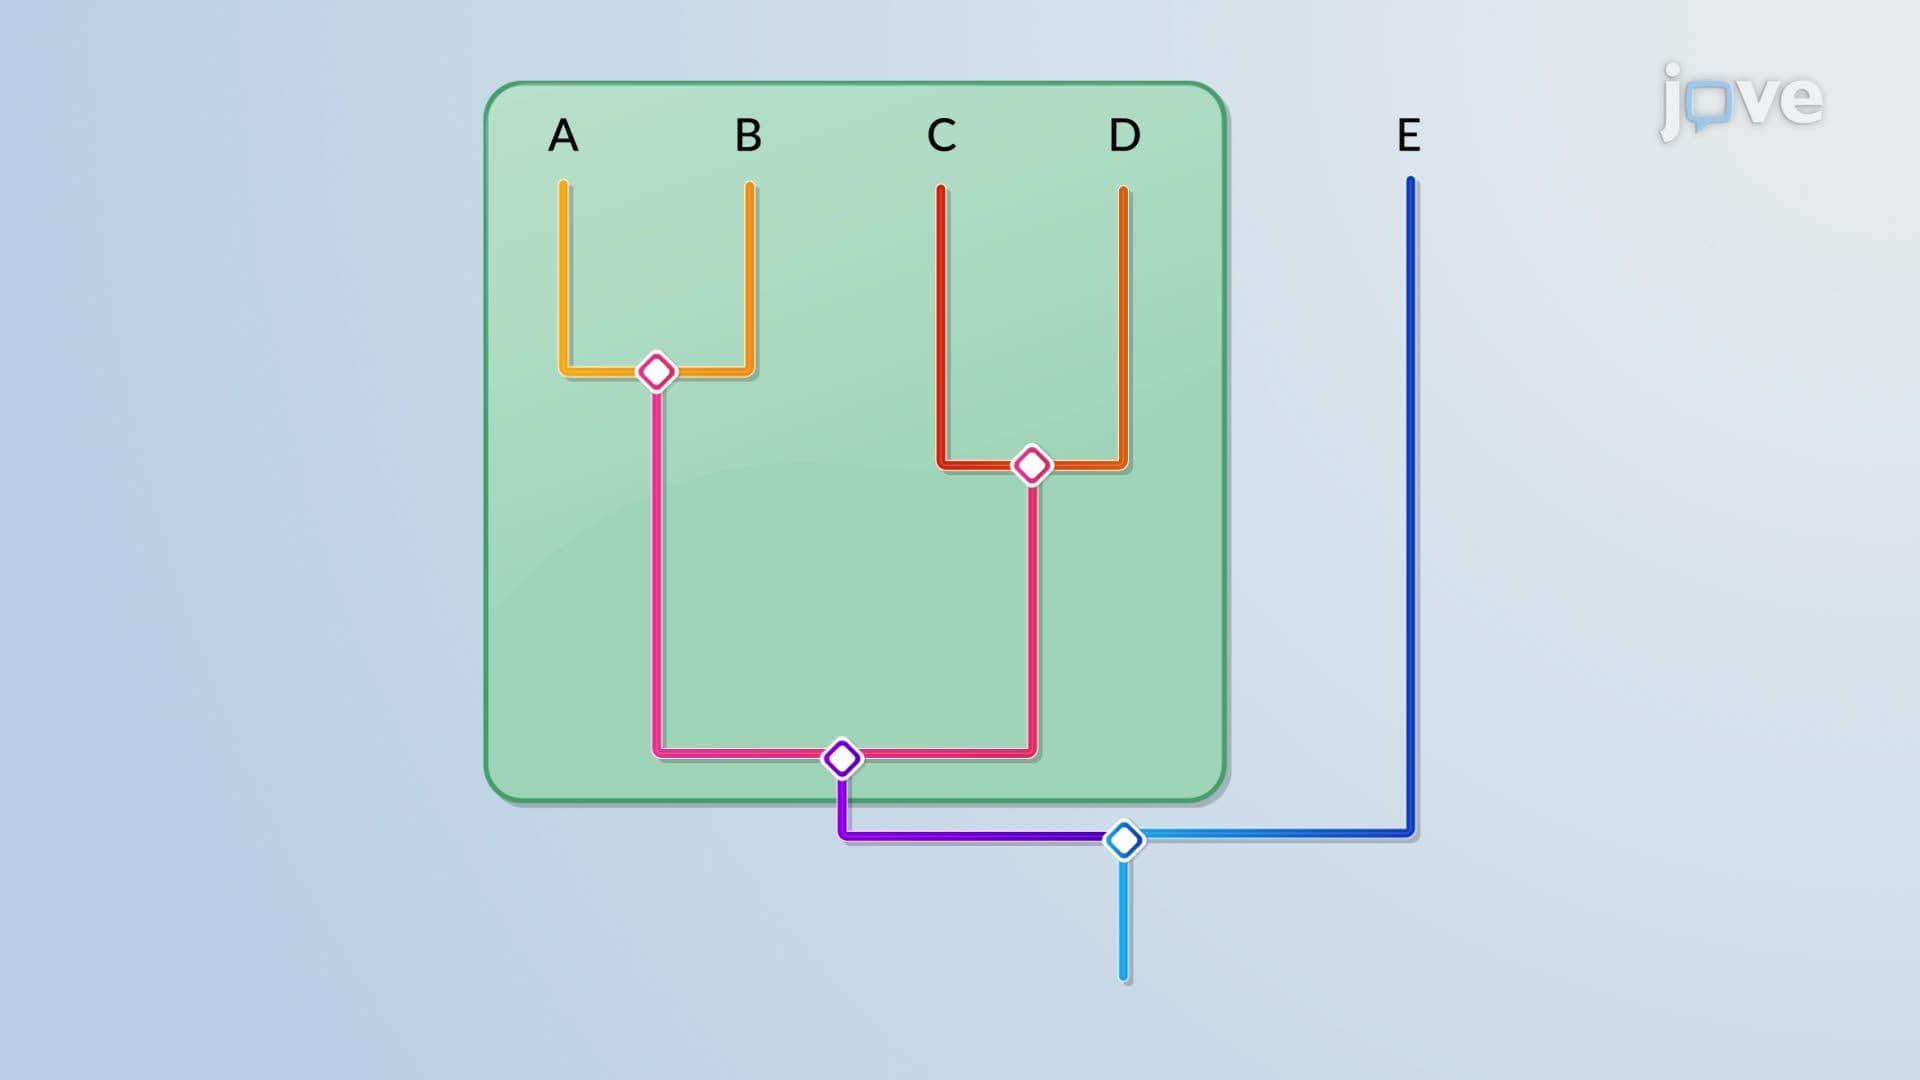
Phylogenetic Trees
03:21Phylogenetic Trees
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?
In contrast, regions which code...
 01:28
Microbial Phylogeny